Related Experiment Video
Updated: May 28, 2026

Basophil Activation Test for Allergy Diagnosis
Published on: May 31, 2021
Structure-based prediction of subtype selectivity of histamine H3 receptor selective antagonists in clinical trials
Soo-Kyung Kim1, Peter Fristrup, Ravinder Abrol
1Materials and Process Simulation Center (MC139-74), California Institute of Technology, 1200 E. California Blvd., Pasadena, California 91125, USA.

Abstract:
Histamine receptors (HRs) are excellent drug targets for the treatment of diseases, such as schizophrenia, psychosis, depression, migraine, allergies, asthma, ulcers, and hypertension. Among them, the human H(3) histamine receptor (hH(3)HR) antagonists have been proposed for specific therapeutic applications, including treatment of Alzheimer's disease, attention deficit hyperactivity disorder (ADHD), epilepsy, and obesity. However, many of these drug candidates cause undesired side effects through the cross-reactivity with other histamine receptor subtypes. In order to develop improved selectivity and activity for such treatments, it would be useful to have the three-dimensional structures for all four HRs. We report here the predicted structures of four HR subtypes (H(1), H(2), H(3), and H(4)) using the GEnSeMBLE (GPCR ensemble of structures in membrane bilayer environment) Monte Carlo protocol, sampling ∼35 million combinations of helix packings to predict the 10 most stable packings for each of the four subtypes. Then we used these 10 best protein structures with the DarwinDock Monte Carlo protocol to sample ∼50 000 × 10(20) poses to predict the optimum ligand-protein structures for various agonists and antagonists. We find that E206(5.46) contributes most in binding H(3) selective agonists (5, 6, 7) in agreement with experimental mutation studies. We also find that conserved E5.46/S5.43 in both of hH(3)HR and hH(4)HR are involved in H(3)/ H(4) subtype selectivity. In addition, we find that M378(6.55) in hH(3)HR provides additional hydrophobic interactions different from hH(4)HR (the corresponding amino acid of T323(6.55) in hH(4)HR) to provide additional subtype bias. From these studies, we developed a pharmacophore model based on our predictions for known hH(3)HR selective antagonists in clinical study [ABT-239 1, GSK-189,254 2, PF-3654746 3, and BF2.649 (tiprolisant) 4] that suggests critical selectivity directing elements are: the basic proton interacting with D114(3.32), the spacer, the aromatic ring substituted with the hydrophilic or lipophilic groups interacting with lipophilic pockets in transmembranes (TMs) 3-5-6 and the aliphatic ring located in TMs 2-3-7. These 3D structures for all four HRs should help guide the rational design of novel drugs for the subtype selective antagonists and agonists with reduced side effects.
Related Concept Videos
 01:28
Acid Suppressive Drugs for Peptic Ulcer Disease: Histamine H2-Receptor Antagonists
 01:25
Dose-Response Relationship: Selectivity and Specificity
 01:17
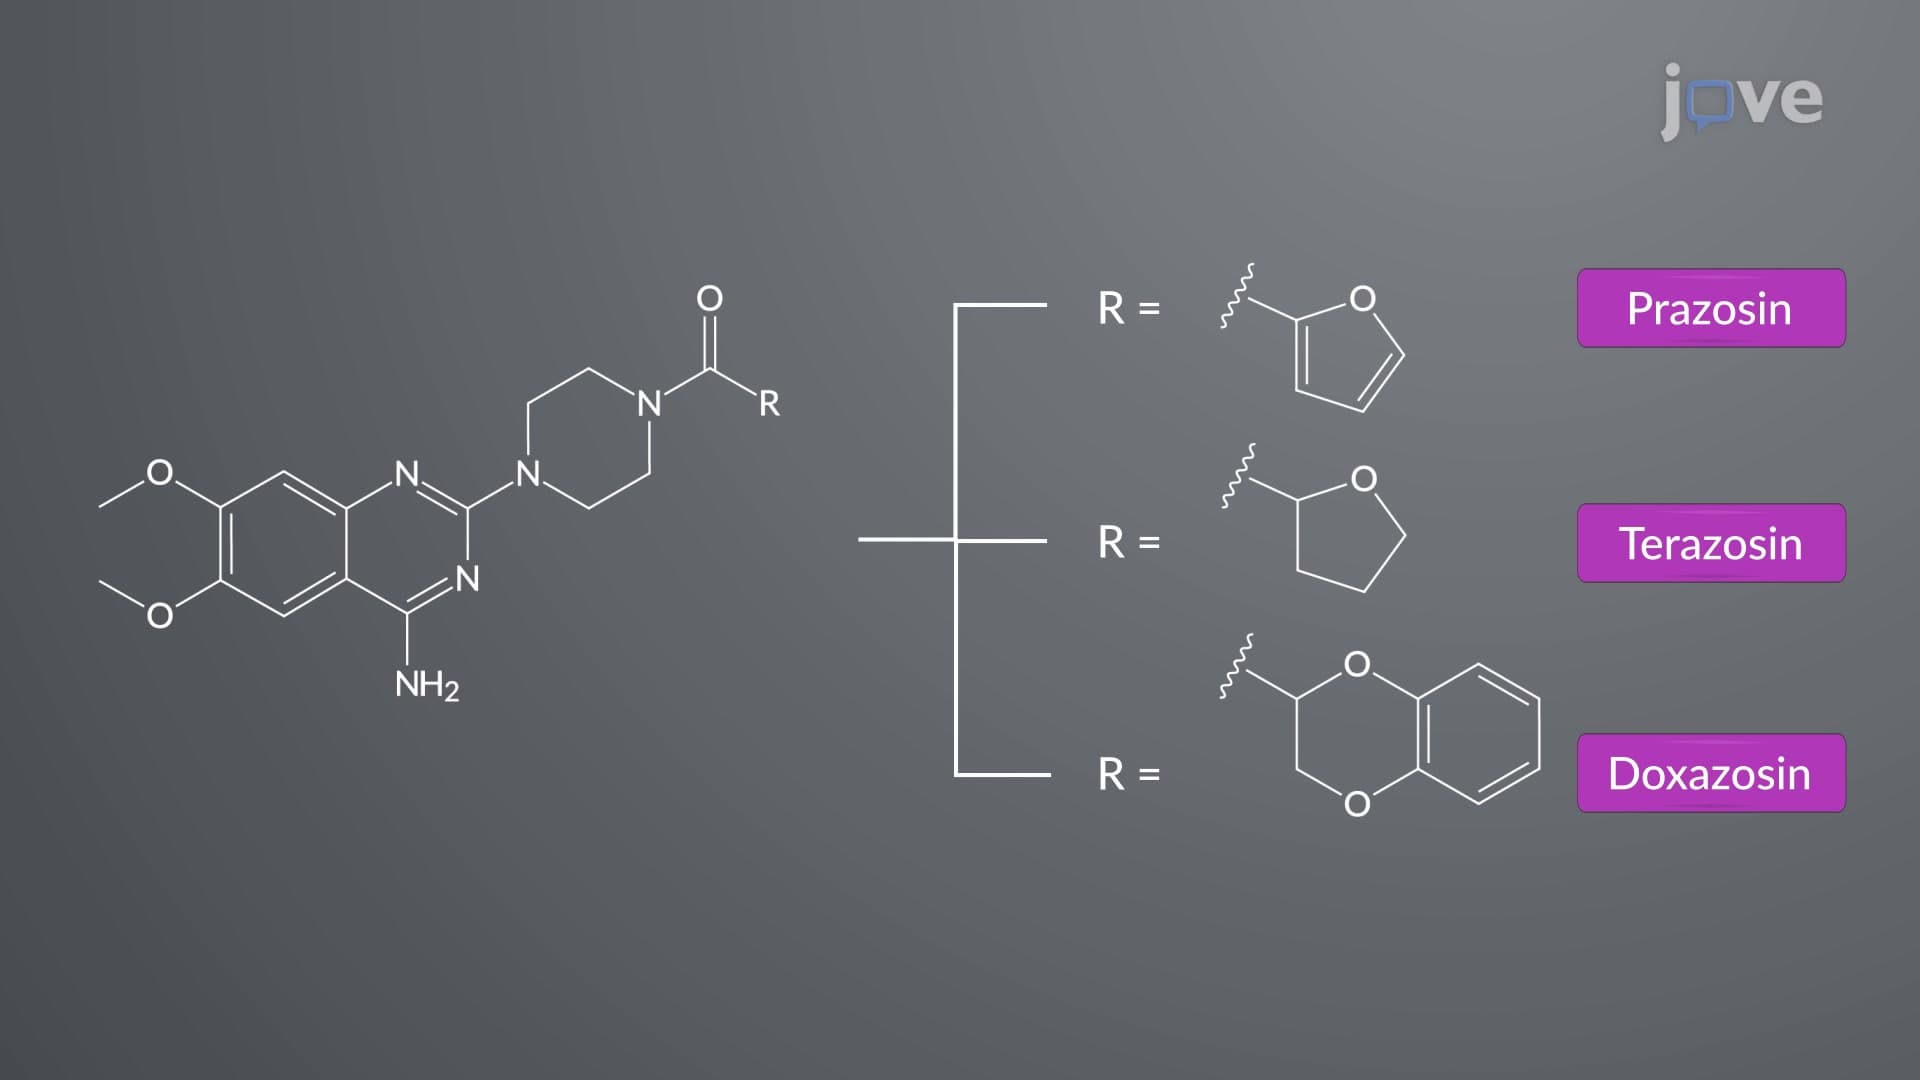
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:25
Adrenergic Antagonists: Chemistry and Classification of β-Receptor Blockers