A catalytic defect in mitochondrial respiratory chain complex I due to a mutation in NDUFS2 in a patient with Leigh
Lock Hock Ngu1, Leo G Nijtmans, Felix Distelmaier
1Nijmegen Center for Mitochondrial Disorders, Department of Pediatrics, Radboud University Nijmegen Medical Center, Nijmegen, The Netherlands.

Abstract:
In this study, we investigated the pathogenicity of a homozygous Asp446Asn mutation in the NDUFS2 gene of a patient with a mitochondrial respiratory chain complex I deficiency. The clinical, biochemical, and genetic features of the NDUFS2 patient were compared with those of 4 patients with previously identified NDUFS2 mutations. All 5 patients presented with Leigh syndrome. In addition, 3 out of 5 showed hypertrophic cardiomyopathy. Complex I amounts in the patient carrying the Asp446Asn mutation were normal, while the complex I activity was strongly reduced, showing that the NDUFS2 mutation affects complex I enzymatic function. By contrast, the 4 other NDUFS2 patients showed both a reduced amount and activity of complex I. The enzymatic defect in fibroblasts of the patient carrying the Asp446Asn mutation was rescued by transduction of wild type NDUFS2. A 3-D model of the catalytic core of complex I showed that the mutated amino acid residue resides near the coenzyme Q binding pocket. However, the K(M) of complex I for coenzyme Q analogs of the Asp446Asn mutated complex I was similar to the K(M) observed in other complex I defects and in controls. We propose that the mutation interferes with the reduction of coenzyme Q or with the coupling of coenzyme Q reduction with the conformational changes involved in proton pumping of complex I.
More Related Videos
 08:07
Analysis of the Expression and Complexes Assembly of the Mitochondrial Respiratory Chain Proteins in the Fission Yeast Schizosaccharomyces pombe
Published on: May 2, 2025
 08:56
Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Related Concept Videos
 01:18
ATP Synthase: Structure
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:48
ATP Synthase: Mechanism
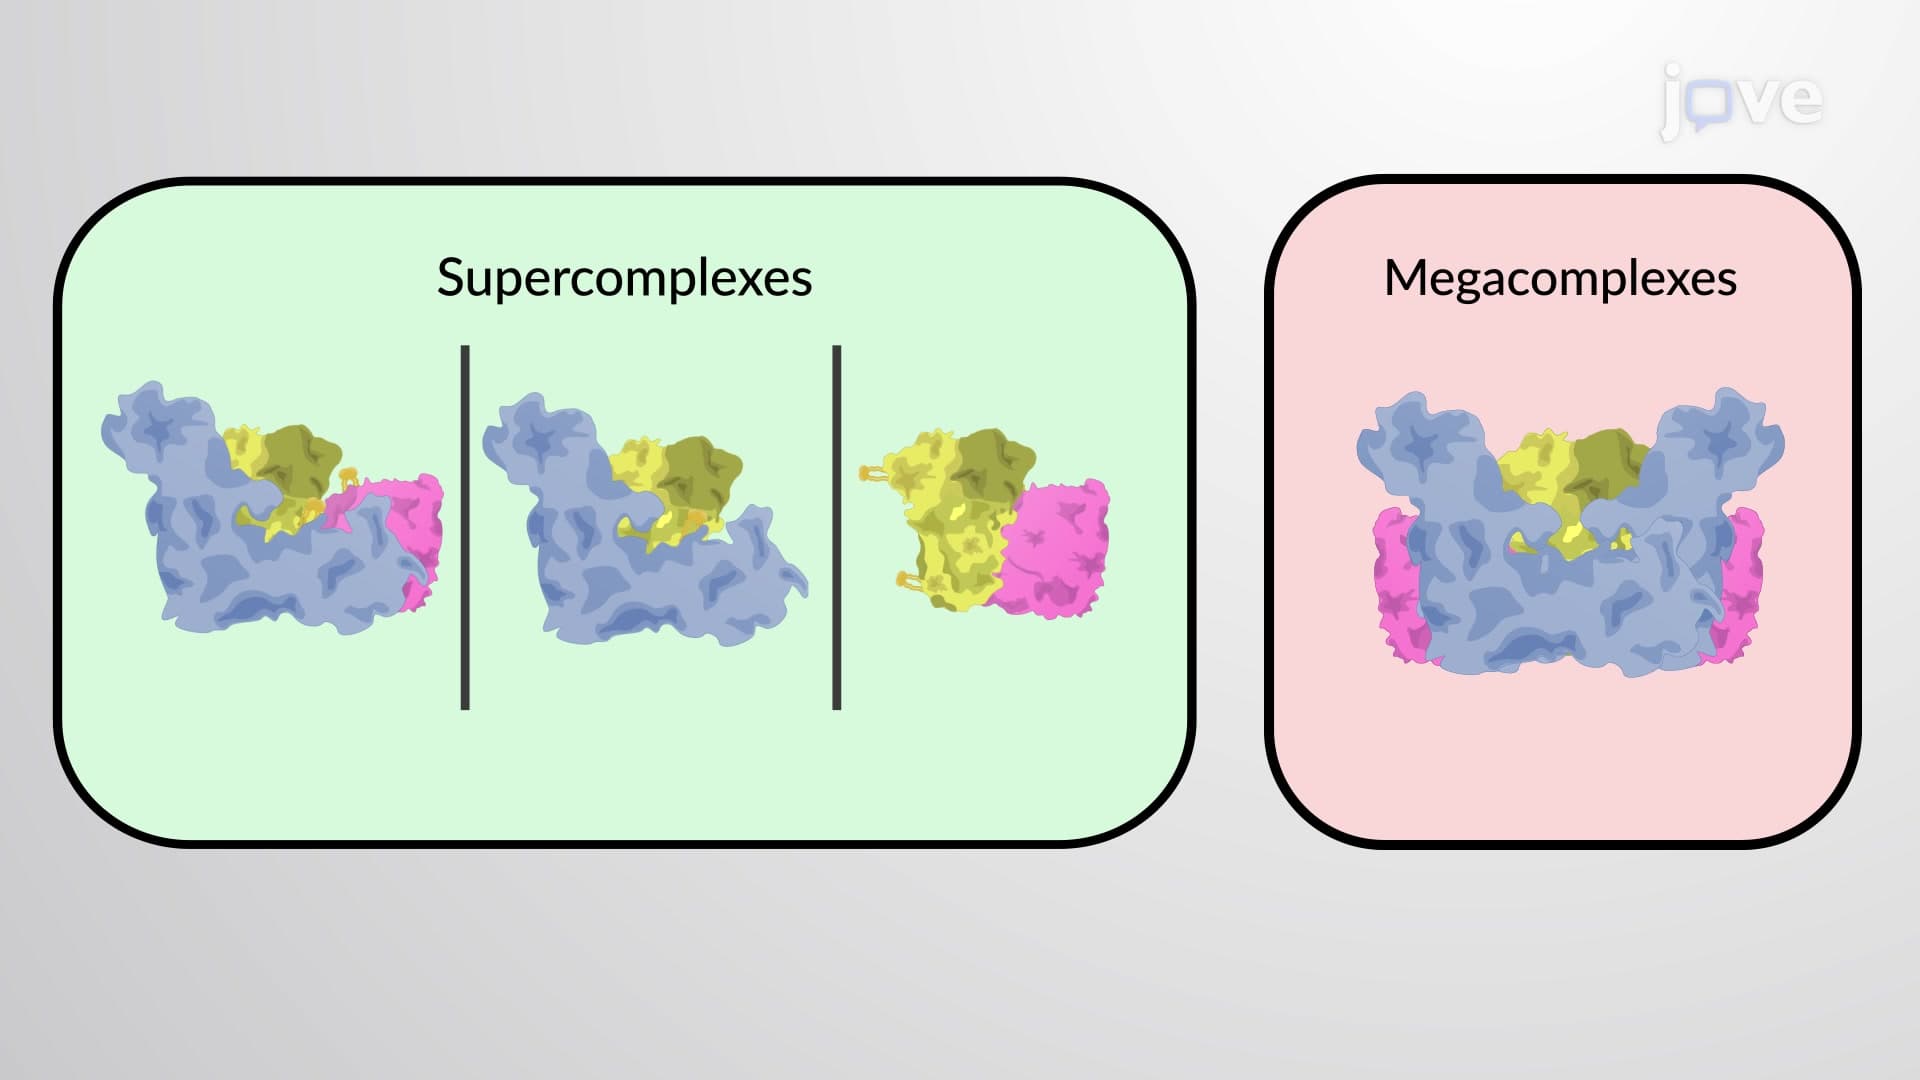 01:41
The Supercomplexes in the Crista Membrane
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life
01:31Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of Life