Related Experiment Video
Updated: May 28, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Calculation of relative free energies for ligand-protein binding, solvation, and conformational transitions using the
Sereina Riniker1, Clara D Christ, Halvor S Hansen
1Laboratory of Physical Chemistry, Swiss Federal Institute of Technology, ETH, 8093 Zürich, Switzerland.

Abstract:
The calculation of the relative free energies of ligand-protein binding, of solvation for different compounds, and of different conformational states of a polypeptide is of considerable interest in the design or selection of potential enzyme inhibitors. Since such processes in aqueous solution generally comprise energetic and entropic contributions from many molecular configurations, adequate sampling of the relevant parts of configurational space is required and can be achieved through molecular dynamics simulations. Various techniques to obtain converged ensemble averages and their implementation in the GROMOS software for biomolecular simulation are discussed, and examples of their application to biomolecules in aqueous solution are given.


Related Concept Videos
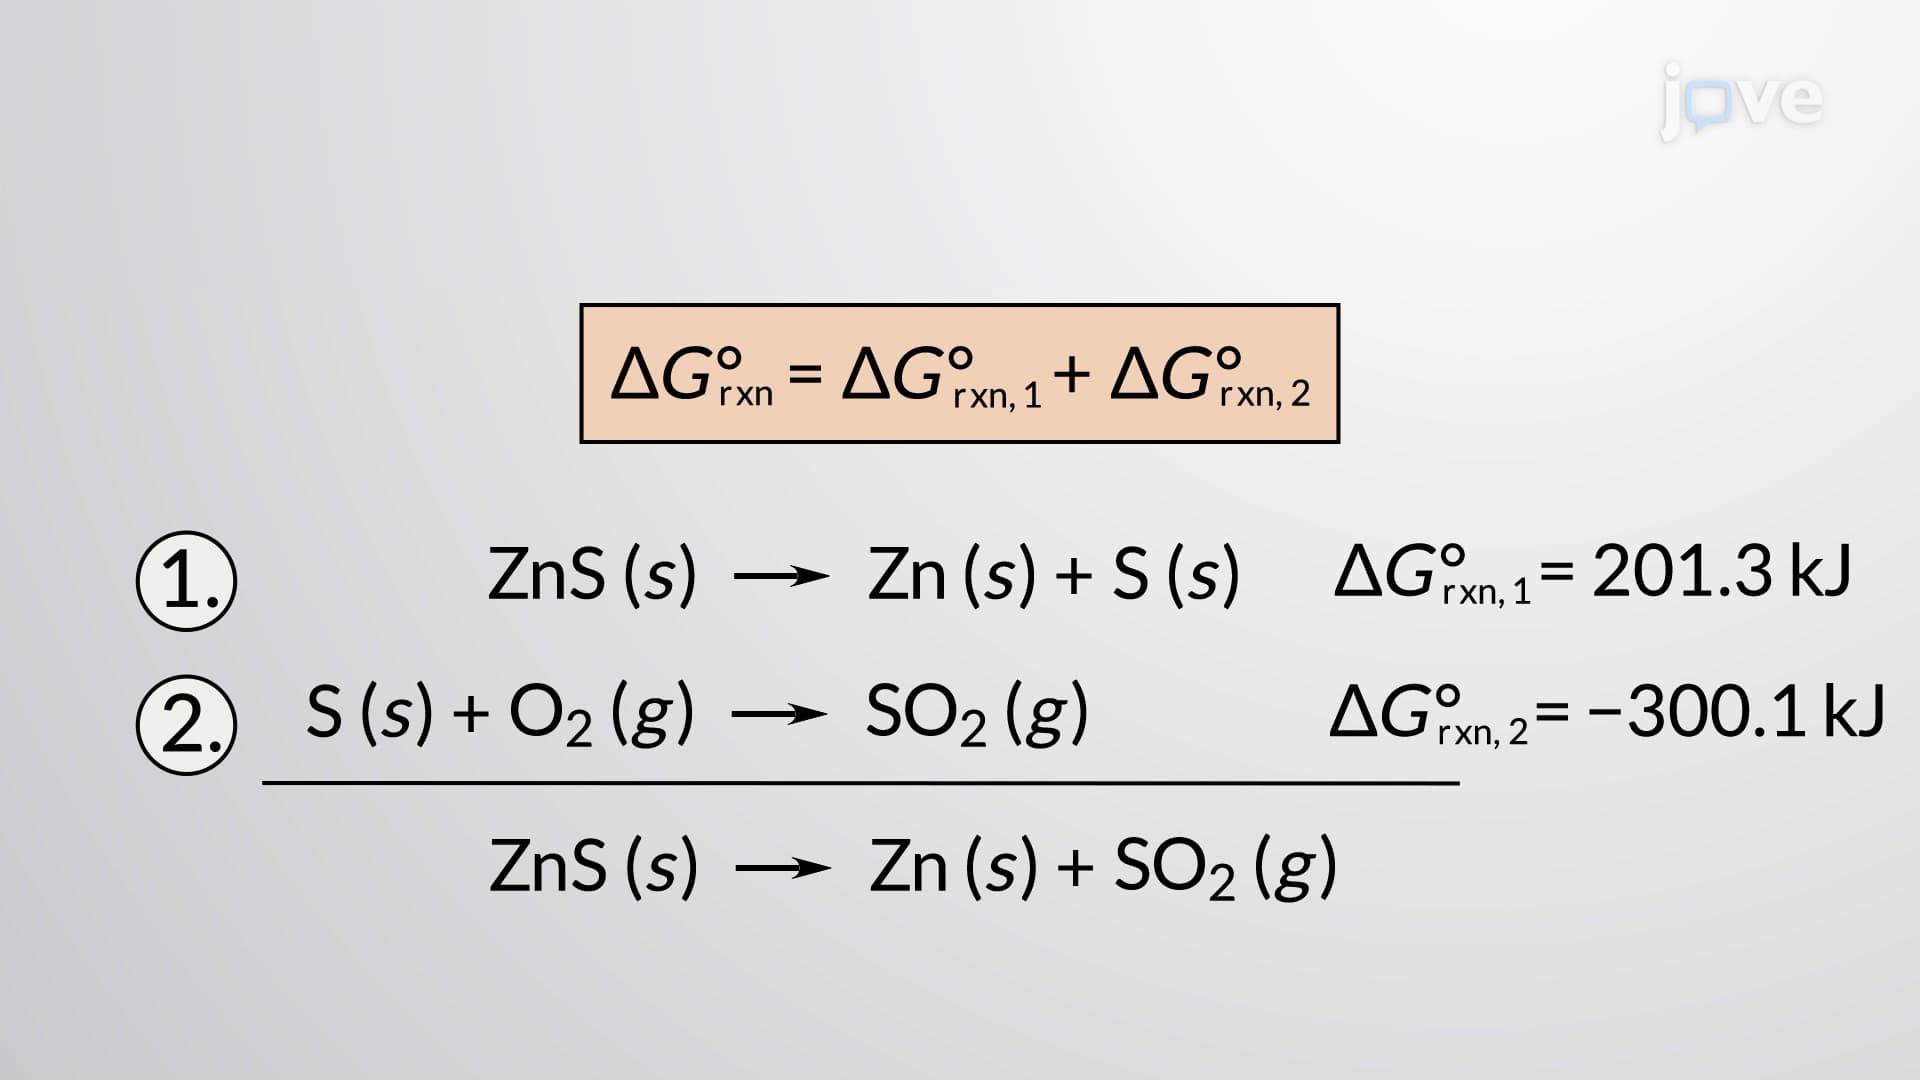 02:49
Calculating Standard Free Energy Changes
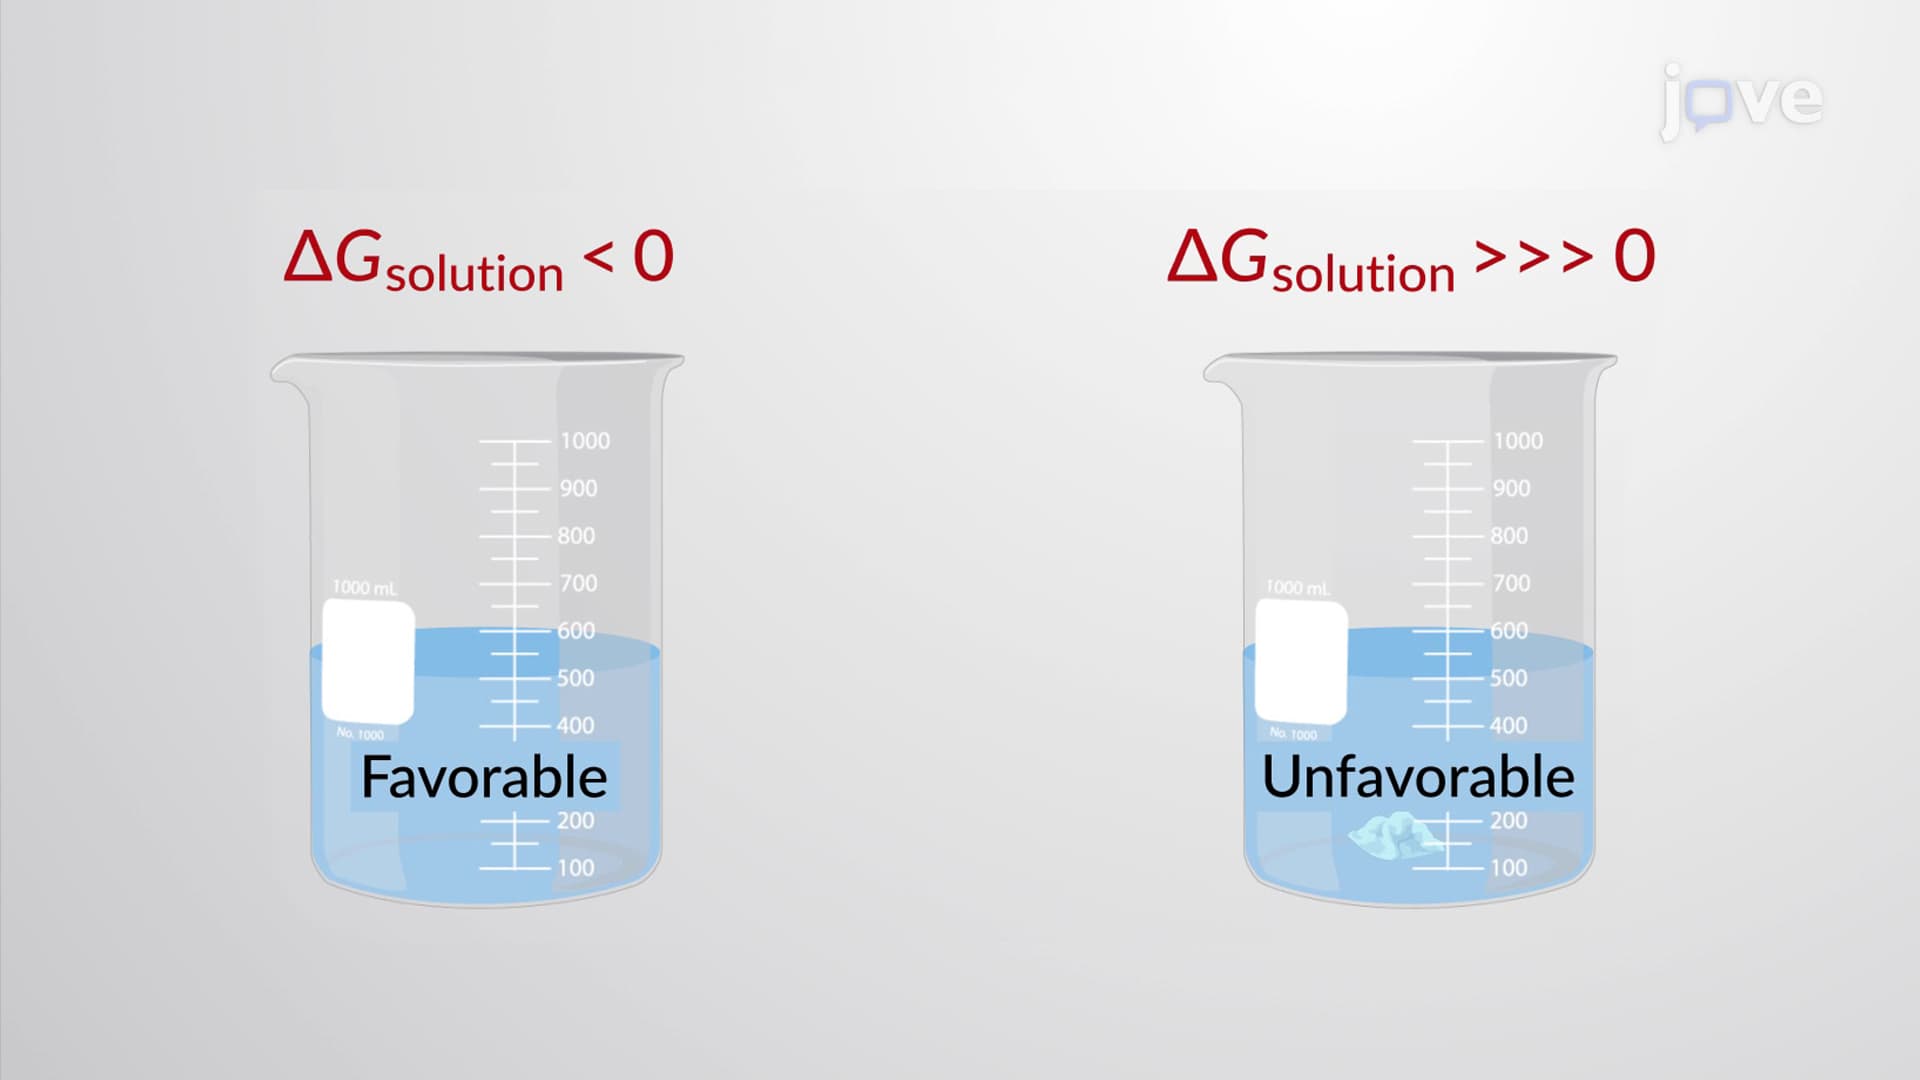 02:21
Chemical and Solubility Equilibria
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
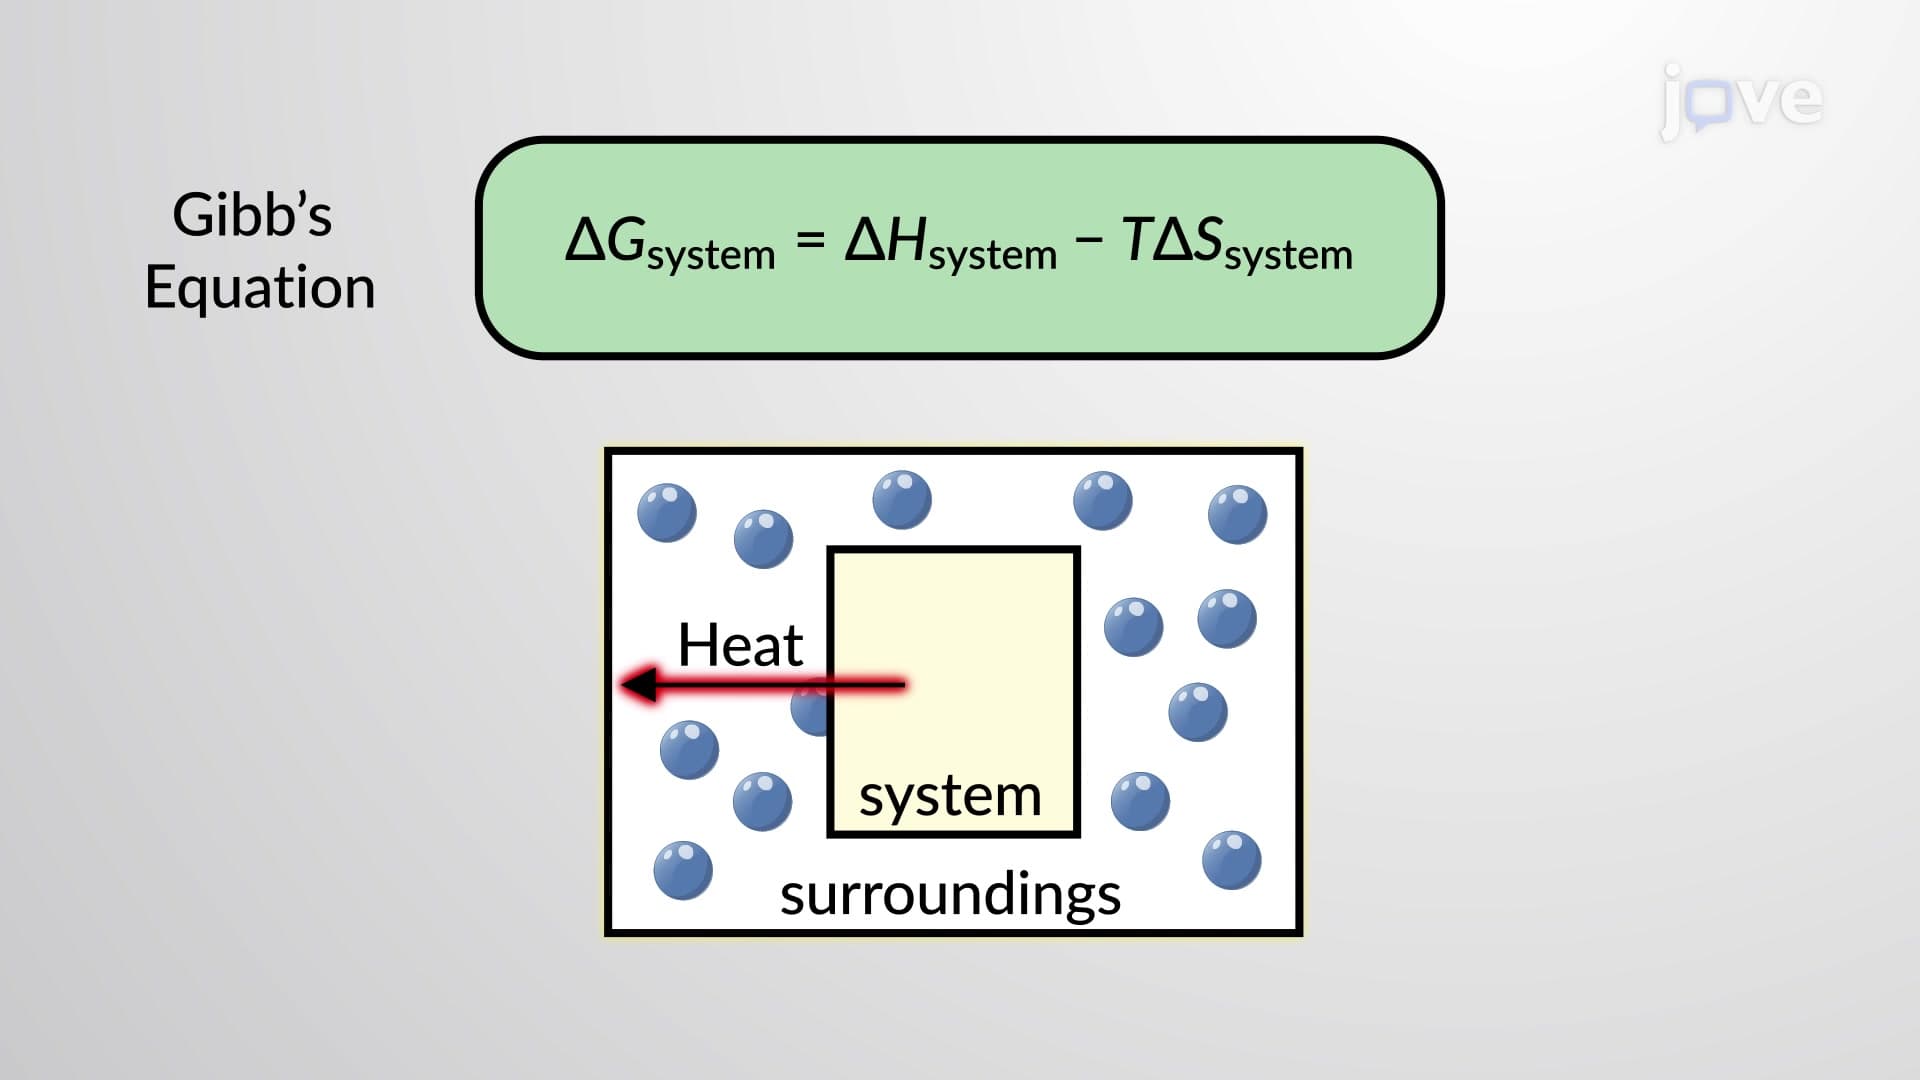 01:05
An Introduction to Free Energy
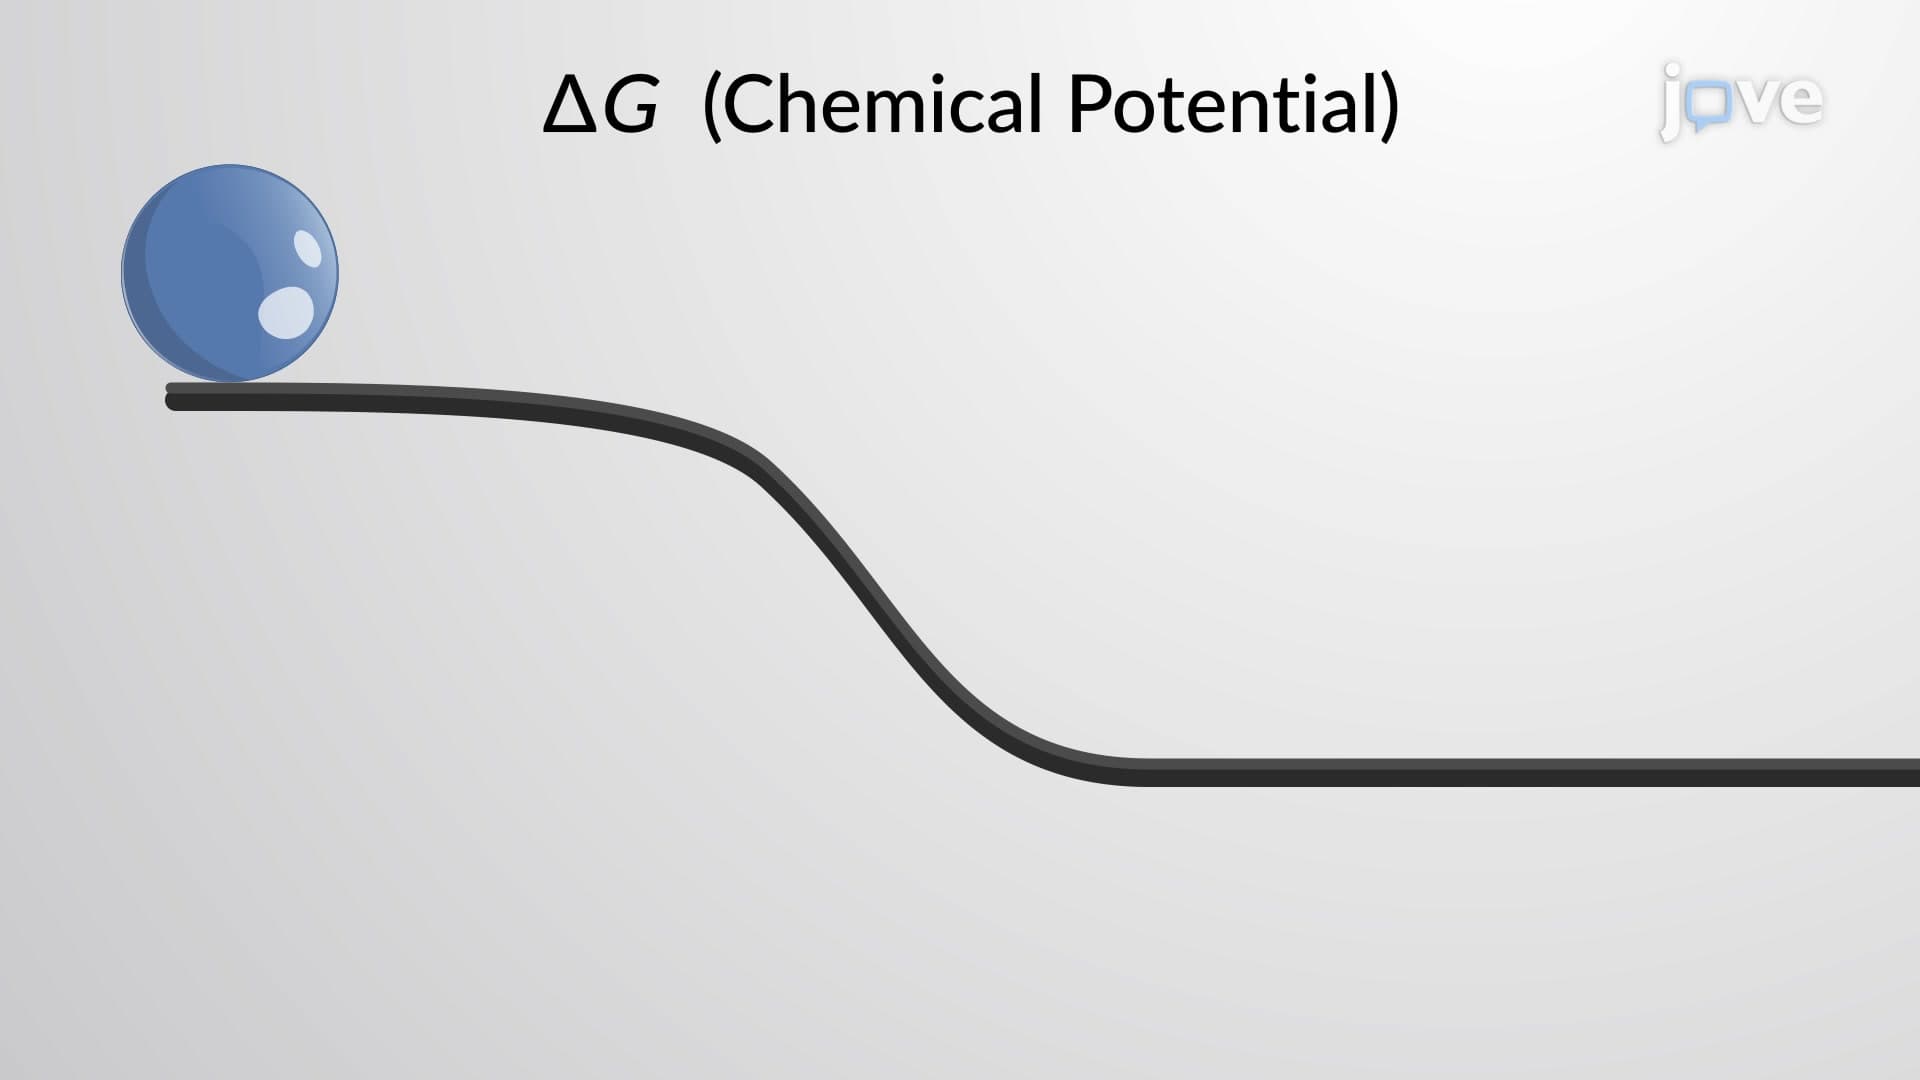 02:39
Gibbs Free Energy