Related Experiment Video
Updated: May 27, 2026

Heuristic Mining of Hierarchical Genotypes and Accessory Genome Loci in Bacterial Populations
Published on: December 7, 2021
Horizontal gene transfer in Histophilus somni and its role in the evolution of pathogenic strain 2336, as determined
Shivakumara Siddaramappa1, Jean F Challacombe, Alison J Duncan
1Center for Molecular Medicine and Infectious Diseases, Virginia-Maryland Regional College of Veterinary Medicine, Virginia Polytechnic Institute and State University, Blacksburg, Virginia 24061, USA.

Background:
Pneumonia and myocarditis are the most commonly reported diseases due to Histophilus somni, an opportunistic pathogen of the reproductive and respiratory tracts of cattle. Thus far only a few genes involved in metabolic and virulence functions have been identified and characterized in H. somni using traditional methods. Analyses of the genome sequences of several Pasteurellaceae species have provided insights into their biology and evolution. In view of the economic and ecological importance of H. somni, the genome sequence of pneumonia strain 2336 has been determined and compared to that of commensal strain 129Pt and other members of the Pasteurellaceae.
Results:
The chromosome of strain 2336 (2,263,857 bp) contained 1,980 protein coding genes, whereas the chromosome of strain 129Pt (2,007,700 bp) contained only 1,792 protein coding genes. Although the chromosomes of the two strains differ in size, their average GC content, gene density (total number of genes predicted on the chromosome), and percentage of sequence (number of genes) that encodes proteins were similar. The chromosomes of these strains also contained a number of discrete prophage regions and genomic islands. One of the genomic islands in strain 2336 contained genes putatively involved in copper, zinc, and tetracycline resistance. Using the genome sequence data and comparative analyses with other members of the Pasteurellaceae, several H. somni genes that may encode proteins involved in virulence (e.g., filamentous haemaggutinins, adhesins, and polysaccharide biosynthesis/modification enzymes) were identified. The two strains contained a total of 17 ORFs that encode putative glycosyltransferases and some of these ORFs had characteristic simple sequence repeats within them. Most of the genes/loci common to both the strains were located in different regions of the two chromosomes and occurred in opposite orientations, indicating genome rearrangement since their divergence from a common ancestor.
Conclusions:
Since the genome of strain 129Pt was ~256,000 bp smaller than that of strain 2336, these genomes provide yet another paradigm for studying evolutionary gene loss and/or gain in regard to virulence repertoire and pathogenic ability. Analyses of the complete genome sequences revealed that bacteriophage- and transposon-mediated horizontal gene transfer had occurred at several loci in the chromosomes of strains 2336 and 129Pt. It appears that these mobile genetic elements have played a major role in creating genomic diversity and phenotypic variability among the two H. somni strains.
Insights
This study sequenced the genomes of two Histophilus somni strains, revealing significant differences in size and gene content that likely contribute to virulence and adaptation. Comparative genomics highlights the role of horizontal gene transfer in shaping bacterial evolution.
Area of Science:
- Microbiology
- Genomics
- Bacterial Pathogenesis
Background:
- Histophilus somni is an opportunistic pathogen causing pneumonia and myocarditis in cattle.
- Limited understanding of H. somni virulence and metabolic functions due to traditional identification methods.
- Genomic analysis of Pasteurellaceae offers insights into bacterial biology and evolution.
Purpose of the Study:
- Determine and compare the genome sequences of H. somni pneumonia strain 2336 and commensal strain 129Pt.
- Identify genes and genomic features associated with H. somni virulence and adaptation.
- Investigate the role of horizontal gene transfer in H. somni genomic diversity.
Main Methods:
- Whole-genome sequencing of H. somni strains 2336 and 129Pt.
- Comparative genomic analysis with other Pasteurellaceae species.
- Identification of protein-coding genes, genomic islands, and prophage regions.
Main Results:
- Strain 2336 genome (2.26 Mbp) has more protein-coding genes (1,980) than strain 129Pt (2.01 Mbp, 1,792 genes).
- Genomic islands in strain 2336 contain genes for copper, zinc, and tetracycline resistance.
- Identified putative virulence factors including adhesins and polysaccharide biosynthesis enzymes.
- Genome rearrangement and horizontal gene transfer (HGT) via phages and transposons were observed.
Conclusions:
- Genome size differences between strains provide a model for studying gene gain/loss in virulence.
- HGT has significantly contributed to genomic diversity and phenotypic variation in H. somni.
- Mobile genetic elements play a crucial role in the evolution of H. somni pathogenicity.
Related Concept Videos
 01:27
Horizontal Gene Transfer
 01:08
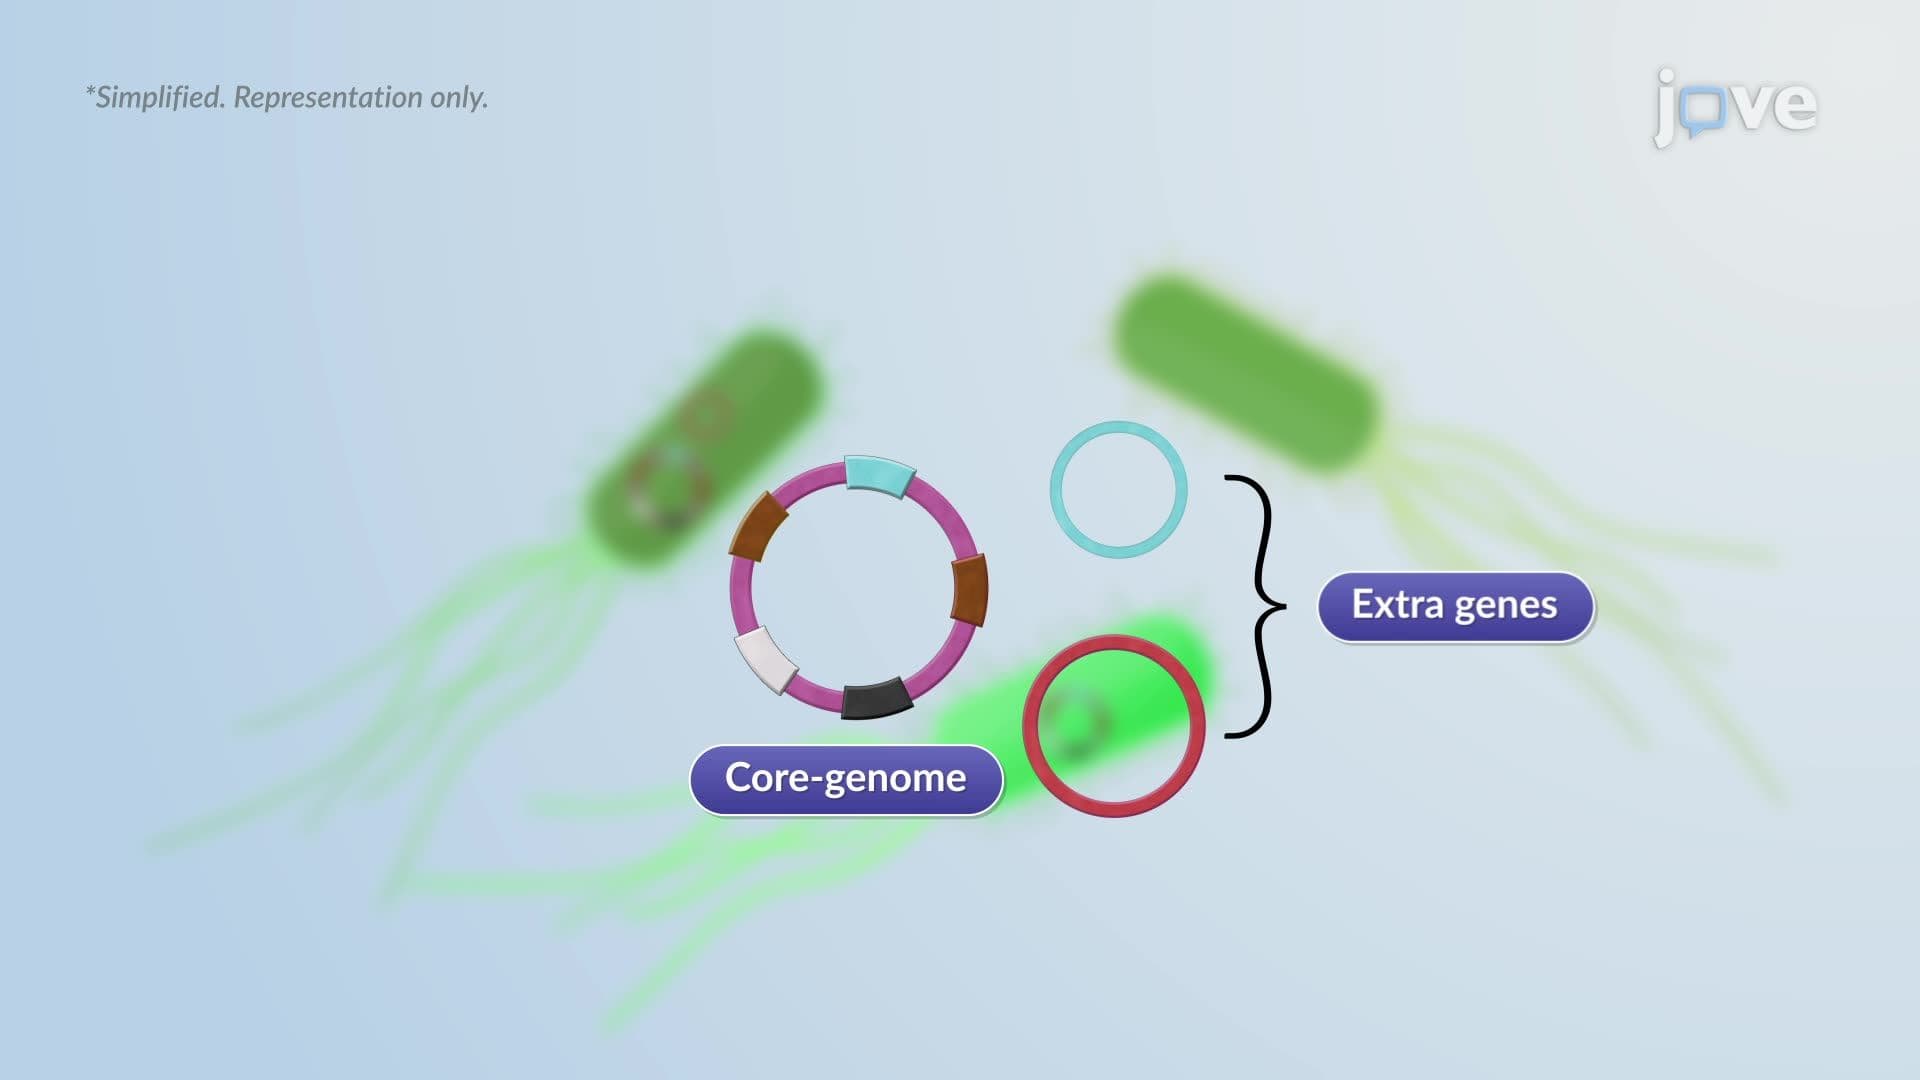
Evolution of Microbial Genome
 02:18
Types of Genetic Transfer Between Organisms
02:18Types of Genetic Transfer Between Organisms
 01:16
Transduction
 01:29
Modern Molecular Taxonomy