Related Experiment Video
Updated: May 26, 2026


Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Assignment of protonation states in proteins and ligands: combining pKa prediction with hydrogen bonding network
Elmar Krieger1, Roland L Dunbrack, Rob W W Hooft
1Center for Molecular and Biomolecular Informatics, Radboud University Nijmegen Medical Centre, Nijmegen, The Netherlands. elmar@cmbi.ru.nl

Abstract:
Among the many applications of molecular modeling, drug design is probably the one with the highest demands on the accuracy of the underlying structures. During lead optimization, the position of every atom in the binding site should ideally be known with high precision to identify those chemical modifications that are most likely to increase drug affinity. Unfortunately, X-ray crystallography at common resolution yields an electron density map that is too coarse, since the chemical elements and their protonation states cannot be fully resolved.This chapter describes the steps required to fill in the missing knowledge, by devising an algorithm that can detect and resolve the ambiguities. First, the pK (a) values of acidic and basic groups are predicted. Second, their potential protonation states are determined, including all permutations (considering for example protons that can jump between the oxygens of a phosphate group). Third, those groups of atoms are identified that can adopt alternative but indistinguishable conformations with essentially the same electron density. Fourth, potential hydrogen bond donors and acceptors are located. Finally, all these data are combined in a single "configuration energy function," whose global minimum is found with the SCWRL algorithm, which employs dead-end elimination and graph theory. As a result, one obtains a complete model of the protein and its bound ligand, with ambiguous groups rotated to the best orientation and with protonation states assigned considering the current pH and the H-bonding network. An implementation of the algorithm has been available since 2008 as part of the YASARA modeling & simulation program.


Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
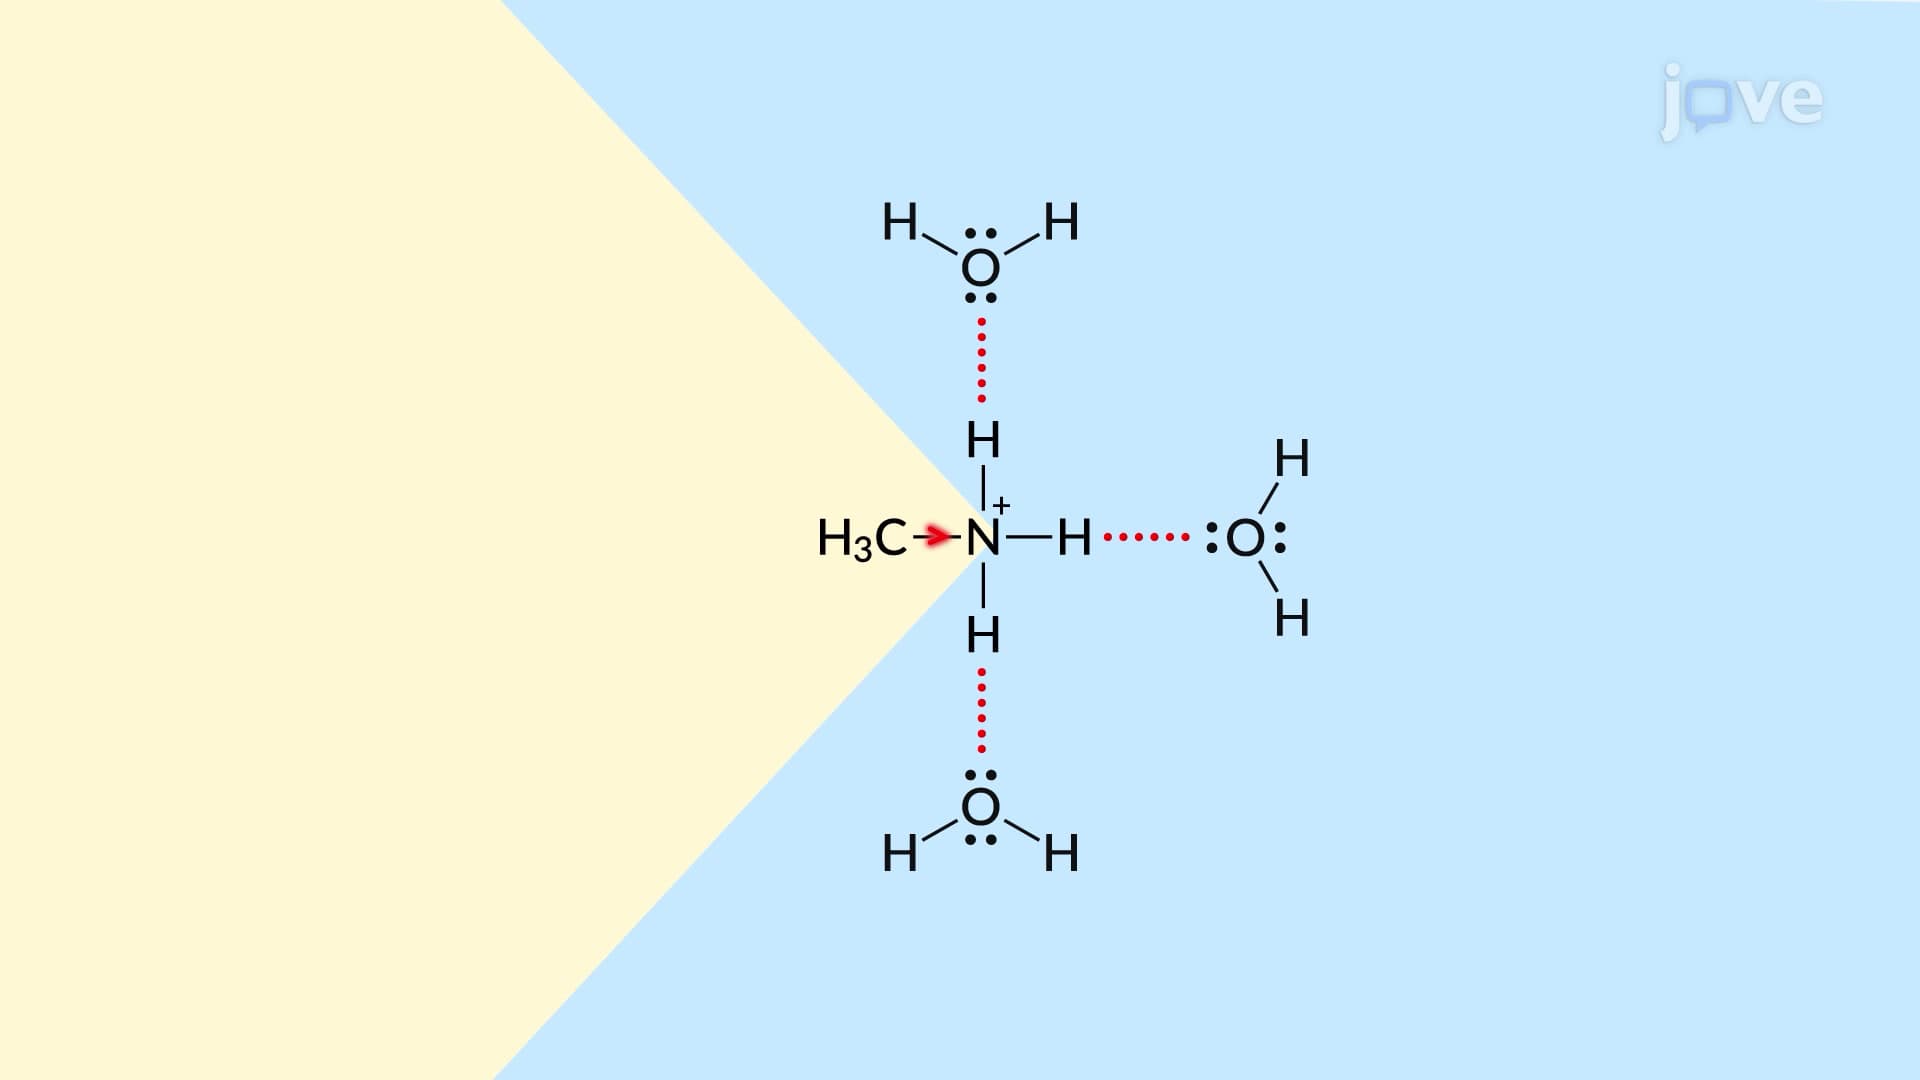 01:21
Basicity of Aliphatic Amines
To measure the basicity of amines, two conventions are generally used. The first defines Kb as the basicity constant for the deprotonation reaction of water by the amine, as presented in Figure 1. Conventionally, lower Kb indicates higher...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
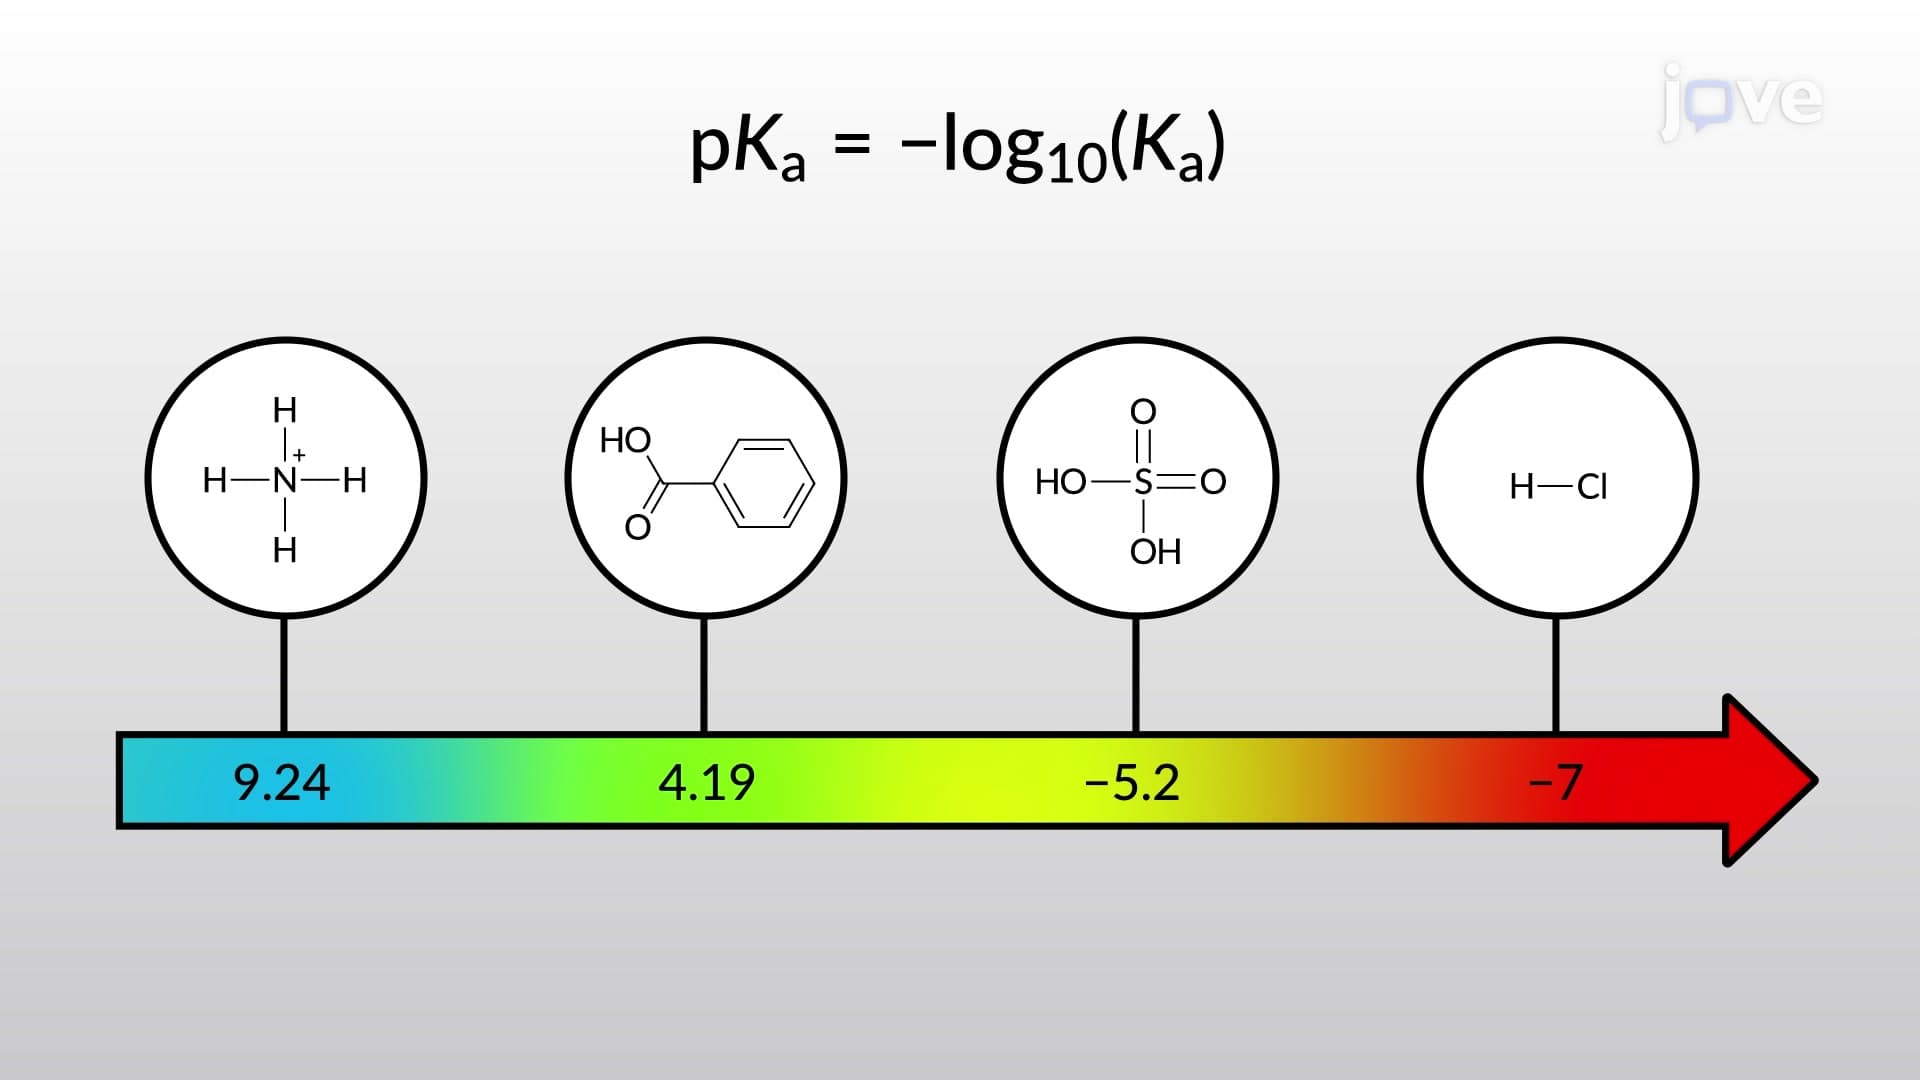 02:35
Acid and Bases: Ka, pKa, and Relative Strengths
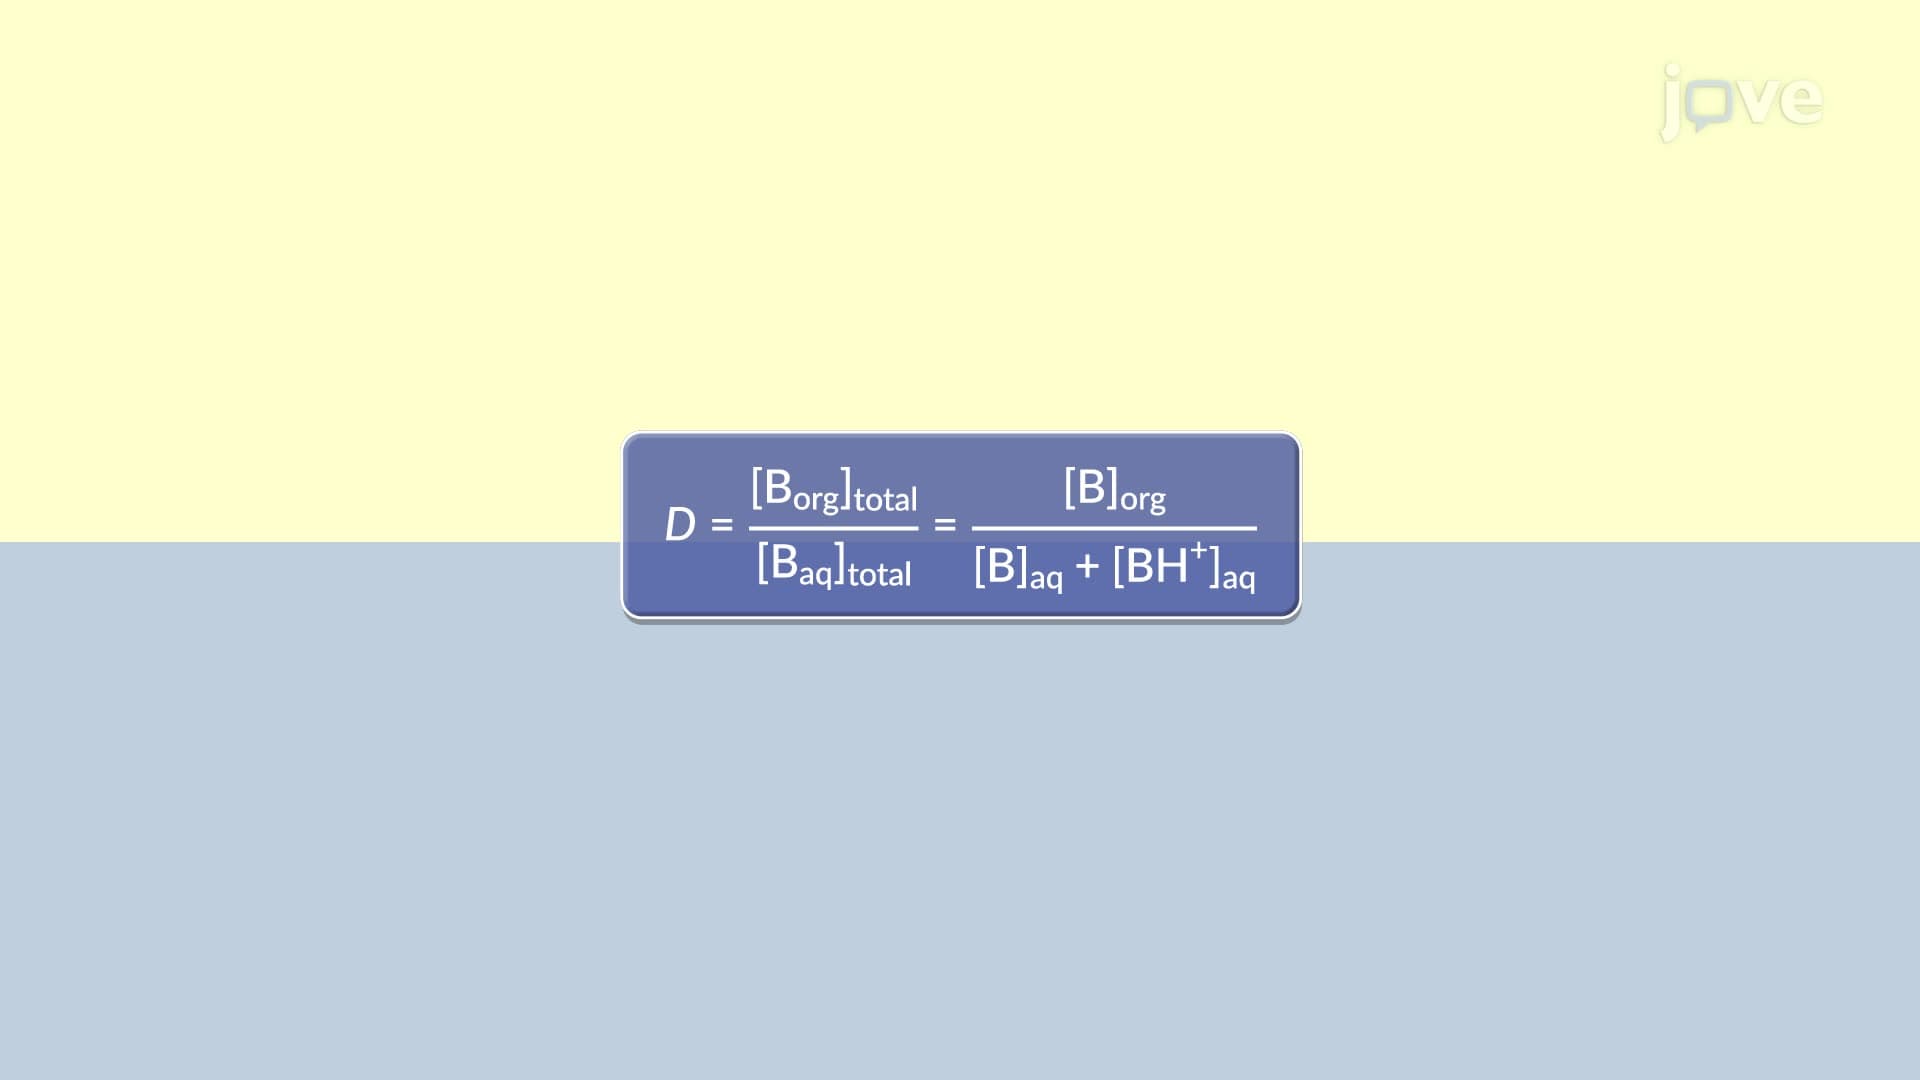 00:53
Extraction: Effects of pH
 02:29
Relative Strengths of Conjugate Acid-Base Pairs