Related Experiment Video
Updated: May 26, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Prediction of hydration free energies for aliphatic and aromatic chloro derivatives using molecular dynamics
Oliver Beckstein1, Bogdan I Iorga
1Structural Bioinformatics and Computational Biochemistry Unit, Department of Biochemistry, University of Oxford, Oxford OX1 3QU, UK.

Abstract:
All-atom molecular dynamics computer simulations were used to blindly predict the hydration free energies of a range of chloro-organic compounds as part of the SAMPL3 challenge. All compounds were parameterized within the framework of the OPLS-AA force field, using an established protocol to compute the absolute hydration free energy via a windowed free energy perturbation approach and thermodynamic integration. Three different approaches to deriving partial charge parameters were pursued: (1) using existing OPLS-AA atom types and charges with minor adjustments of partial charges on equivalent connecting atoms; (2) calculation of quantum mechanical charges via geometry optimization, followed by electrostatic potential (ESP) fitting, using Jaguar at the LMP2/cc-pVTZ(-F) level; and (3) via geometry optimization and CHelpG charges (Gaussian03 at the HF/6-31G* level), followed by two-stage RESP fitting. Protocol 3 generated the most accurate predictions with a root mean square (RMS) error of 1.2 kcal mol(-1) for the entire data set. It was found that the deficiency of the standard OPLS-AA parameters, protocol 1 (RMS error 2.4 kcal mol(-1) overall), was mostly due to compounds with more than three chlorine substituents on an aromatic ring. For this latter subset, the RMS errors were 1.4 kcal mol(-1) (protocol 3) and 4.3 kcal mol(-1) (protocol 1), respectively. We propose new OPLS-AA atom types for aromatic carbon and chlorine atoms in rings with ≥4 Cl-substituents that perform better than the best QM-based approach, resulting in an RMS error of 1.2 kcal mol(-1) for these difficult compounds.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Related Concept Videos
 01:20
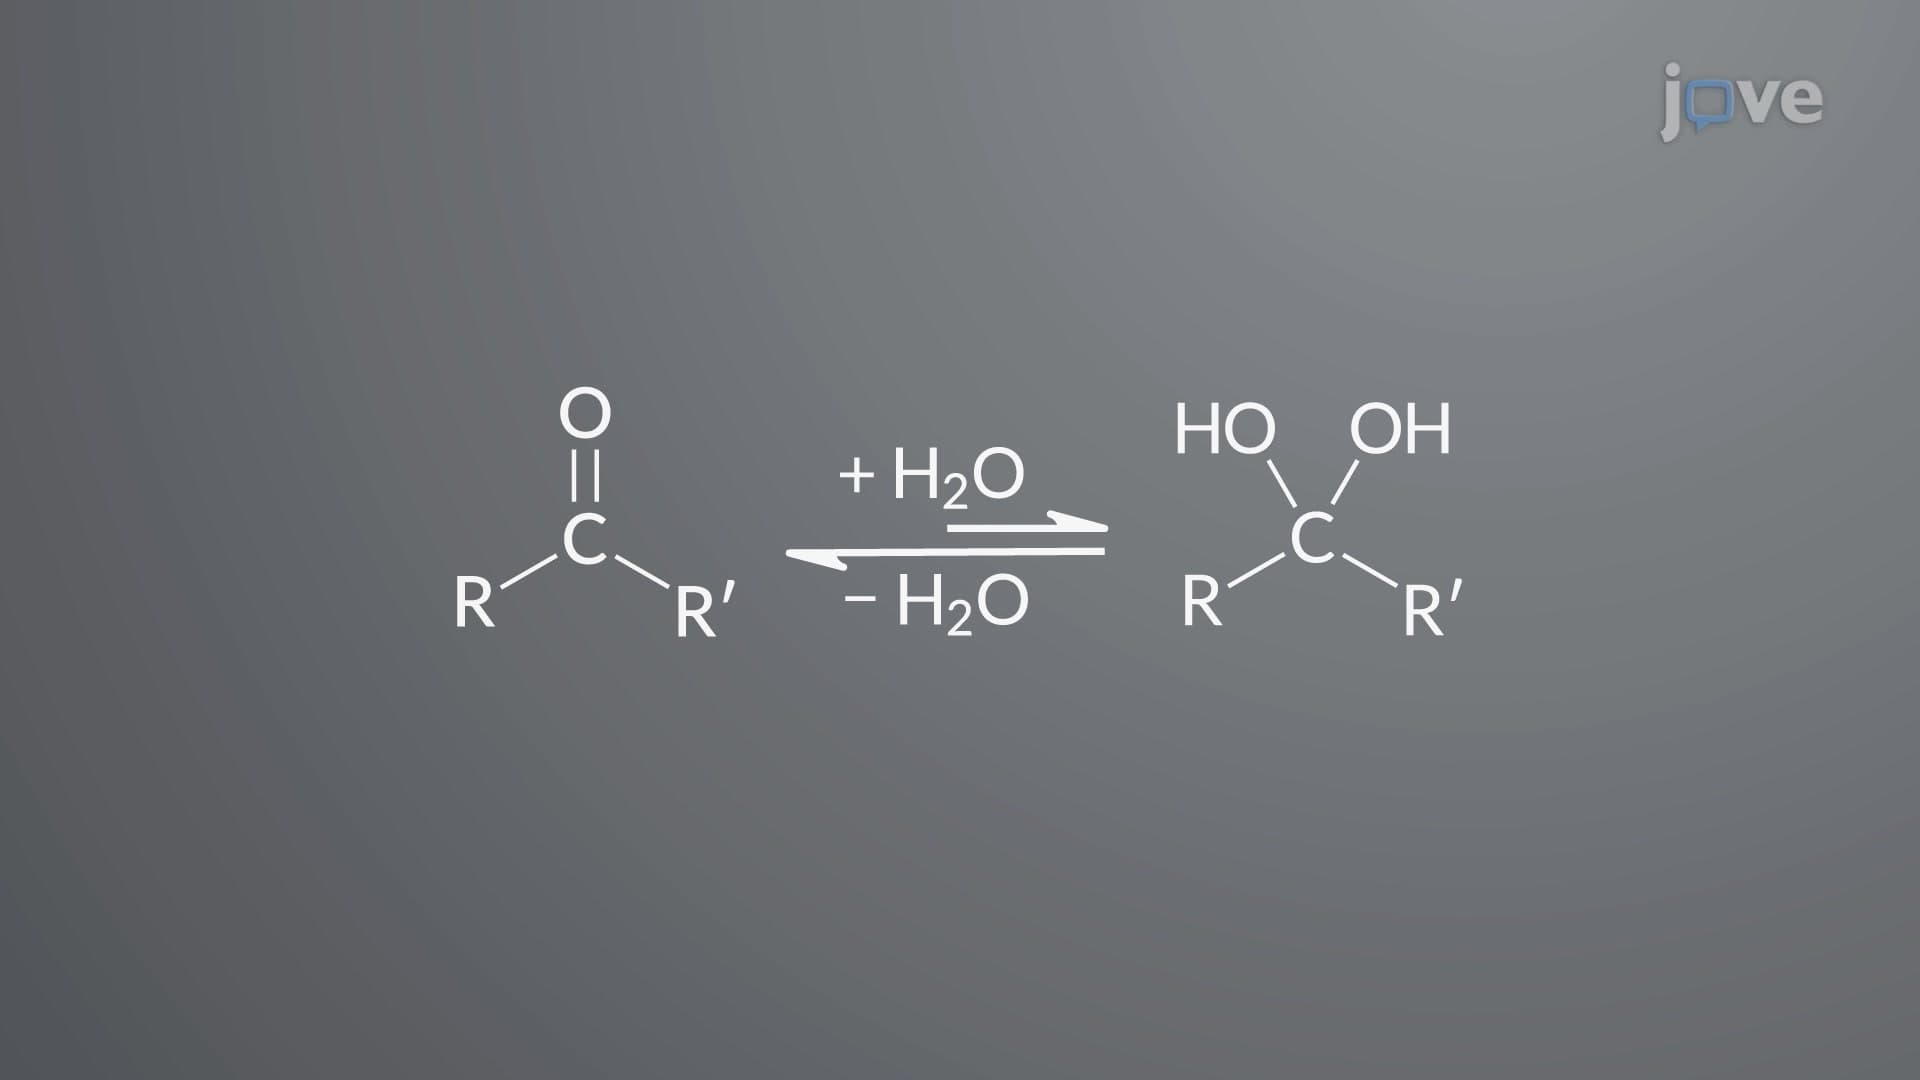
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:45
Acid-Catalyzed Hydration of Alkenes
 02:27
Predicting Molecular Geometry
 01:27
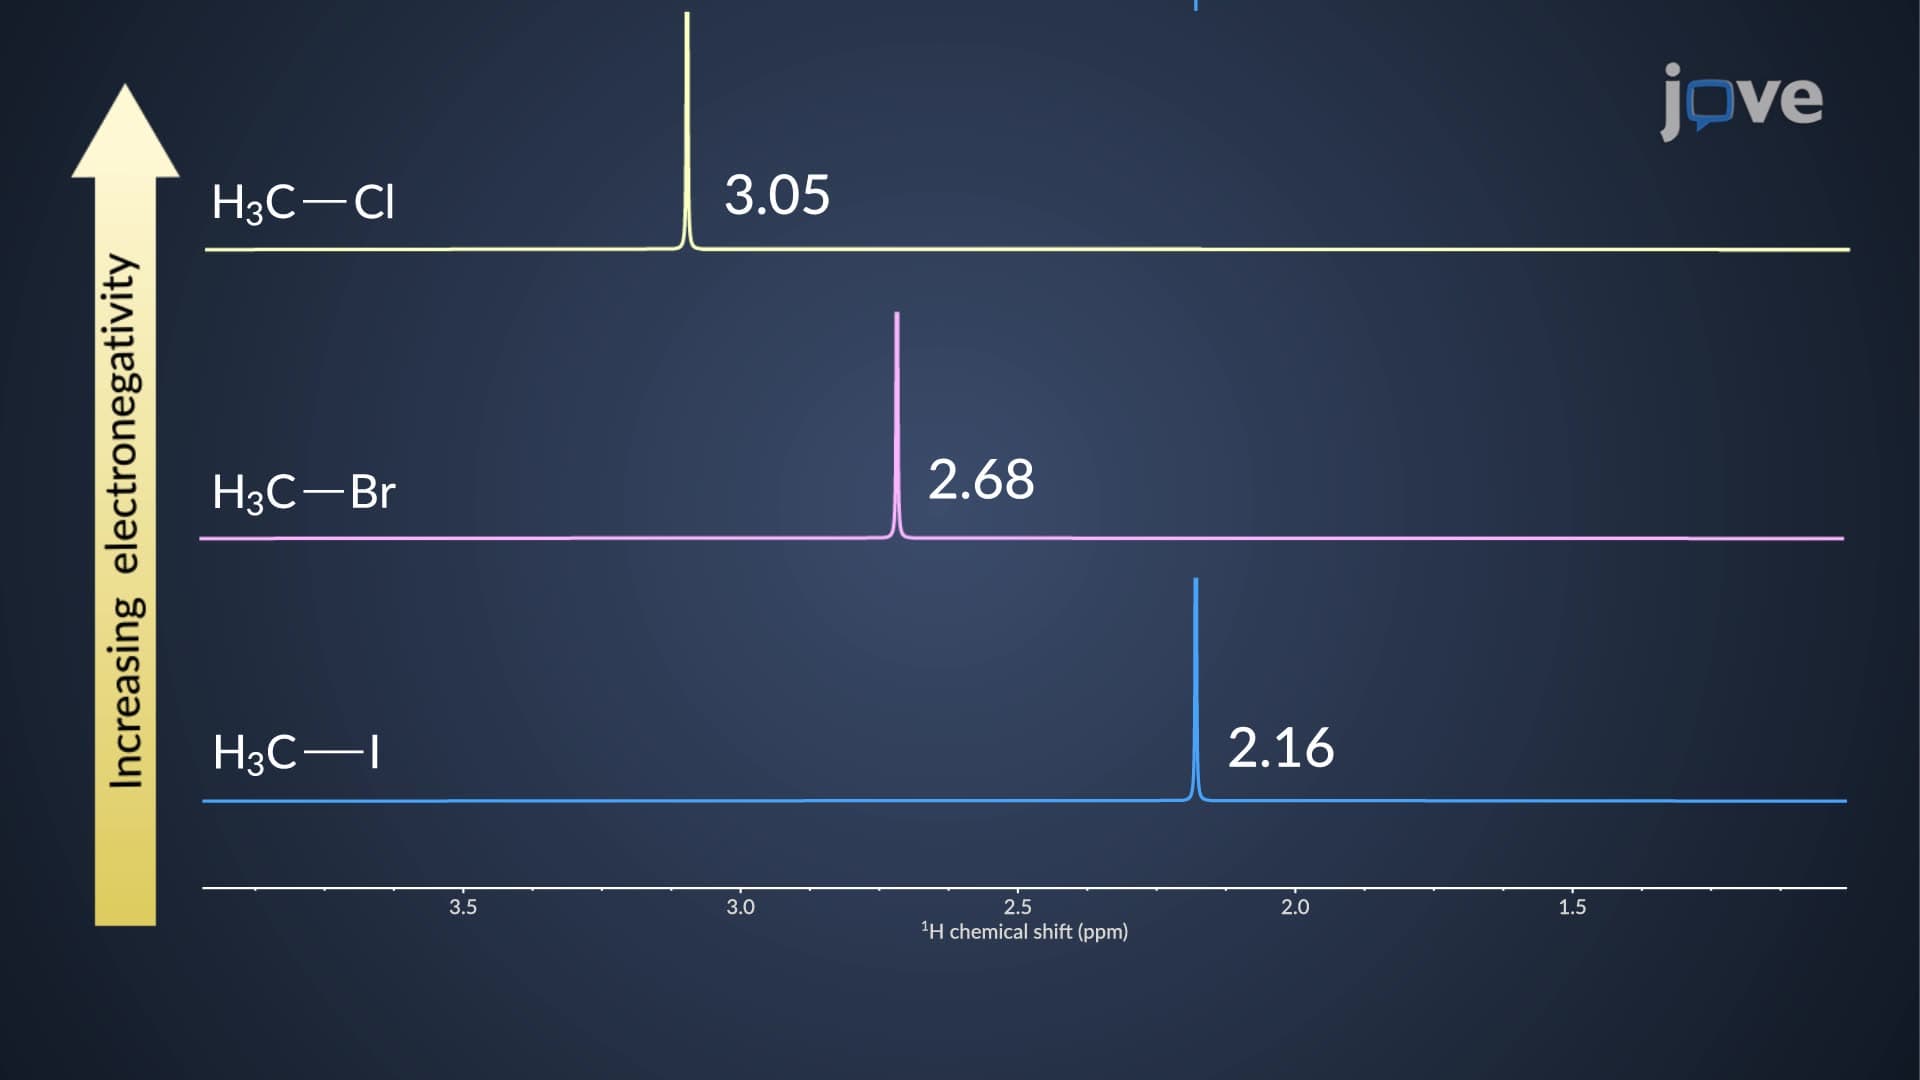
Inductive Effects on Chemical Shift: Overview
 01:29
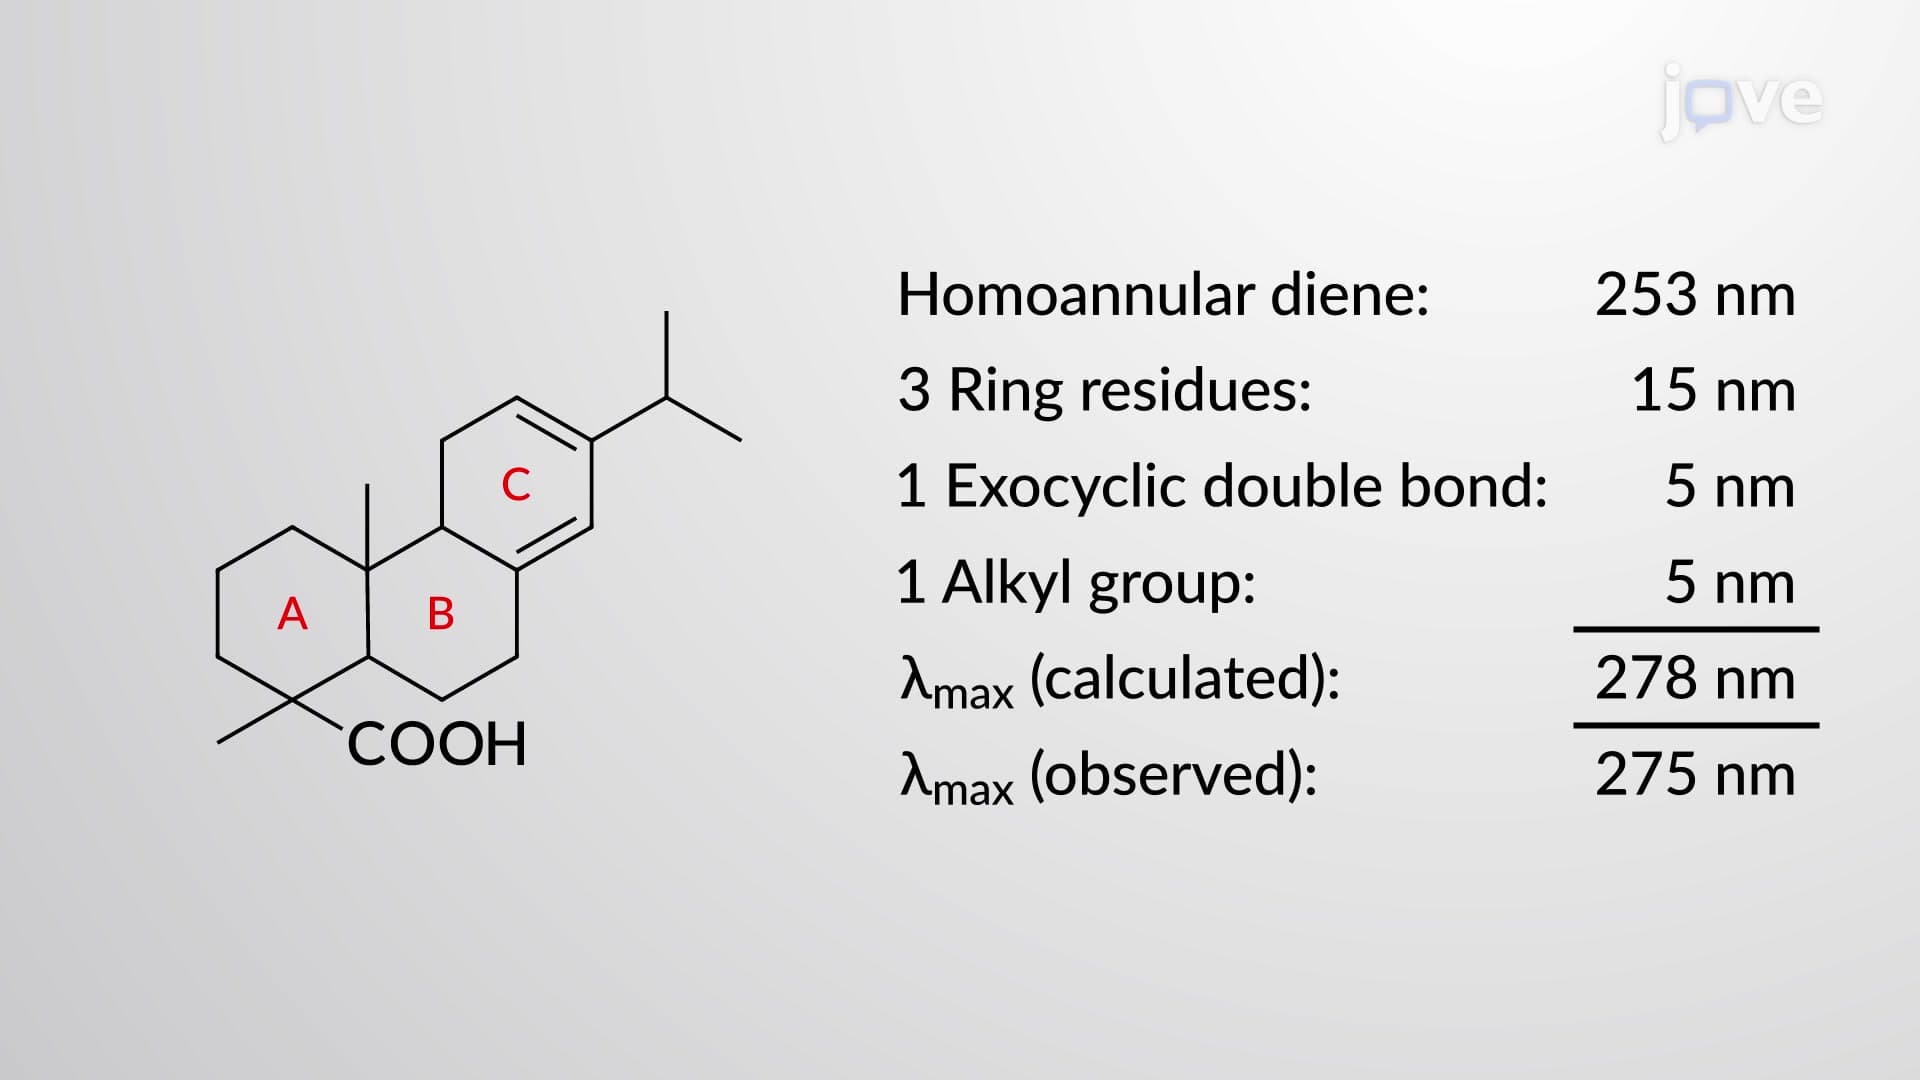
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:40
Alkynes to Aldehydes and Ketones: Acid-Catalyzed Hydration
Analogous to alkenes, alkynes also undergo acid-catalyzed hydration. While the addition of water to an alkene gives an alcohol, hydration of alkynes produces different products such as aldehydes and ketones.