Related Experiment Video
Updated: May 26, 2026
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Intermolecular interaction between CO or CO2 and ethylene oxide or ethylene sulfide in a complex, investigated by
Yoshiyuki Kawashima1, Akinori Sato, Yukari Orita
1Department of Applied Chemistry, Faculty of Engineering, Kanagawa Institute of Technology, Atsugi, Kanagawa 240-0292, Japan. kawasima@chem.kanagawa-it.ac.jp

Abstract:
The rotational spectra of the CO-ethylene oxide (EO), CO-ethylene sulfide (ES), CO(2)-EO, and CO(2)-ES complexes were measured by Fourier transform microwave spectroscopy in the frequency region from 4 up to 31 GHz. The isotopologues with a single (13)C atom in the EO or ES, (18)O in the EO, (34)S in the ES, and (13)C in the CO(2) moiety, respectively, were observed in natural abundance, and enriched samples, (13)CO or C(18)O in the CO-EO or CO-ES complex and C(18)OO and C(18)O(2) in the CO(2)-EO or CO(2)-ES complex, were employed to record respective rotational transitions. The rotational spectra observed for the CO-EO, CO-ES, CO(2)-EO, and CO(2)-ES complexes were analyzed by using an asymmetric-rotor S-reduced Hamiltonian to determine rotational and centrifugal distortion constants. The r(s) coordinates of the atoms in the four complexes, which were calculated from the observed rotational constants, led to a structure in which the CO or CO(2) moiety is located in a plane perpendicular to the EO or ES skeletal plane and bisecting the COC or CSC angle. We have also carried out ab initio molecular orbital calculations at the level of MP2 with basis sets 6-311++G(d,p) and aug-cc-pVDZ using the Gaussian 09 package. The MP2/6-311++G(d,p) calculations yield rotational constants in better agreement with the experimental values than with the other basis set; in other words, the molecular structures calculated using this basis set are close to those experimentally found for the ground state. The estimated dissociation energies of the complexes, including the zero-point vibrational energy corrections ΔZPV and the basis set superposition errors (BSSE) calculated with the counterpoise correction (CP), are in good agreement with the experimentally obtained binding energies E(B). We have applied an NBO analysis to the complexes to calculate the stabilization energy CT (=ΔE(σσ*)), which we found are closely correlated with the binding energies E(B). We have thus achieved a consistent overview on the intermolecular interaction in the complexes under consideration. It is to be noted that the spectral intensities of the inner OC(18)O-EO and OC(18)O-ES complexes were larger by a factor of 2 than those of the outer (18)OCO-EO/ES complexes. This observation was explained by the zero-point energy of the inner conformer being a little smaller than that of the outer one.
Related Concept Videos
 01:12
Cycloaddition Reactions: MO Requirements for Photochemical Activation
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene π orbitals.
 01:08
Spin–Spin Coupling Constant: Overview
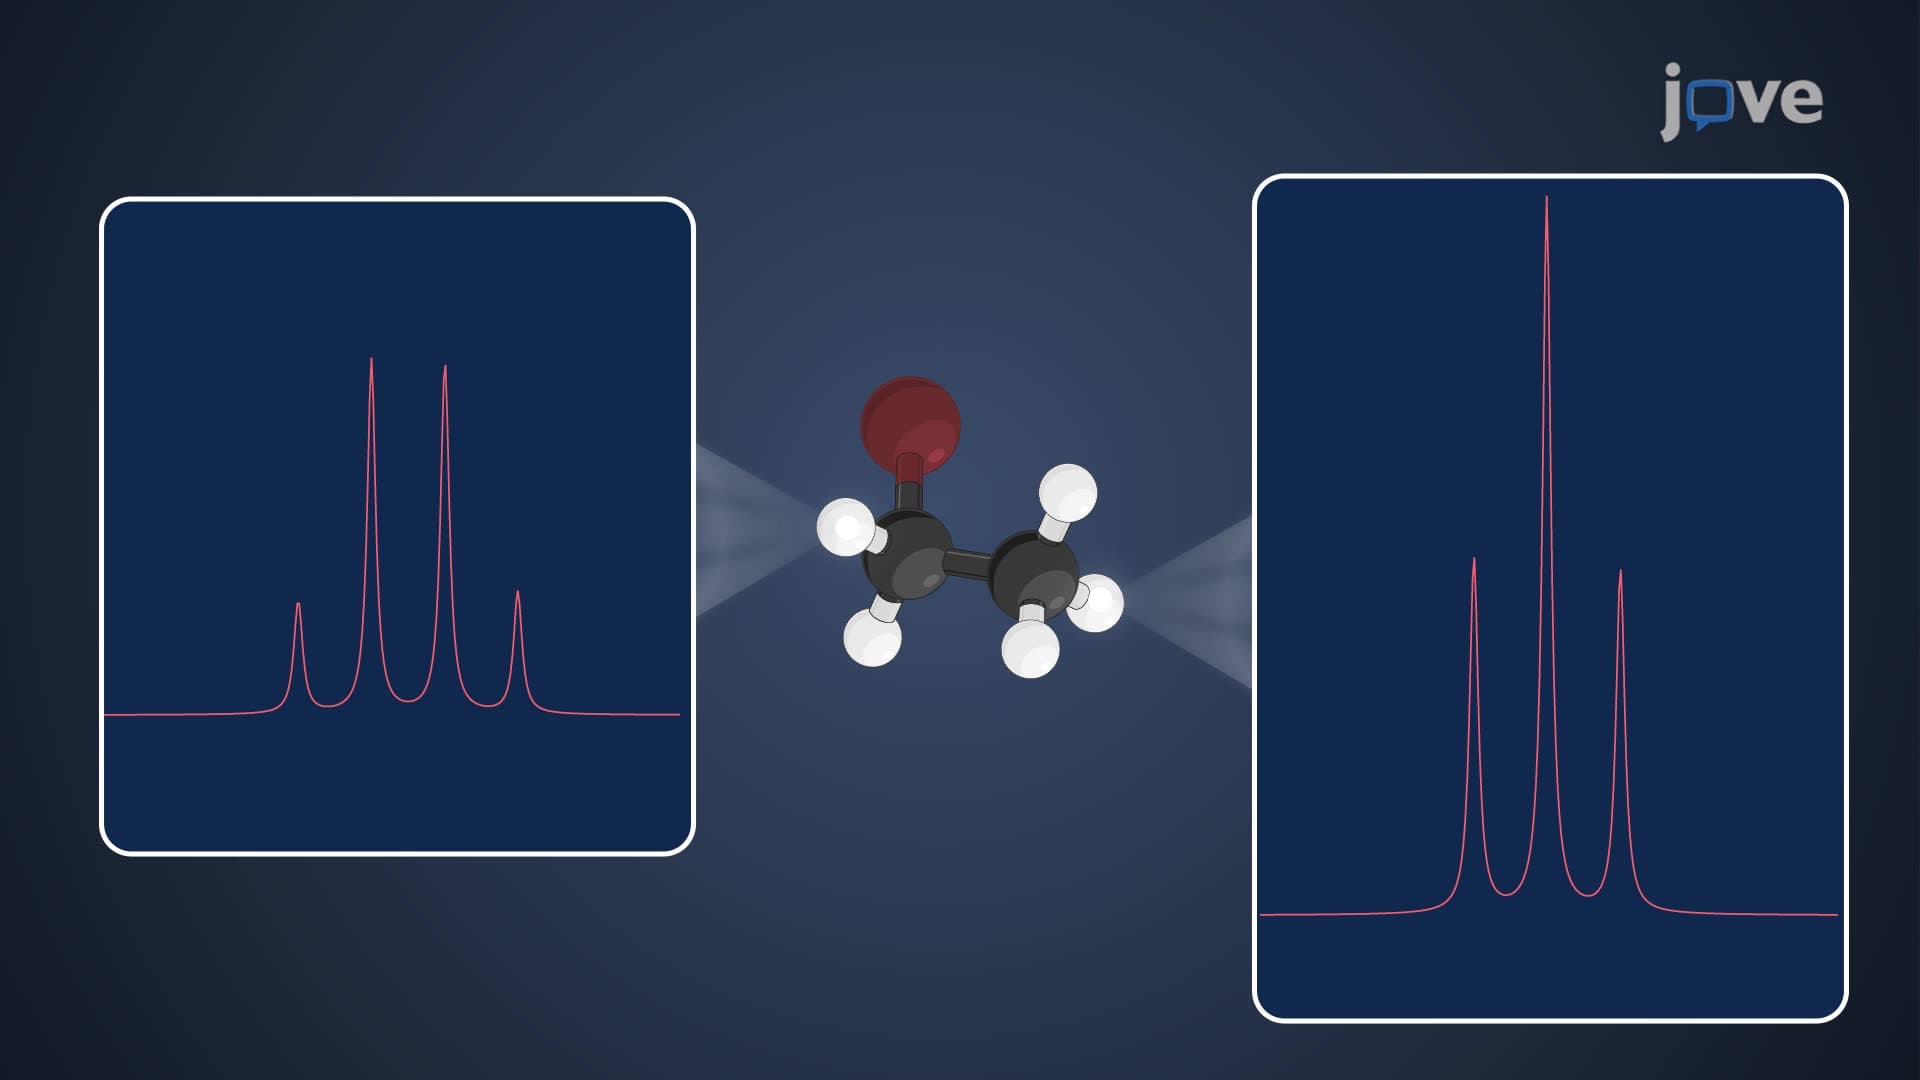
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)