Dispersive interactions in water bilayers at metallic surfaces: a comparison of the PBE and RPBE functional including semiempirical dispersion corrections
Related Experiment Video
Updated: May 26, 2026
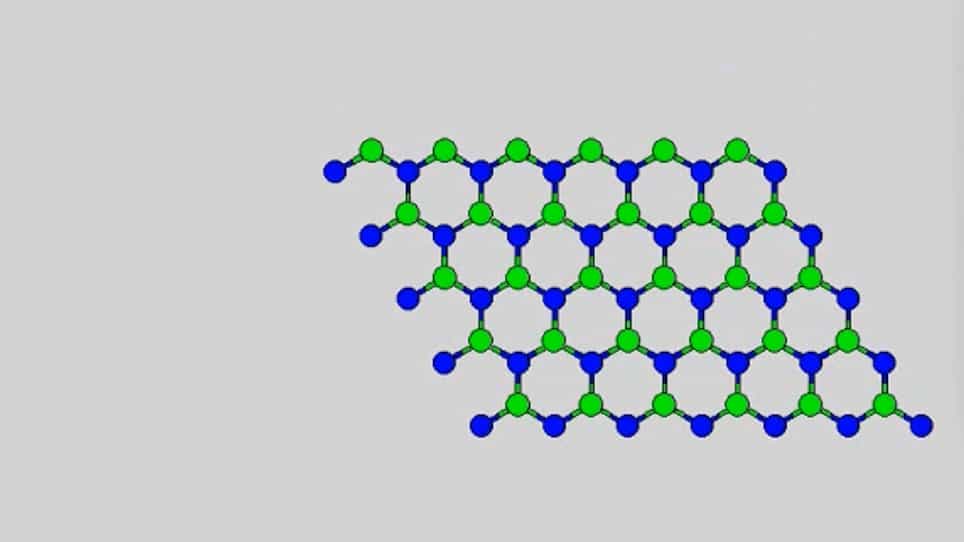
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
1Institute of Theoretical Chemistry, Ulm University, Ulm D-89069, Germany.

Abstract:
The accuracy and reliability of the density functional theory (DFT)-D approach to account for dispersion effects in first-principles studies of water-metal interfaces has been addressed by studying several water-metal systems. In addition to performing periodic DFT calculations for semi-infinite substrates using the popular PBE and RPBE functionals, the water dimer and water-metal atom systems have also been treated by coupled-cluster calculations. We show that indeed semiempirical dispersion correction schemes can be used to yield thermodynamically stable water bilayers at surfaces. However, the actual density functional needs to be chosen carefully. Whereas the dispersion-corrected RPBE functional yields a good description of both the water-water and the water-metal interaction, the dispersion-corrected PBE functional overestimates the energies of both systems. In contrast thereto, the adsorption distances predicted by the PBE functional is hardly changed due to the additional dispersion interaction, explaining the good performance of previous DFT-PBE studies of water-metal systems.
More Related Videos

Related Concept Videos
 01:24
Van der Waals Interactions
 02:56
Intermolecular Forces and Physical Properties
 03:13
Intermolecular Forces
03:13Intermolecular Forces
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...
 01:30
The Electrical Double Layer