Related Experiment Video
Updated: May 25, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Progress in time-dependent density-functional theory
1Laboratoire de Chimie Théorique, Département de Chimie Moléculaire, Institut de Chimie Moléculaire de Grenoble, Université Joseph Fourier, France. mark.casida@ujf-grenoble.fr

Abstract:
The classic density-functional theory (DFT) formalism introduced by Hohenberg, Kohn, and Sham in the mid-1960s is based on the idea that the complicated N-electron wave function can be replaced with the mathematically simpler 1-electron charge density in electronic structure calculations of the ground stationary state. As such, ordinary DFT cannot treat time-dependent (TD) problems nor describe excited electronic states. In 1984, Runge and Gross proved a theorem making TD-DFT formally exact. Information about electronic excited states may be obtained from this theory through the linear response (LR) theory formalism. Beginning in the mid-1990s, LR-TD-DFT became increasingly popular for calculating absorption and other spectra of medium- and large-sized molecules. Its ease of use and relatively good accuracy has now brought LR-TD-DFT to the forefront for this type of application. As the number and the diversity of applications of TD-DFT have grown, so too has our understanding of the strengths and weaknesses of the approximate functionals commonly used for TD-DFT. The objective of this article is to continue where a previous review of TD-DFT in Volume 55 of the Annual Review of Physical Chemistry left off and highlight some of the problems and solutions from the point of view of applied physical chemistry. Because doubly-excited states have a particularly important role to play in bond dissociation and formation in both thermal and photochemistry, particular emphasis is placed on the problem of going beyond or around the TD-DFT adiabatic approximation, which limits TD-DFT calculations to nominally singly-excited states.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:26
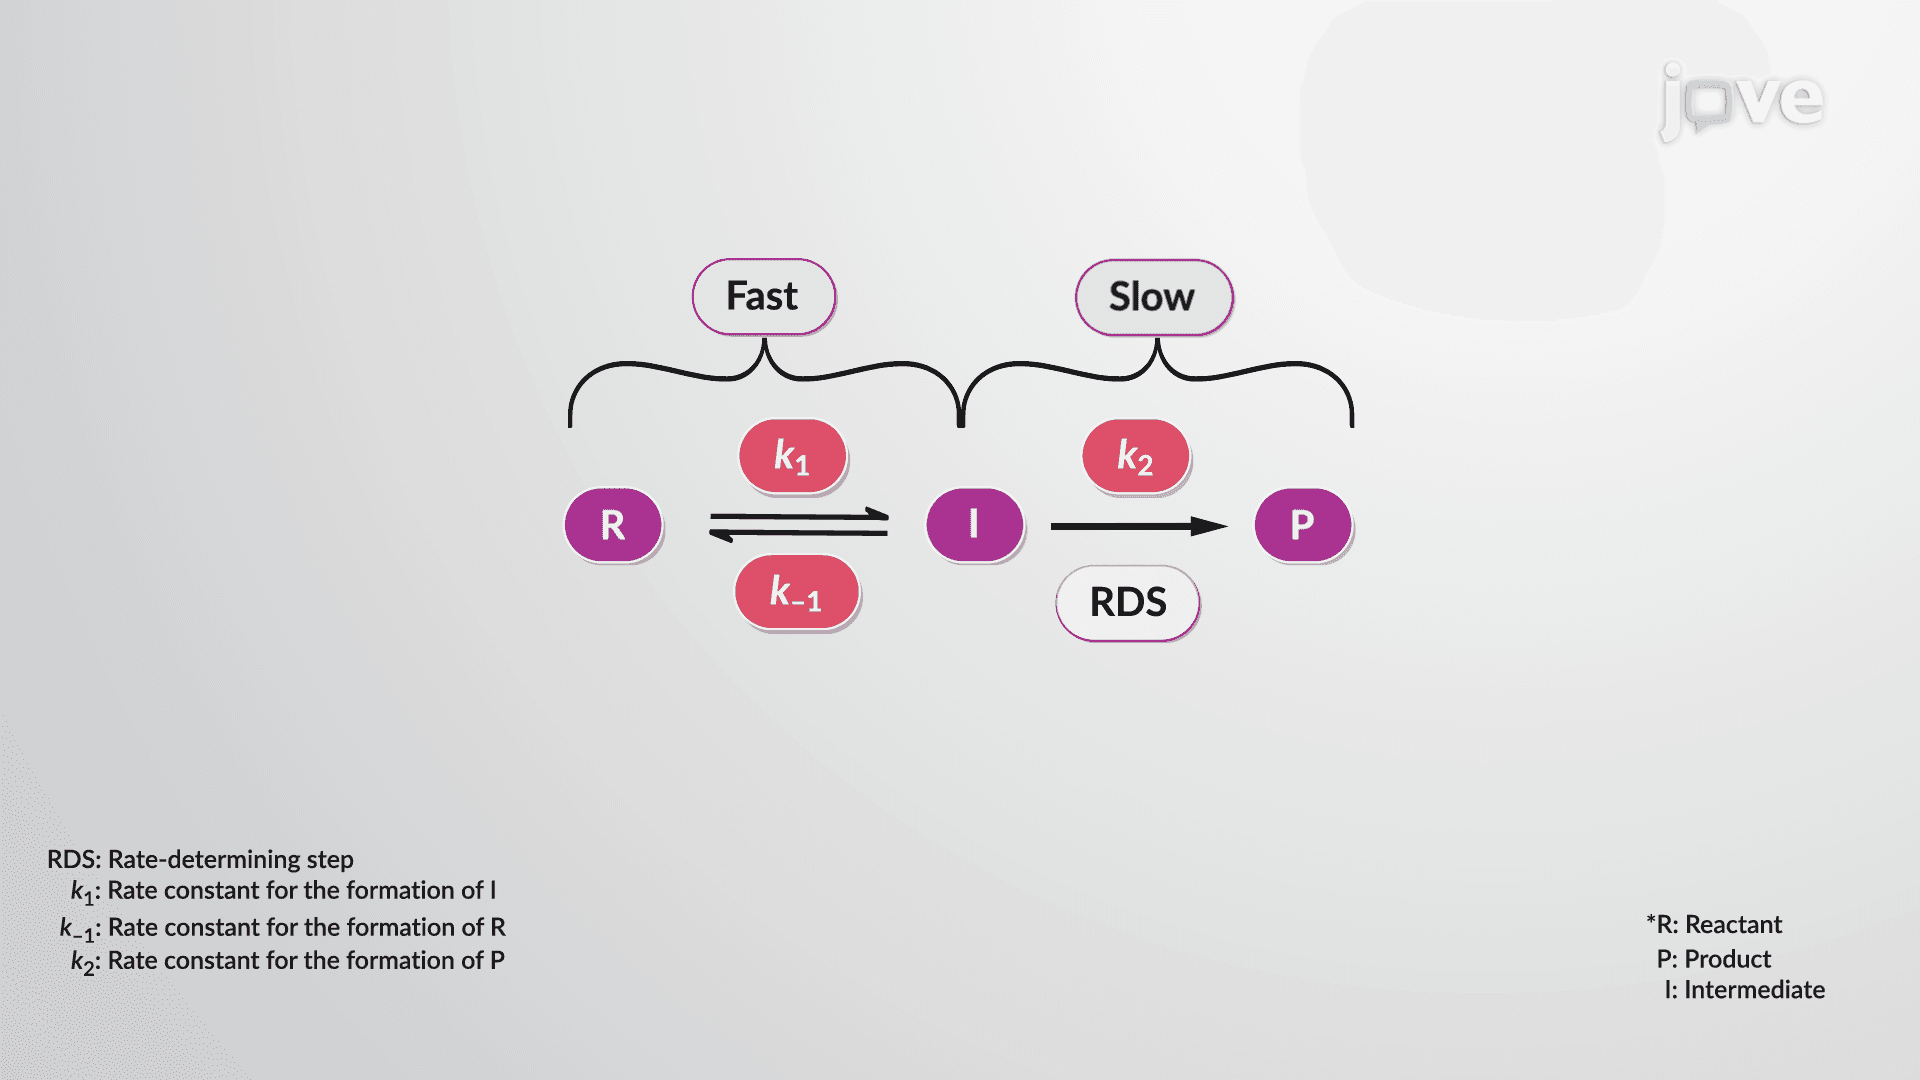
Reaction Mechanisms: The Steady-State Approximation
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 01:12
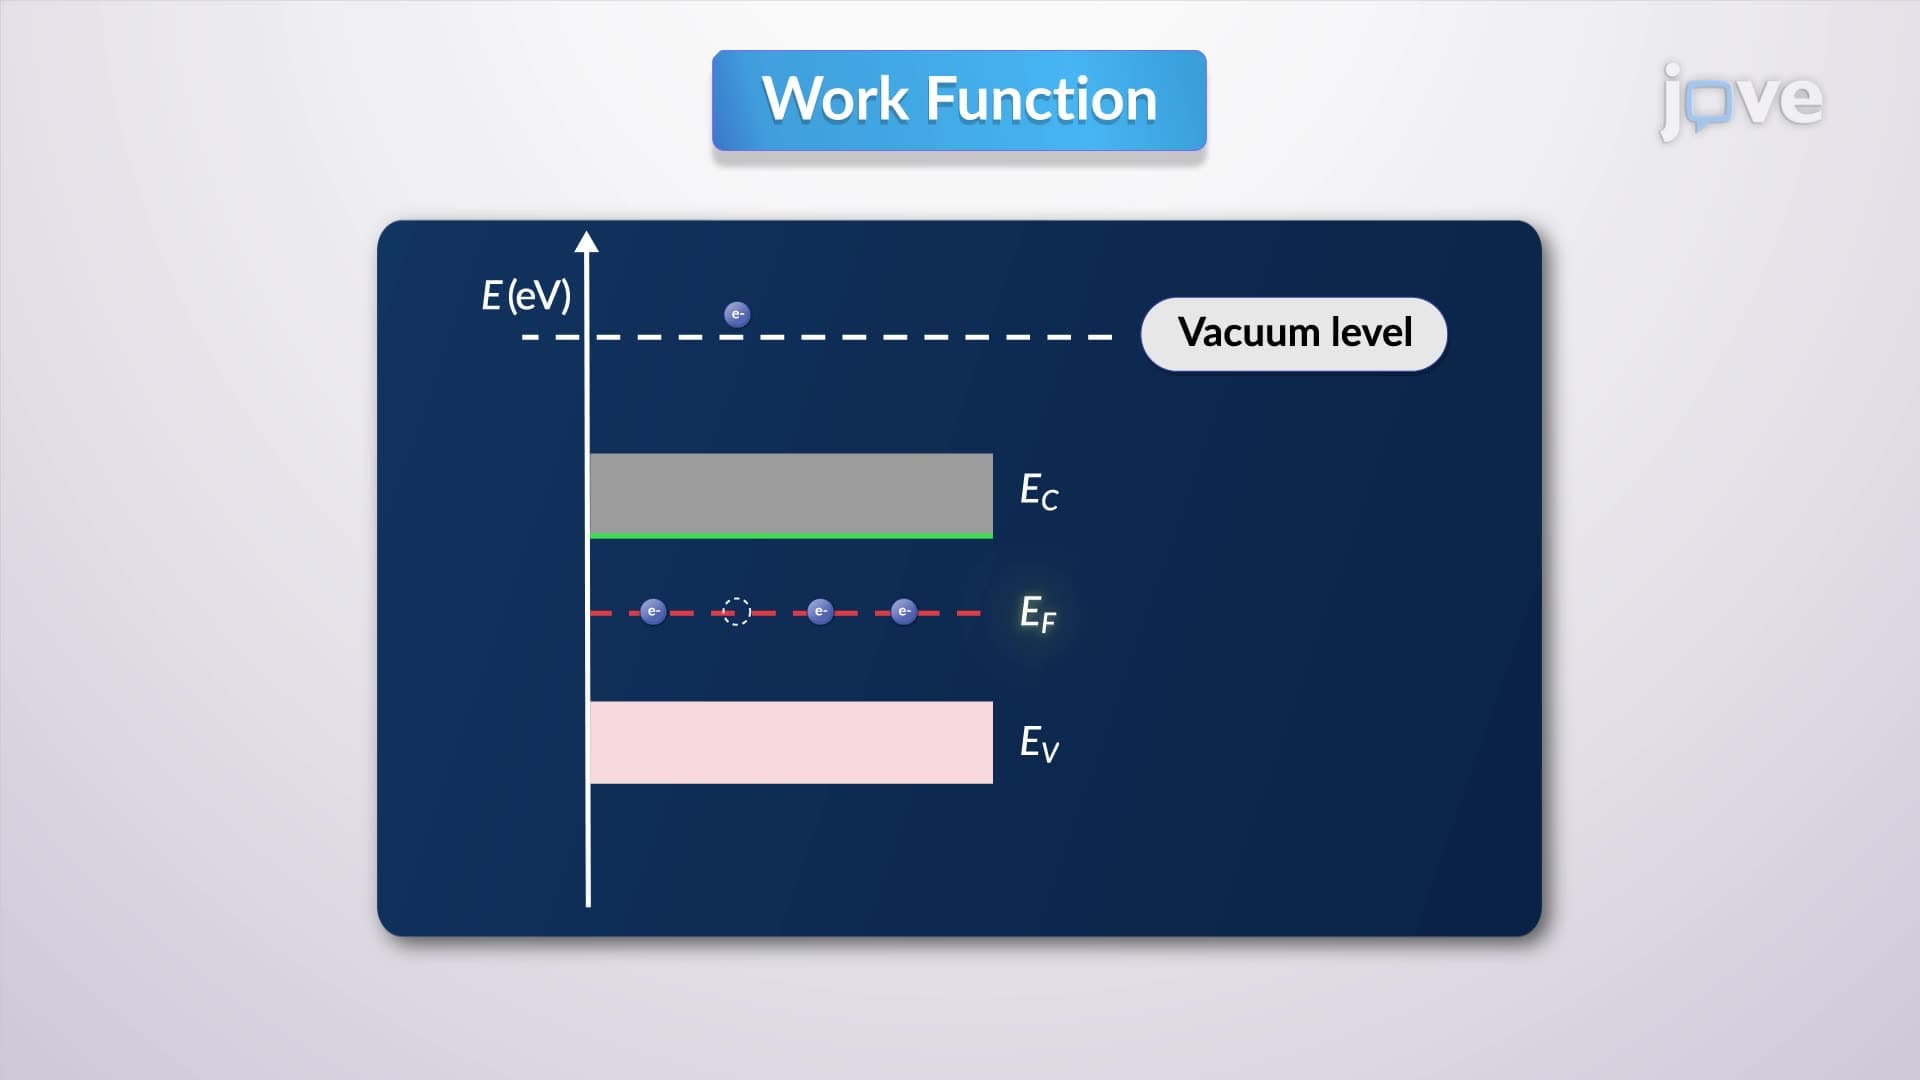
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:39
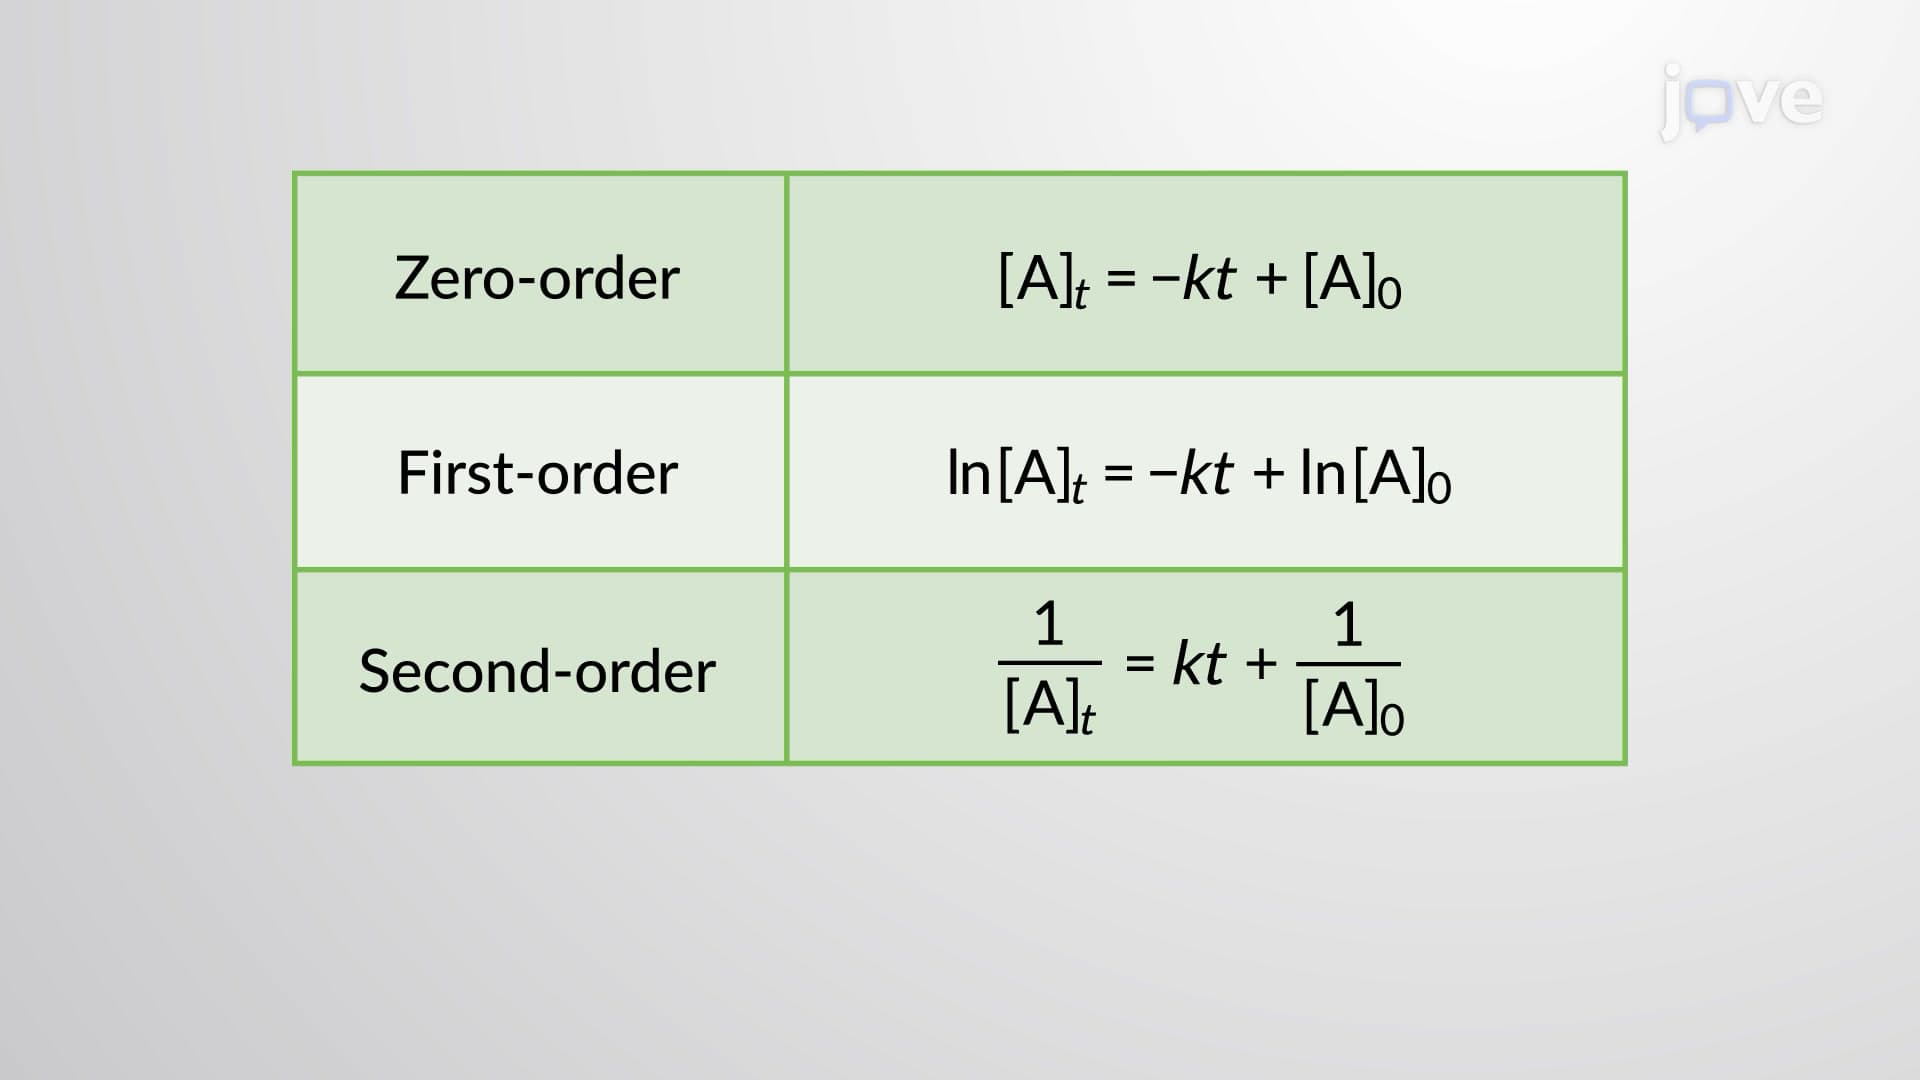
The Integrated Rate Law: The Dependence of Concentration on Time
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...