Related Experiment Video
Updated: May 25, 2026
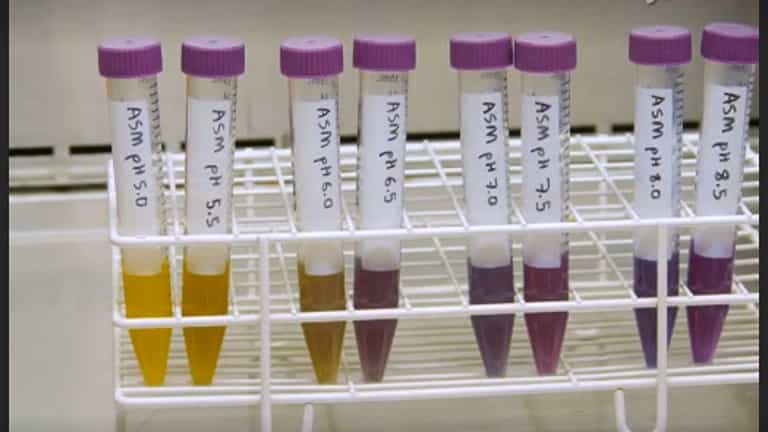

The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
An advanced stochastic model for mucociliary particle clearance in cystic fibrosis lungs
1Division of Physics and Biophysics, Department of Material Science and Physics, University of Salzburg, Salzburg, Austria.

Background:
A mathematical model describing mucociliary clearance in cystic fibrosis (CF) patients and its development with progressing course of the disease was developed. The approach should support the prediction of the disease state on the basis of measured bronchial clearance efficiencies.
Methods:
The approach is based on the assumption of a steady-state steady-flow mucus transport through the tracheobronchial tree which enables the determination of airway generation-specific mucus velocities by using a measured tracheal mucus velocity and a realistic morphometric dataset of the human lung. Architecture of the tracheobronchial tree was approximated by a stochastic model, reflecting the intra-subject variability of geometric parameters within a given lung generation.
Results:
As predicted by the appropriately validated mathematical approach, mucociliary clearance efficiency in CF patients is partly significantly decreased with respect to healthy controls. 24-h retention of patients with mild CF (FEV(1) >70% of predicted) is reduced by 10% compared to healthy subjects, whilst 24-h retention of patients with moderate to severe CF (FEV(1) <70% of predicted) differs by 25% from that of the healthy controls. These discrepancies are further enhanced with continuation of the clearance process.
Conclusions:
The theoretical results lead to the conclusion that CF patients have a higher risk of inhaled particle accumulation and related particle overload in specific lung compartments than healthy subjects.
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
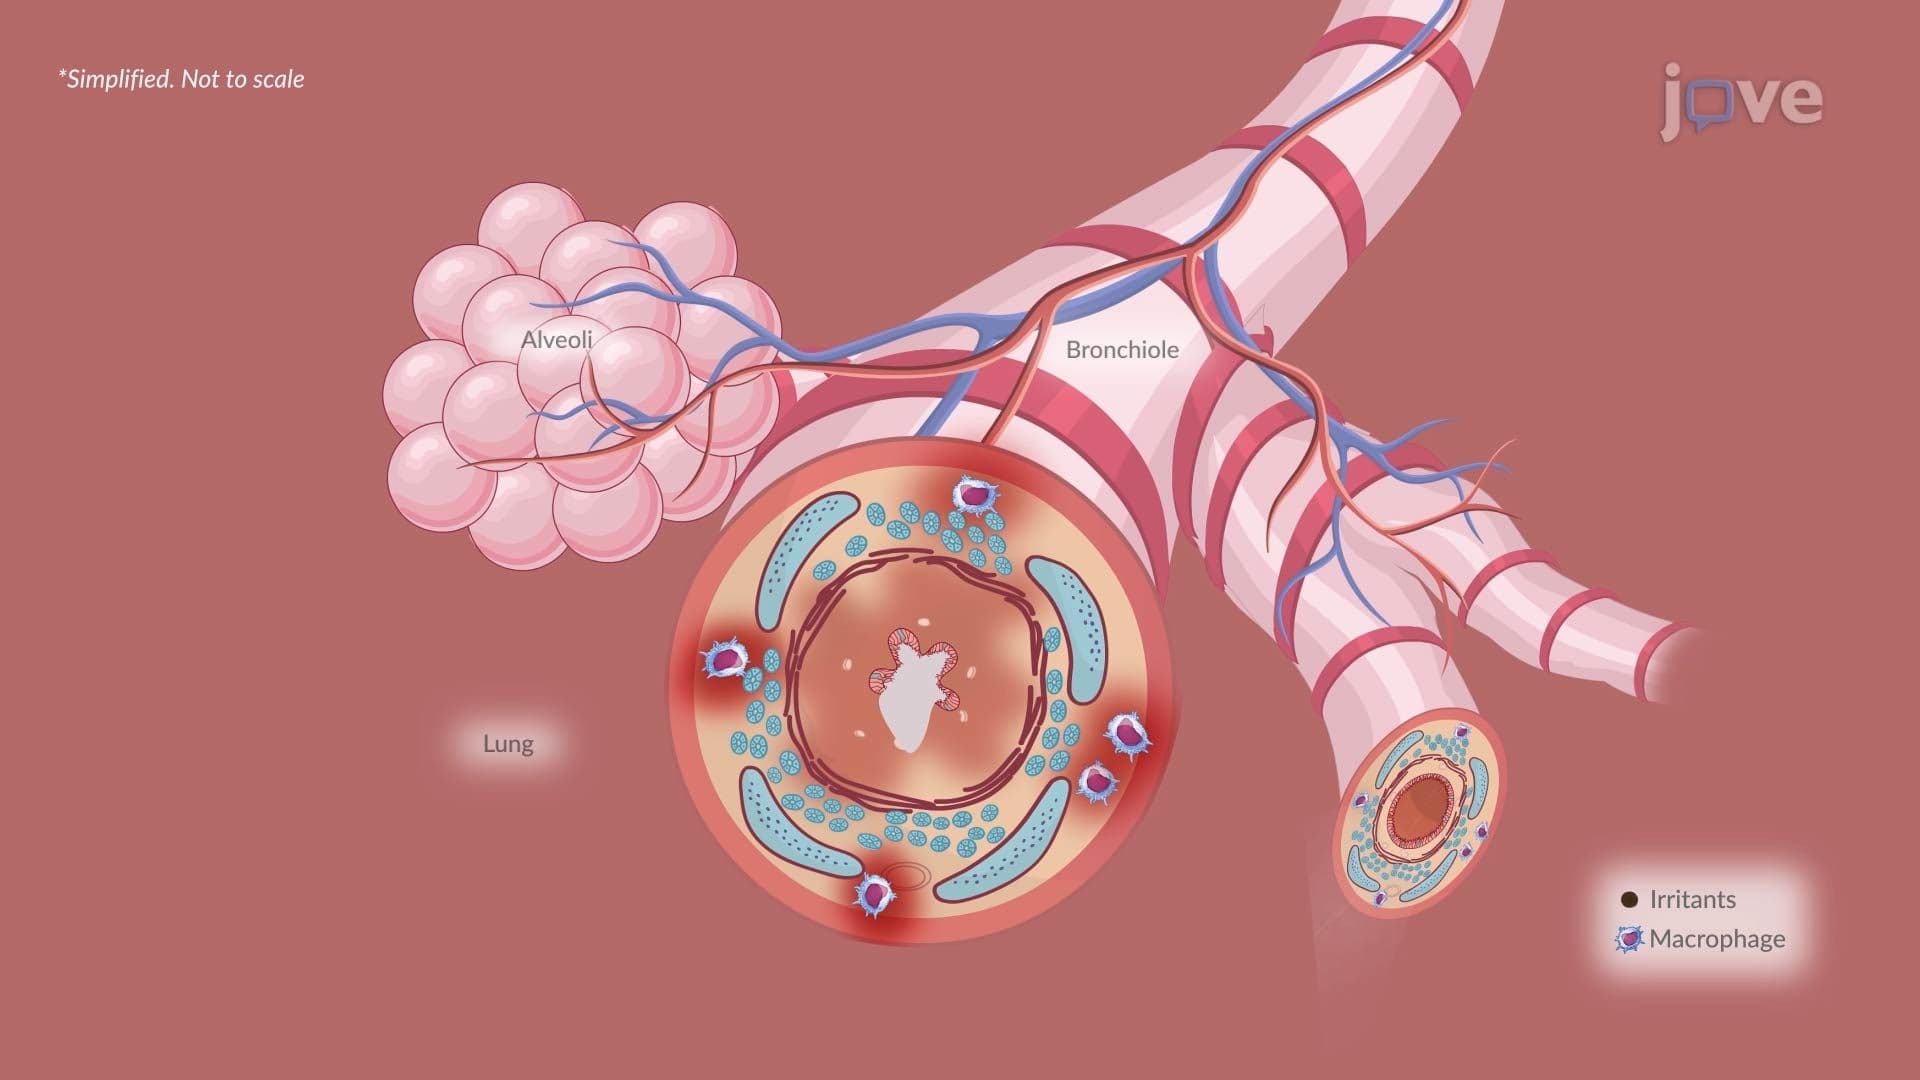
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features