Communication: improving the density functional theory+U description of CeO2 by including the contribution of the O
José J Plata1, Antonio M Márquez, Javier Fdez Sanz
1Departamento de Química Física, Facultad de Química, Universidad de Sevilla, 41012 Sevilla, Spain.

Abstract:
Density functional theory (DFT) based approaches within the local-density approximation or generalized gradient approximation frameworks fail to predict the correct electron localization in strongly correlated systems due to the lack of cancellation of the Coulomb self-interaction. This problem might be circumvented either by using hybrid functionals or by introducing a Hubbard-like term to account for the on site interactions. This latter DFT+U approach is less expensive and therefore more practical for extensive calculations in solid-state computational simulations. By and large, the U term only affects the metal electrons, in our case the Ce 4f ones. In the present work, we report a systematic analysis of the effect of adding such a U term also to the oxygen 2p electrons. We find that using a set of U(f) = 5 eV and U(p) = 5eV effective terms leads to improved description of the lattice parameters, band gaps, and formation and reduction energies of CeO(2).
More Related Videos

Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:51
Molecular Orbital Theory II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
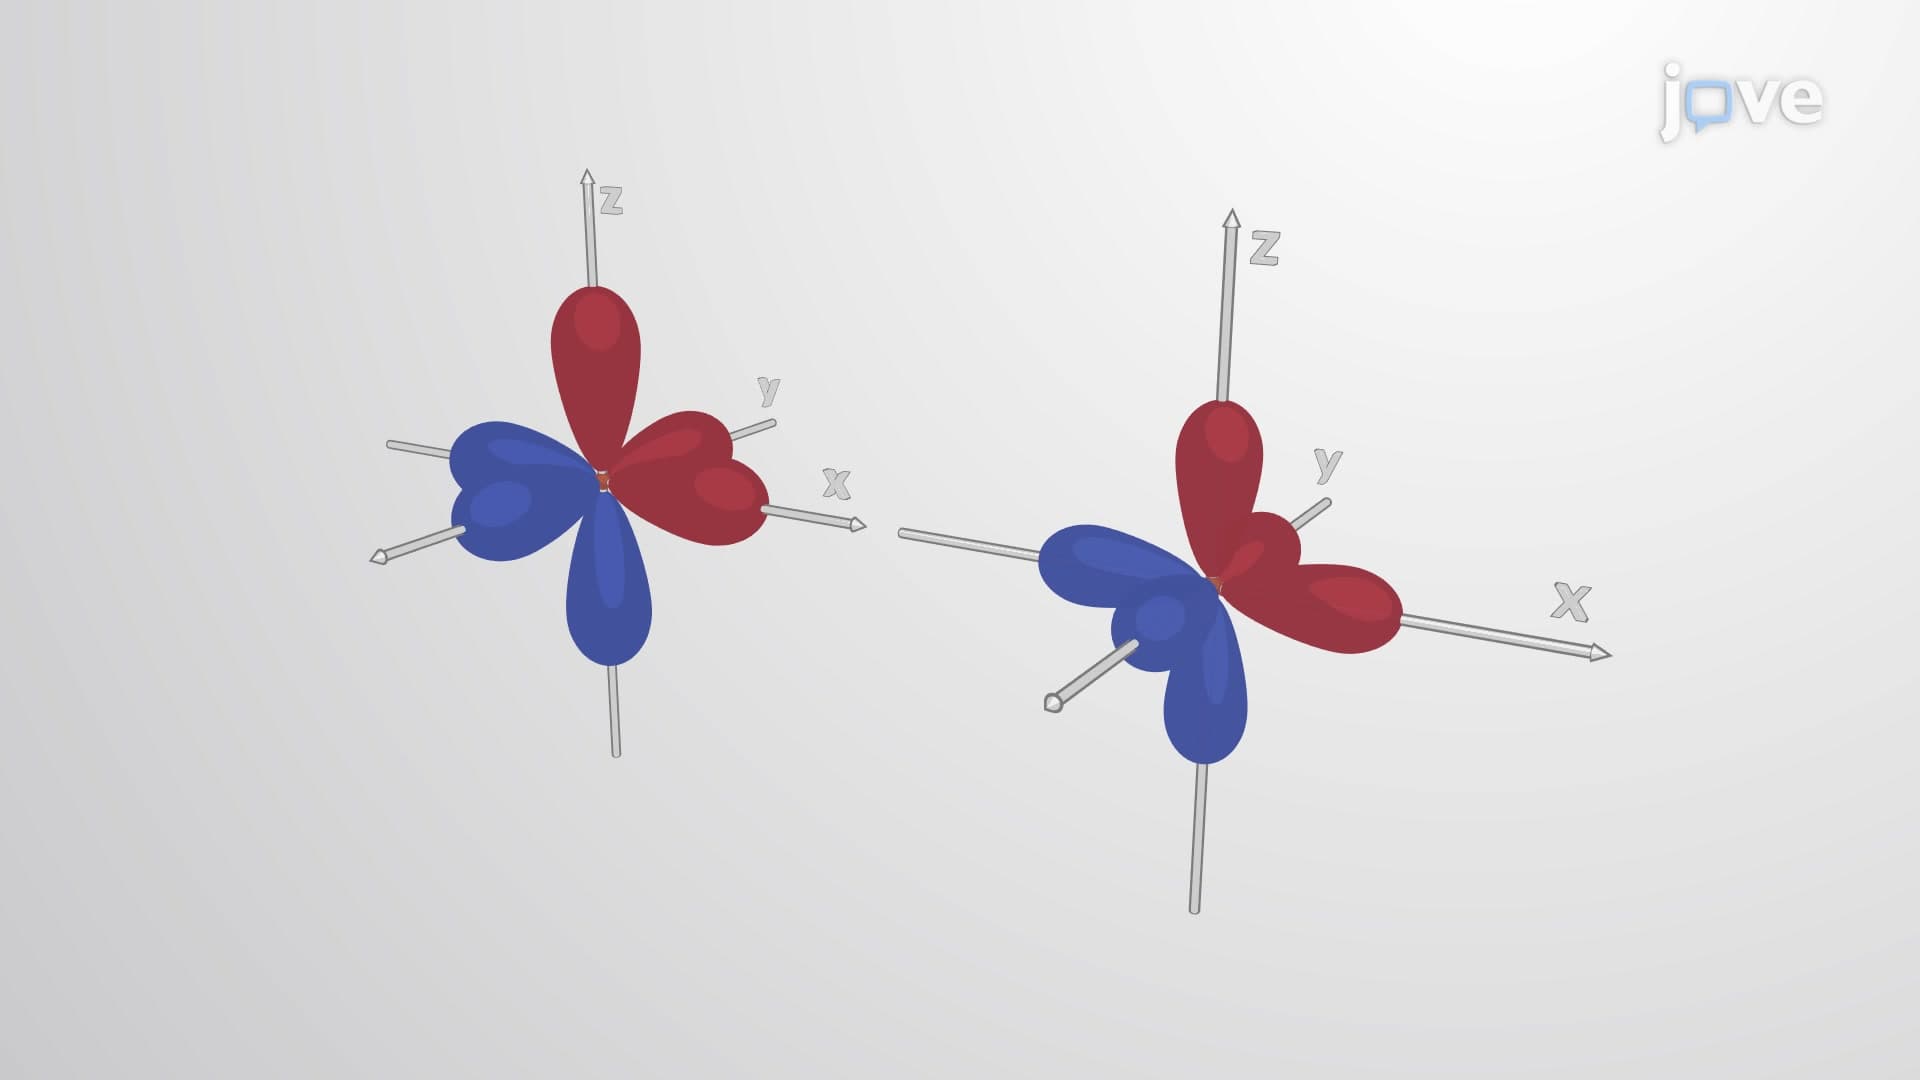
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:35
Molecular Orbital Theory I