Molecular simulation and macroscopic modeling of the diffusion of hydrogen, carbon monoxide and water in heavy n-alkane mixtures
Related Experiment Video
Updated: May 24, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Zoi A Makrodimitri1, Dominik J M Unruh, Ioannis G Economou
1National Center for Scientific Research Demokritos, Institute of Physical Chemistry, Molecular Thermodynamics and Modelling of Materials Laboratory, GR-153 10 Aghia Paraskevi Attikis, Greece.

Abstract:
The self-diffusion coefficient of hydrogen (H(2)), carbon monoxide (CO) and water (H(2)O) in n-alkanes was studied by molecular dynamics simulation. Diffusion in a few pure n-alkanes (namely n-C(8), n-C(20), n-C(64) and n-C(96)) was examined. In addition, binary n-C(12)-n-C(96) mixtures with various compositions as well as more realistic five- and six-n-alkane component mixtures were simulated. In all cases, the TraPPE united atom force field was used for the n-alkane molecules. The force field for the mixture of n-alkanes was initially validated against experimental density values and was shown to be accurate. Moreover, macroscopic correlations for predicting diffusion coefficient of H(2), CO and H(2)O in n-alkanes and mixtures of n-alkanes were developed. The functional form of the correlation was based on the rough hard sphere theory (RHS). The correlation was applied to simulation data and an absolute average deviation (AAD) of 5.8% for pure n-alkanes and 3.4% for n-alkane mixtures was obtained. Correlation parameters vary in a systematic way with carbon number and so they can be used to provide predictions in the absence of any experimental or molecular simulation data. Finally, in order to reduce the number of adjustable parameters, for the n-alkane mixtures the "pseudo-carbon number" approach was used. This approach resulted in relatively higher deviation from MD simulation data (AAD of 18.2%); however, it provides a convenient and fast method to predict diffusion coefficients. The correlations developed here are expected to be useful for engineering calculations related to the design of the Gas-to-Liquid process.
Related Concept Videos
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 02:00
Molecular Models
 02:26
Molecular Comparison of Gases, Liquids, and Solids
 02:56
Intermolecular Forces and Physical Properties
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:33
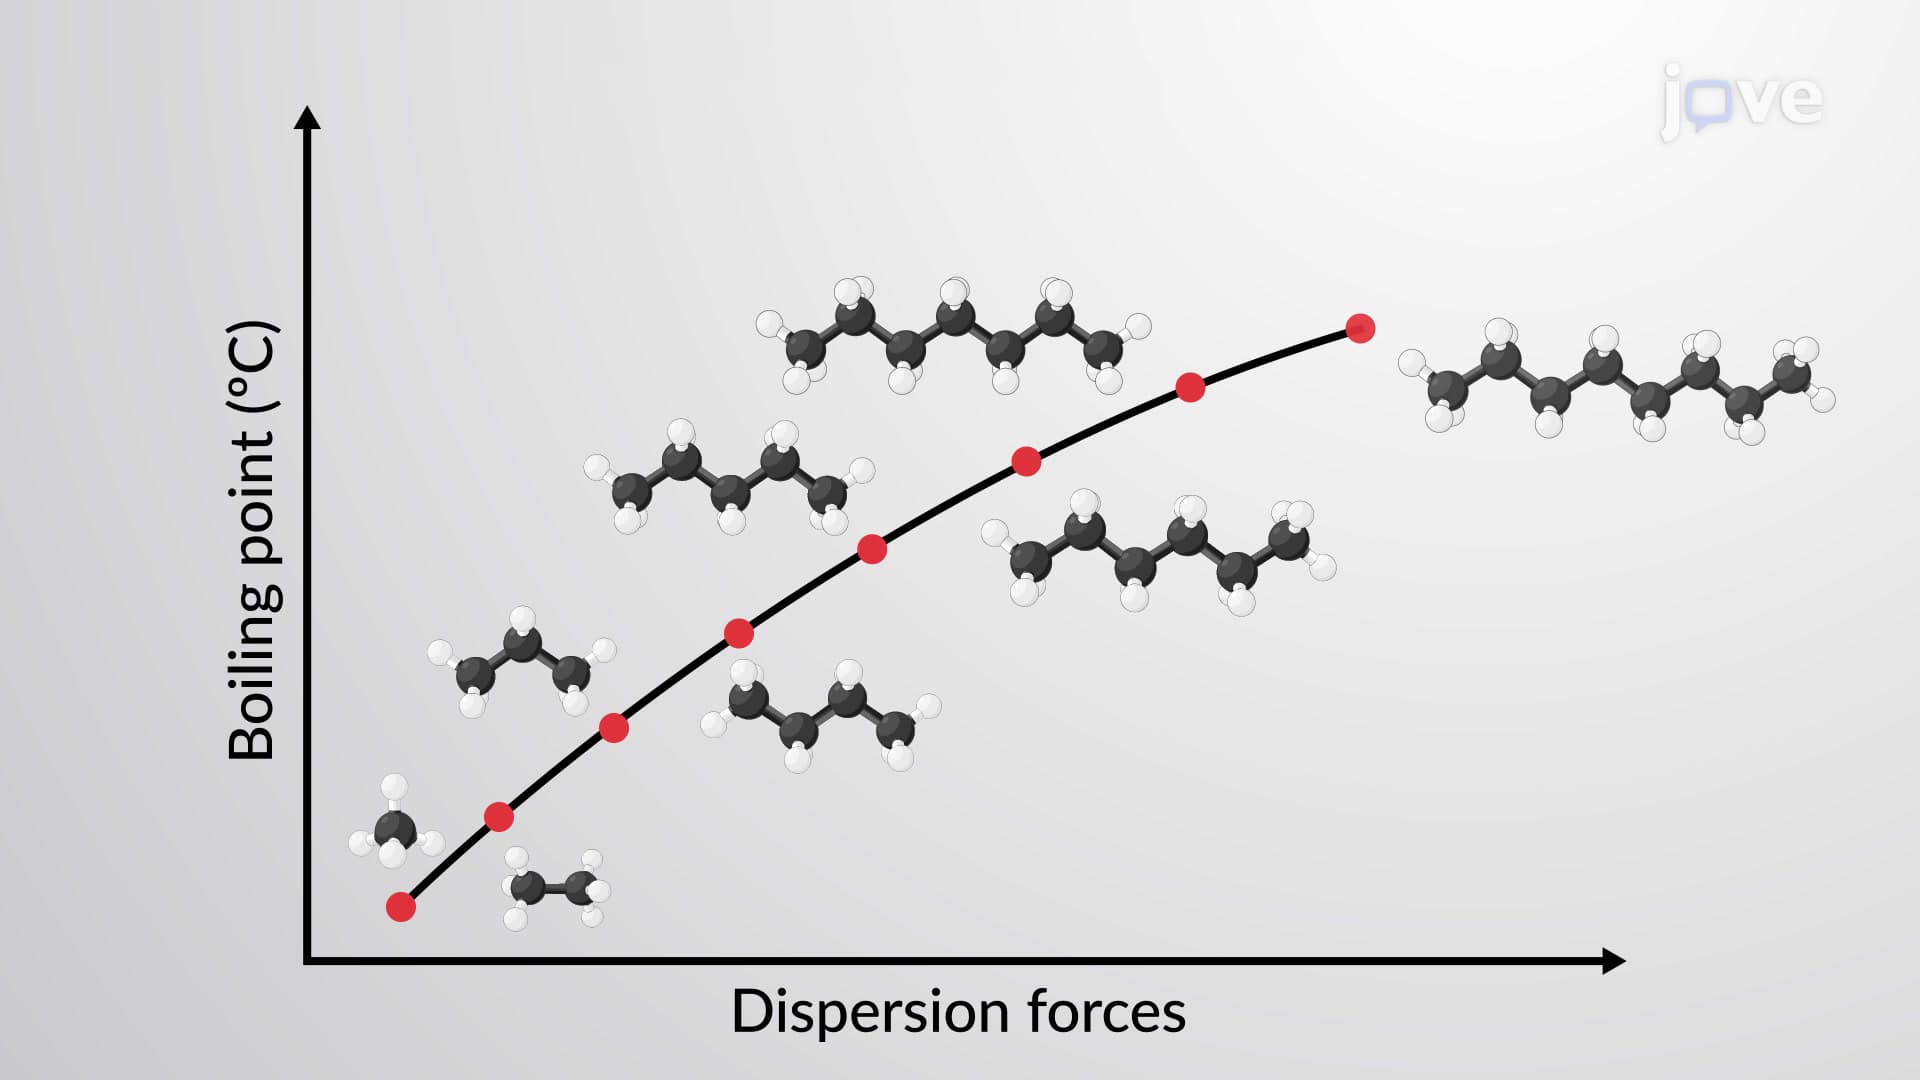
Physical Properties of Alkanes