Related Experiment Video
Updated: May 24, 2026

Construction of Model Lipid Membranes Incorporating G-protein Coupled Receptors (GPCRs)
Published on: February 5, 2022
Evidence for modulatory sites at the lipid-protein interface of the human multidrug transporter P-glycoprotein
Debjani Mandal1, Karobi Moitra, Debabrata Ghosh
1Department of Biochemistry, Uniformed Services University School of Medicine, Bethesda, Maryland 20814, United States.

Abstract:
The human multidrug transporter P-glycoprotein (Pgp or ABCB1) sets up pharmacological barriers to many clinically important drugs, a therapeutic remedy for which has yet to be formulated. For the rational design of mechanism-based inhibitors (or modulators), it is necessary to map the potential sites for modulator interaction and understand their modes of communication with the other functional domains of Pgp. In this study, combining directed mutagenesis with homology modeling, we provide evidence of two modulator-specific sites at the lipid protein interface of Pgp. Targeting 21 variant positions in the COOH-terminal transmembrane (TM) regions, we find residues M948 (in TM11) and F983, M986, V988, and Q990 (all four in TM12) critically involved in substrate-site modulation by a thioxanthene-based allosteric modulator cis-(Z)-flupentixol. Interestingly, for ATP-site modulation by the same modulator, only two (M948 and Q990) of those four residues appear indispensable, together with two additional residues, T837 and I864 in TM9 and TM10, respectively, suggesting independent modes of communication linking the allosteric site with the substrate binding and ATPase domains. None of the seven residues identified prove to be critical for modulation of the substrate or ATP sites by Pgp modulators that are transported by the pump, such as cyclosporin A or verapamil, indicating their specificity for cis-(Z)-flupentixol. On the other hand, ATP-site modulation by verapamil proves to be highly sensitive to replacement at positions F716 (in TM7) and I765 (in TM8), and to a more moderate extent at I764 and L772 (both in TM8). Homology modeling based on the known crystal structures of the bacterial multidrug transporter SAV1866 and the mouse Pgp homologue maps the identified residues primarily at the lipid-protein interface of Pgp, in two spatially distinct modulator-specific clusters. The two modulatory sites demonstrate negative synergism in influencing ATP hydrolysis, consolidating their spatial distinctness. Because Pgp is known to recruit drug molecules directly from the lipid bilayer, identification of modulatory sites at the lipid-protein interface and at the same time outside the conventional central drug binding cavity is mechanistically revealing.
Related Concept Videos
 01:16
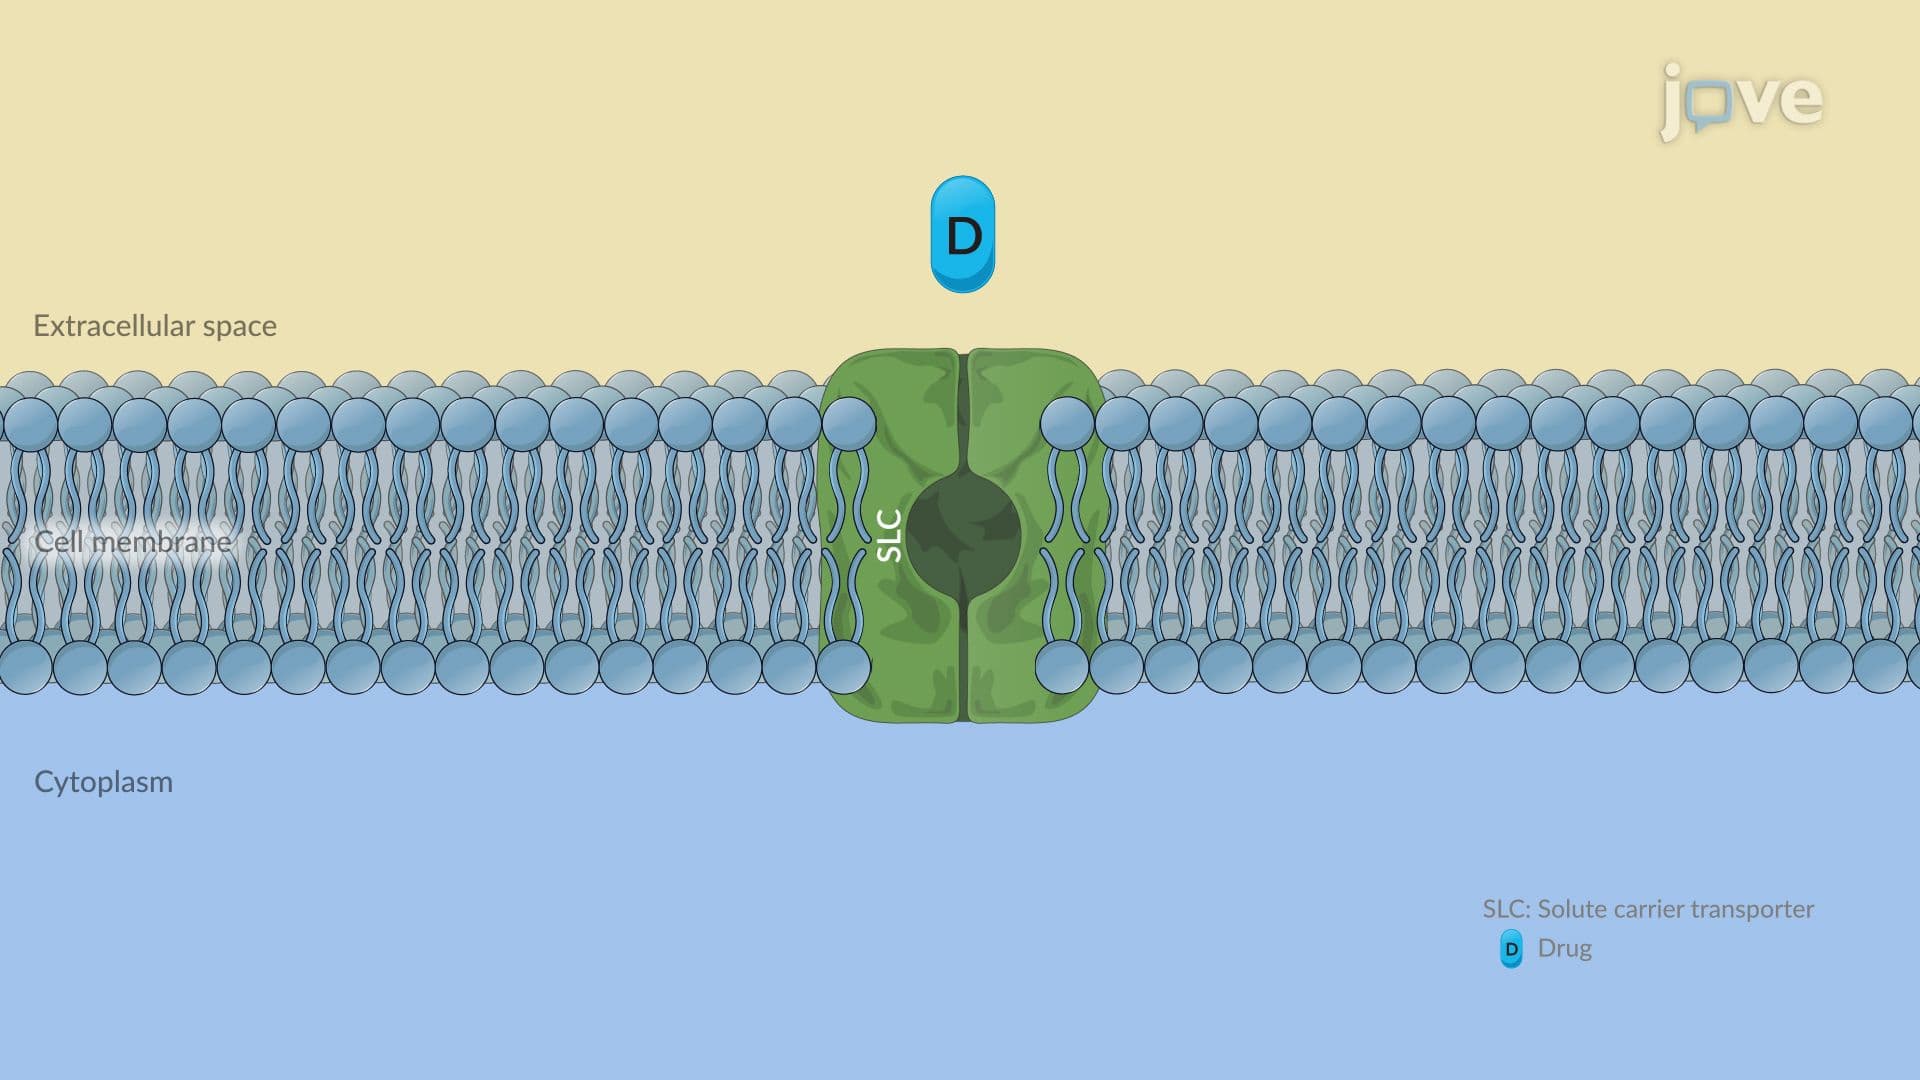
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:06
Carrier-Mediated Transport
Active transport involves two types of membrane-spanning transporters: uptake and efflux. Uptake transporters are expressed in the small...
 01:44
The Significance of Membrane Transport
Transporters facilitate either an active or passive movement of solutes. They can allow a single-molecule transport down its...
 01:24
Cellular Membranes and Drug Transport
Phospholipids arrange themselves into a bilayer, with hydrophilic heads oriented outward and hydrophobic tails facing inward.
 01:19
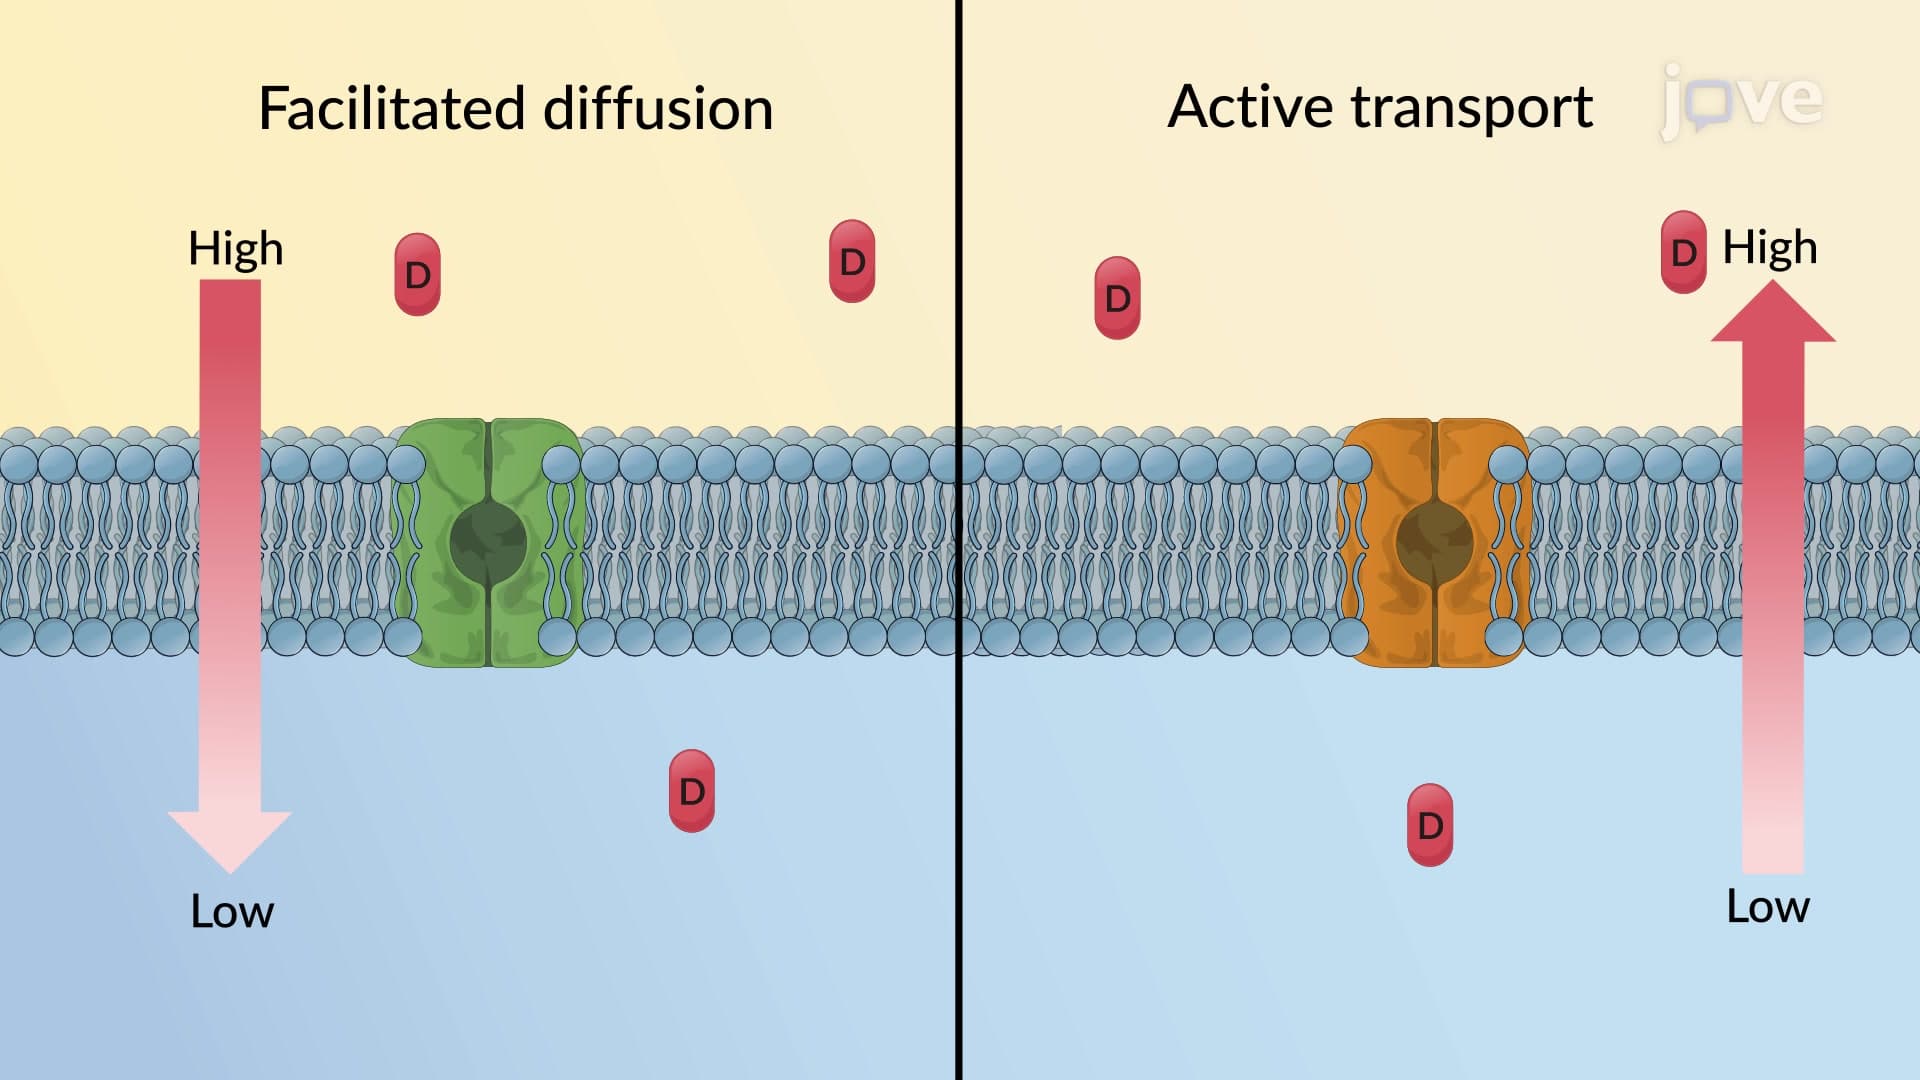
Drug Absorption Mechanism: Carrier-Mediated Membrane Transport
Facilitated diffusion is a passive process that utilizes human Solute Carrier (SLC) transporters. These transporters bind to the drug, undergo structural...
 01:17
Pore Transport and Ion-Pair Transport
Pore transport, also known as convective transport, is a process where small molecules like urea, water, and sugars rapidly cross cell membranes as though there were channels or pores in the membrane. Although direct microscopic evidence is limited but the concept of pores or channels is widely accepted based on physiological evidence. Despite the lack of direct microscopic...