Related Experiment Video
Updated: May 24, 2026
Prion Safety Laboratory Swipe Test
Published on: February 14, 2025
The contribution of different prion protein types and host polymorphisms to clinicopathological variations in
Mark W Head1, James W Ironside
1The National CJD Research & Surveillance Unit, School of Molecular & Clinical Medicine, University of Edinburgh, Edinburgh, UK. m.w.head@ed.ac.uk

Abstract:
Creutzfeldt-Jakob disease is a fatal neurodegenerative disease that primarily affects the central nervous system. In this respect, it can be considered alongside the more frequently occurring neurodegenerative diseases, such as Alzheimer's disease. Creutzfeldt-Jakob disease is perhaps the paradigmatic protein misfolding disorder, so comparisons between the mechanisms involved in Creutzfeldt-Jakob disease and other neurodegenerative diseases associated with protein misfolding (such as the tauopathies and synucleinopathies) may also be informative. Like many of these diseases, Creutzfeldt-Jakob disease occurs sporadically or can, more rarely, be associated with mutations. However, Creutzfeldt-Jakob disease can also be acquired and is experimentally transmissible. These properties have had profound public health implications and made the disease of interest to virologists, in addition to those interested in protein misfolding disorders and neurodegeneration. The possible causes for the pronounced phenotypic variation among different forms of Creutzfeldt-Jakob disease are beginning to become understood, and these appear to depend in large measure on the genetics of the host (specifically the sequence of the prion protein gene, PRNP) and the epigenetic aspects of the agent (thought to be a misfolded and aggregated form of the PRNP gene product, termed a prion). This review will examine whether this model in its present form has sufficient complexity and subtlety to account for the clinicopathological variation evident in Creutzfeldt-Jakob disease and will outline the ways in which a more complete and informative molecular definition of human prions are currently being sought.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
Parkinson Disease ll: Pathophysiology
 01:27
Principles of Pharmacogenetics: Types of Genetic Variants
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 01:23
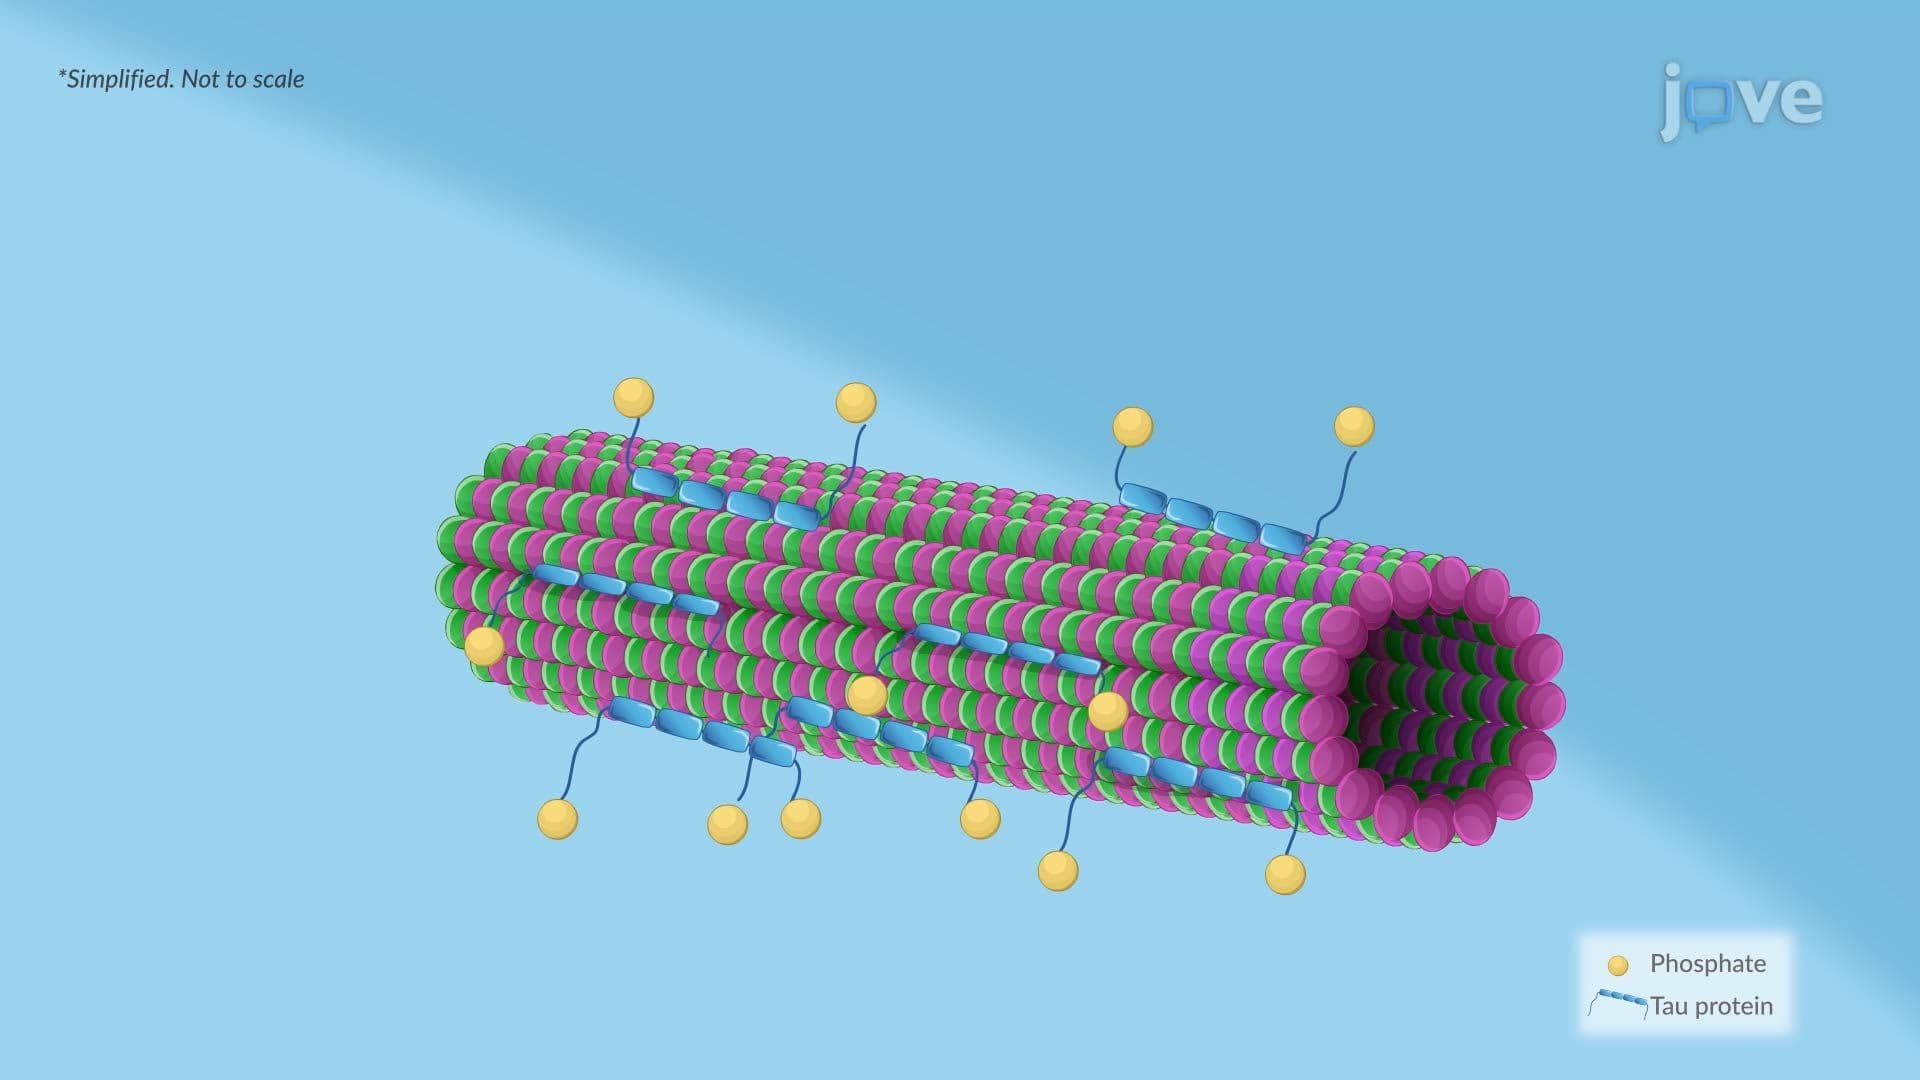
Alzheimer Disease ll: Pathophysiology
 01:05
Single Nucleotide Polymorphisms-SNPs