The spinocerebellar ataxias: clinical aspects and molecular genetics

Antoni Matilla-Dueñas1, Marc Corral-Juan, Victor Volpini

  • 1Department of Neurosciences, Universitat Autònoma de Barcelona, Barcelona, Spain. amatilla@igtp.cat

Related Concept Videos

Animal Mitochondrial Genetics02:59

Animal Mitochondrial Genetics

Among all the organelles in an animal cell, only mitochondria have their own independent genomes. Animal mitochondrial DNA is a double-stranded, closed-circular molecule with around 20,000 base pairs. Mitochondrial DNA is unique in that one of its two strands, the heavy, or H, -strand is guanine rich, whereas the complementary strand is cytosine rich and called the light, or L, -strand. Compared to nuclear DNA, mitochondrial DNA has a very low percentage of non-coding regions and is marked by...
Parkinson Disease ll: Pathophysiology01:24

Parkinson Disease ll: Pathophysiology

Parkinson disease (PD) is a progressive neurodegenerative disorder primarily affecting movement, with additional non-motor features. Its pathophysiology involves complex interactions among genetic susceptibility, environmental exposures, and cellular dysfunction, including dopaminergic neuron loss, protein aggregation, and mitochondrial impairment.Selective NeurodegenerationA key feature is the degeneration of dopaminergic neurons in the substantia nigra pars compacta, leading to reduced...
Principles of Pharmacogenetics: Types of Genetic Variants01:27

Principles of Pharmacogenetics: Types of Genetic Variants

The human genome is over 99.9% identical between individuals, yet genetic differences exist at millions of bases. The human genome contains approximately 3 million variant positions per individual, many of which are heterozygous, contributing to genetic diversity and individual traits. Genetic variations include single-nucleotide polymorphisms (SNPs), insertions, deletions, and copy number variations (CNVs).SNPs, the most common variation, involve single-base changes in DNA. These can be...
Huntington Disease l: Introduction01:21

Huntington Disease l: Introduction
Huntington disease or HD is a progressive, fatal neurodegenerative disorder inherited in an autosomal dominant pattern.PathophysiologyIt is caused by expansion of the CAG trinucleotide repeat in the HTT gene on chromosome 4 (4p16.3), producing an abnormal huntingtin protein with an expanded polyglutamine tract. This misfolded protein disrupts cellular function, leading to neuronal death. Normal alleles have ≤26 repeats, 27–35 are intermediate (risk of expansion), 36–39 show reduced penetrance,...
Alzheimer Disease ll: Pathophysiology01:23

Alzheimer Disease ll: Pathophysiology

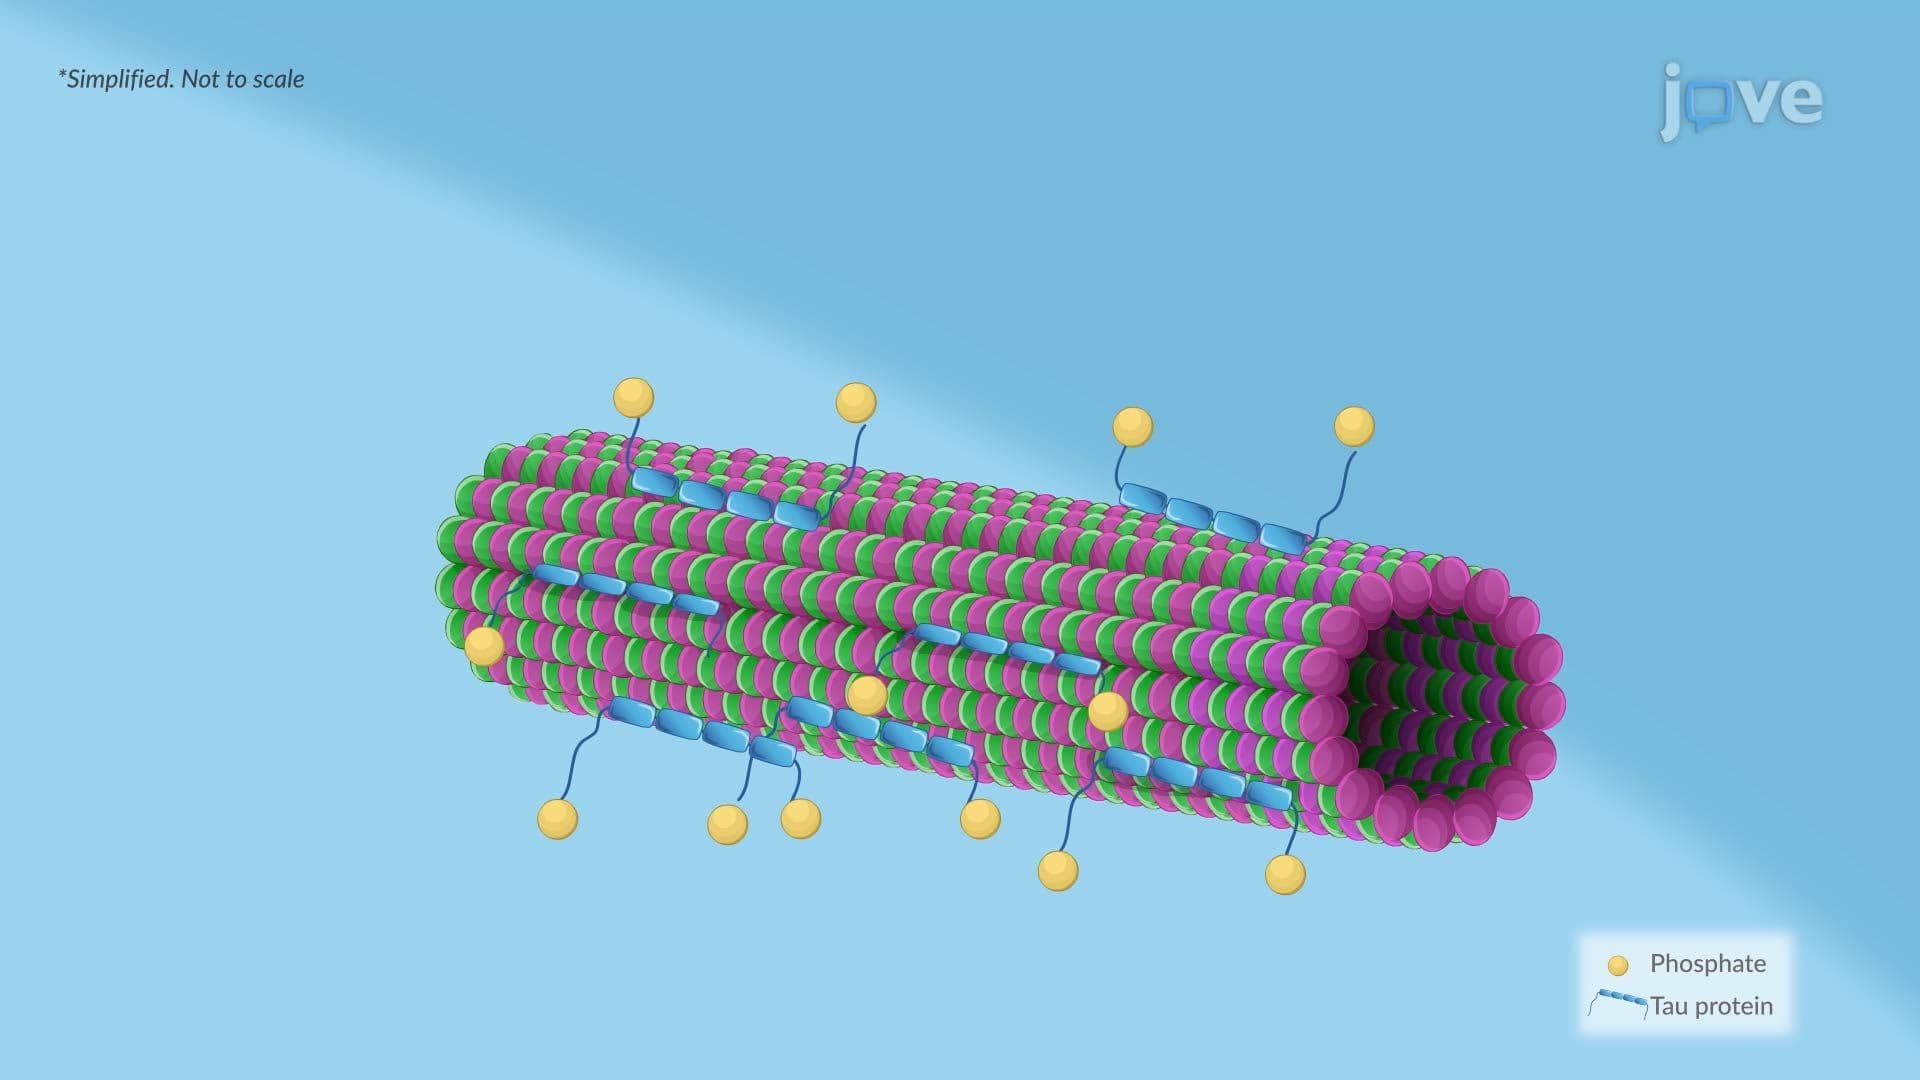
Alzheimer disease involves structural changes in the brain that begin long before symptoms appear. The most distinctive features are extracellular neuritic plaques and intracellular neurofibrillary tangles.Neuritic plaques form in the cerebral cortex and around blood vessels. These plaques contain a dense core of beta-amyloid (Aβ)—a toxic protein fragment that clumps outside neurons. The core is surrounded by damaged neuronal extensions, as well as reactive astrocytes and microglia. Abnormal...
Sex-linked Disorders01:43

Sex-linked Disorders

Like autosomes, sex chromosomes contain a variety of genes necessary for normal body function. When a mutation in one of these genes results in biological deficits, the disorder is considered sex-linked.