Analysis of structure-based virtual screening studies and characterization of identified active compounds
Related Experiment Video
Updated: May 23, 2026
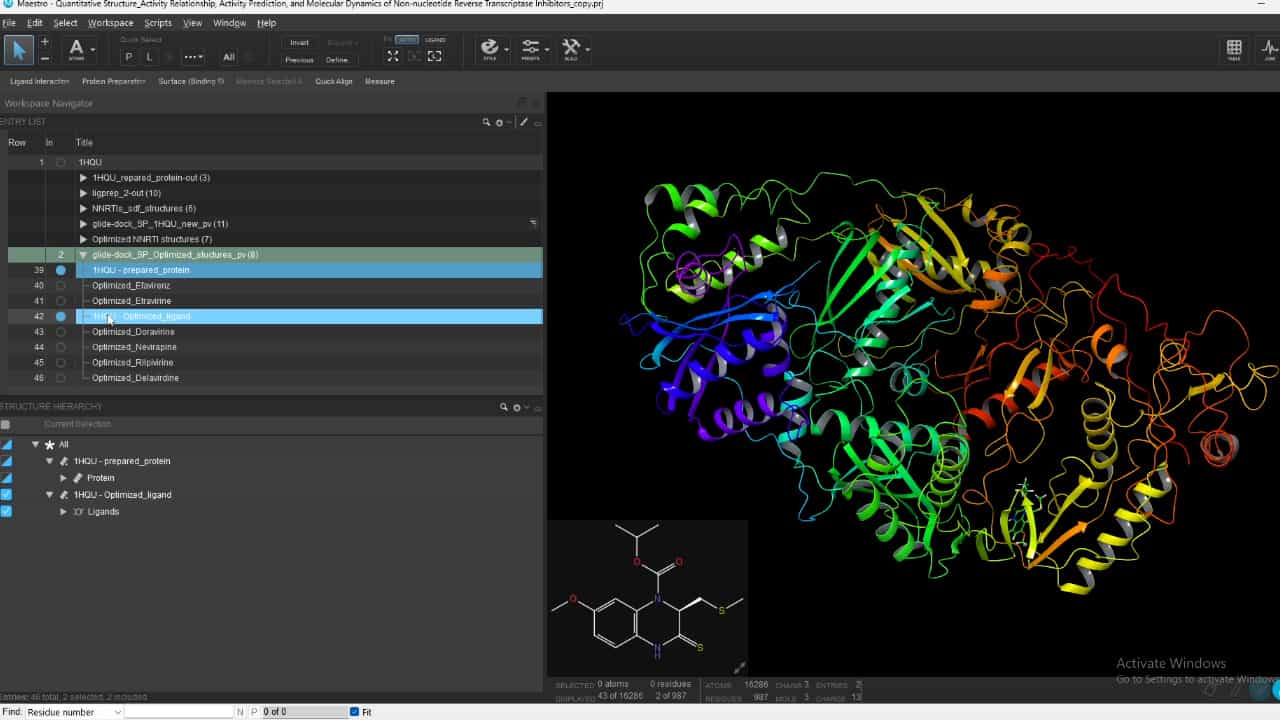
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Peter Ripphausen1, Dagmar Stumpfe, Jürgen Bajorath
1Department of Life Science Informatics, B-IT, LIMES Program Unit Chemical Biology and Medicinal Chemistry, Rheinische Friedrich-Wilhelms-Universität, Dahlmannstr. 2, D-53113 Bonn, Germany.

Abstract:
Structure-based virtual screening makes explicit or implicit use of 3D target structure information to detect novel active compounds. Results of nearly 300 currently available original applications have been analyzed to characterize the state-of-the-art in this field. Compound selection from docking calculations is much influenced by subjective criteria. Although submicromolar compounds are identified, the majority of docking hits are only weakly potent. However, only a small percentage of docking hits can be reproduced by ligand-based methods. When docking calculations identify potent hits, they often originate from specialized compound sources (e.g., pharmaceutical compound decks or target-focused libraries) and also display a notable bias towards kinase targets. Structure-based virtual screening is the dominant approach to computational hit identification. Docking calculations frequently identify active compounds. Limited accuracy of compound scoring and ranking currently presents a major caveat of the approach that is often compensated for by chemical intuition and knowledge.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:26
Drug Discovery: Overview