Related Experiment Video
Updated: May 23, 2026

Tuning Oxide Properties by Oxygen Vacancy Control During Growth and Annealing
Published on: June 9, 2023
First-principles calculations of interfacial and segregation energies in α-Cr2O3
1Department of Materials Science and Engineering, Pennsylvania State University, University Park, PA 16802, USA.

Abstract:
The interfacial energies of three twin boundaries with low-index boundary planes: prismatic (101⁻0), basal O-terminated (0001), and basal Cr-terminated (0001), and the segregation energies of five doping elements (Ce, Hf, La, Y and Zr) have been calculated as a function of temperature. The static energies at 0 K were obtained through first-principles calculations and the energies at finite temperatures were derived based on the Debye model. The calculation results show that both the interfacial and segregation energies decrease as temperature increases and the segregation energies are found to be proportional to the ionic size mismatch and the interfacial energy. Our combined approaches suggest an efficient and less computationally intensive way to derive grain boundary energetics at finite temperatures.
More Related Videos
Related Concept Videos
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 01:29
Imperfections in Crystal Structure: Non-Stoichiometric Defects
 01:27
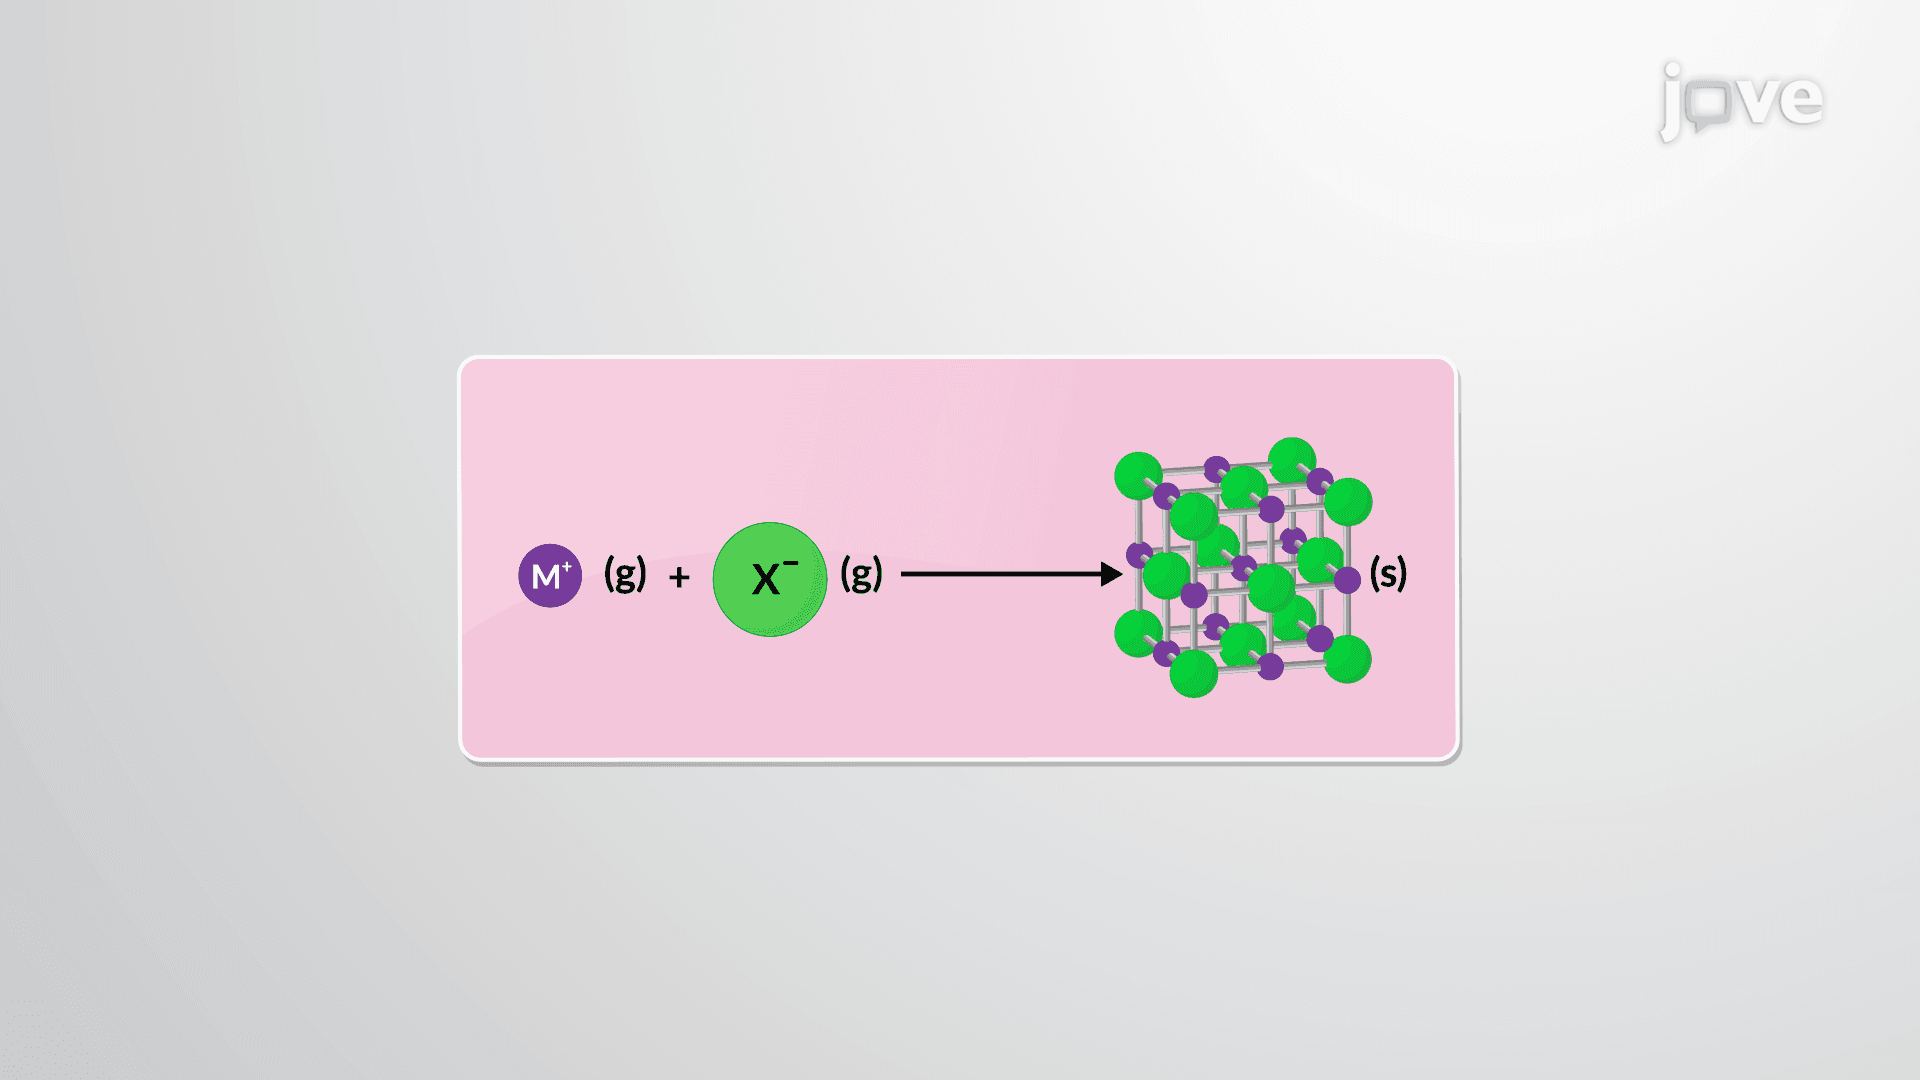
Lattice Energies of Ionic Crystals
 01:10
Recrystallization: Solid–Solution Equilibria
 02:54
Trends in Lattice Energy: Ion Size and Charge