Related Experiment Video
Updated: May 23, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A geometrical correction for the inter- and intra-molecular basis set superposition error in Hartree-Fock and density
1Theoretische Organische Chemie, Organisch-Chemisches Institut der Universität Münster, Corrensstr. 40, D-48149 Münster, Germany.

Abstract:
A semi-empirical counterpoise-type correction for basis set superposition error (BSSE) in molecular systems is presented. An atom pair-wise potential corrects for the inter- and intra-molecular BSSE in supermolecular Hartree-Fock (HF) or density functional theory (DFT) calculations. This geometrical counterpoise (gCP) denoted scheme depends only on the molecular geometry, i.e., no input from the electronic wave-function is required and hence is applicable to molecules with ten thousands of atoms. The four necessary parameters have been determined by a fit to standard Boys and Bernadi counterpoise corrections for Hobza's S66×8 set of non-covalently bound complexes (528 data points). The method's target are small basis sets (e.g., minimal, split-valence, 6-31G*), but reliable results are also obtained for larger triple-ζ sets. The intermolecular BSSE is calculated by gCP within a typical error of 10%-30% that proves sufficient in many practical applications. The approach is suggested as a quantitative correction in production work and can also be routinely applied to estimate the magnitude of the BSSE beforehand. The applicability for biomolecules as the primary target is tested for the crambin protein, where gCP removes intramolecular BSSE effectively and yields conformational energies comparable to def2-TZVP basis results. Good mutual agreement is also found with Jensen's ACP(4) scheme, estimating the intramolecular BSSE in the phenylalanine-glycine-phenylalanine tripeptide, for which also a relaxed rotational energy profile is presented. A variety of minimal and double-ζ basis sets combined with gCP and the dispersion corrections DFT-D3 and DFT-NL are successfully benchmarked on the S22 and S66 sets of non-covalent interactions. Outstanding performance with a mean absolute deviation (MAD) of 0.51 kcal/mol (0.38 kcal/mol after D3-refit) is obtained at the gCP-corrected HF-D3/(minimal basis) level for the S66 benchmark. The gCP-corrected B3LYP-D3/6-31G* model chemistry yields MAD=0.68 kcal/mol, which represents a huge improvement over plain B3LYP/6-31G* (MAD=2.3 kcal/mol). Application of gCP-corrected B97-D3 and HF-D3 on a set of large protein-ligand complexes prove the robustness of the method. Analytical gCP gradients make optimizations of large systems feasible with small basis sets, as demonstrated for the inter-ring distances of 9-helicene and most of the complexes in Hobza's S22 test set. The method is implemented in a freely available FORTRAN program obtainable from the author's website.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:14
NMR Spectrometers: Resolution and Error Correction
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:08
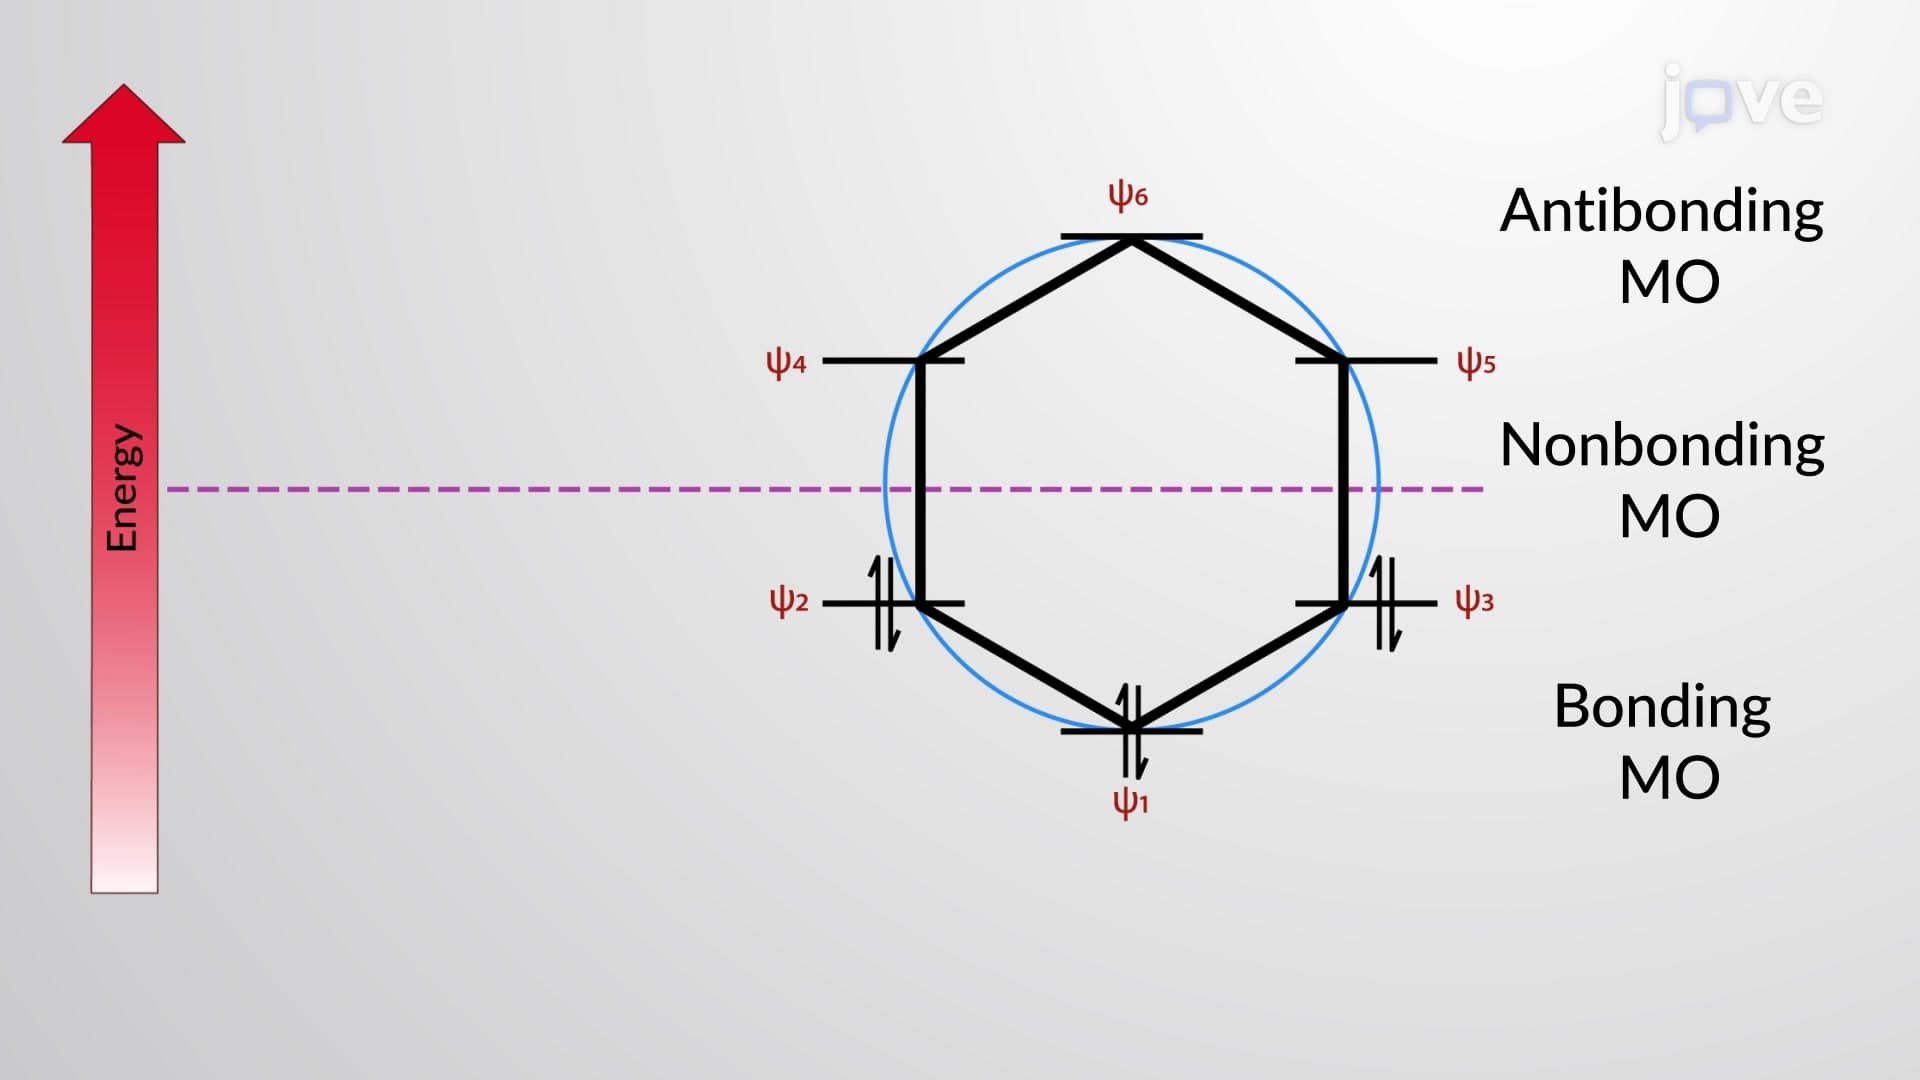
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...