Related Experiment Video
Updated: May 22, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Accurate extension of multiple sequence alignments using a phylogeny-aware graph algorithm
Ari Löytynoja1, Albert J Vilella, Nick Goldman
1EMBL-European Bioinformatics Institute, Hinxton, CB10 1SD, UK. ari.loytynoja@helsinki.fi

Motivation:
Accurate alignment of large numbers of sequences is demanding and the computational burden is further increased by downstream analyses depending on these alignments. With the abundance of sequence data, an integrative approach of adding new sequences to existing alignments without their full re-computation and maintaining the relative matching of existing sequences is an attractive option. Another current challenge is the extension of reference alignments with fragmented sequences, as those coming from next-generation metagenomics, that contain relatively little information. Widely used methods for alignment extension are based on profile representation of reference sequences. These do not incorporate and use phylogenetic information and are affected by the composition of the reference alignment and the phylogenetic positions of query sequences.
Results:
We have developed a method for phylogeny-aware alignment of partial-order sequence graphs and apply it here to the extension of alignments with new data. Our new method, called PAGAN, infers ancestral sequences for the reference alignment and adds new sequences in their phylogenetic context, either to predefined positions or by finding the best placement for sequences of unknown origin. Unlike profile-based alternatives, PAGAN considers the phylogenetic relatedness of the sequences and is not affected by inclusion of more diverged sequences in the reference set. Our analyses show that PAGAN outperforms alternative methods for alignment extension and provides superior accuracy for both DNA and protein data, the improvement being especially large for fragmented sequences. Moreover, PAGAN-generated alignments of noisy next-generation sequencing (NGS) sequences are accurate enough for the use of RNA-seq data in evolutionary analyses.
Availability:
PAGAN is written in C++, licensed under the GPL and its source code is available at http://code.google.com/p/pagan-msa.
Related Concept Videos
 02:54
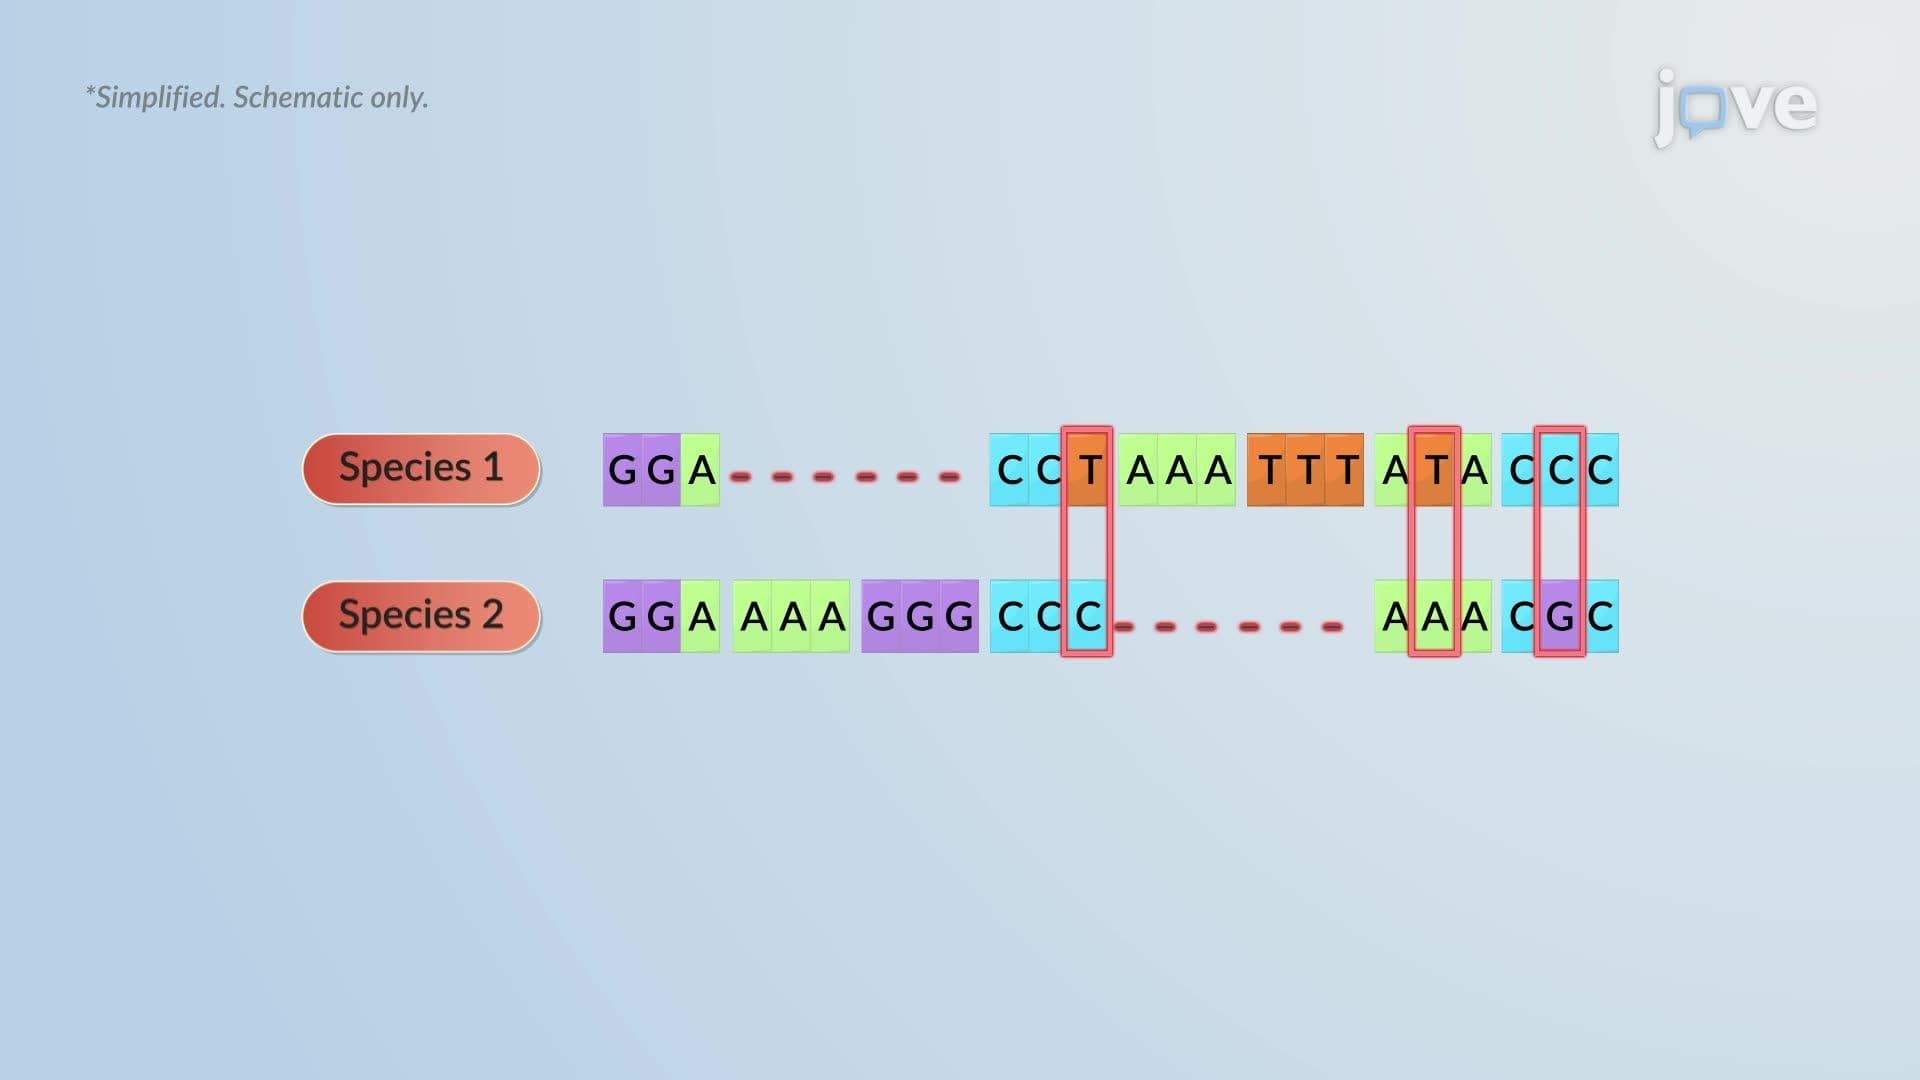
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 03:21
Phylogenetic Trees
03:21Phylogenetic Trees
 01:23
Phylogeny
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved DNA...