Related Experiment Video
Updated: May 22, 2026

Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
An accurate study of the dynamics of the C+OH reaction on the second excited 1(4)A" potential energy surface
A Zanchet1, T González-Lezana, O Roncero
1Instituto de Física Fundamental (CSIC), Serrano 123, 28006 Madrid, Spain.

Abstract:
The dynamics of the C((3)P)+OH(X(2)Π) → CO(a(3)Π)+H((2)S) on its second excited potential energy surface, 1(4)A", have been investigated in detail by means of an accurate quantum mechanical (QM) time-dependent wave packet (TDWP) approach. Reaction probabilities for values of the total angular momentum J up to 50 are calculated and integral cross sections for a collision energy range which extends up to 0.1 eV are shown. The comparison with quasi-classical trajectory (QCT) and statistical methods reveals the important role played by the double well structure existing in the potential energy surface. The TDWP differential cross sections exhibit a forward-backward symmetry which could be interpreted as indicative of a complex-forming mechanism governing the dynamics of the process. The QM statistical method employed in this study, however, is not capable to reproduce the main features of the possible insertion nature in the reactive collision. The ability to stop individual trajectories selectively at specific locations inside the potential energy surface makes the QCT version of the statistical approach a better option to understand the overall dynamics of the process.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:22
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Related Concept Videos
 02:45
E2 Reaction: Kinetics and Mechanism
 01:26
Reaction Mechanisms: The Steady-State Approximation
 02:46
E1 Reaction: Kinetics and Mechanism
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement
 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:18
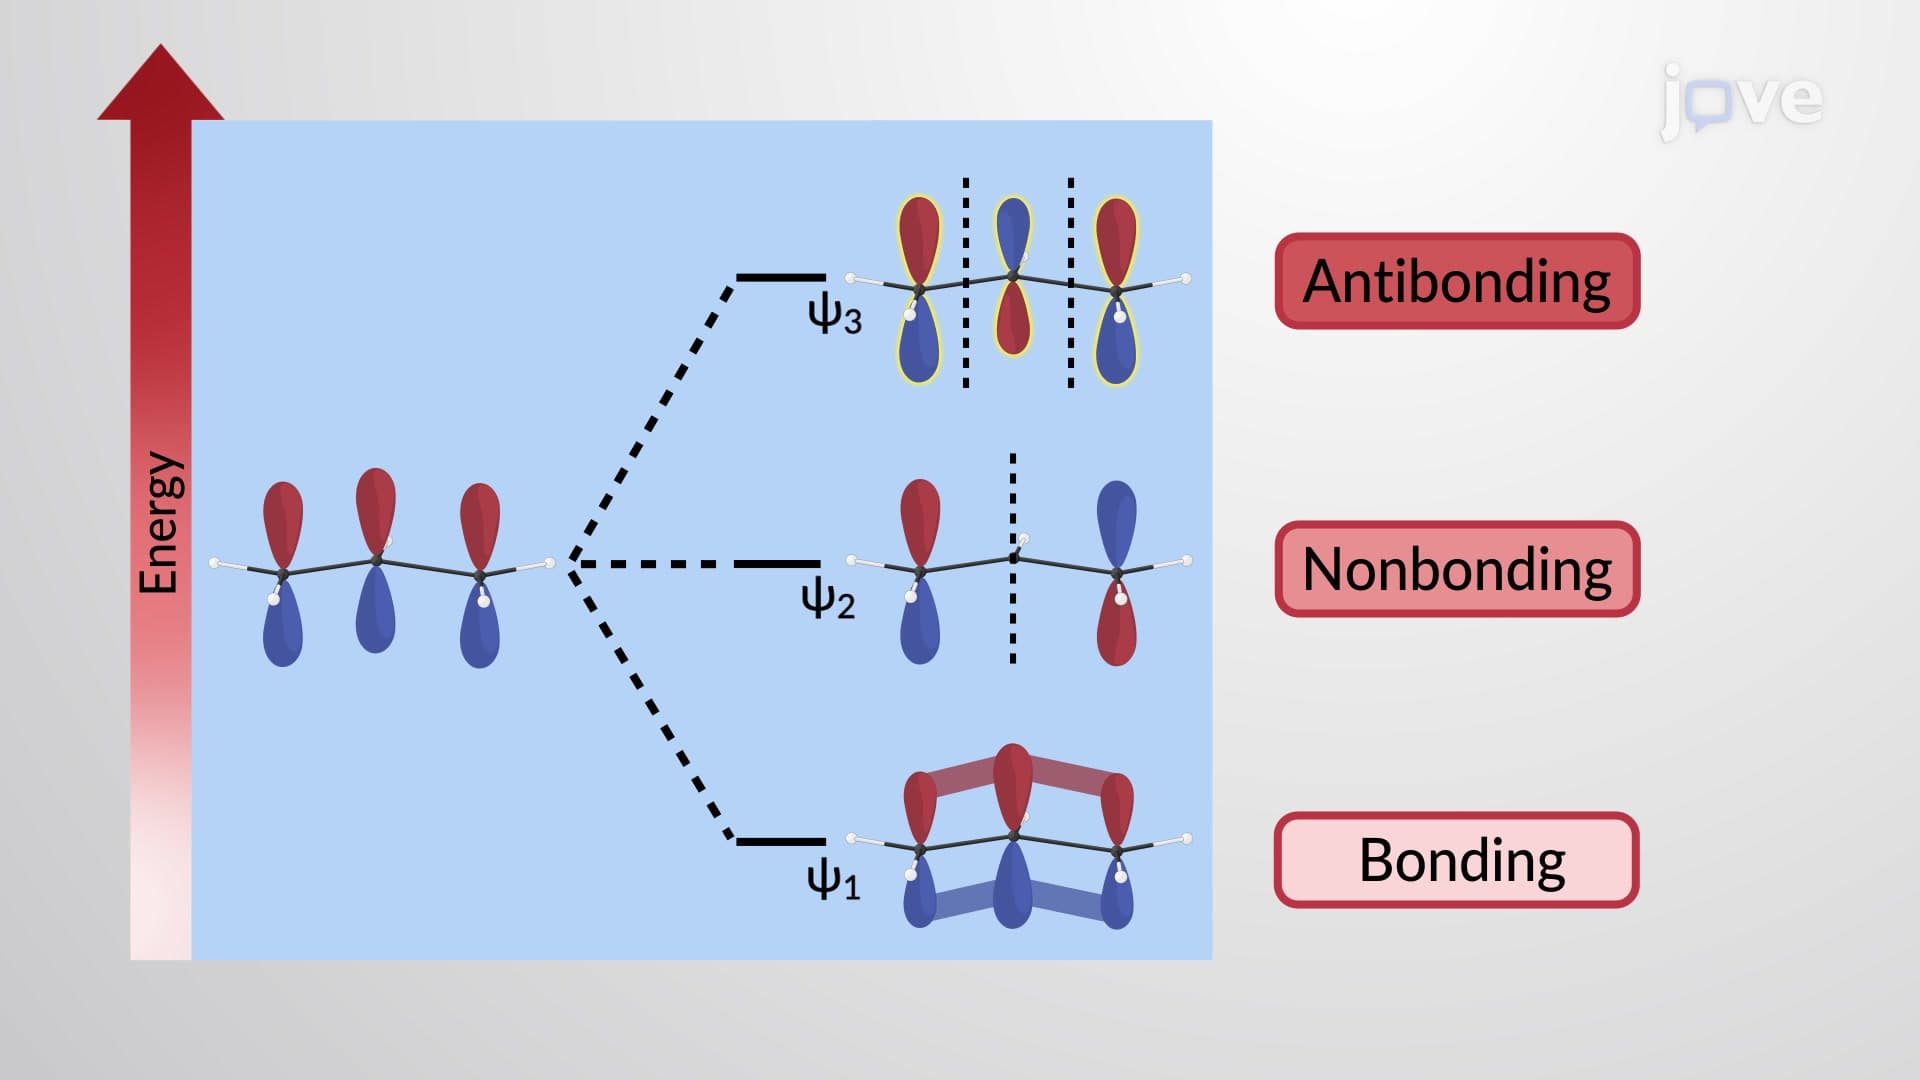
π Molecular Orbitals of the Allyl Cation and Anion