Comment on "A minimal implementation of the AMBER-GAUSSIAN interface for ab initio QM/MM-MD simulation"
Related Experiment Video
Updated: May 22, 2026
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Benjamin P Roberts1, Gustavo M Seabra, Adrian E Roitberg
1Quantum Theory Project, Department of Physics, University of Florida, Gainesville, Florida 32611-8435, USA.

Abstract:
We comment upon the recent critique of use of the Program for User Package Interfacing and Linking (PUPIL) system for linking AMBER and GAUSSIAN in a multiscale quantum mechanical/molecular mechanics (QM/MM) simulation (Okamoto et al., J. Comput. Chem. 2011, 32, 932). Specifically, their method for computing forces on the MM particles from the QM region via the GAUSSIAN-03 electrical field was already implemented in PUPIL version 1.3, publicly available beginning December 2009. Some other doubtful characterizations of PUPIL are discussed briefly in the context of current awareness of open-source codes more generally.
Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 03:35
Hybridization of Atomic Orbitals II
 01:25
Atomic Absorption Spectroscopy: Atomization Methods
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II
 01:19
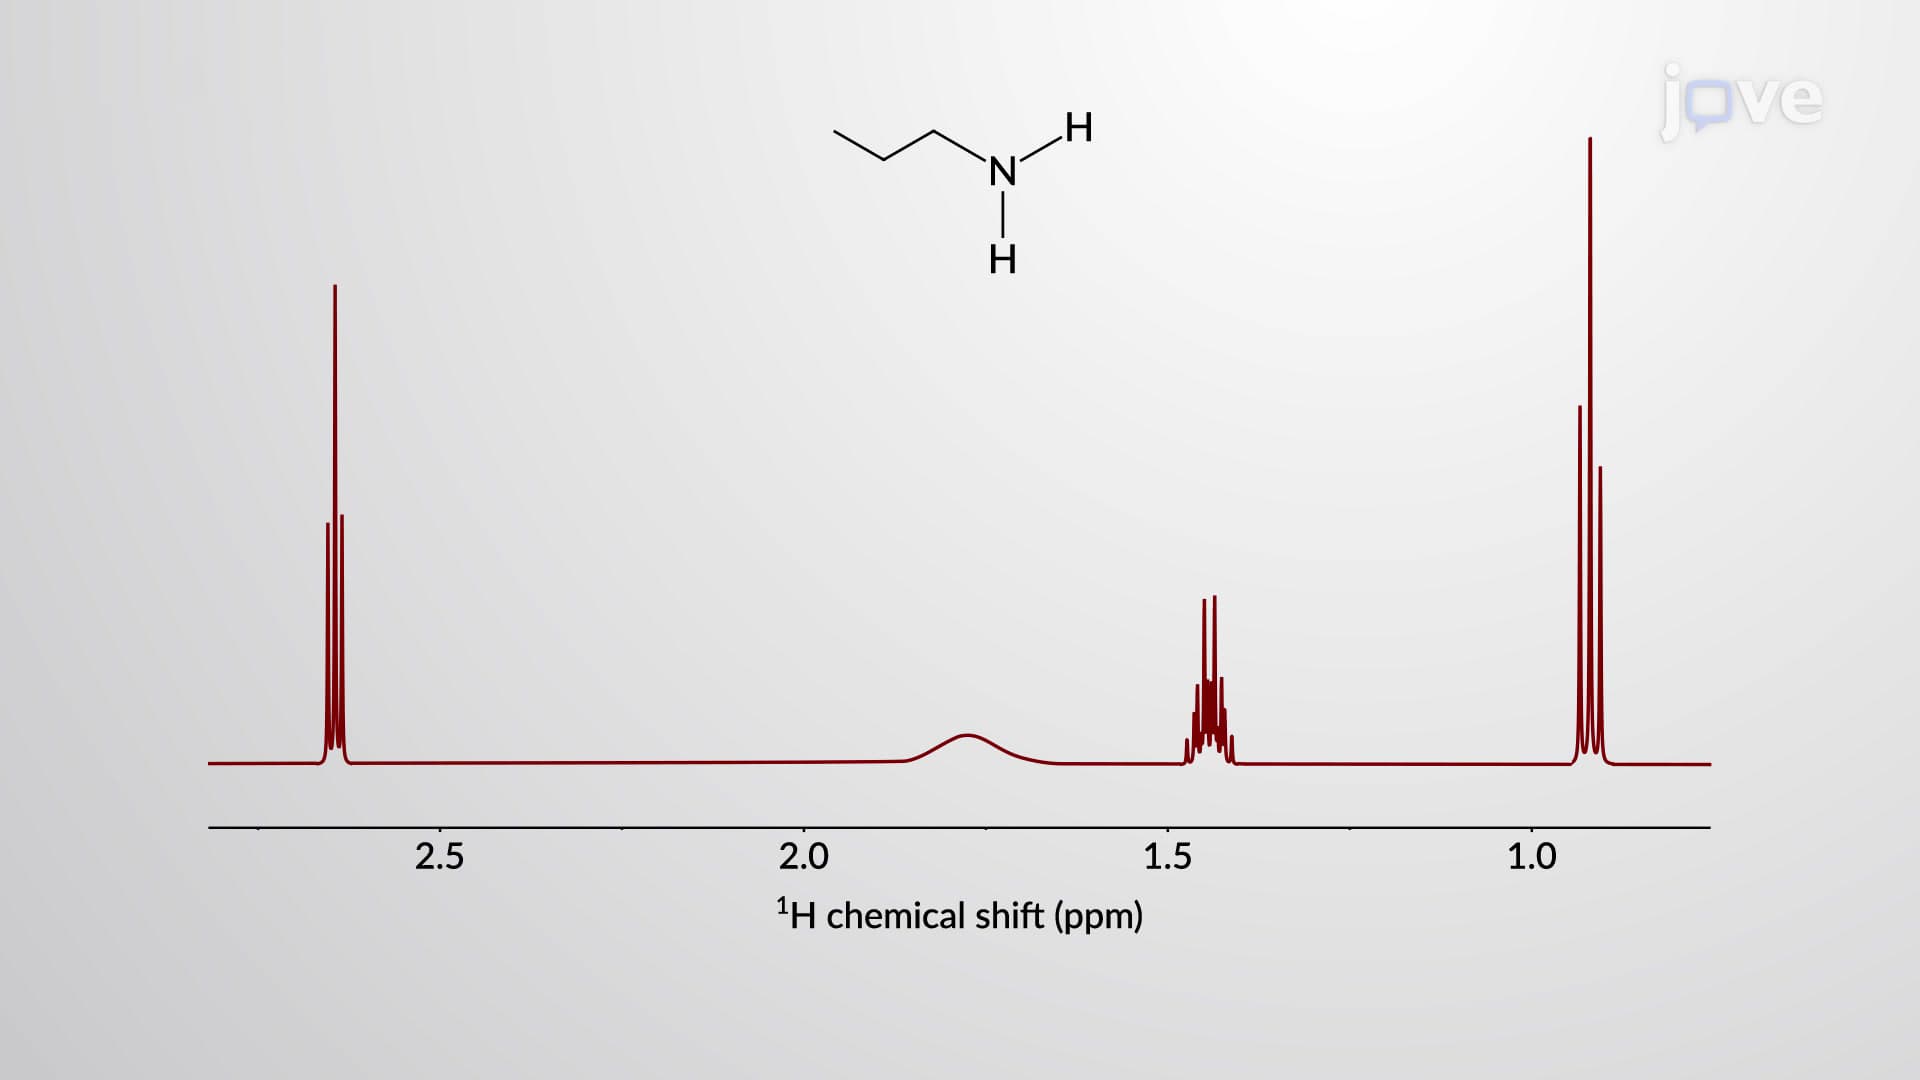
NMR Spectroscopy Of Amines