Related Experiment Video
Updated: May 22, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quasi-classical trajectory study of the HO + CO → H + CO2 reaction on a new ab initio based potential energy surface
Jun Li1, Changjian Xie, Jianyi Ma
1Department of Chemistry and Chemical Biology, University of New Mexico, Albuquerque, New Mexico 87131, USA.

Abstract:
We report extensive quasi-classical trajectory calculations of the HO + CO → H + CO(2) reaction on a newly developed potential energy surface based on a large number of UCCSD(T)-F12/AVTZ calculations. This complex-forming reaction is known for its unusual kinetics and dynamics because of its unique potential energy surface, which is dominated by the HOCO wells flanked by an entrance channel bottleneck and a transition state leading to the H + CO(2) products. It was found that the thermal rate coefficients are in reasonably good agreement with known experimental data in both low and high pressure limits. Excitation of the OH vibration is shown to enhance reactivity, due apparently to its promoting effect over the transition state between the HOCO intermediate and the H + CO(2) product. On the other hand, neither CO vibrational excitation nor rotational excitation in either CO or OH has a significant effect on reactivity, in agreement with experiment. However, significant discrepancies have been found between theory and the available molecular beam experiments. For example, the calculated translational energy distribution of the products substantially underestimates the experiment. In addition, the forward bias in the differential cross section observed in the experiment was not reproduced theoretically. While the origin of the discrepancies is still not clear, it is argued that a quantum mechanical treatment of the dynamics might be needed.


Related Concept Videos
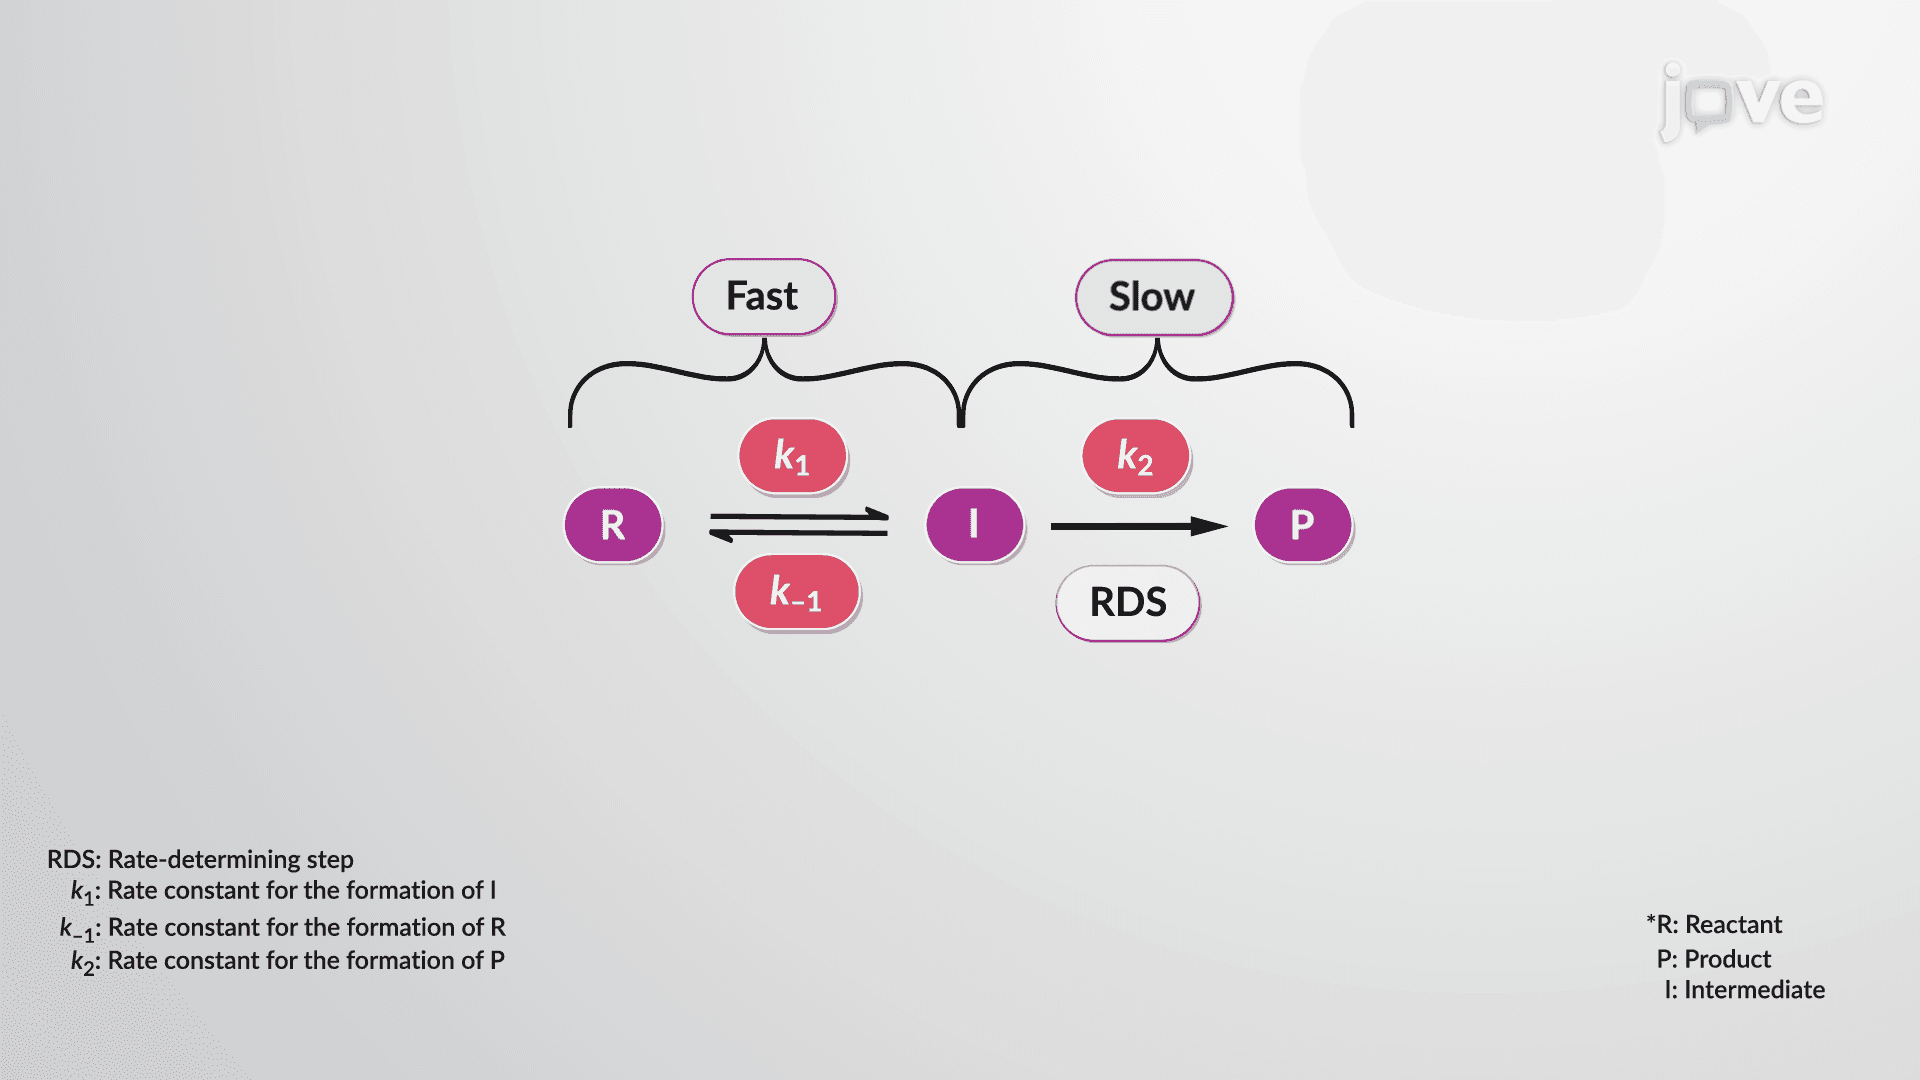 01:26
Reaction Mechanisms: The Steady-State Approximation
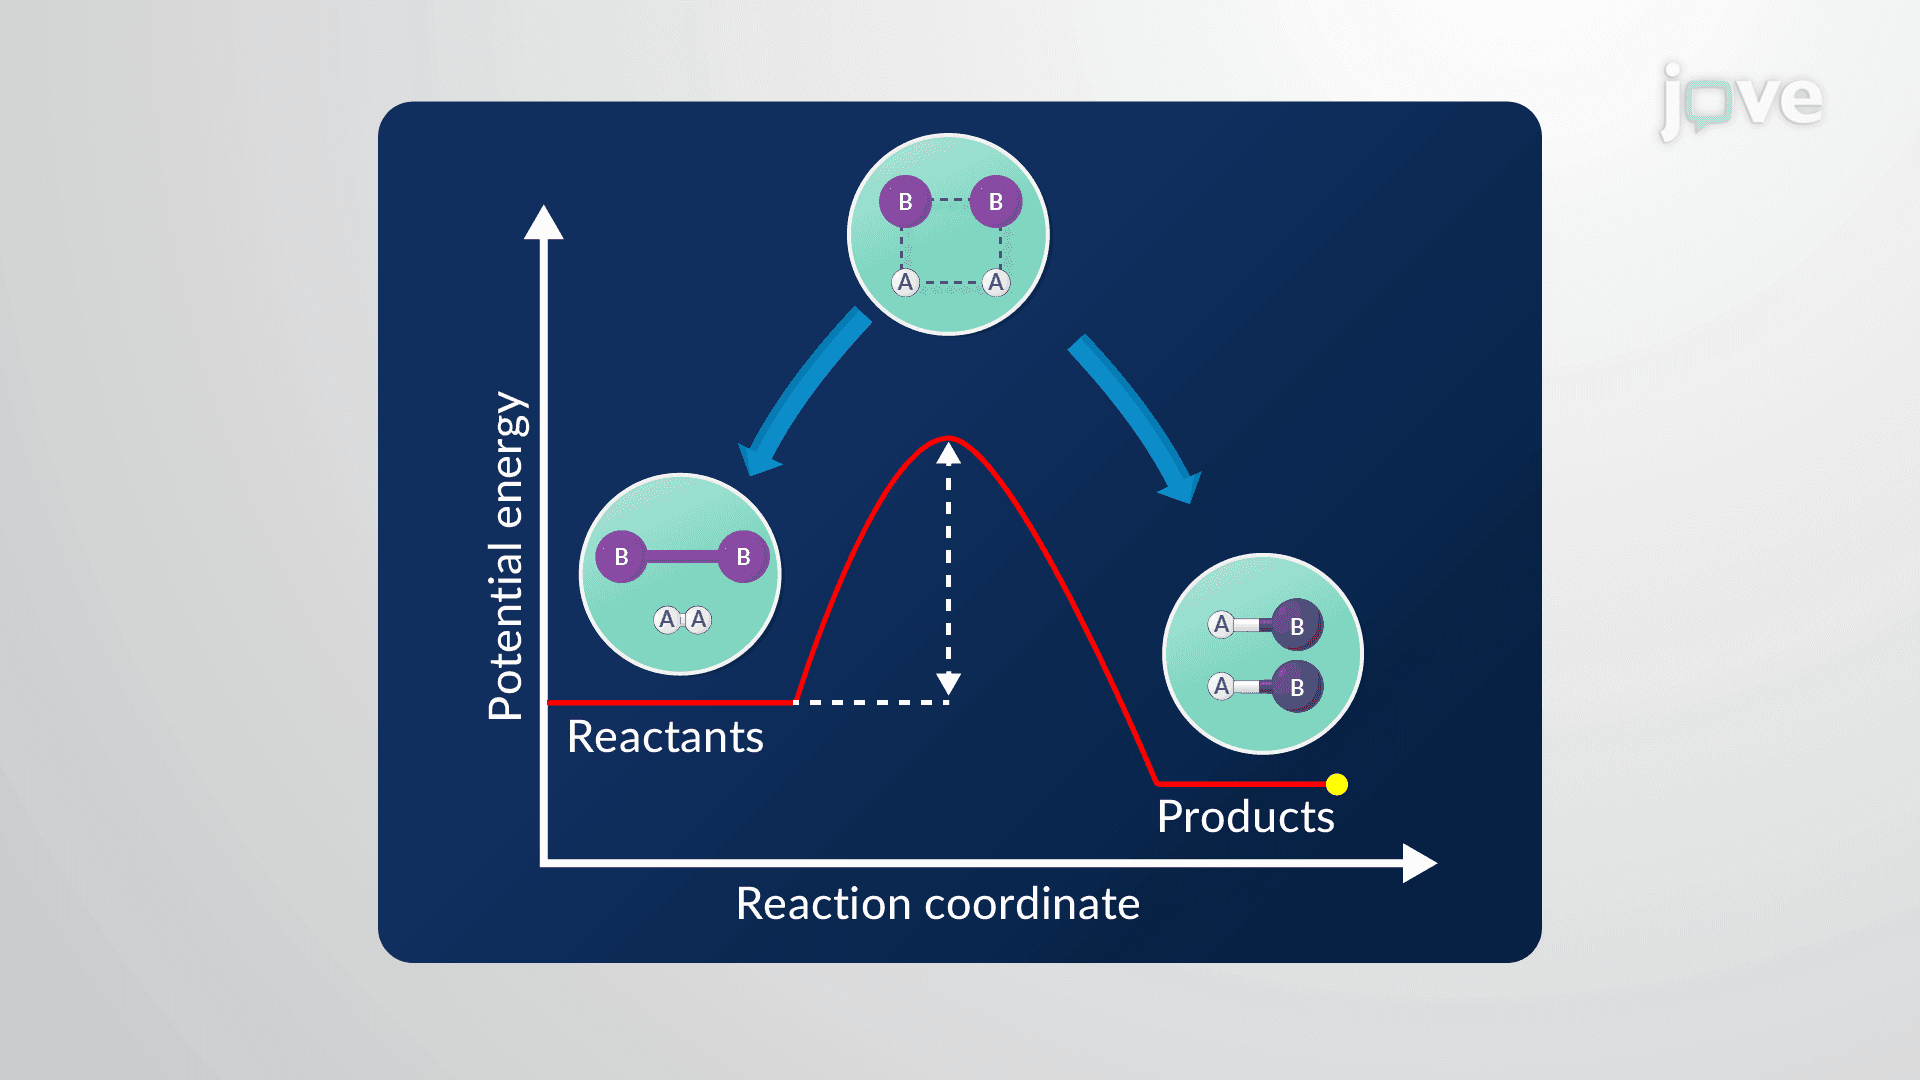 01:25
Transition State Theory
 01:17
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions. Cells...
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:45
E2 Reaction: Kinetics and Mechanism
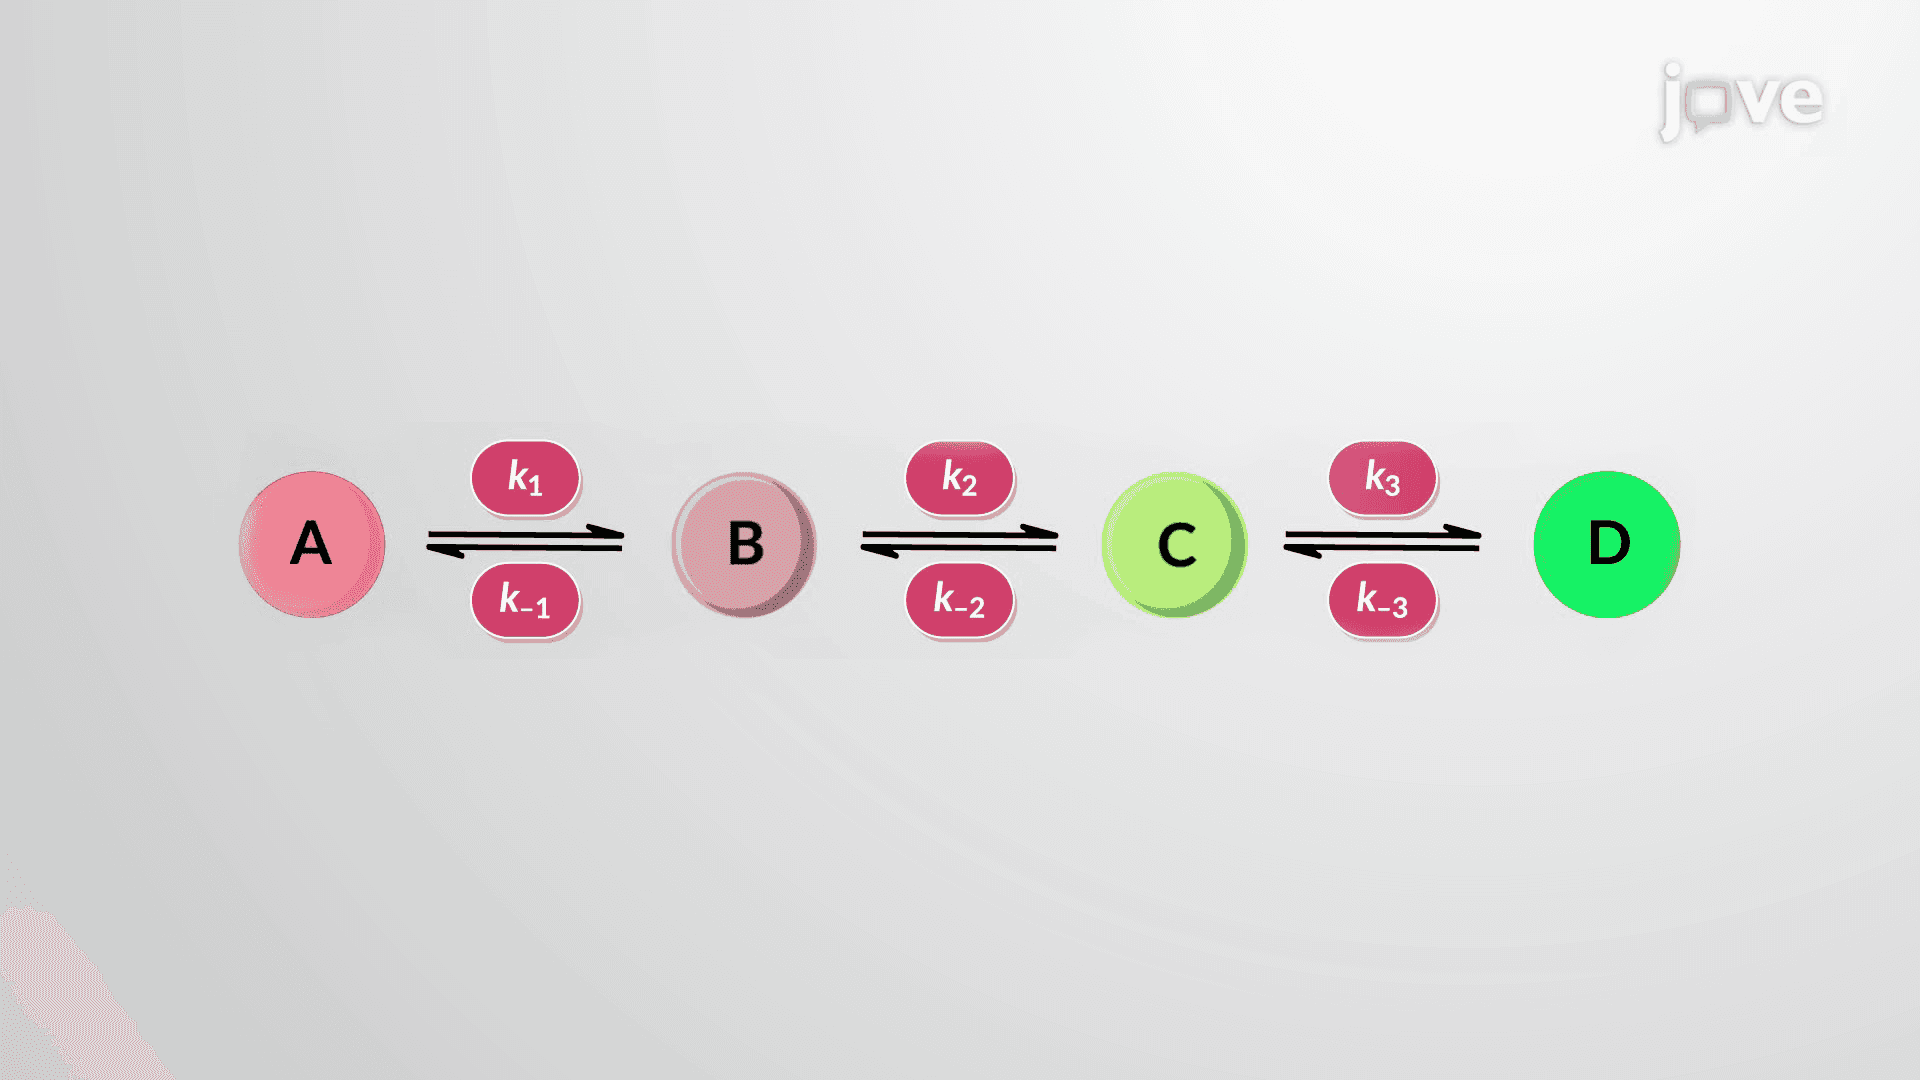 01:29
Reaction Mechanisms: Rate-limiting Step Approximation