Related Experiment Video
Updated: May 21, 2026
Synthesis and Performance Characterizations of Transition Metal Single Atom Catalyst for Electrochemical CO2 Reduction
Published on: April 10, 2018
A quantum chemical calculation on Fe(CO)5 revealing the operation of the Dewar-Chatt-Duncanson model
1Max-Planck-Institut für Bioanorganische Chemie, Stiftstrasse 34-36, Postfach 101365, D-45413 Mülheim an der Ruhr, Germany. bachler@mpi-muelheim.mpg.de

Abstract:
A nonstandard computational scheme has been applied to calculate Fe(CO)(5) with the aim to illustrate the operation of the Dewar-Chatt-Duncanson model by computation. A full configuration interaction (CI) calculation in an active space has been performed. The active space is built from naturally localized molecular orbitals (NLMOs) localized in bond regions or forming lone pairs. For selecting this active space, Weinhold's perturbation theory formulated in the natural bond orbital (NBO) space has been applied. Bonding, lone pair, and antibond NBOs exhibiting large interaction energies serve to define the active space. The actually applied active space, however, comprises NLMOs that are close in shape to the NBOs indicated by perturbation theory. Thus, a CI calculation with localized orbitals has been performed meeting the classical reasoning of chemists that is often based on local bonding concepts. The computational scheme yields the Lewis structure for Fe(CO)(5) whose energy is identical to the Hartree-Fock energy. The Lewis energy comprises CO → Fe σ-electron transfer (ET) and CO ← Fe electron back donation (BD). This Lewis energy gets lowered by localized correlation energy contributions caused by ET processes where electrons are back donated from the Fe d-lone pairs into the CO ligands. Thus, electron correlation within the selected active space is dominated by electron BD. Energies and electron populations of the NBOs support the notion that electrons are preferentially back donated into the equatorial CO ligands. Weights of local Slater determinants, determining the correlation energy, also point to a predominant BD into the equatorial CO ligands. Correlation energy increments resulting from electron BD into single antibond orbitals of the CO ligands have been calculated. These energy quantities also demonstrate that BD into the equatorial CO ligands is more energy lowering than BD into the axial CO ligands.
More Related Videos
 08:55
Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
 06:53
Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:26
The Van der Waals Equation
 01:10
Van der Waals Equation
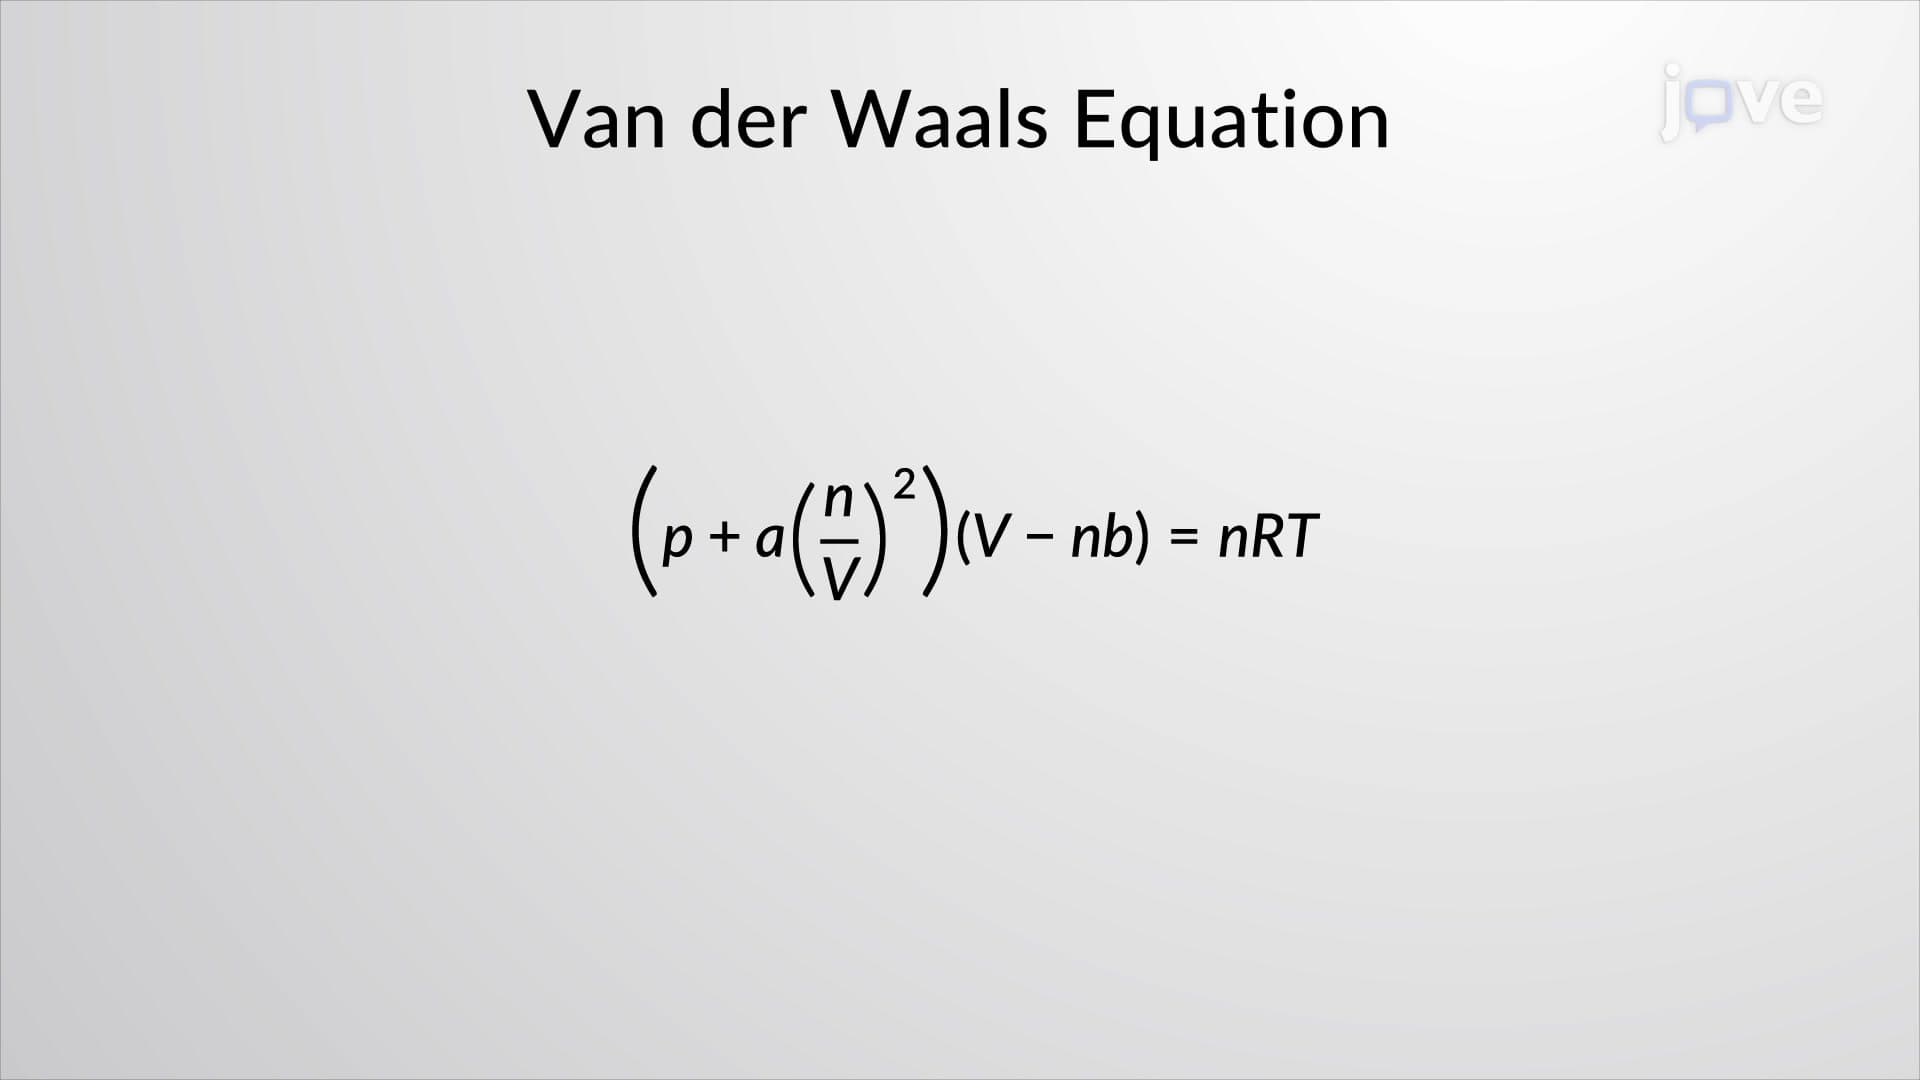
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:28
Debye–Huckel–Onsager Conductance Equation
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 02:35
Molecular Orbital Theory I