Related Experiment Video
Updated: May 21, 2026

In Vitro Aggregation Assays Using Hyperphosphorylated Tau Protein
Published on: January 2, 2015
Clinical, neuropathological, and biochemical characterization of the novel tau mutation P332S
Vincent Deramecourt1, Florence Lebert, Claude-Alain Maurage
1Univ Lille Nord de France, UDSL, Lille, France.

Abstract:
MAPT mutations cause autosomal dominant frontotemporal lobar degeneration. These diseases are characterized by considerable heterogeneity in their clinical, neuropathological, and biochemical presentations. We describe the full characterization of a family with autosomal dominant frontotemporal lobar degeneration caused by a novel MAPT mutation. Clinical, imaging, neuropathological, and biochemical data are presented. The proband was a woman who died at 85 years old, 25 years after the onset of a slowly progressive and isolated anarthria and opercular syndrome. The pathological examination of her brain showed marked atrophy of primary motor and premotor cortices, associated with predominant neuronal tau-positive lesions mimicking Pick bodies. At the biochemical level, the six tau isoforms aggregate to display a pathological triplet at 60, 64, and 69 kDa. Two of her sons presented at 48 and 50 years old with a right temporal variant of frontotemporal degeneration characterized by severe prosopagnosia, semantic impairment, and behavioral modifications. In these three patients, the molecular analysis of MAPT showed the c.1945C>T mutation on exon 11 resulting in the P332S substitution in tau sequence. This mutation changes the PGGG motif of the third repeat domain of the protein and therefore reduces the ability of tau to bind microtubule. From a clinical point of view, this mutation is associated with considerable intrafamilial phenotypic variation.
More Related Videos
Related Concept Videos
 01:23
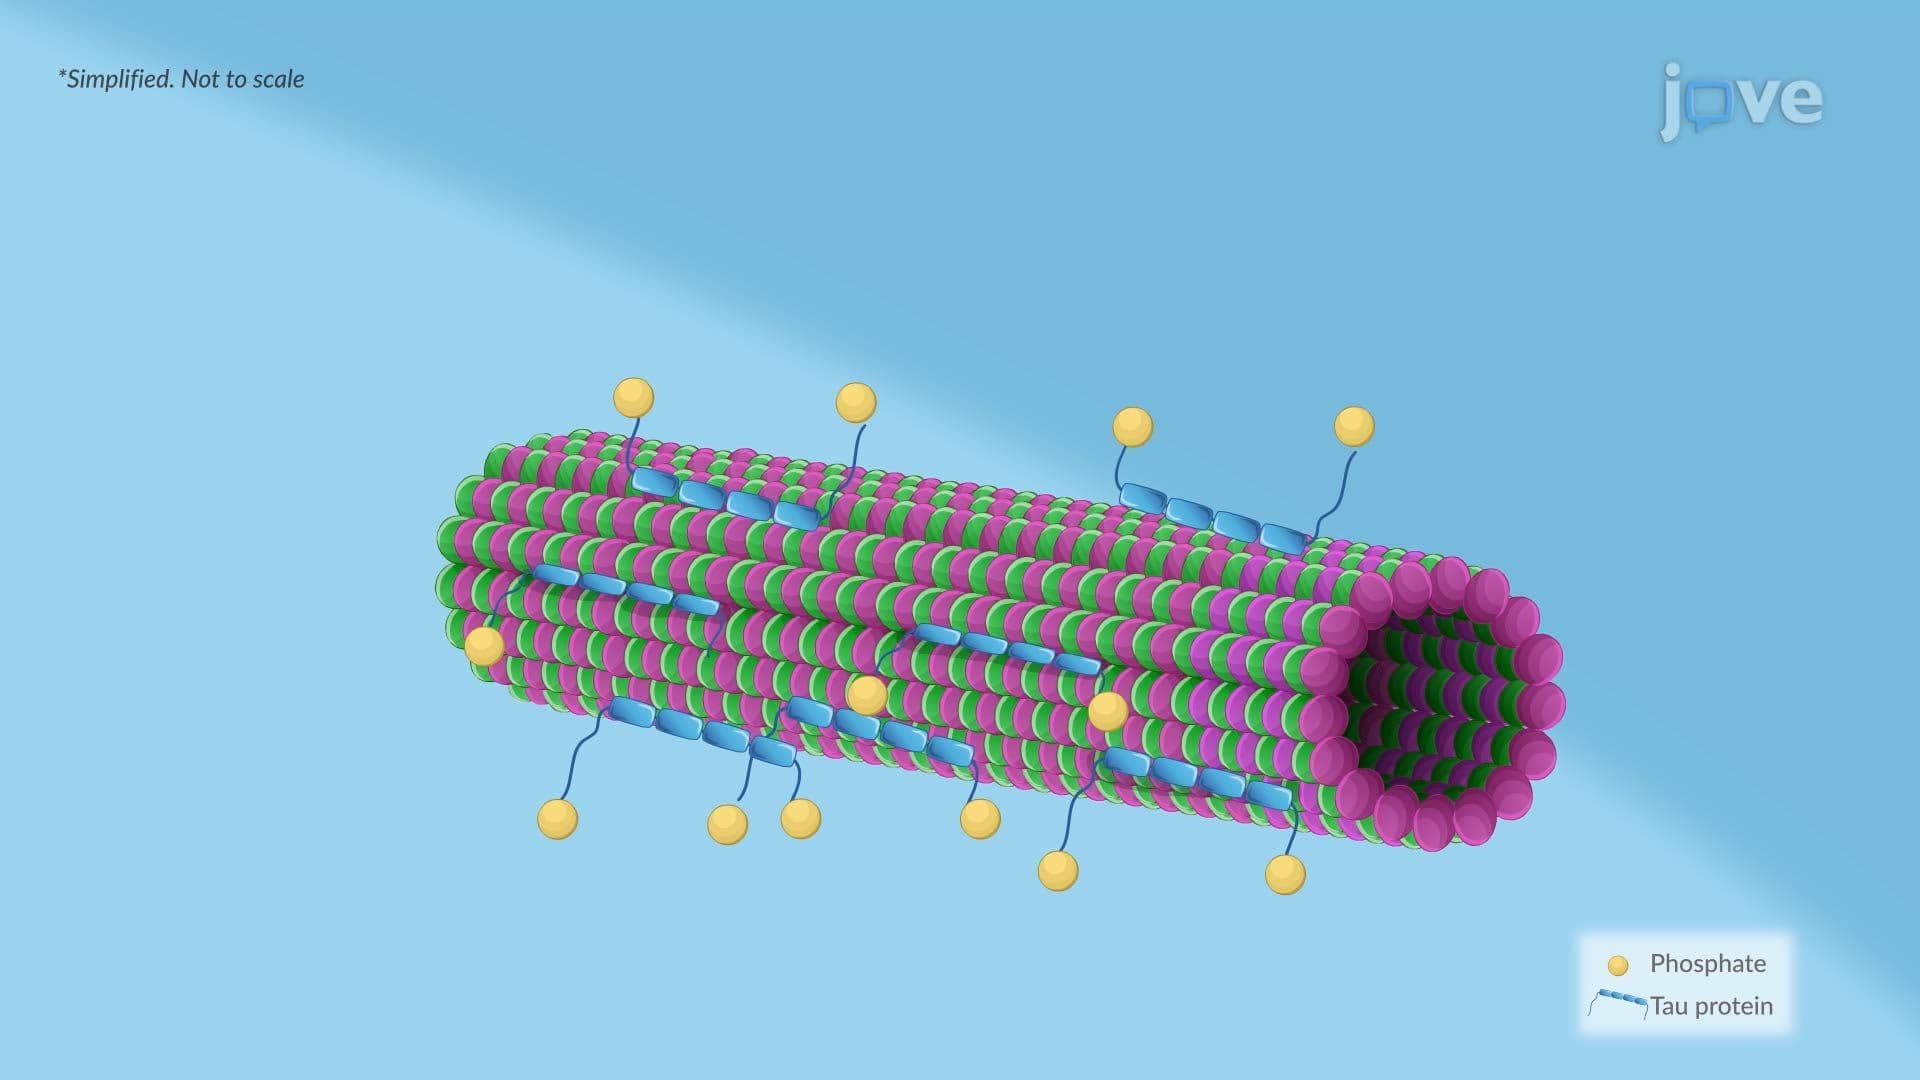
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 01:24
Parkinson Disease ll: Pathophysiology