Interfacing the GROMOS (bio)molecular simulation software to quantum-chemical program packages
Related Experiment Video
Updated: May 21, 2026
New Features in Visual Dynamics 3.0
Published on: August 9, 2024
Katharina Meier1, Nathan Schmid, Wilfred F van Gunsteren
1Laboratory of Physical Chemistry, Swiss Federal Institute of Technology Zürich ETH, 8093 Zürich, Switzerland.

Abstract:
The newly implemented quantum-chemical/molecular-mechanical (QM/MM) functionality of the Groningen molecular simulation (GROMOS) software for (bio)molecular simulation is described. The implementation scheme is based on direct coupling of the GROMOS C++ software to executables of the quantum-chemical program packages MNDO and TURBOMOLE, allowing for an independent further development of these packages. The new functions are validated for different test systems using program and model testing techniques. The effect of truncating the QM/MM electrostatic interactions at various QM/MM cutoff radii is discussed and the application of semiempirical versus density-functional Hamiltonians for a solute molecule in aqueous solution is compared.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 10:29
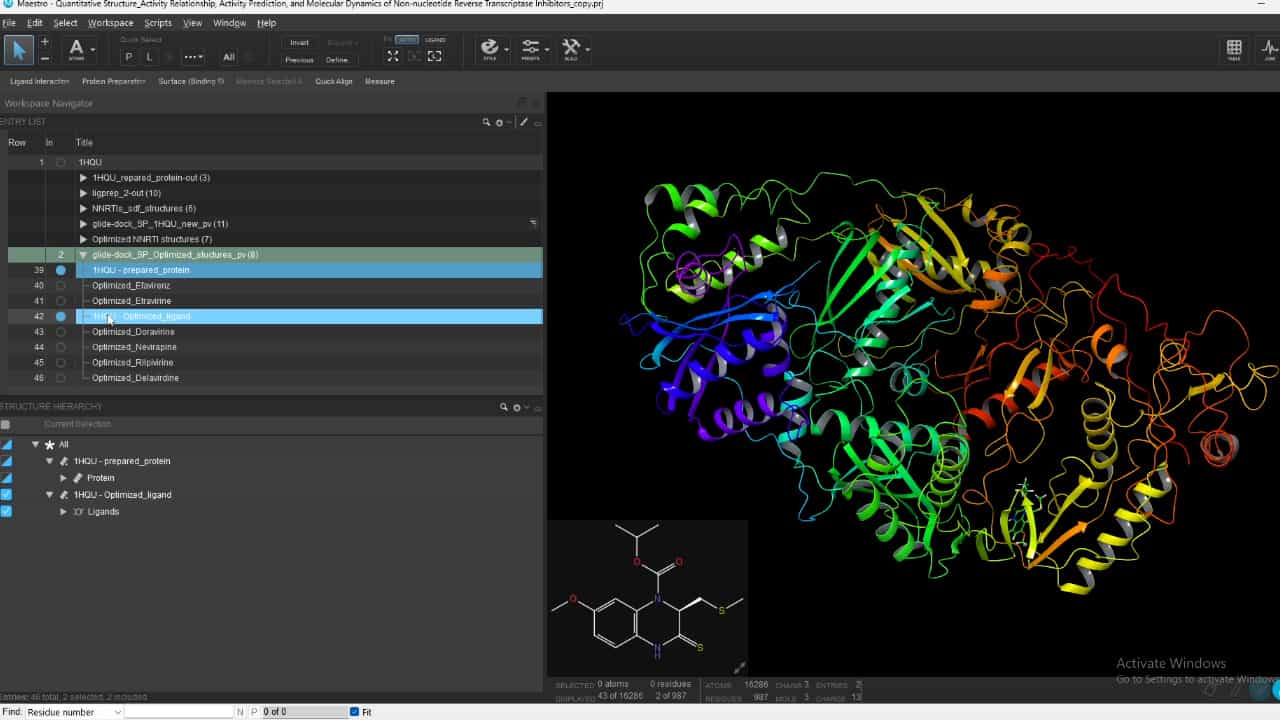
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:00
Molecular Models
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:26
Molecular Comparison of Gases, Liquids, and Solids
 01:17
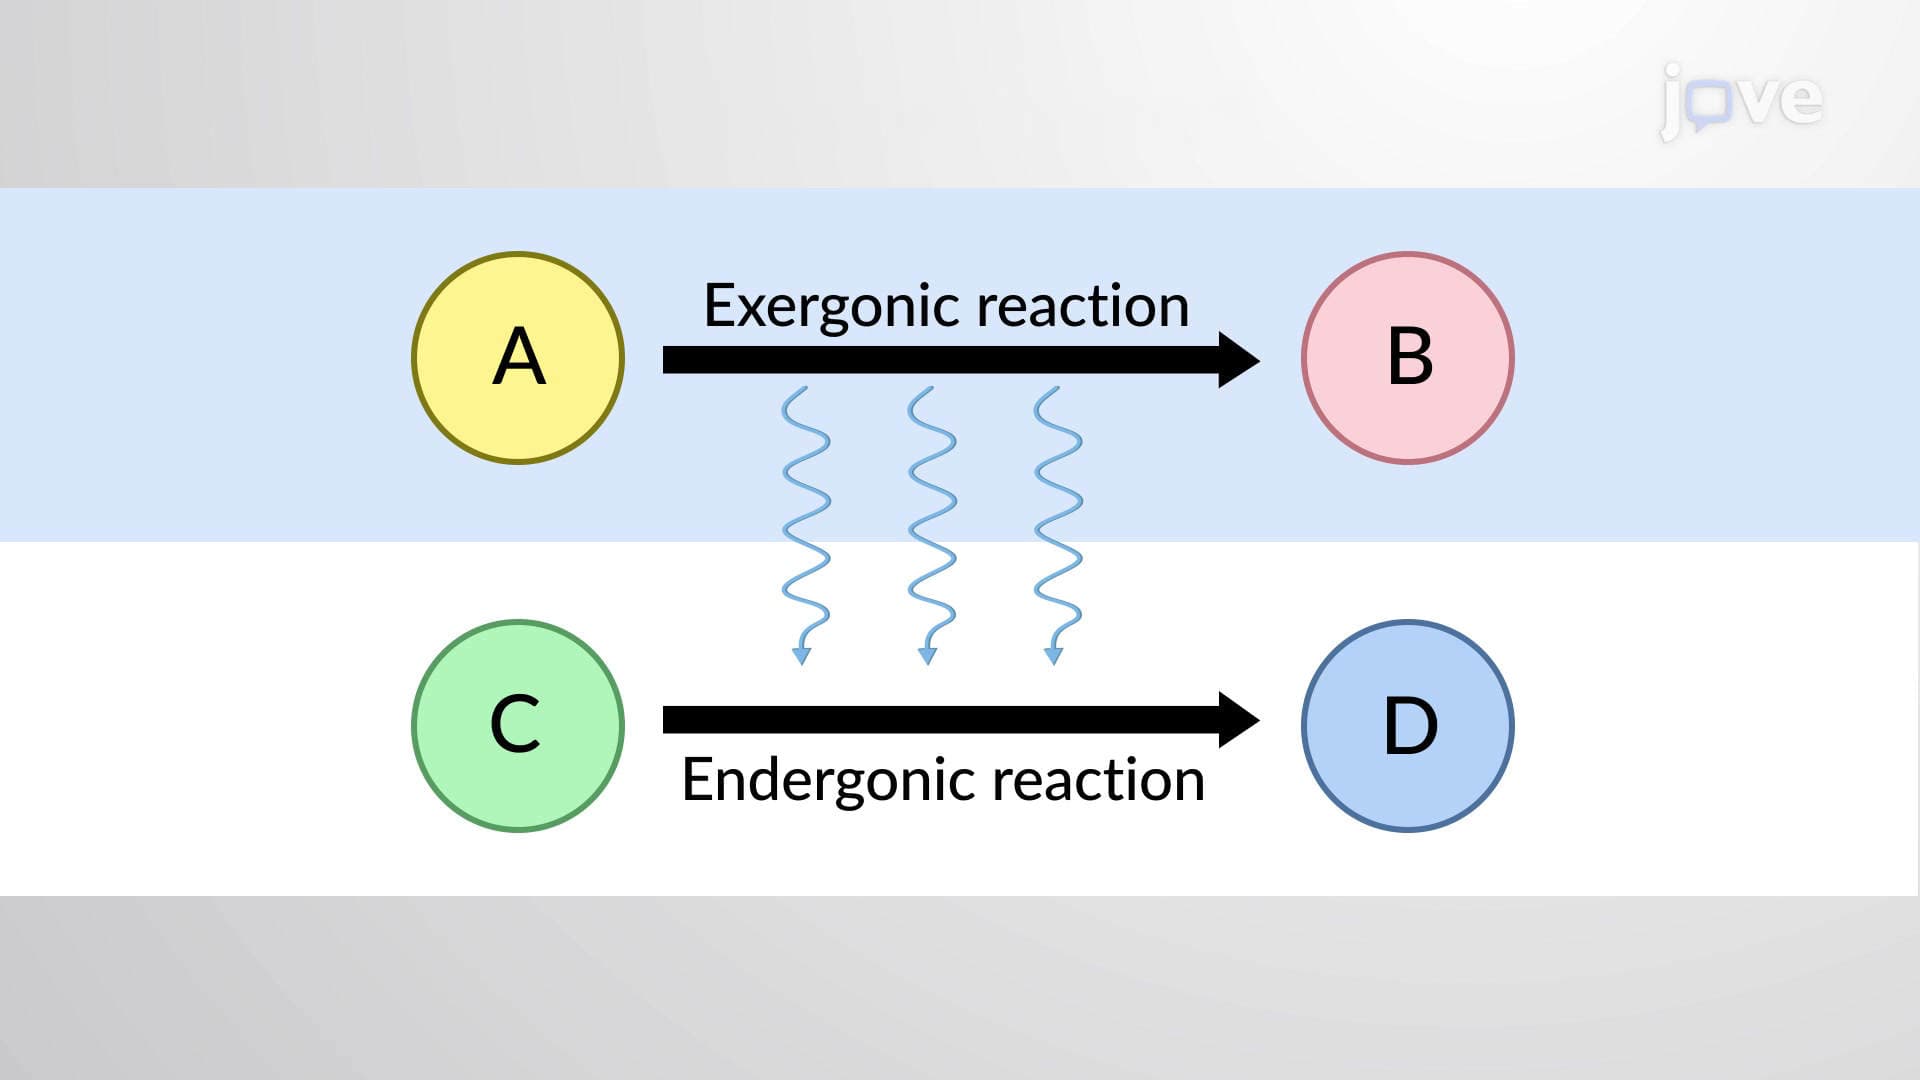
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions. Cells...