Related Experiment Video
Updated: May 20, 2026
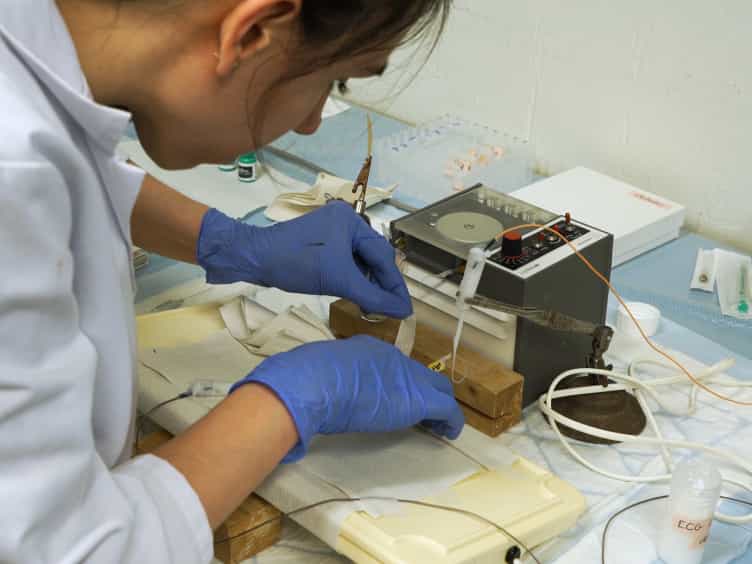
Nasal Potential Difference to Quantify Trans-epithelial Ion Transport in Mice
Published on: July 4, 2018
A pharmacologic approach to acquired cystic fibrosis transmembrane conductance regulator dysfunction in smoking
Peter A Sloane1, Suresh Shastry, Andrew Wilhelm
1Cystic Fibrosis Research Center, University of Alabama at Birmingham, Birmingham, Alabama, United States of America.

Background:
Mucus stasis in chronic obstructive pulmonary disease (COPD) is a significant contributor to morbidity and mortality. Potentiators of cystic fibrosis transmembrane conductance regulator (CFTR) activity pharmacologically enhance CFTR function; ivacaftor is one such agent approved to treat CF patients with the G551D-CFTR gating mutation. CFTR potentiators may also be useful for other diseases of mucus stasis, including COPD.
Methods And Findings:
In primary human bronchial epithelial cells, exposure to cigarette smoke extract diminished CFTR-mediated anion transport (65.8±0.2% of control, P<0.005) and mucociliary transport (0.17±0.05 µm/sec vs. 2.4±0.47 µm/sec control, P<0.05) by reducing airway surface liquid depth (7.3±0.6 µm vs. 13.0±0.6 µm control, P<0.005) and augmenting mucus expression (by 64%, P<0.05) without altering transepithelial resistance. Smokers with or without COPD had reduced CFTR activity measured by nasal potential difference compared to age-matched non-smokers (-6.3±1.4 and -8.0±2.0 mV, respectively vs. -15.2±2.7 mV control, each P<0.005, n = 12-14/group); this CFTR decrement was associated with symptoms of chronic bronchitis as measured by the Breathlessness Cough and Sputum Score (r = 0.30, P<0.05) despite controlling for smoking (r = 0.31, P<0.05). Ivacaftor activated CFTR-dependent chloride transport in non-CF epithelia and ameliorated the functional CFTR defect induced by smoke to 185±36% of non-CF control (P<0.05), thereby increasing airway surface liquid (from 7.3±0.6 µm to 10.1±0.4 µm, P<0.005) and mucociliary transport (from 0.27±0.11 µm/s to 2.7±0.28 µm/s, P<0.005).
Conclusions:
Cigarette smoking reduces CFTR activity and is causally related to reduced mucus transport in smokers due to inhibition of CFTR dependent fluid transport. These effects are reversible by the CFTR potentiator ivacaftor, representing a potential therapeutic strategy to augment mucociliary clearance in patients with smoking related lung disease.
More Related Videos
Related Concept Videos
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:29
Chronic Obstructive Pulmonary Disease-V: Management
Smoking Cessation
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:24
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:23
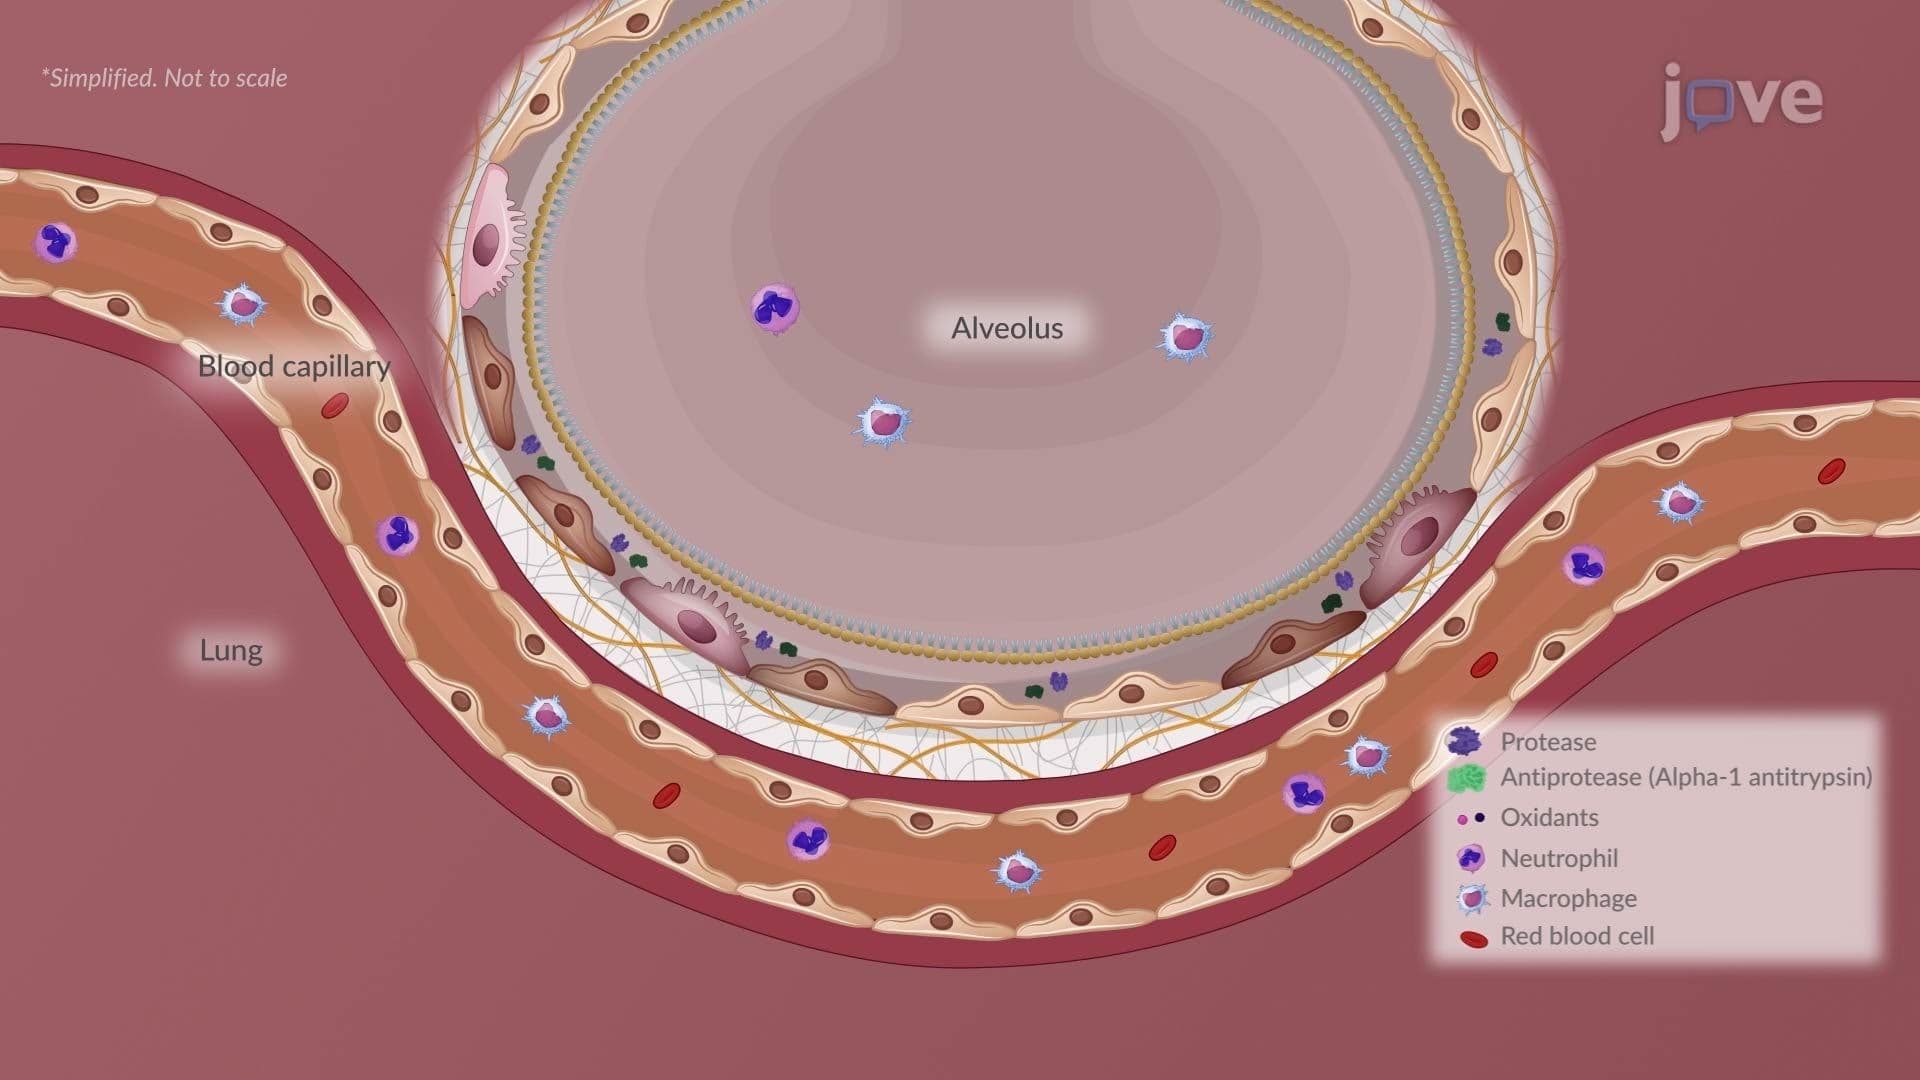
Chronic Obstructive Pulmonary Disease II: Emphysema