Related Experiment Video
Updated: May 20, 2026
A Plate-Based Assay for the Measurement of Endogenous Monoamine Release in Acute Brain Slices
Published on: August 11, 2021
Pharmacokinetics of quinacrine efflux from mouse brain via the P-glycoprotein efflux transporter
Misol Ahn1, Sina Ghaemmaghami, Yong Huang
1Institute for Neurodegenerative Diseases, University of California San Francisco, San Francisco, California, USA.

Abstract:
The lipophilic cationic compound quinacrine has been used as an antimalarial drug for over 75 years but its pharmacokinetic profile is limited. Here, we report on the pharmacokinetic properties of quinacrine in mice. Following an oral dose of 40 mg/kg/day for 30 days, quinacrine concentration in the brain of wild-type mice was maintained at a concentration of ∼1 µM. As a substrate of the P-glycoprotein (P-gp) efflux transporter, quinacrine is actively exported from the brain, preventing its accumulation to levels that may show efficacy in some disease models. In the brains of P-gp-deficient Mdr1(0/0) mice, we found quinacrine reached concentrations of ∼80 µM without any signs of acute toxicity. Additionally, we examined the distribution and metabolism of quinacrine in the wild-type and Mdr1(0/0) brains. In wild-type mice, the co-administration of cyclosporin A, a known P-gp inhibitor, resulted in a 6-fold increase in the accumulation of quinacrine in the brain. Our findings argue that the inhibition of the P-gp efflux transporter should improve the poor pharmacokinetic properties of quinacrine in the CNS.
Related Concept Videos
 01:16
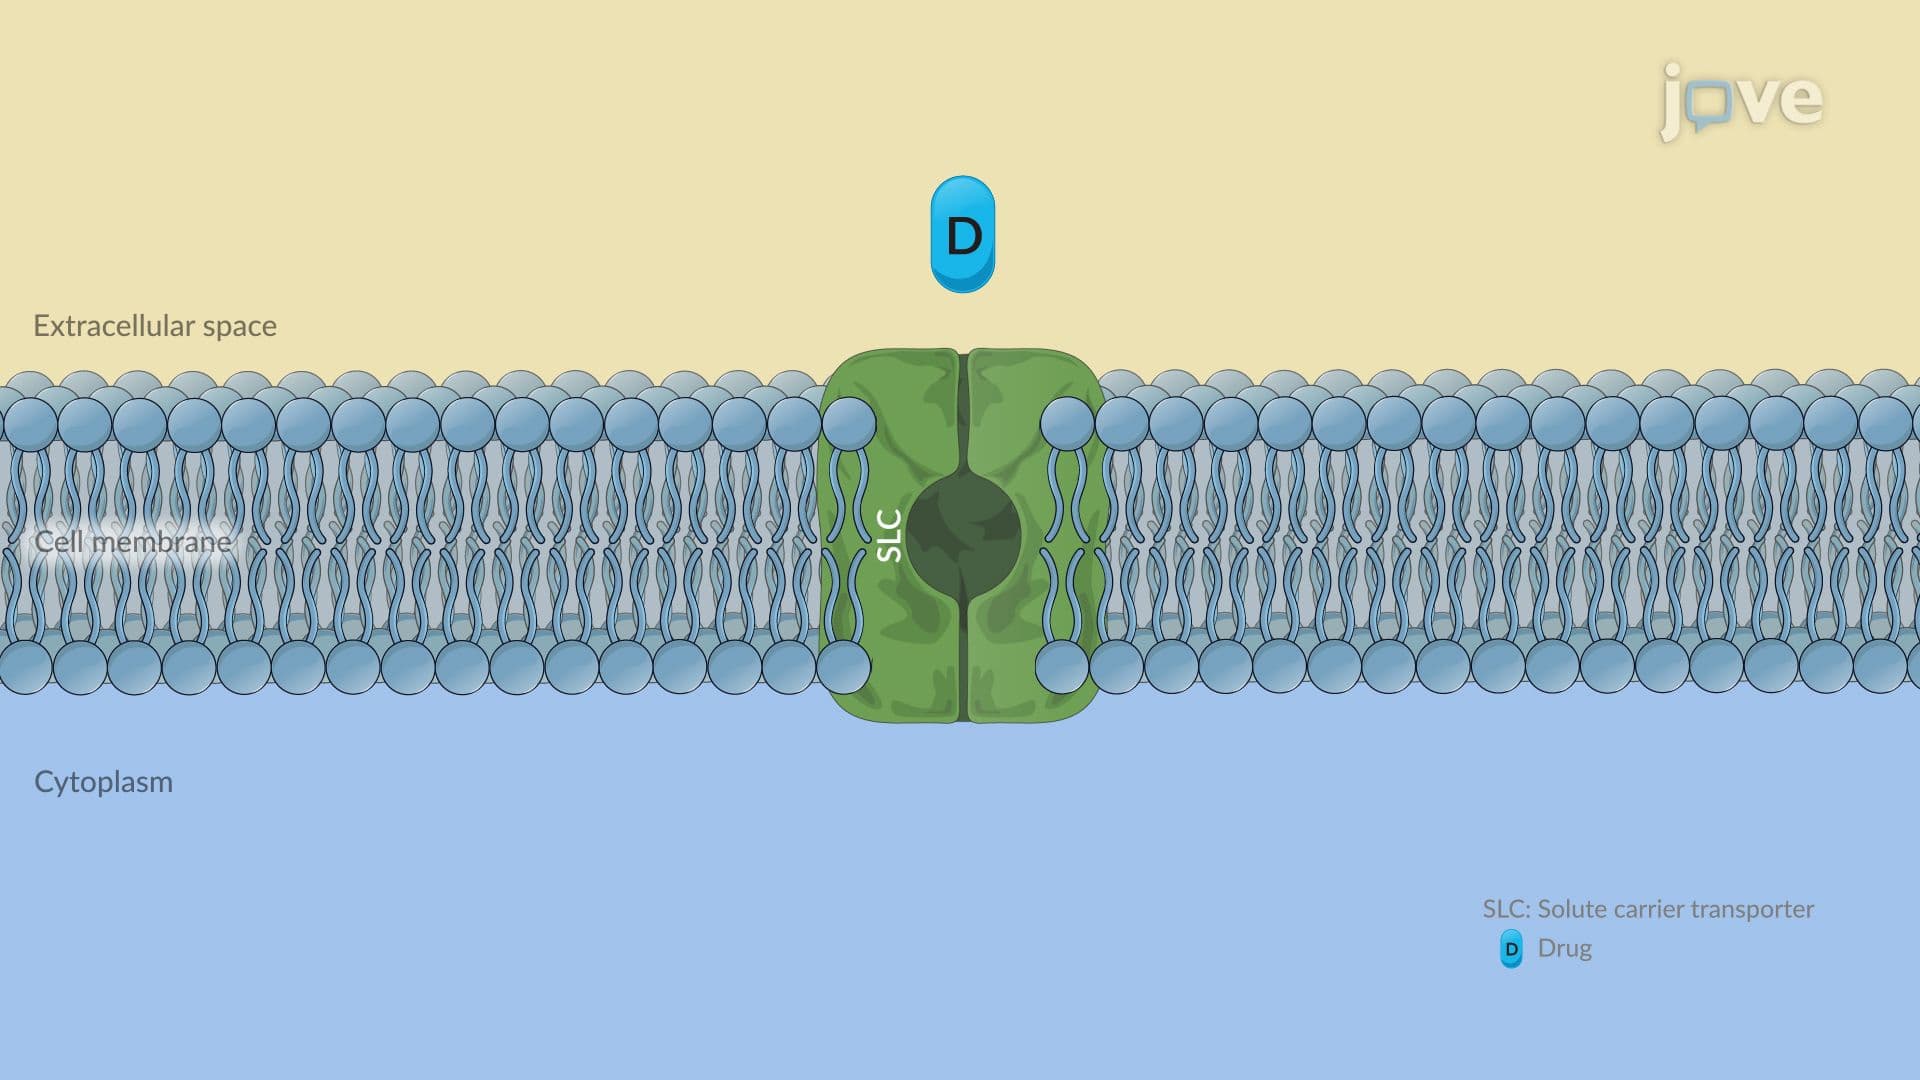
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:24
Cholinergic Antagonists: Pharmacokinetics
 01:22
Indirect-Acting Cholinergic Agonists: Pharmacokinetics
Reversible agents containing quaternary amines, such as neostigmine and edrophonium, are not easily absorbed orally because they are...
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model