Related Experiment Video
Updated: May 20, 2026

A Multiplexed Luciferase-based Screening Platform for Interrogating Cancer-associated Signal Transduction in Cultured Cells
Published on: July 3, 2013
Differential involvement of RalA and RalB in colorectal cancer
Timothy D Martin1, Channing J Der
1Deparment of Pharmacology, Lineberger Comprehensive Cancer Center, University of North Carolina at Chapel Hill, Chapel Hill, NC, USA.

Abstract:
Mutationally activated K-Ras can utilize a multitude of downstream effector proteins to promote oncogenesis. While the Raf and phosphoinositol 3-kinase effector pathways are the best-studied and validated, recent studies have established the critical importance of Ral guanine nucleotide exchange factor (RalGEF) activation of the RalA and RalB small GTPases in cancer biology. Due to recent evidence that the RalGEF-Ral pathway is necessary for the tumorigenic and metastatic potential of KRAS mutant pancreatic ductal adenocarcinoma (PDAC) tumor cells, we investigated whether or not Ral signaling was necessary for KRAS mutant colorectal cancer (CRC) tumor cell growth. As in PDAC, we found upregulated RalA and RalB activation in CRC tumor cell lines and tumors. Surprisingly we found antagonistic roles for RalA and RalB in the regulation of CRC tumor cell anchorage-independent growth. This observation contrasts with PDAC, where RalA but not RalB is necessary for PDAC tumor cell anchorage-independent growth. Our results emphasize cancer cell type differences in Ral function and hence the need for distinct Ral targeted therapeutic approaches in the treatment of CRC vs. PDAC.
Insights
KRAS-mutant colorectal cancer cells rely on RalA and RalB signaling for growth, but these proteins have opposing roles. This differs from pancreatic cancer, highlighting the need for tailored therapies targeting Ral signaling in different cancers.
Area of Science:
- Oncology
- Molecular Biology
- Cancer Signaling Pathways
Background:
- K-Ras mutations drive oncogenesis through various downstream effectors.
- The Ral guanine nucleotide exchange factor (RalGEF)-Ral pathway is increasingly recognized for its role in cancer.
- Previous studies linked RalGEF-Ral pathway activation to KRAS-mutant pancreatic ductal adenocarcinoma (PDAC) tumorigenesis.
Purpose of the Study:
- To investigate the necessity of Ral signaling for KRAS-mutant colorectal cancer (CRC) tumor cell growth.
- To compare the roles of RalA and RalB in CRC with their known functions in PDAC.
Main Methods:
- Analysis of RalA and RalB activation in CRC cell lines and tumors.
- Assessment of RalA and RalB roles in CRC cell anchorage-independent growth.
Main Results:
- Upregulated RalA and RalB activation was observed in CRC, similar to PDAC.
- RalA and RalB exhibited antagonistic roles in regulating CRC cell anchorage-independent growth.
- This contrasts with PDAC, where only RalA is essential for anchorage-independent growth.
Conclusions:
- Ral signaling plays a crucial role in KRAS-mutant CRC growth.
- Distinct roles of RalA and RalB in CRC versus PDAC necessitate cancer type-specific therapeutic strategies.
- Targeted therapies for Ral signaling should be tailored for CRC and PDAC treatment.
More Related Videos
Related Concept Videos
 02:38
The Ras Gene
Ras is a superfamily...
 01:24
Small GTPases - Ras and Rho
Three regulatory proteins control their activity:
 01:21
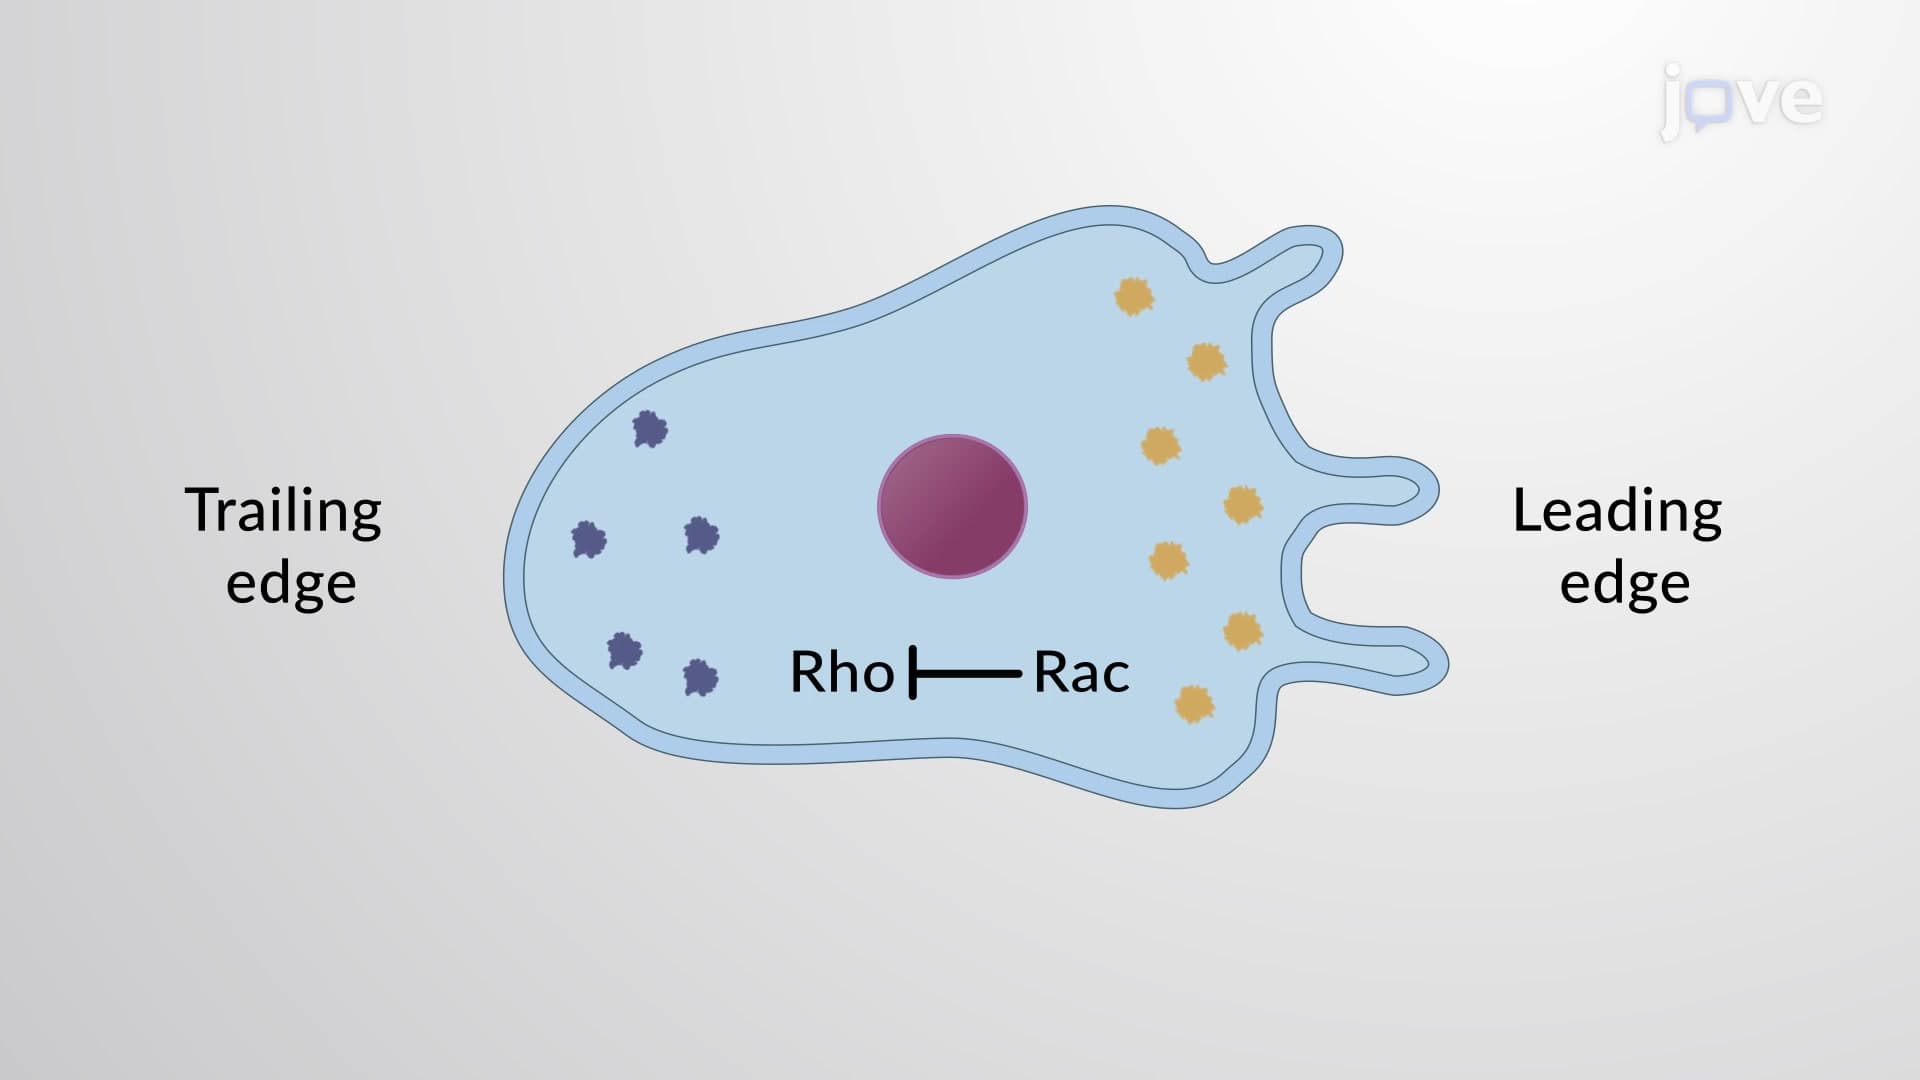
Cell Polarization by Rho Proteins
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...
 02:07
Tumor Progression
Colon cancer is one of the best-documented examples of tumor progression. Early mutation in the APC gene in colon cells causes a small growth on the colon wall called a polyp. With time, this polyp grows into a benign, pre-cancerous tumor. Further...
 01:03
Rous Sarcoma Virus (RSV) and Cancer
RSV is a retrovirus that contains two copies of a plus-strand RNA genome. Its genome consists of four main open...