Related Experiment Video
Updated: May 20, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Parametric effects of the potential energy function on the geometrical features of ternary Lennard-Jones clusters
Xia Wu1, Yan Sun, Chunsheng Li
1School of Chemistry and Chemical Engineering, Anqing Normal University, Anqing 246011, P. R. China. xiawu@aqtc.edu.cn

Abstract:
The impact of parameters in potential function for describing atomic or molecular clusters is complex due to the complicated potential energy surface. Ternary Lennard-Jones (TLJ) A(l)B(m)C(n) clusters with two-body potential are investigated to study the effect of parameters. In the potential, the size parameter (σ(AA)) of A atoms is fixed, and corresponding parameters of B and C atoms (relative to A atoms), i.e., σ(BB)/σ(AA) and σ(CC)/σ(AA) > 1.00, are used to control the atomic interaction among A, B, and C atoms in TLJ clusters. The minimum energy configurations of A(l)B(m)C(n) clusters with different species are optimized by adaptive immune optimization algorithm. Ternary cluster structures, bonds, and energies of the putative minima are studied. The results show that two different structures based on double-icosahedra are found in 30-atom TLJ clusters. Furthermore, with increasing potential size parameters of B and C atoms, A atoms tend to be more compact for the increasing numbers of A-A bonds, but the short-range attractive part in TLJ clusters becomes weaker. To lower the potential energy, B and C atoms grow around the A atoms in pursuit of a compact configuration. The results are also approved in A(l)B(m)C(n) (l + m + n = 9-55) clusters and A(l)B(m)C(n) (l = 13, m + n = 42) clusters.

Related Concept Videos
 01:04
Force and Potential Energy in Three Dimensions
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:36
Molecular Geometry and Dipole Moments
 01:28
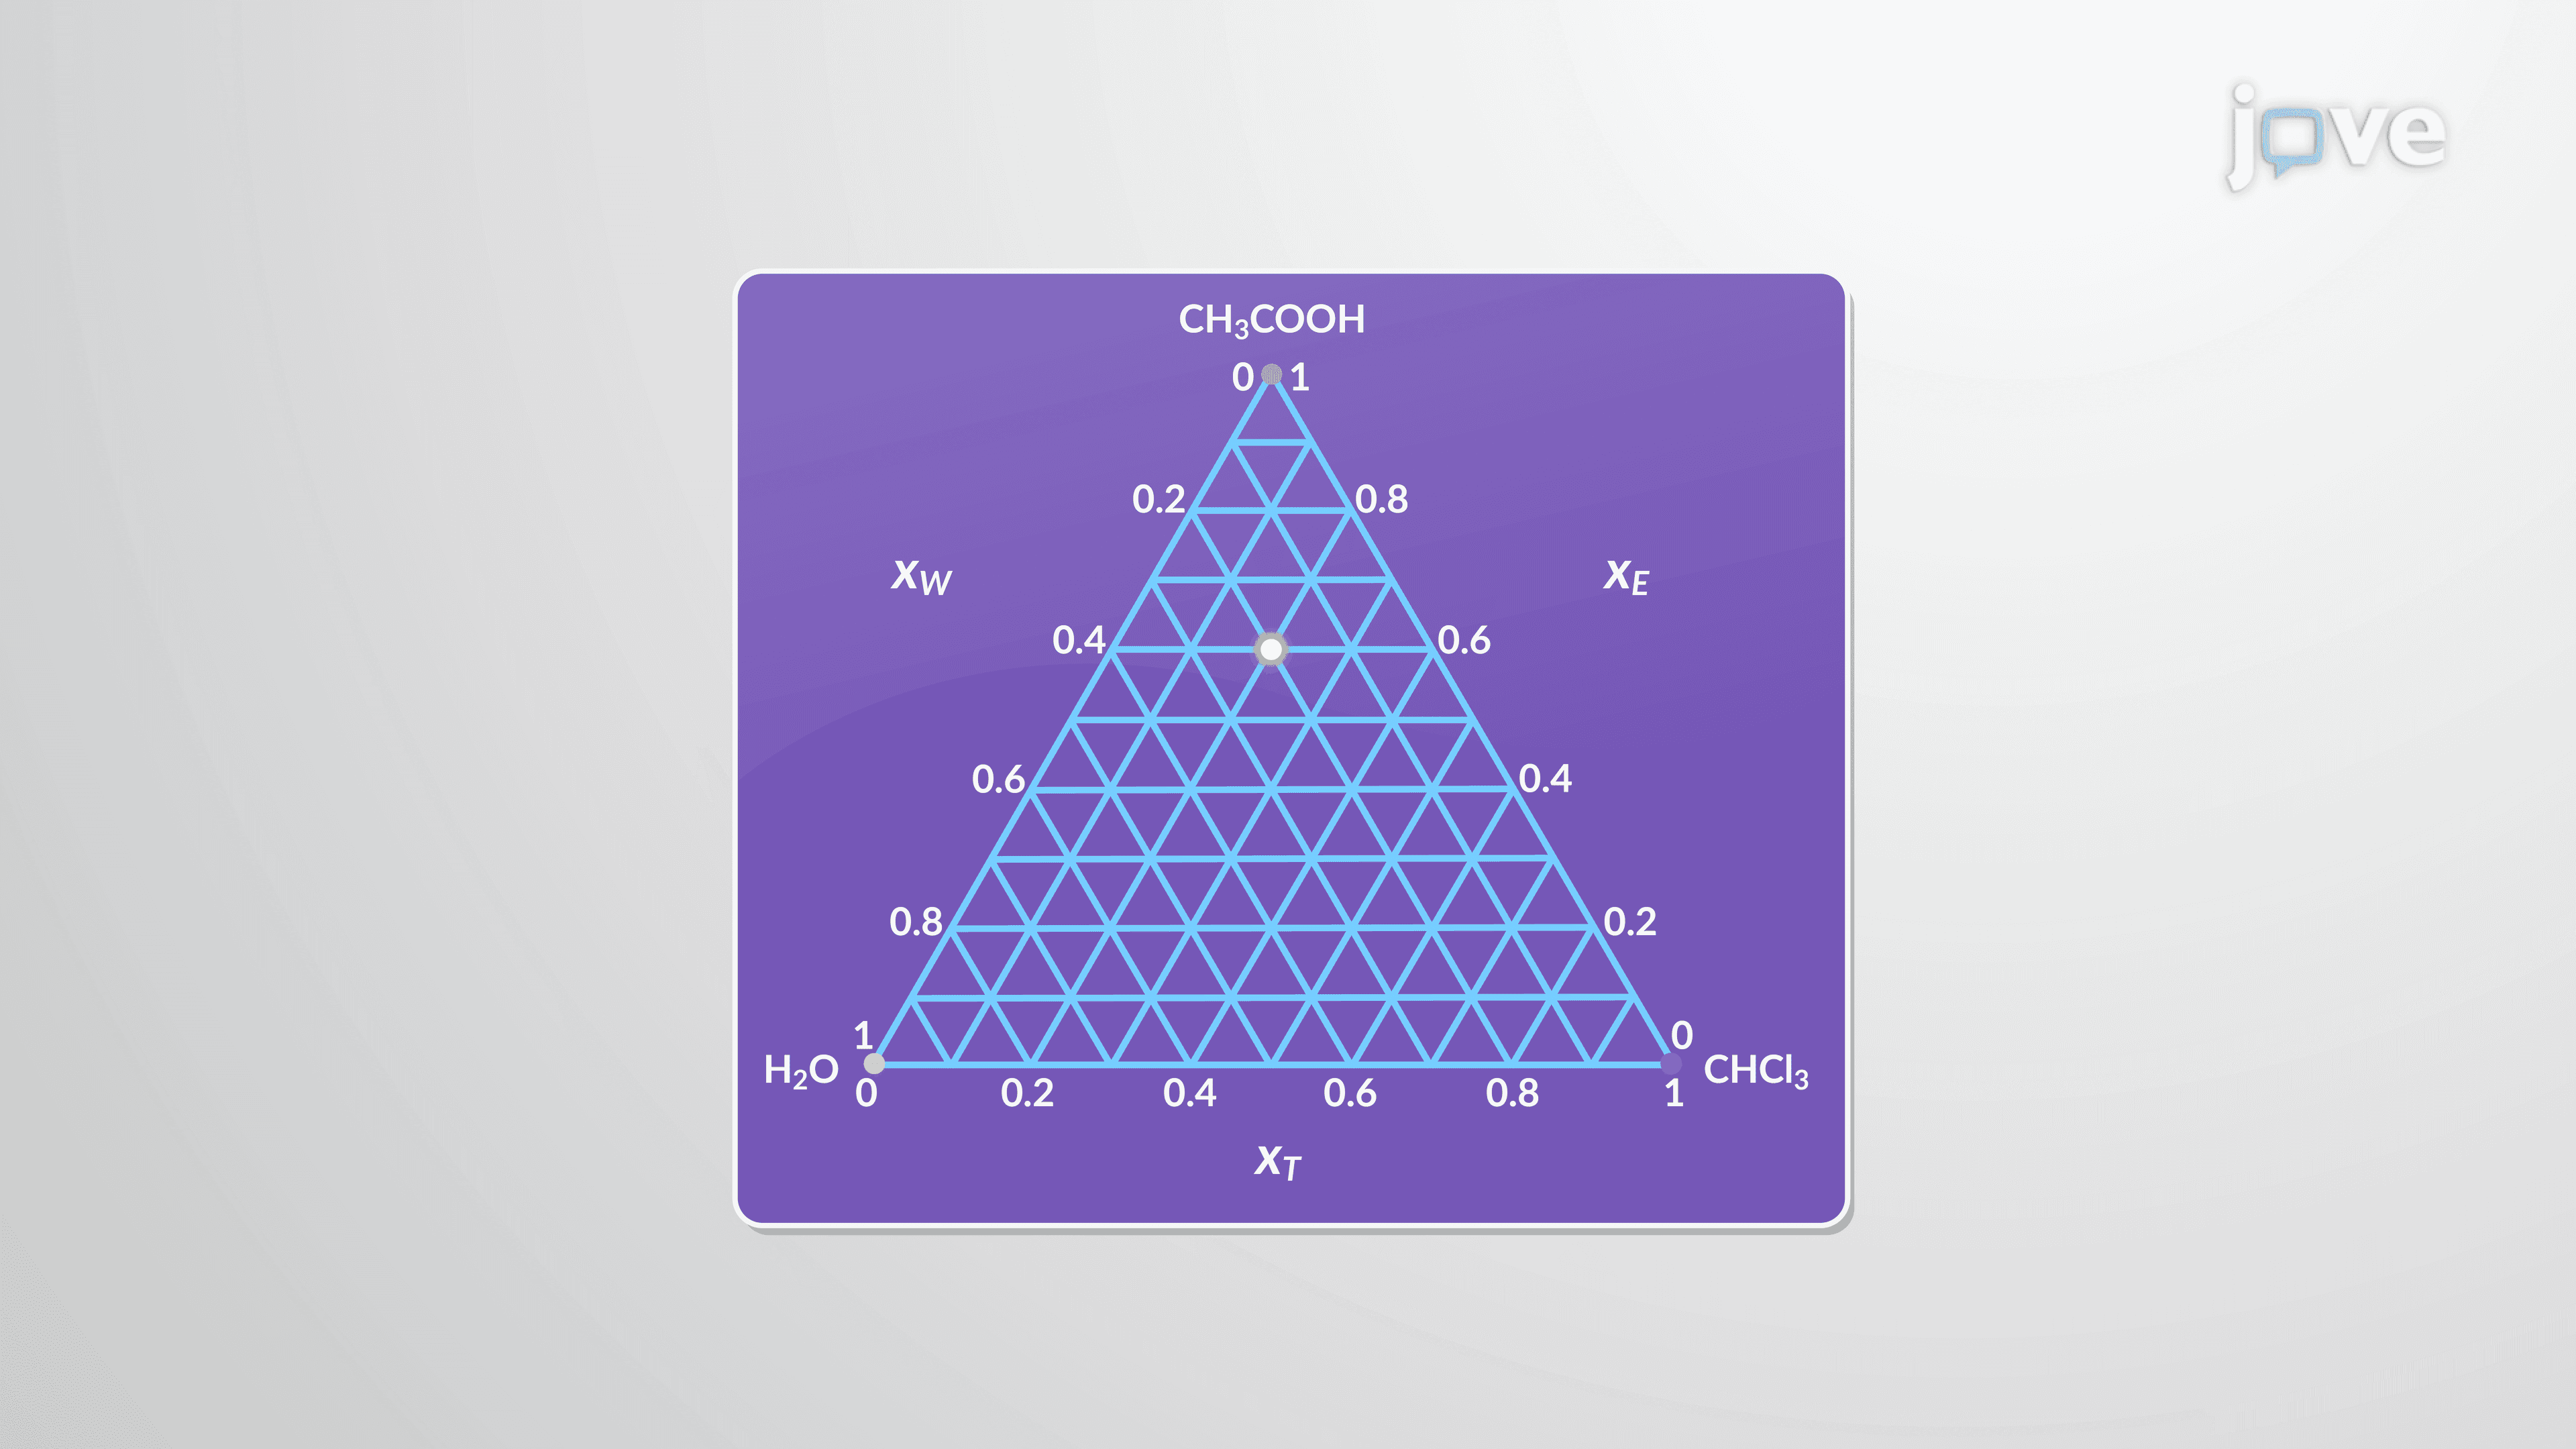
Phase Diagrams of Ternary Systems
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...