Related Experiment Video
Updated: May 20, 2026

Autoradiography as a Simple and Powerful Method for Visualization and Characterization of Pharmacological Targets
Published on: March 12, 2019
Rebinding: or why drugs may act longer in vivo than expected from their in vitro target residence time
1Free University Brussels (VUB), Department of Molecular and Biochemical Pharmacology, Building E.5.10, Pleinlaan 2, B-1050 Brussels, Belgium +32 2 6291955 ; +32 2 6291358 ; gvauquel@vub.ac.be.

Importance Of The Field:
It is well established that the in vivo duration of drug action not only depends on macroscopic pharmacokinetic properties like its plasma half-life, but also on the residence time of the drug-target complexes. However, drug 'rebinding' (i.e., the consecutive binding of dissociated drug molecules to the original target and/or targets nearby) can be influential in vivo as well.
Areas Covered In This Review:
Information about rebinding is available since the 1980s but it is dispersed in the life sciences literature. This review compiles this information. In this respect, neurochemists and biopohysicians advance the same equations to describe drug rebinding.
What The Reader Will Gain:
The rebinding mechanism is explained according to the prevailing viewpoint in different life science disciplines. There is a general consensus that high target densities, high association rates and local phenomena that hinder the diffusion of free drug molecules away from their target all promote rebinding.
Take Home Message:
Simulations presented here for the first time suggest that rebinding may increase the duration and even the constancy of the drug's clinical action. Intact cell radioligand dissociation and related ex vivo experiments offer useful indications about a drug's aptitude to experience target rebinding.
Related Concept Videos
 01:21
Drug Distribution: Tissue Binding
For...
 01:23
Pharmacokinetic–Pharmacodynamic Relationship: Influence of Elimination Half-Life on Effect Duration
 01:22
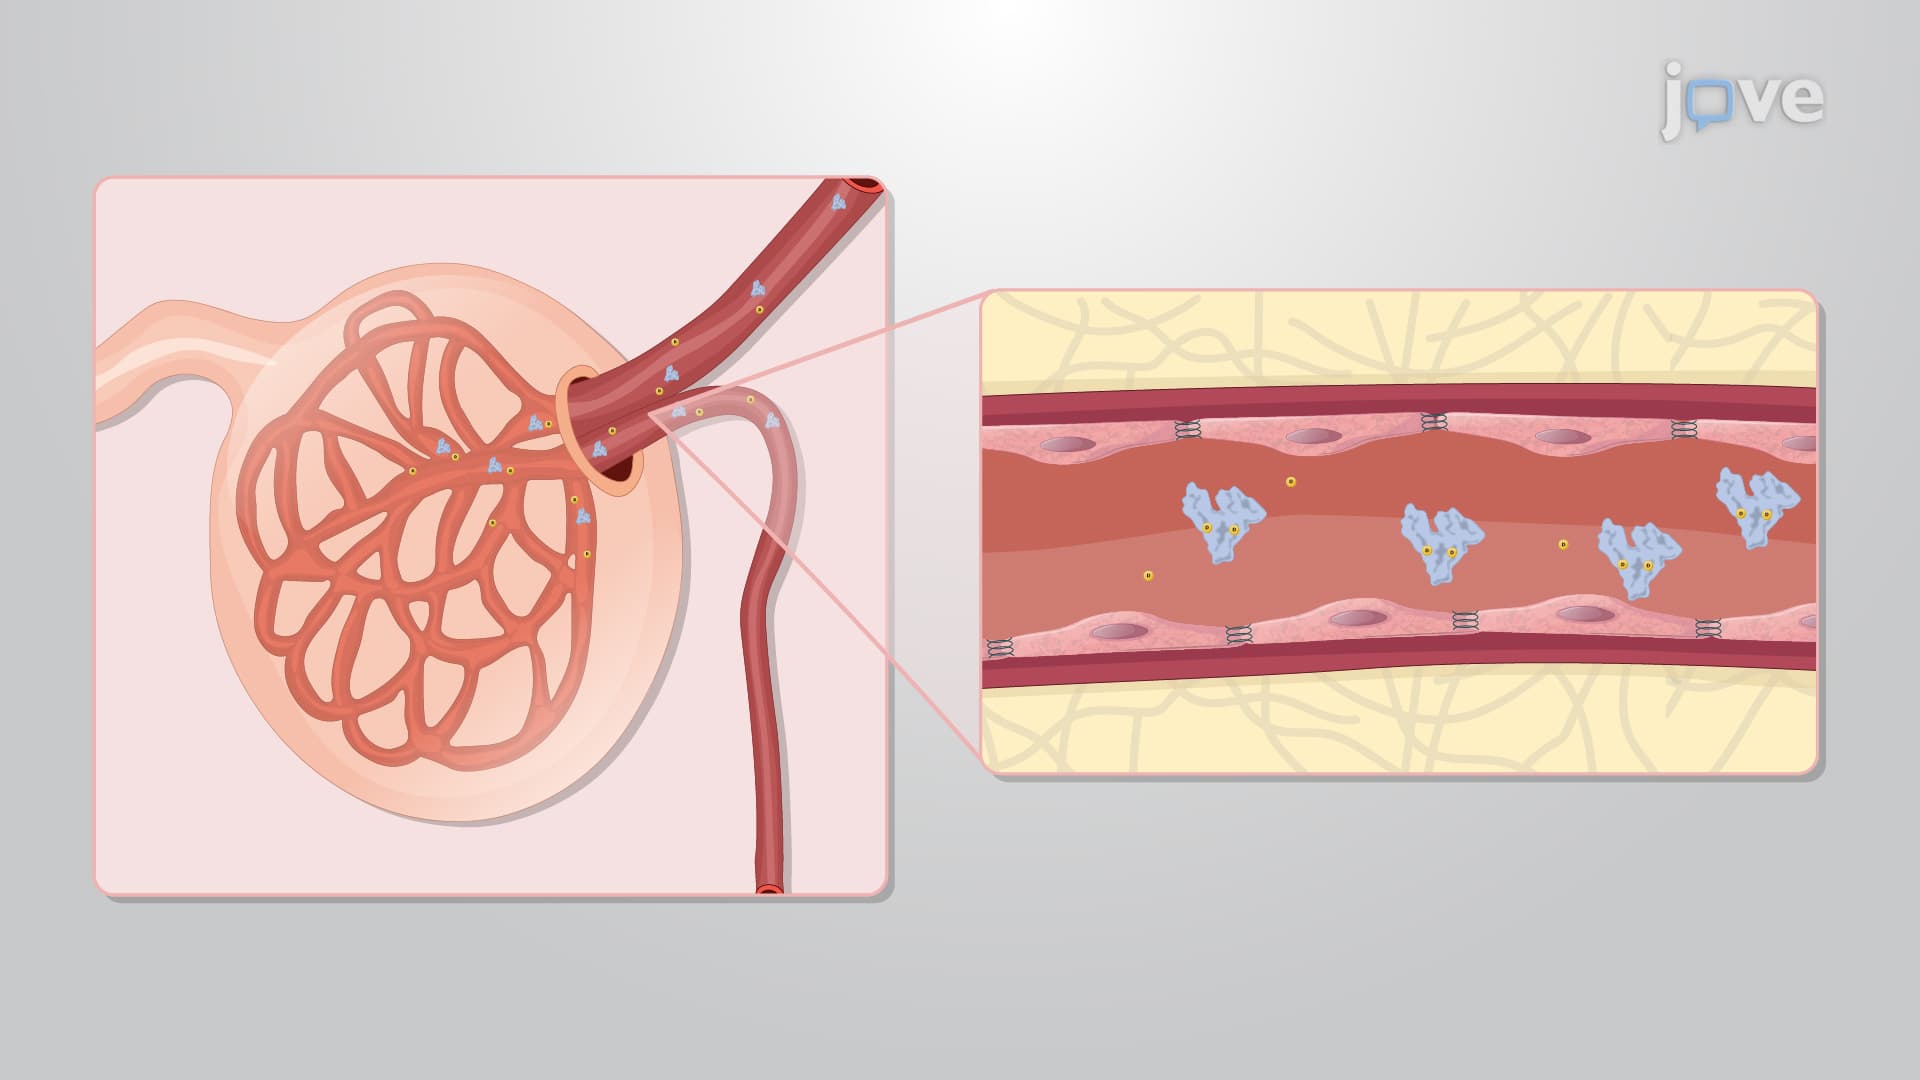
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
 01:16
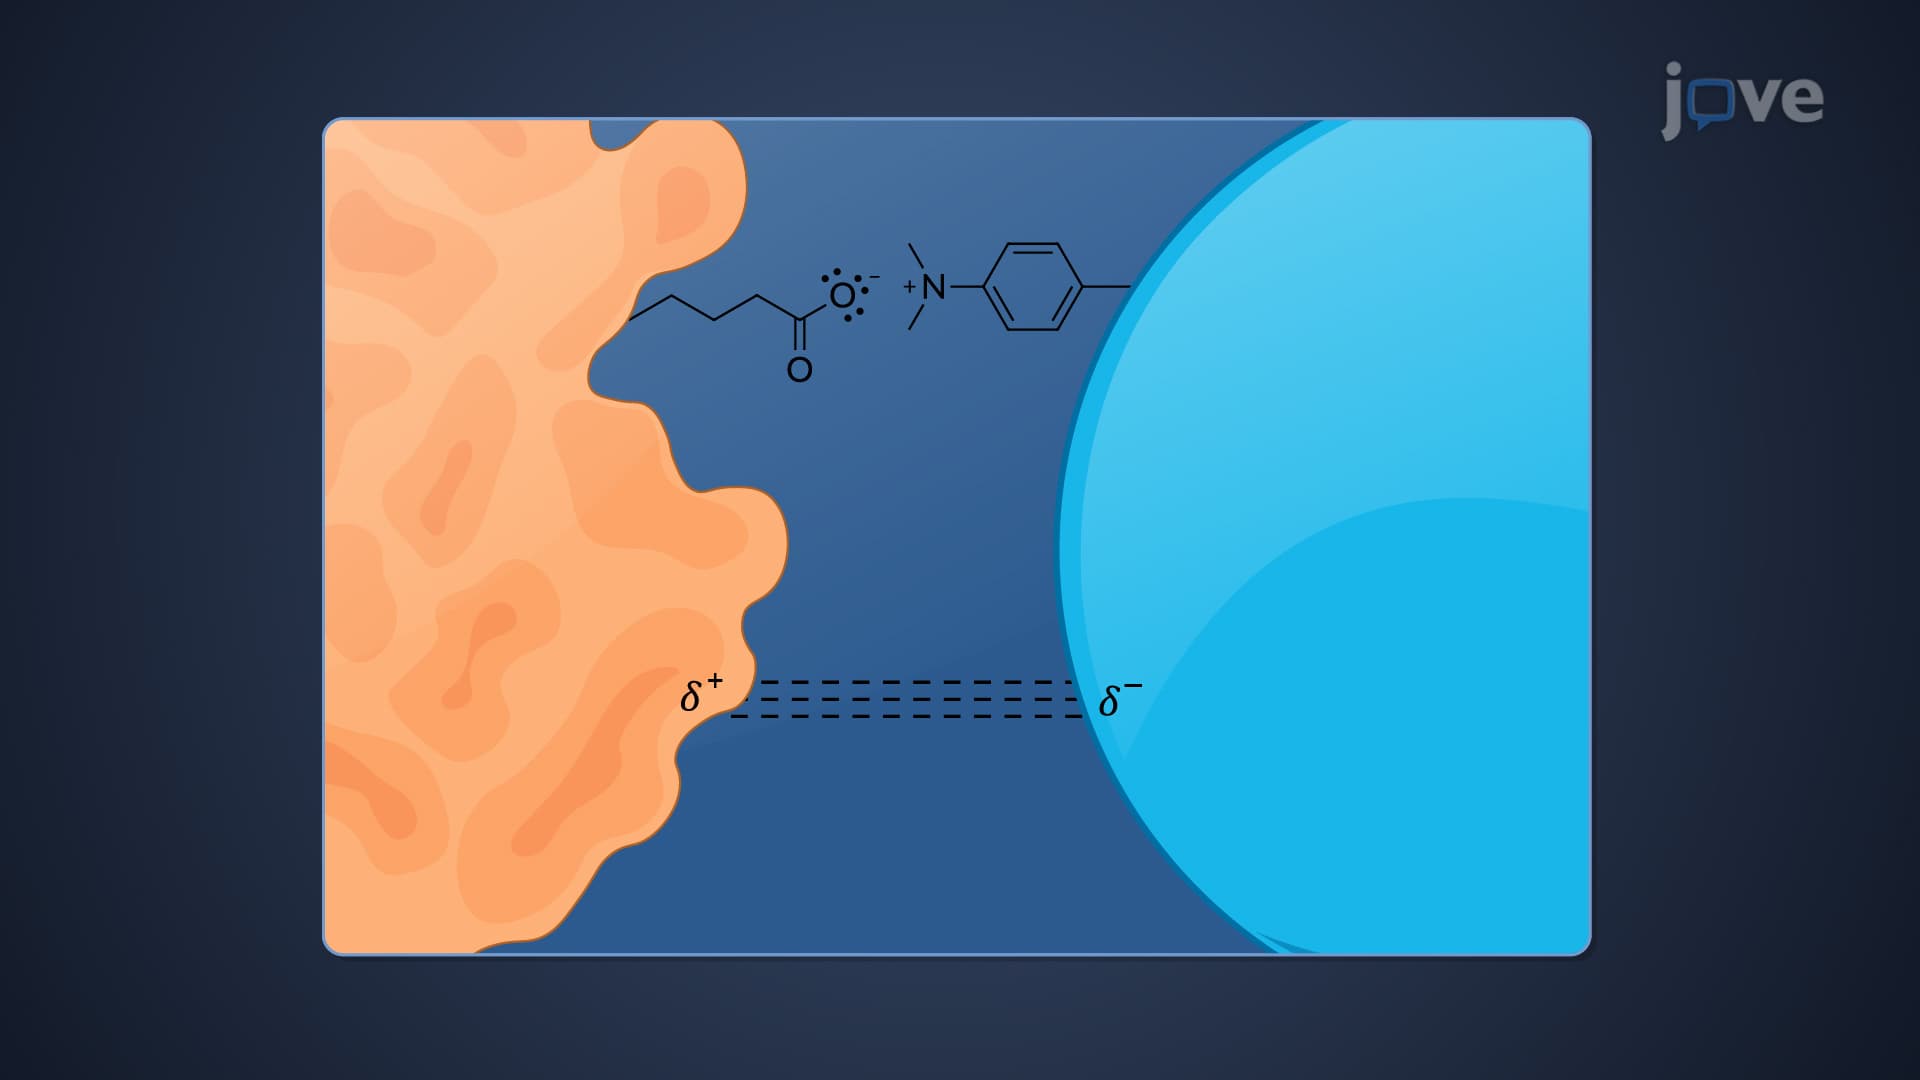
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:29
Drug Distribution: Plasma Protein Binding
 01:30
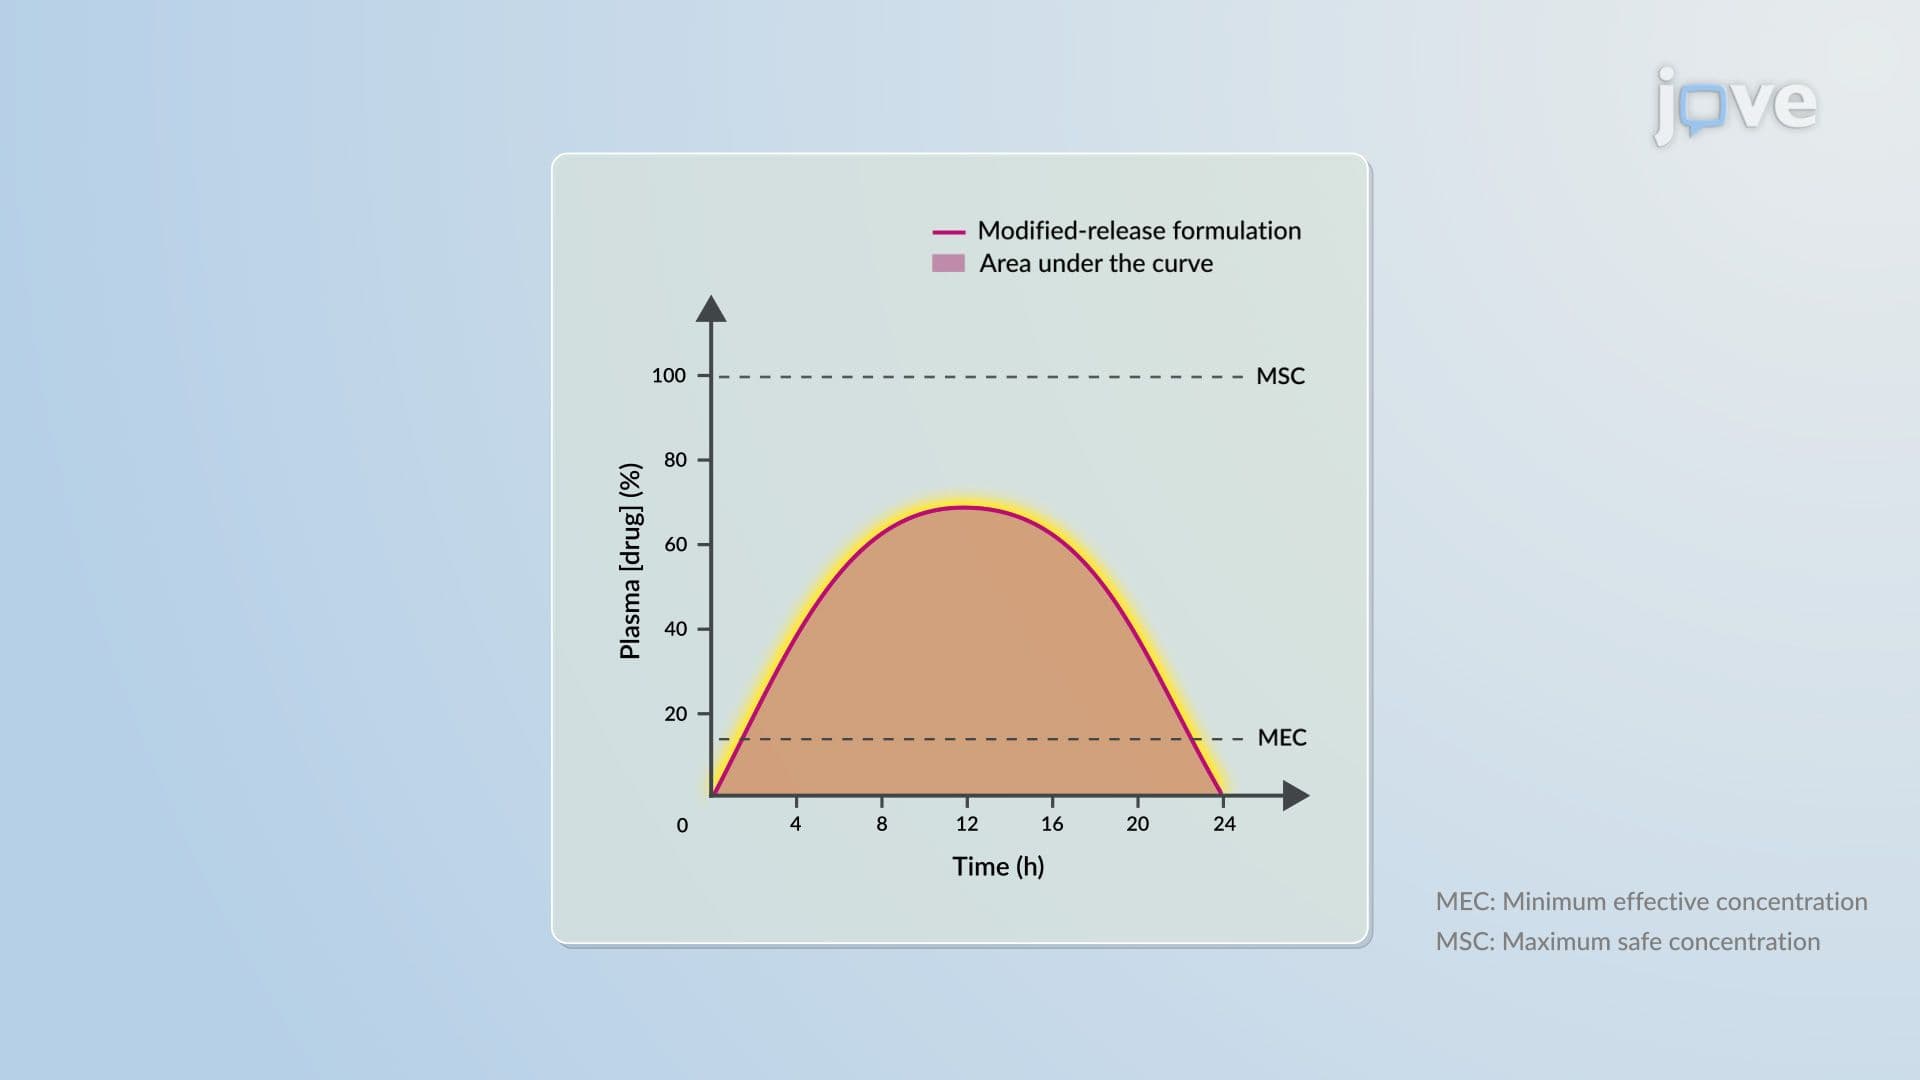
Modified-Release Drug Delivery Systems: Bioavailability