Related Experiment Video
Updated: May 20, 2026
Preparation of a Corannulene-functionalized Hexahelicene by Copper(I)-catalyzed Alkyne-azide Cycloaddition of Nonplanar Polyaromatic Units
Published on: September 18, 2016
(1) H NMR spectral analysis and conformational behavior of n-alkanes in different chemical environments
Tuulia Tynkkynen1, Tommi Hassinen, Mika Tiainen
1School of Pharmacy, Faculty of Health Sciences, University of Eastern Finland, PO Box 1627, 70211, Kuopio, Finland.

Abstract:
Alkyl chains are common structural units, for example in lipids, and their (1) H NMR spectral parameters offer valuable information about their conformational behavior in solvent environment. Even the spectra of short n-alkanes are complex, which is obviously a reason why their accurate spectral analyses have not been reported before. The present study reports the quantum mechanical analysis of (1) H NMR spectra of n-butane, n-pentane, n-hexane, and n-heptane. The spectral parameters were used to characterize the conformational behavior of n-alkanes. The temperature dependence analysis of coupling constants suggests that the enthalpy difference between the gauche (g) and trans (t) conformations (ΔH(g) ) of n-butane in chloroform is 2.55-2.85 kJ mol(-1) . The difference between the trans-gauche (tg) and all-trans (tt) conformers of n-pentane (ΔH(tg) ) seems to be 0.1-0.2 kJ mol(-1) higher. The coupling constant information shows that the t(n) conformations become more favored with longer chains, although not only for energetic reasons but also partly because the g(+) g(-) arrangements become sterically unfavorable, which decreases the number of favorable g(n) -type conformations. The analysis of the (1) H NMR spectra of n-pentane and n-hexane in solvents representing different chemical environments indicates that polar and spherical dimethyl sulfoxide favors clearly the g conformations, whereas n-hexane-d(14) favors slightly the extended t(n) conformation. In addition to the intrinsic scientific importance for NMR spectral parameter prediction and molecular modeling in solution, the results provide some insights to behavior of hydrocarbon chains and their spectra in different chemical environments.
Related Concept Videos
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:22
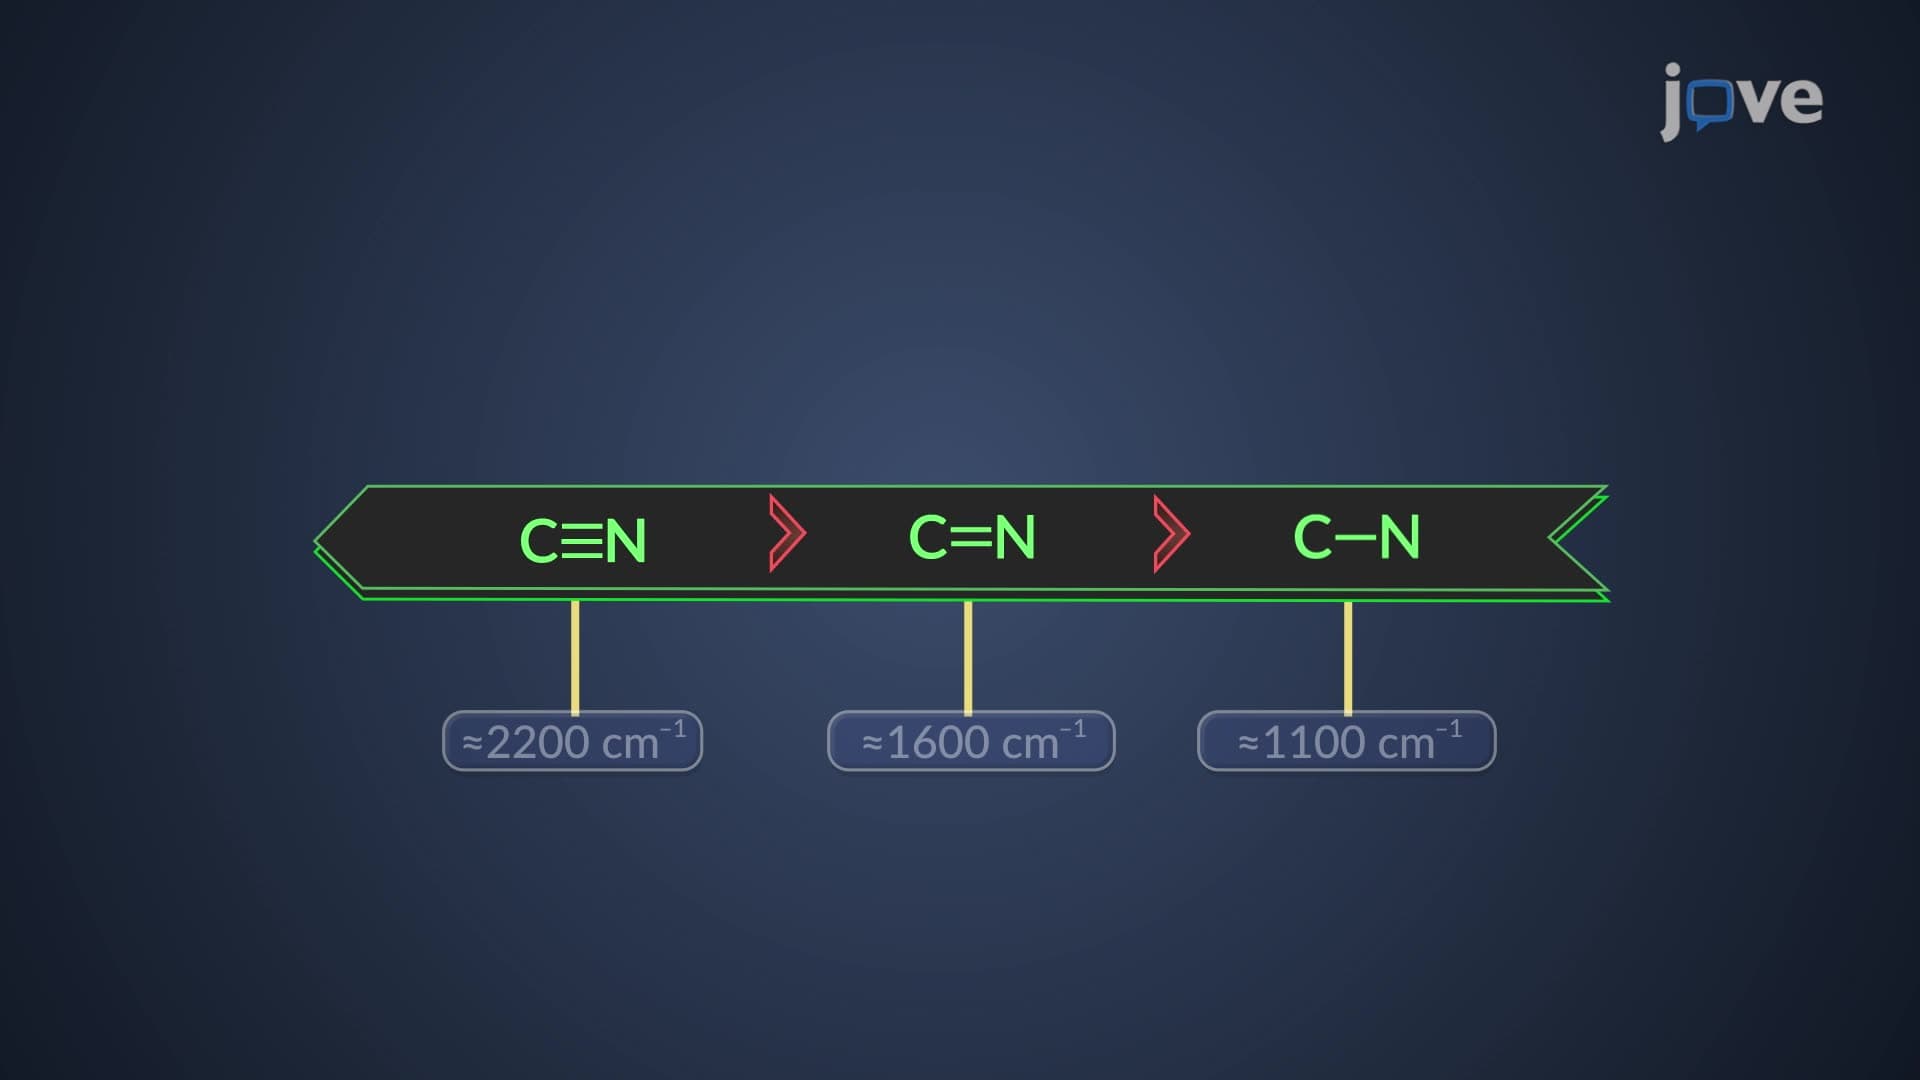
IR Frequency Region: Alkyne and Nitrile Stretching
Comparing the stretching vibrational frequency of C≡C triple bonds with that of double and single bonds, it is evident that C≡C triple bonds exhibit a higher stretching frequency than C=C double and C–C single bonds. Similarly, the C≡N triple bond exhibits higher stretching absorption than the C=N...
 01:19
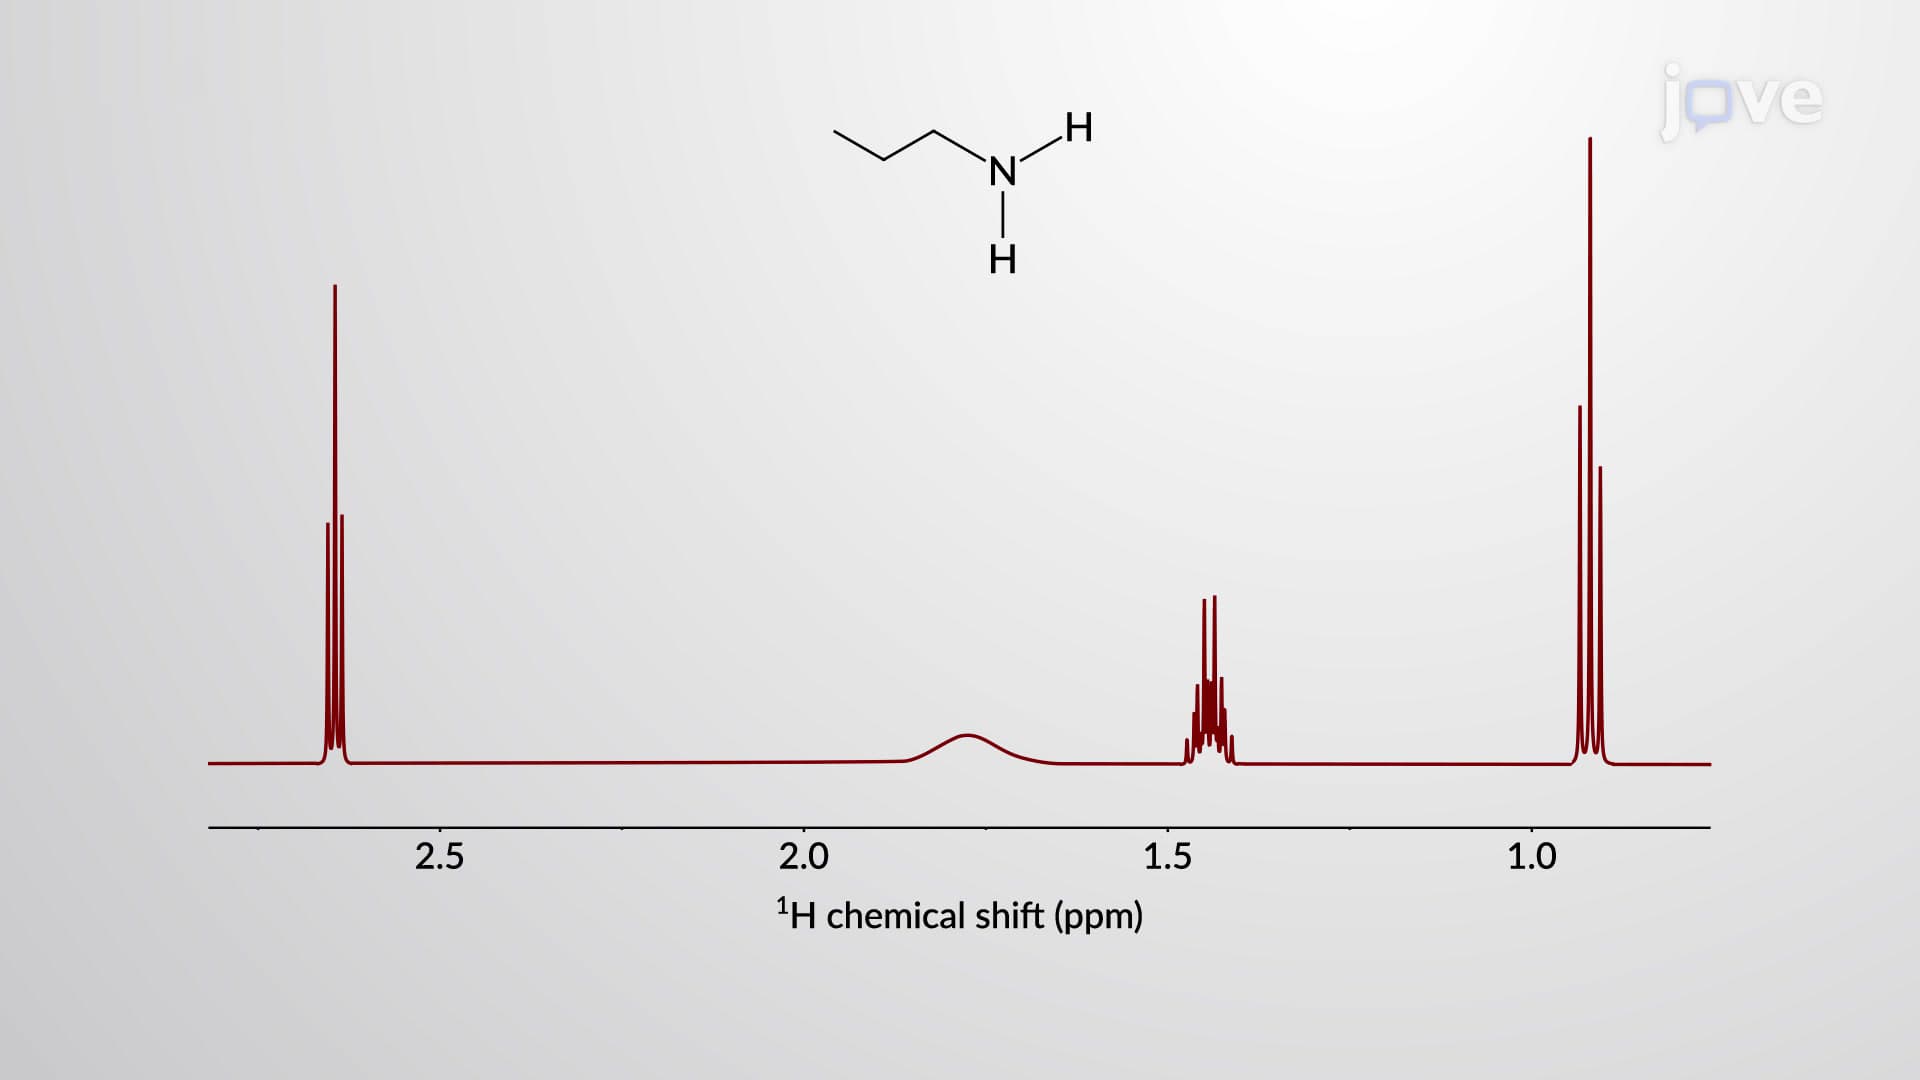
NMR Spectroscopy Of Amines
 00:58
¹H NMR Chemical Shift Equivalence: Enantiotopic and Diastereotopic Protons
In chiral compounds such as 2-butanol, replacing the methylene hydrogens at C3 produces a pair of...
 01:15
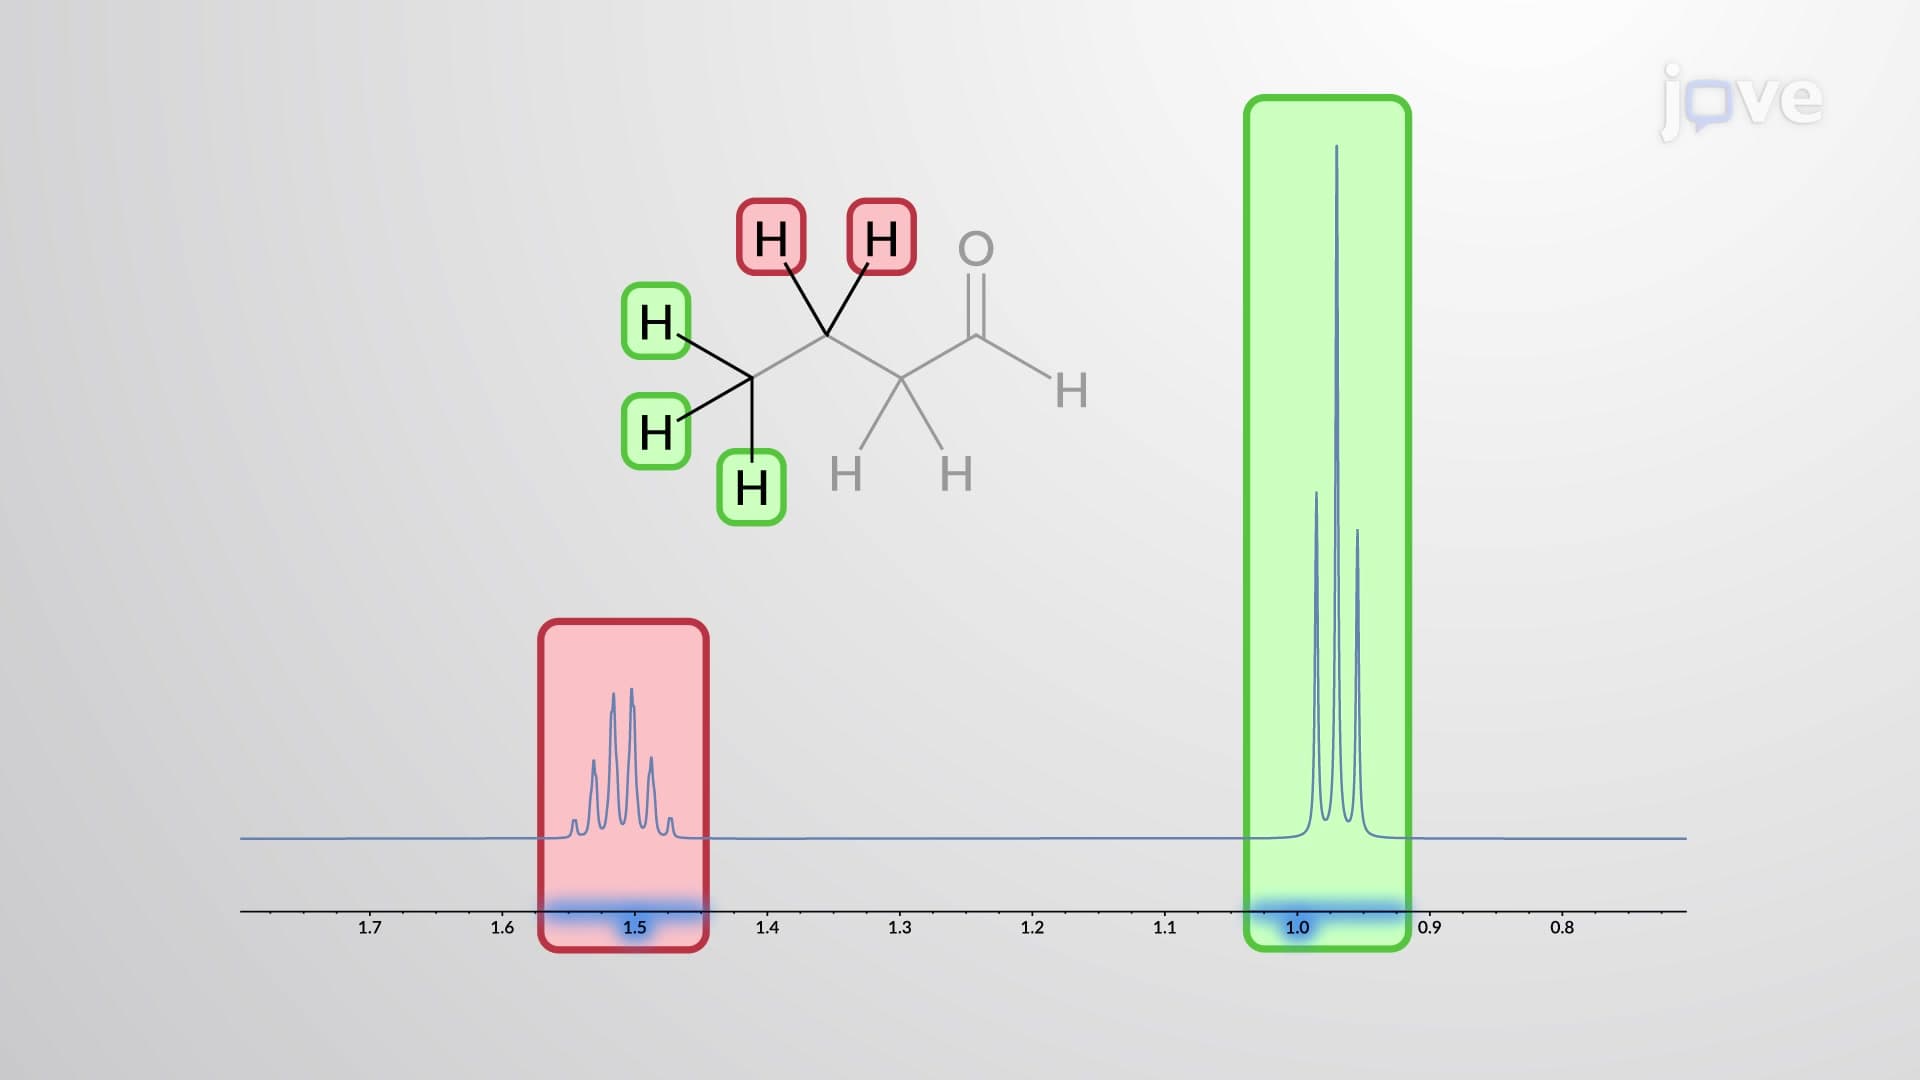
NMR Spectroscopy and Mass Spectrometry of Aldehydes and Ketones