Related Experiment Video
Updated: May 19, 2026
Unraveling Entropic Rate Acceleration Induced by Solvent Dynamics in Membrane Enzymes
Published on: January 16, 2016
Entropic and surprisingly small intramolecular polarization effects in the mechanism of cyclophilin A
Safieh Tork Ladani1, Donald Hamelberg
1Department of Chemistry and Center for Biotechnology and Drug Design, Georgia State University, Atlanta, Georgia 30302-4098, United States.

Abstract:
The precise catalytic mechanism of peptidyl-prolyl cis-trans isomerases (PPIases) has been elusive, despite many experimental and computational studies. The more than 5 orders of magnitude speedup achieved in catalysis by cyclophilin A (CypA) has been attributed to several factors, including substrate desolvation, enzyme dynamics, and preferential binding of the transition state. Here, we explore the conformational space of a substrate analogue of CypA using accelerated molecular dynamics, free in solution and in the active site of CypA, in order to probe its conformational interconversion during catalysis. We show that the undemanding exchange of the free substrate between β- and α-helical regions is lost in the active site of the enzyme, where it is mainly in the β-region. Our results suggest that the loss in conformational entropy at the transition state relative to the cis and trans states in the free substrate is decreased in the complex. This relative change in conformational entropy contributes favorable to the free energy of stabilizing the transition state by CypA. We also show that the ensuing intramolecular polarization, as a result of the loss in pseudo double bond character of the peptide bond at the transition state, contributes only about -1.0 kcal/mol to stabilizing the transition state. This relatively small contribution demonstrates that routinely used fixed charge classical force fields can reasonably describe these types of biological systems. Our results provide further insights into the mechanism of CypA, a member of a poorly understood family of enzymes that are central to many biological processes.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 02:29
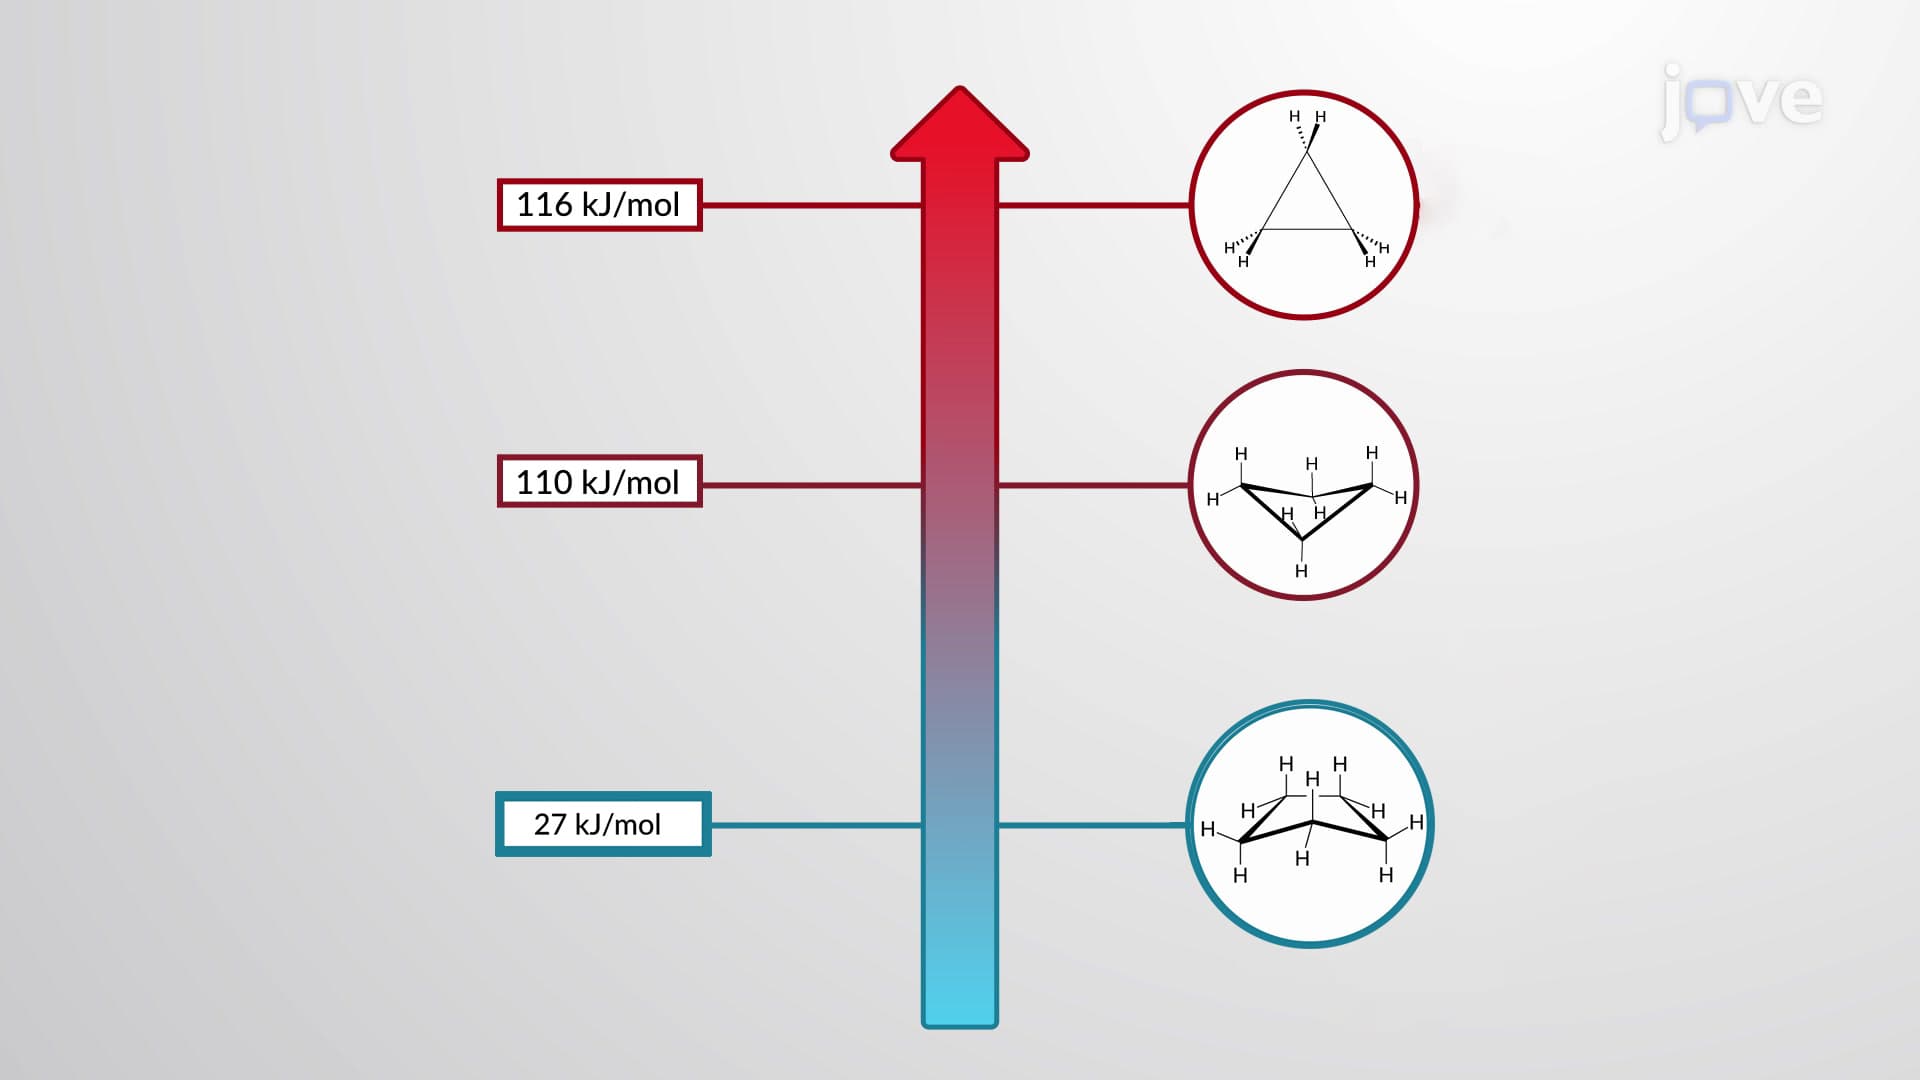
Conformations of Cycloalkanes
 01:22
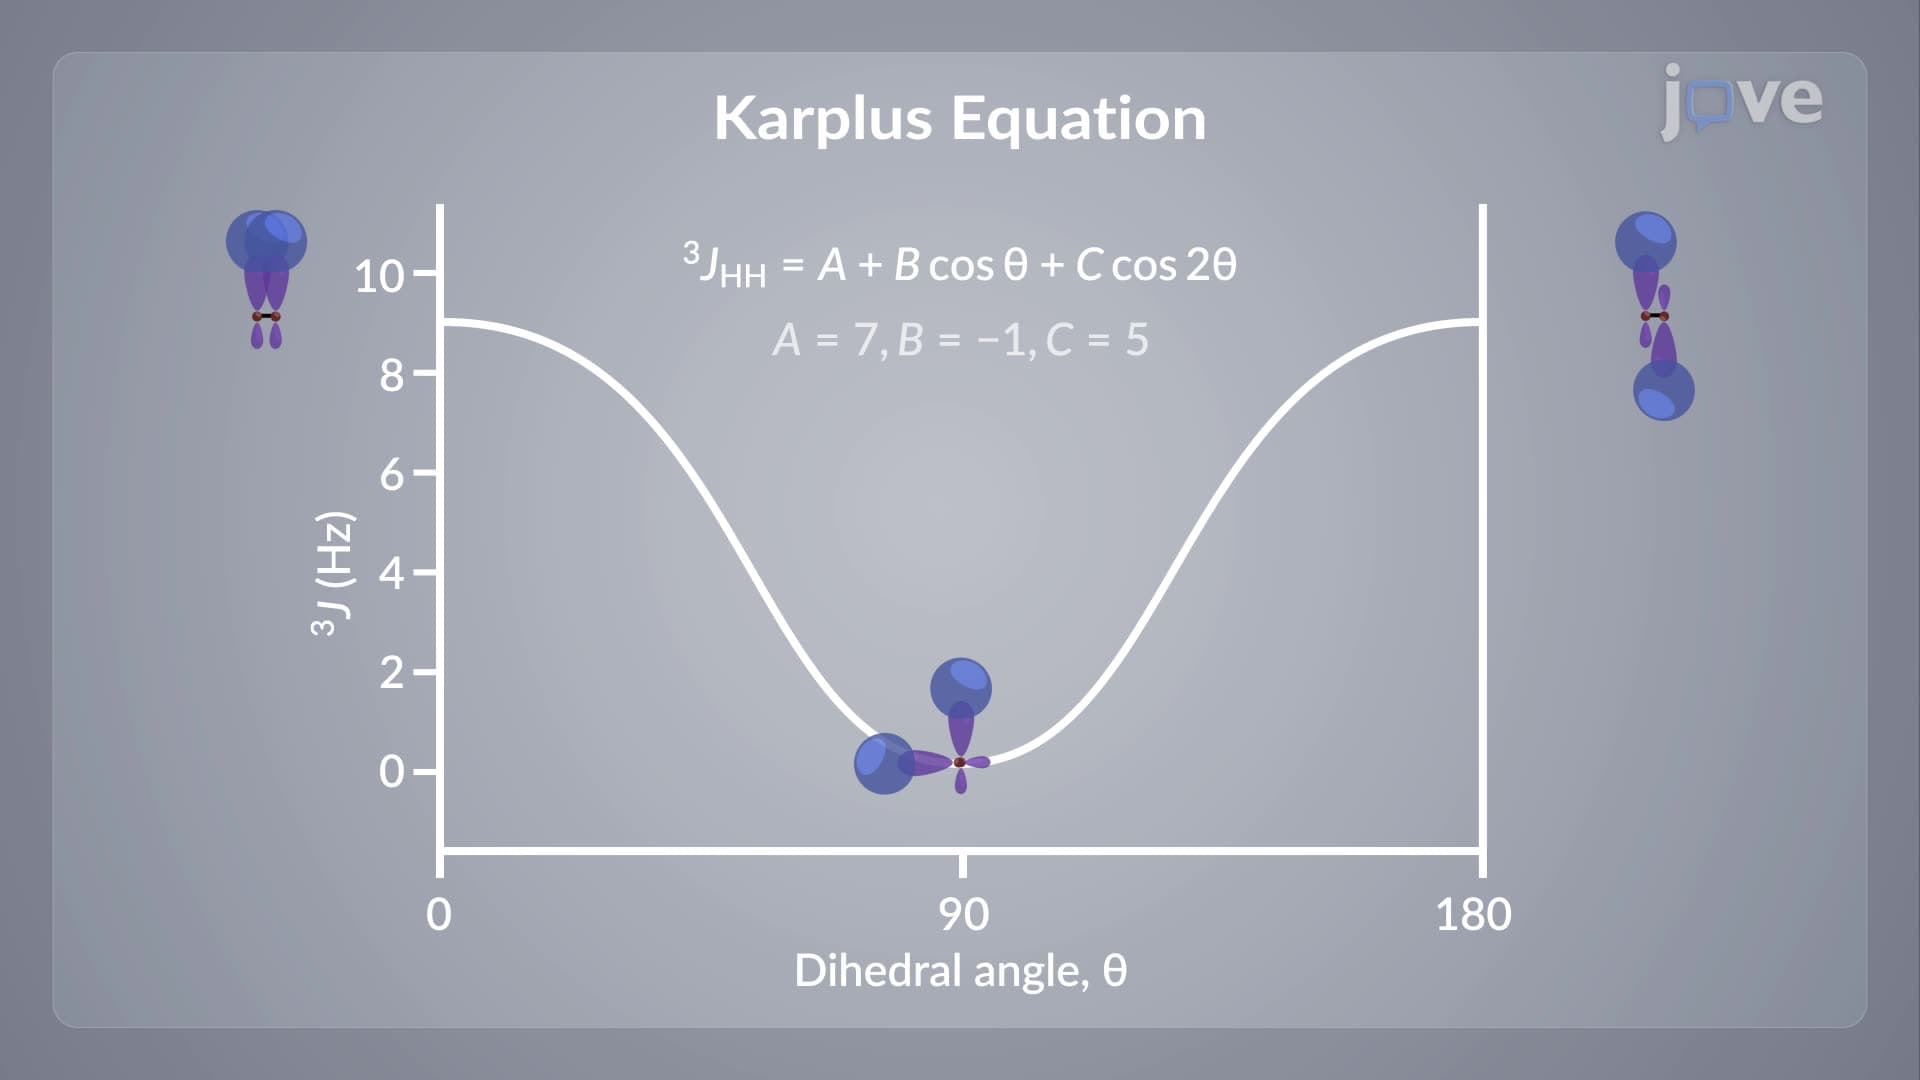
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:17
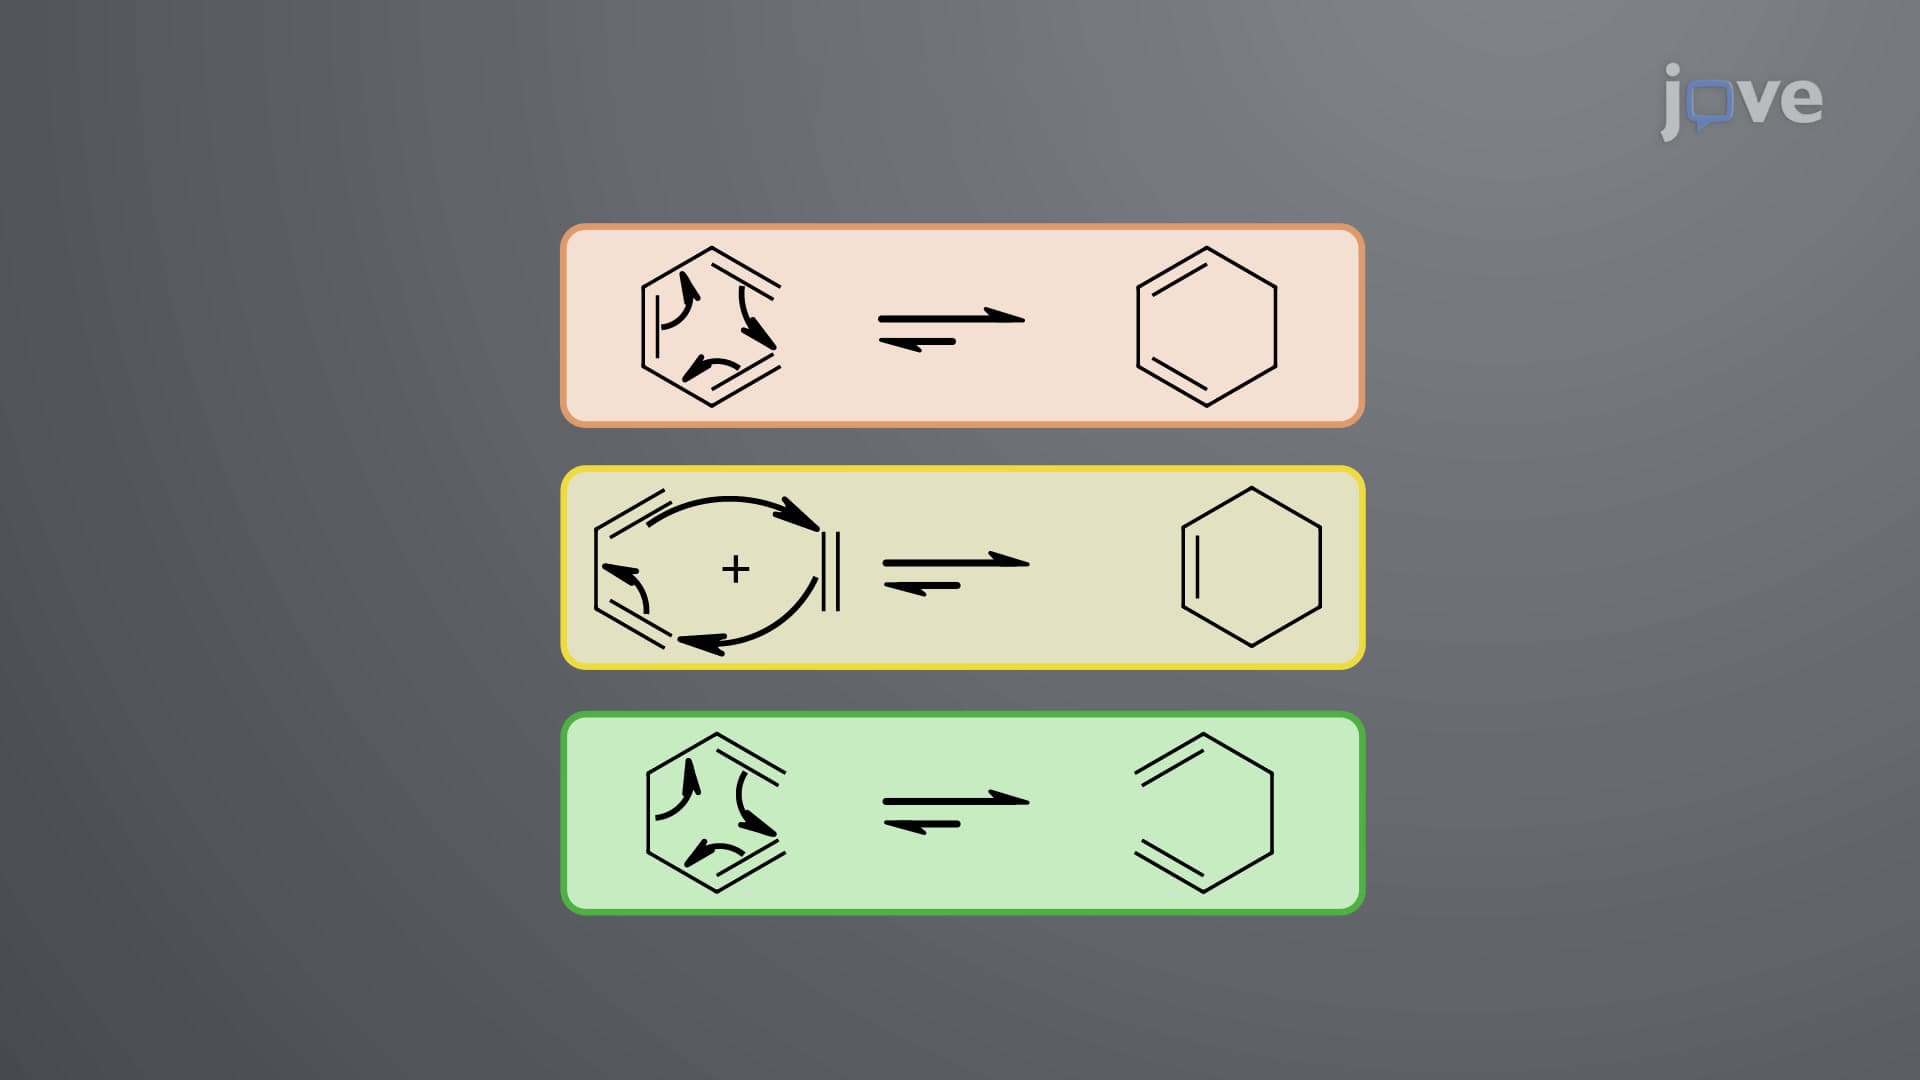
Pericyclic Reactions: Introduction
Pericyclic reactions can be classified into three categories: electrocyclic reactions, cycloaddition reactions, and sigmatropic rearrangements. Electrocyclic reactions and sigmatropic rearrangements are...
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...