

Molecular modeling studies of the structural, electronic, and UV absorption properties of benzophenone derivatives
Bianca A M Corrêa1, Arlan S Gonçalves, Alessandra M T de Souza
1CCS, Faculty of Pharmacy, ModMolQSAR, Federal University of Rio de Janeiro (UFRJ), Rio de Janeiro, RJ, Brazil.

Abstract:
Benzophenone derivatives (BZP), an important class of organic UV filters, are widely used in sunscreen products due to their ability to absorb in the UVA and UVB ranges. The structural, electronic, and spectral properties of BZP derivatives have been studied by density functional theory (DFT) and time-dependent DFT (TD-DFT) methods. DFT/B3LYP with the 6-31G(d) basis set is an accurate method for optimizing the geometry of BZPs. The absorption maxima obtained from the TD-DFT calculations in a vacuum were in agreement with the experimental absorption bands and showed that the main electronic transitions in the UVA/UVB range present π → π* character, the major transition being HOMO → LUMO. The oscillator strength seems to increase in the presence of disubstitution at the para position. For protic substituents, the position appears to be related to the absorption band. Absorption in the UVB range occurs in the presence of para substitution, whereas ortho substitution leads to absorption in the UVA spectral region. The obtained results provide some features for BZP derivatives that can be useful for customizing absorption properties (wavelengths and intensities) and designing new BZP derivatives as sunscreens.

Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:07
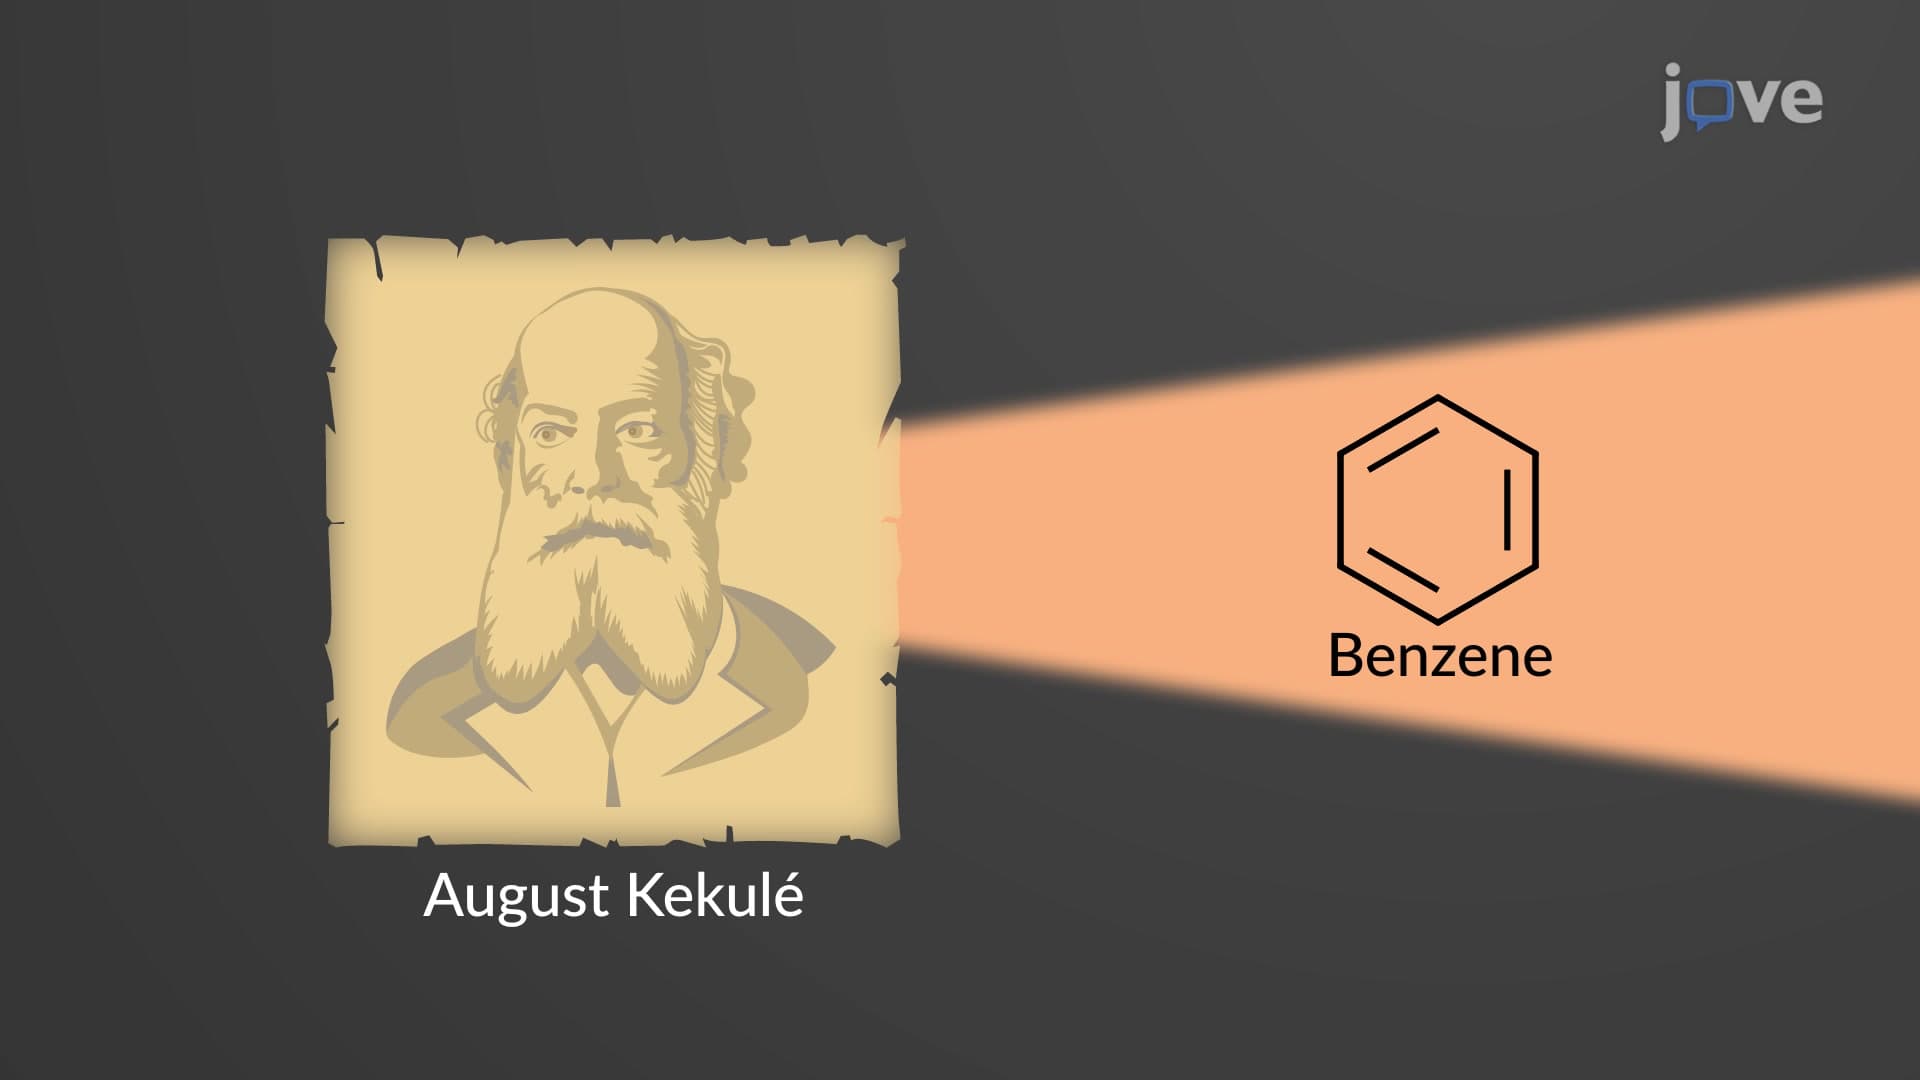
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:37
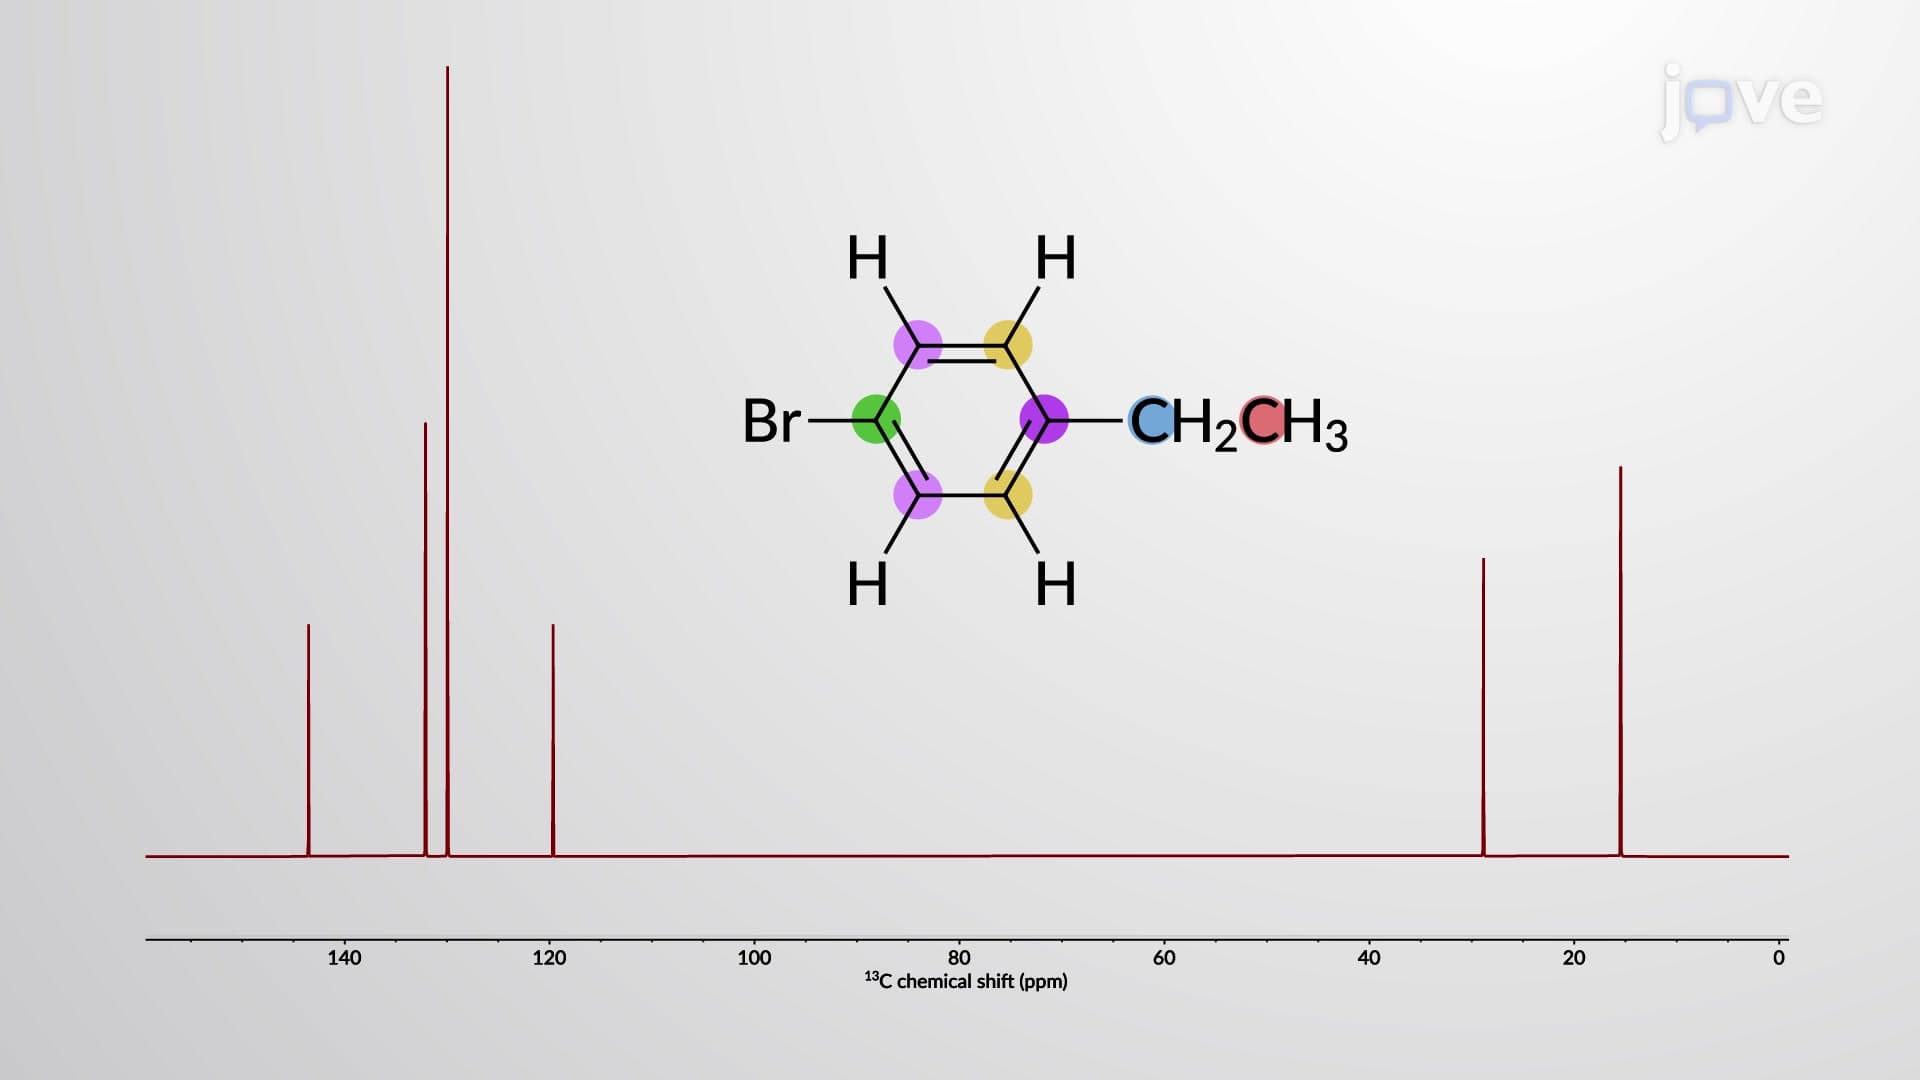
NMR Spectroscopy of Benzene Derivatives
 03:51
Molecular Orbital Theory II
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:18
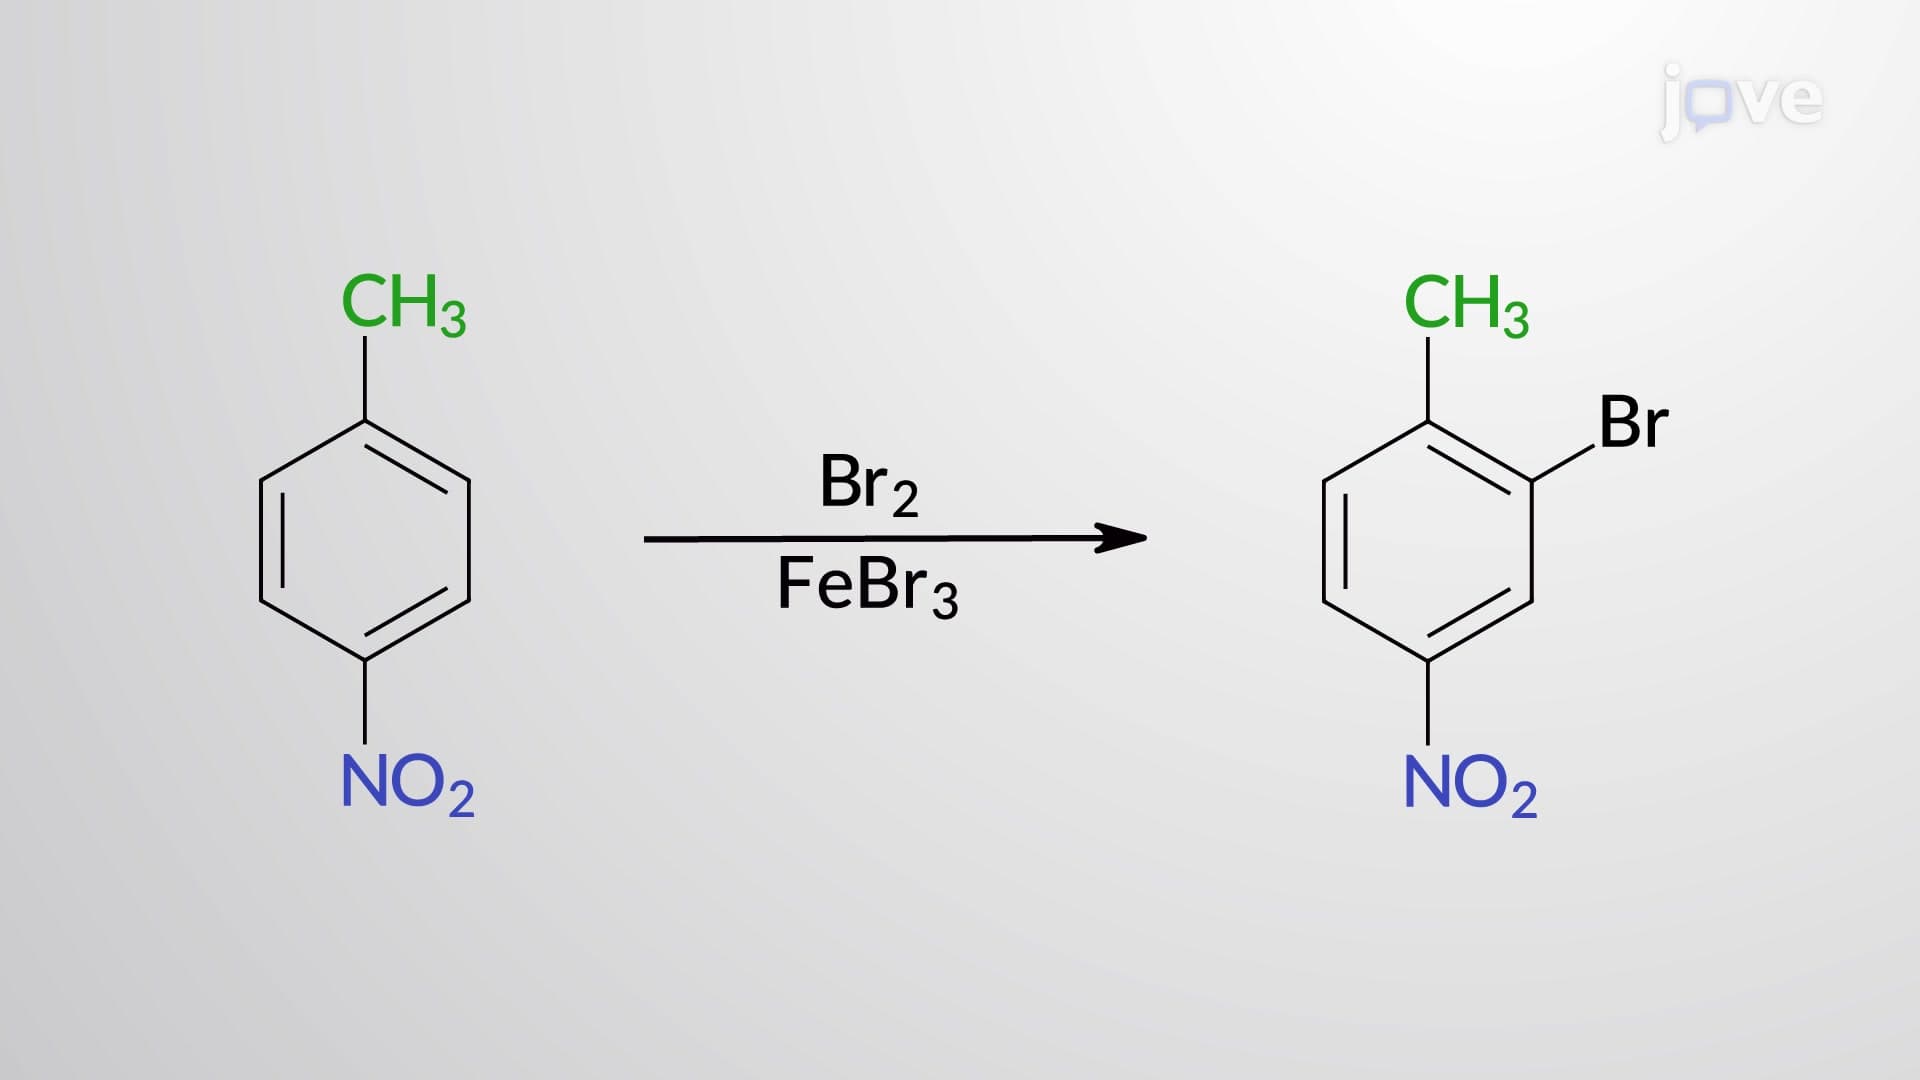
Directing and Steric Effects in Disubstituted Benzene Derivatives