Related Experiment Video
Updated: May 18, 2026

In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
CFTR, mucins, and mucus obstruction in cystic fibrosis
Silvia M Kreda1, C William Davis, Mary Callaghan Rose
1Cystic Fibrosis/Pulmonary Research and Treatment Center, University of North Carolina, Chapel Hill, NC 27517-7248, USA.

Abstract:
Mucus pathology in cystic fibrosis (CF) has been known for as long as the disease has been recognized and is sometimes called mucoviscidosis. The disease is marked by mucus hyperproduction and plugging in many organs, which are usually most fatal in the airways of CF patients, once the problem of meconium ileus at birth is resolved. After the CF gene, CFTR, was cloned and its protein product identified as a cAMP-regulated Cl(-) channel, causal mechanisms underlying the strong mucus phenotype of the disease became obscure. Here we focus on mucin genes and polymeric mucin glycoproteins, examining their regulation and potential relationships to a dysfunctional cystic fibrosis transmembrane conductance regulator (CFTR). Detailed examination of CFTR expression in organs and different cell types indicates that changes in CFTR expression do not always correlate with the severity of CF disease or mucus accumulation. Thus, the mucus hyperproduction that typifies CF does not appear to be a direct cause of a defective CFTR but, rather, to be a downstream consequence. In organs like the lung, up-regulation of mucin gene expression by inflammation results from chronic infection; however, in other instances and organs, the inflammation may have a non-infectious origin. The mucus plugging phenotype of the β-subunit of the epithelial Na(+) channel (βENaC)-overexpressing mouse is proving to be an archetypal example of this kind of inflammation, with a dehydrated airway surface/concentrated mucus gel apparently providing the inflammatory stimulus. Data indicate that the luminal HCO(3)(-) deficiency recently described for CF epithelia may also provide such a stimulus, perhaps by causing a mal-maturation of mucins as they are released onto luminal surfaces. In any event, the path between CFTR dysfunction and mucus hyperproduction has proven tortuous, and its unraveling continues to offer its own twists and turns, along with fascinating glimpses into biology.
Insights
Cystic fibrosis (CF) mucus hyperproduction is a downstream consequence of defective CFTR, not a direct cause. Inflammation, infection, or ion channel defects may trigger this complex mucus pathology.
Area of Science:
- Pulmonary Medicine
- Genetics
- Cell Biology
Background:
- Mucus pathology, termed mucoviscidosis, is a hallmark of cystic fibrosis (CF).
- This pathology involves mucus hyperproduction and plugging, primarily affecting CF patient airways.
- The cloning of the CFTR gene and identification of its protein product as a Cl(-) channel obscured the direct mechanisms of CF mucus phenotype.
Purpose of the Study:
- To investigate mucin gene regulation and its relationship with a dysfunctional CFTR.
- To clarify the causal relationship between CFTR dysfunction and mucus hyperproduction in CF.
Main Methods:
- Examined CFTR expression across various organs and cell types.
- Analyzed mucin gene regulation in response to inflammation and potential non-infectious stimuli.
- Utilized a mouse model overexpressing the beta-subunit of the epithelial sodium channel (βENaC).
Main Results:
- CFTR expression levels do not consistently correlate with CF disease severity or mucus accumulation.
- Mucus hyperproduction in CF appears to be a downstream consequence of CFTR defects, not a direct cause.
- Inflammation, triggered by infection or non-infectious factors like airway dehydration or luminal HCO(3)(-) deficiency, up-regulates mucin gene expression.
Conclusions:
- The link between CFTR dysfunction and CF mucus pathology is complex and multifactorial.
- Inflammatory and ion channel dysregulation play significant roles in driving mucus hyperproduction.
- Understanding these pathways offers insights into CF pathogenesis and potential therapeutic targets.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:24
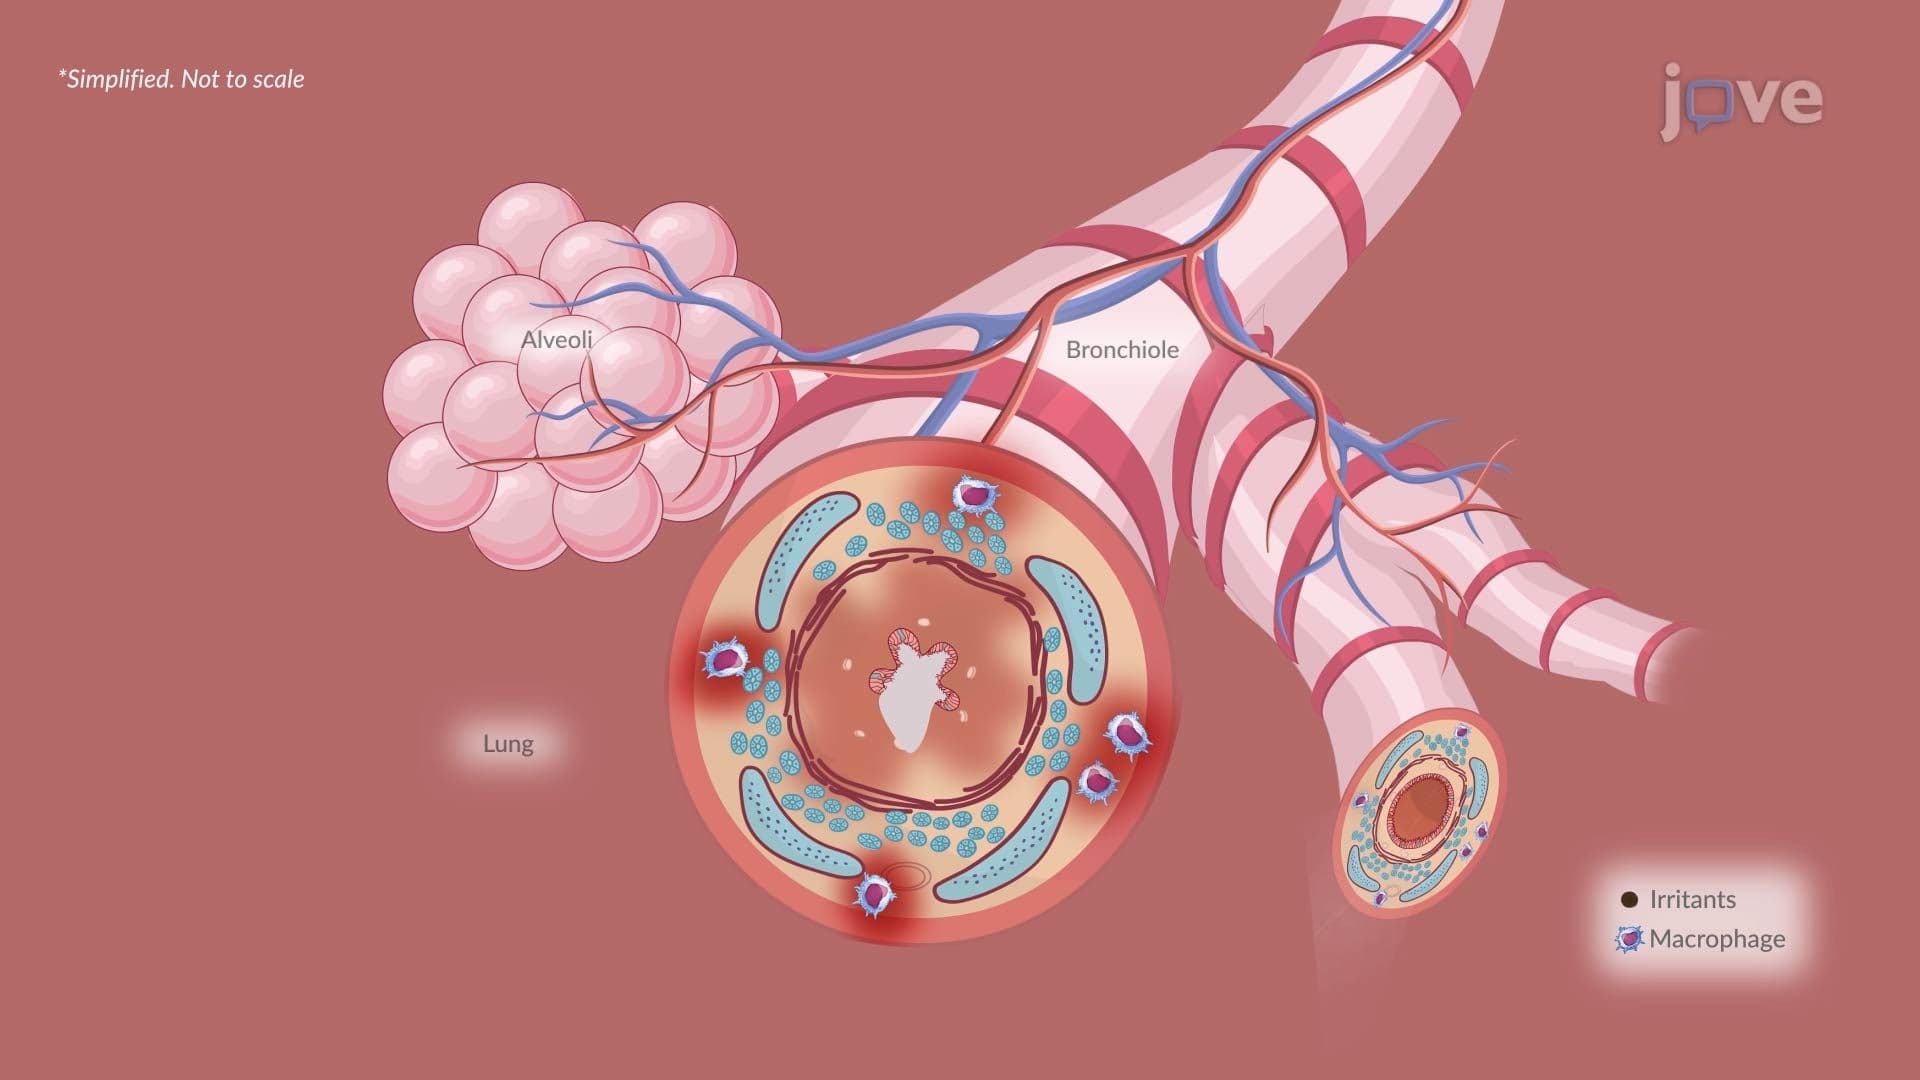
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features
 01:16
Drugs Used in Upper Respiratory Disorders: Overview
Antihistamines (e.g., Benadryl) block histamines from binding. Histamines are chemicals released during an allergic reaction in the body. As a...
 01:21
Drugs for Treatment of Constipation-Predominant IBS
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation