Related Experiment Video
Updated: May 18, 2026

Monitoring Cell-to-cell Transmission of Prion-like Protein Aggregates in Drosophila Melanogaster
Published on: March 12, 2018
Failure of prion protein oxidative folding guides the formation of toxic transmembrane forms
Silvia Lisa1, Beatriz Domingo, Javier Martínez
1Instituto Química-Física Rocasolano, Consejo Superior de Investigaciones Científicas, Serrano 119, 28006 Madrid, Spain.

Abstract:
The mechanism by which pathogenic mutations in the globular domain of the cellular prion protein (PrP(C)) increase the likelihood of misfolding and predispose to diseases is not yet known. Differences in the evidences provided by structural and metabolic studies of these mutants suggest that in vivo folding could be playing an essential role in their pathogenesis. To address this role, here we use the single or combined M206S and M213S artificial mutants causing labile folds and express them in cells. We find that these mutants are highly toxic, fold as transmembrane PrP, and lack the intramolecular disulfide bond. When the mutations are placed in a chain with impeded transmembrane PrP formation, toxicity is rescued. These results suggest that oxidative folding impairment, as on aging, can be fundamental for the genesis of intracellular neurotoxic intermediates key in prion neurodegenerations.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 03:00
Molecular Chaperones and Protein Folding
The...
03:00Molecular Chaperones and Protein Folding
The...
 02:11
Proteins: From Genes to Degradation
Transcription is the synthesis of RNA molecules by RNA...
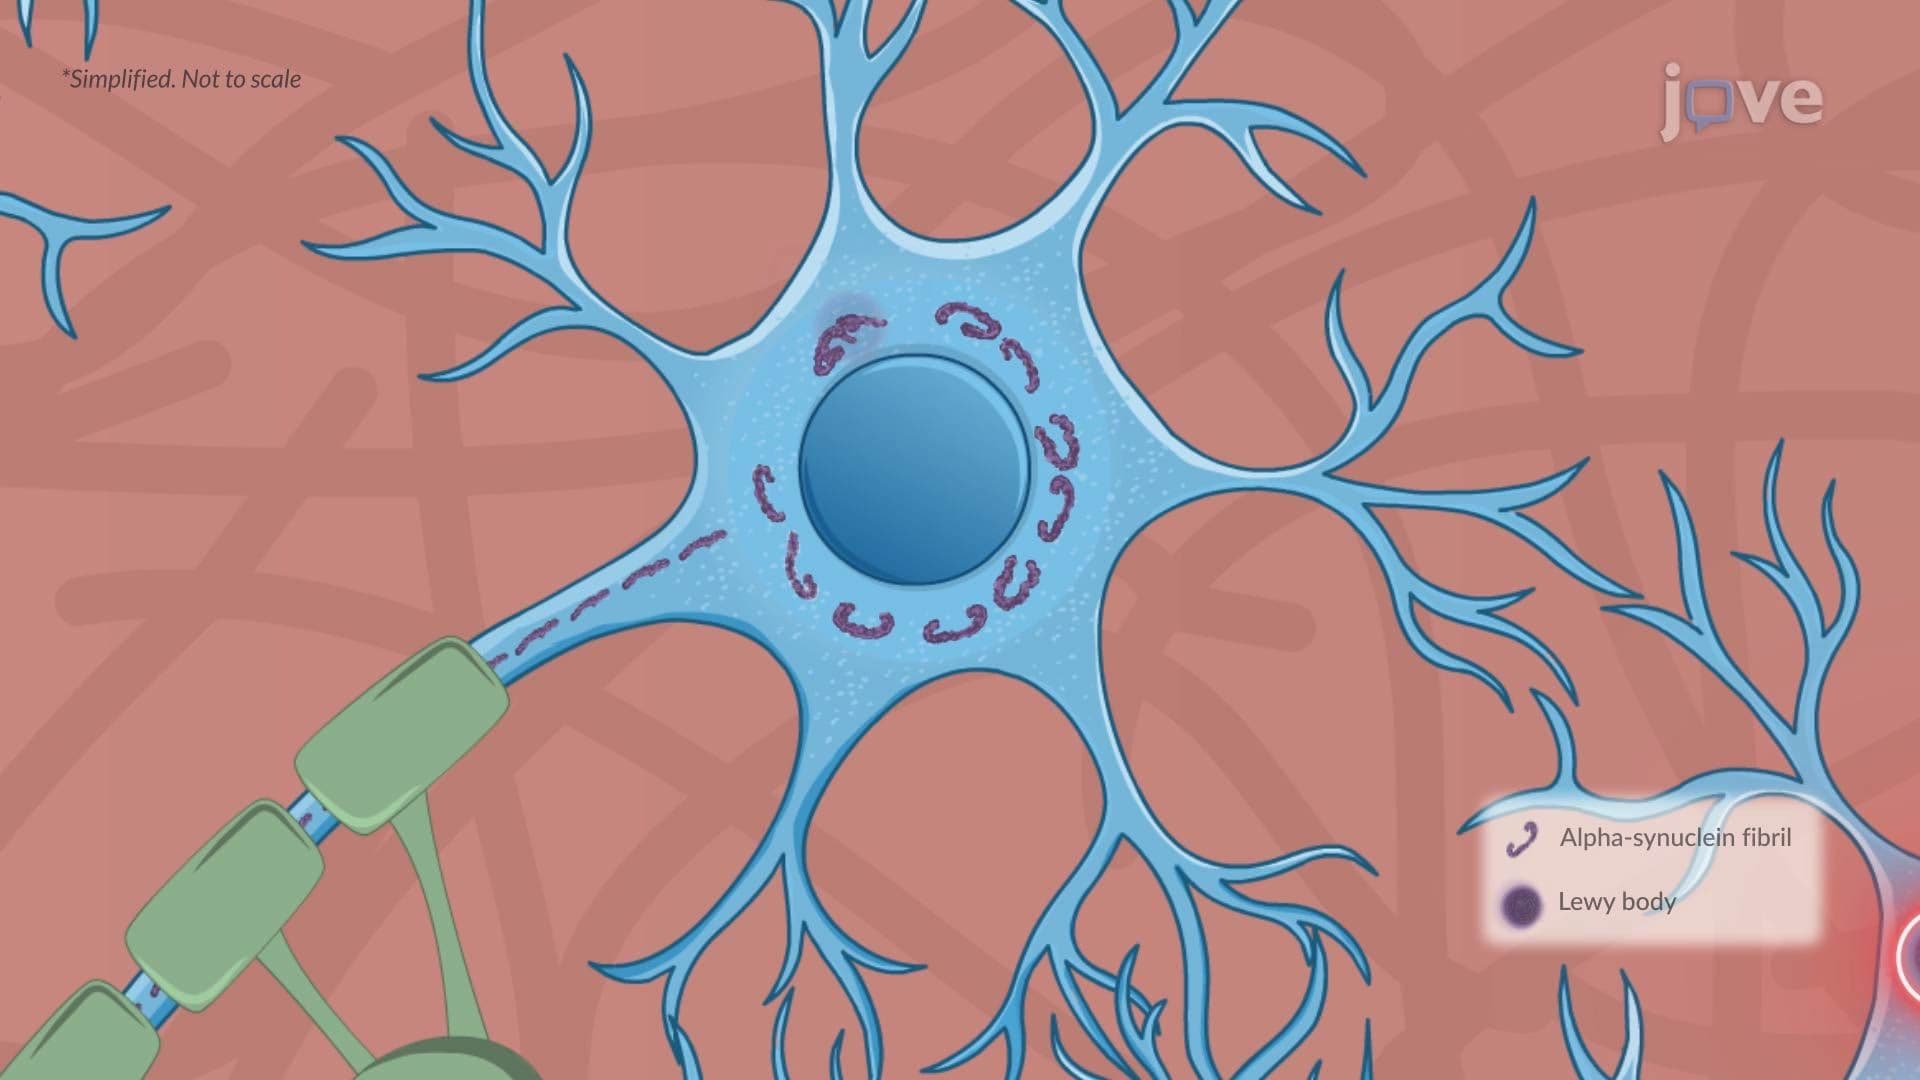 01:24
Parkinson Disease ll: Pathophysiology