Related Experiment Video
Updated: May 18, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
An ab initio based full-dimensional global potential energy surface for FH2O(X2A') and dynamics for the F + H2O → HF
Jun Li1, Richard Dawes, Hua Guo
1Department of Chemistry and Chemical Biology, University of New Mexico, Albuquerque, New Mexico 87131, USA.

Abstract:
A global potential energy surface (PES) for the ground electronic state of FH(2)O is constructed based on more than 30,000 ab initio points at the multi-reference configuration interaction level. The PES features a pre-reaction van der Waals well and two post-reaction hydrogen-bonded complexes, as well as a "reactant-like" transition state with a classical barrier of 3.8 kcal/mol. The adiabatic F + H(2)O → HF + OH reaction dynamics on this PES was investigated using a standard quasi-classical trajectory method. In agreement with experiment, the HF product contains significant vibrational excitation with limited rotational excitation, while the OH product is internally cold, reflecting its spectator role in the reaction. The products are primarily scattered in the backward direction, consistent with a direct abstraction mechanism.

Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:45
Valence Bond Theory
 03:35
Hybridization of Atomic Orbitals II
 01:13
Force and Potential Energy in One Dimension
 02:38
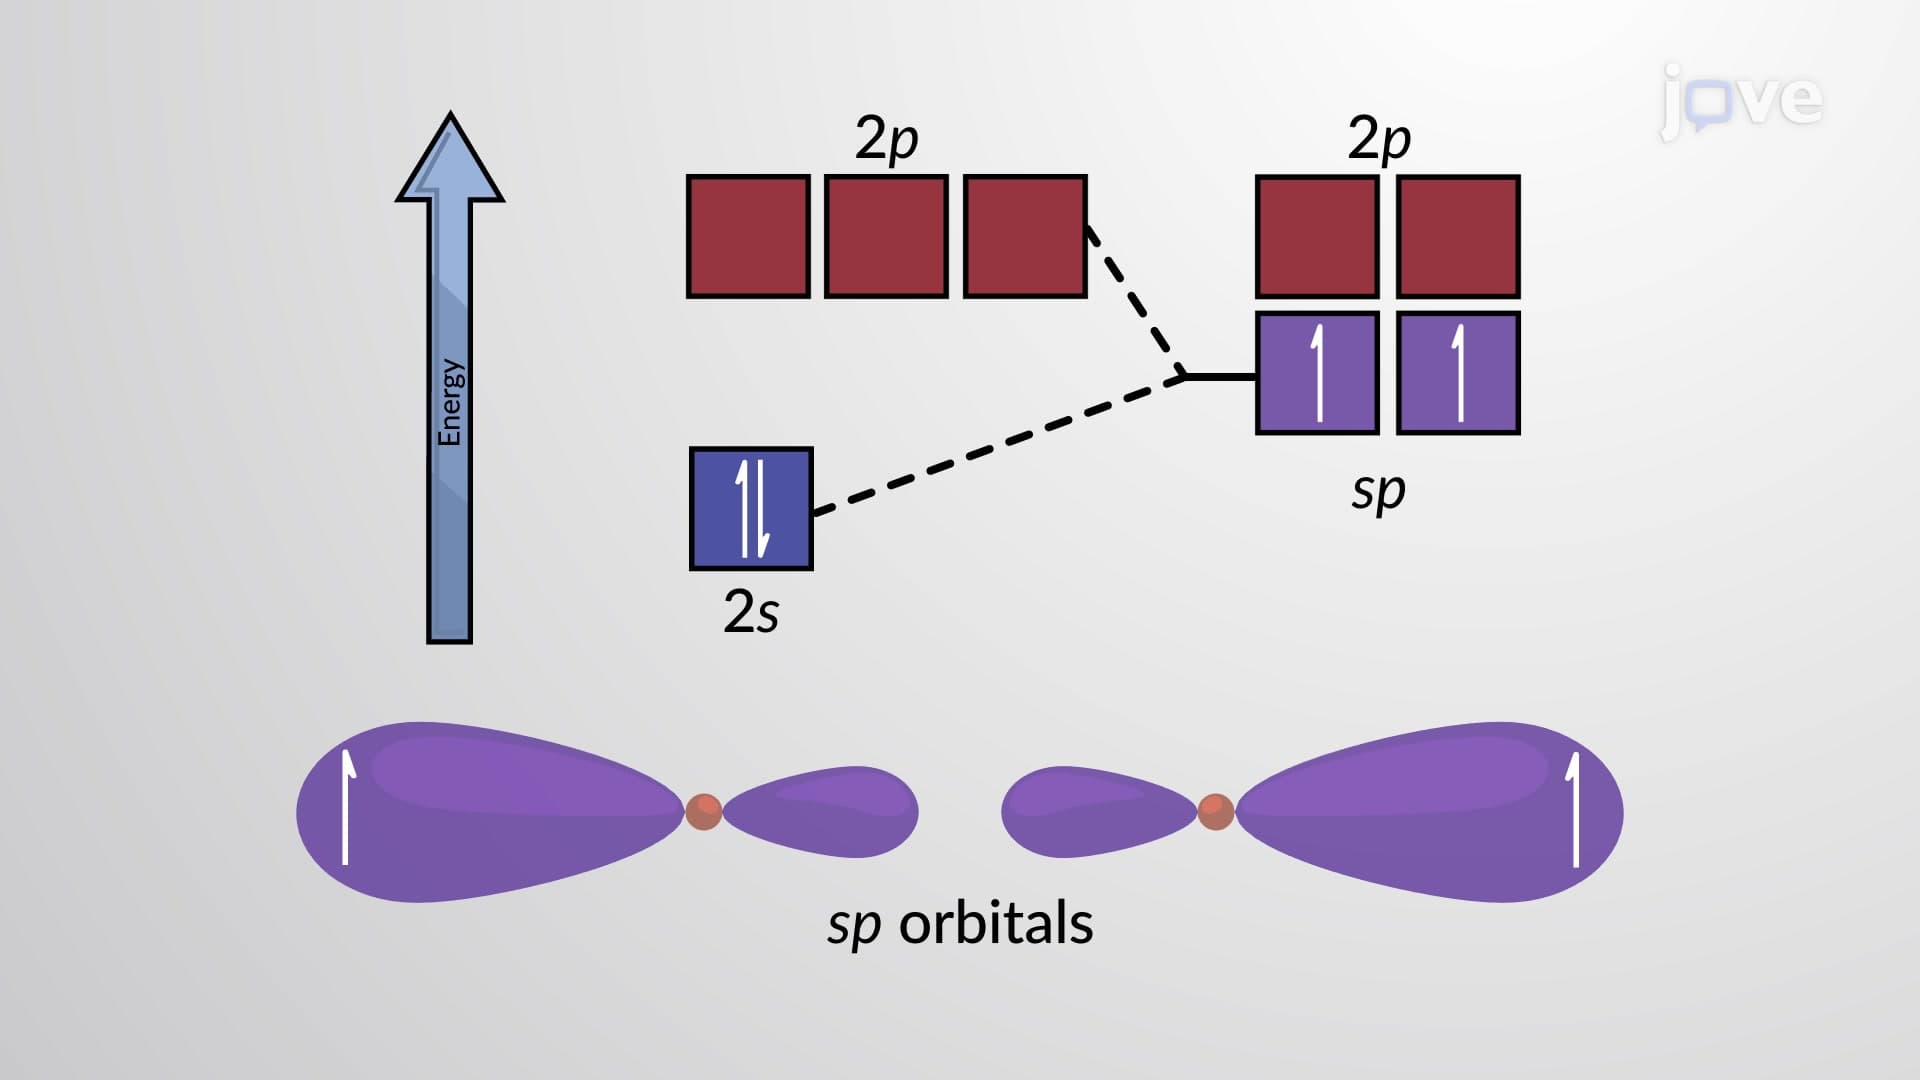
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:51
Molecular Orbital Theory II