Rotamer decomposition and protein dynamics: efficiently analyzing dihedral populations from molecular dynamics
Related Experiment Video
Updated: May 18, 2026
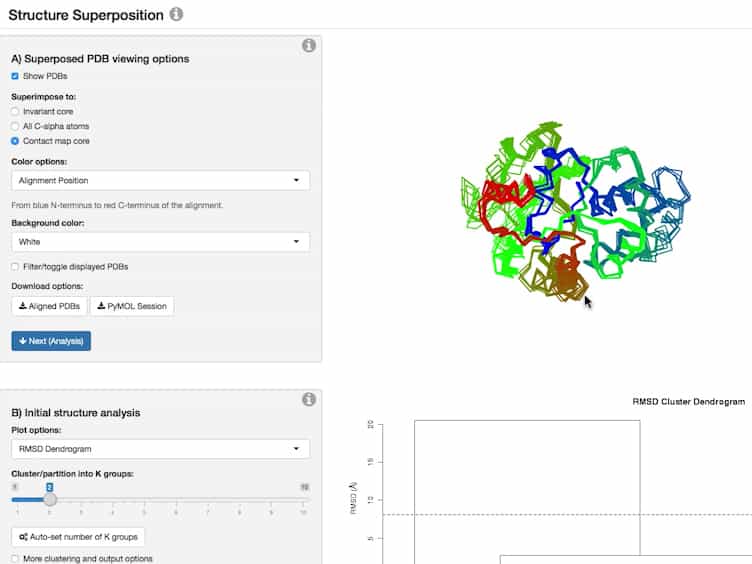
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Hiroshi Watanabe1, Marcus Elstner, Thomas Steinbrecher
1Inst. f. Phys. Chem., KIT, Kaiserstr. 12, 76131 Karlsruhe, Germany.

Abstract:
Molecular mechanics methods have matured into powerful methods to understand the dynamics and flexibility of macromolecules and especially proteins. As multinanosecond to microsecond length molecular dynamics (MD) simulations become commonplace, advanced analysis tools are required to generate scientifically useful information from large amounts of data. Some of the key degrees of freedom to understand protein flexibility and dynamics are the amino acid residue side chain dihedral angles. In this work, we present an easily automated way to summarize and understand the relevant dihedral populations. A tremendous reduction in complexity is achieved by describing dihedral timeseries in terms of histograms decomposed into Gaussians. Using the familiar and widely studied protein lysozyme, it is demonstrated that our approach captures essential properties of protein structure and dynamics. A simple classification scheme is proposed that indicates the rotational state population for each dihedral angle of interest and allows a decision if a given side chain or peptide backbone fragment remains rigid during the course of an MD simulation, adopts a converged distribution between conformational substates or has not reached convergence yet.
More Related Videos
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
