Related Experiment Video
Updated: May 17, 2026

Laboratory Production of Biofuels and Biochemicals from a Rapeseed Oil through Catalytic Cracking Conversion
Published on: September 2, 2016
Ab initio simulations reveal that reaction dynamics strongly affect product selectivity for the cracking of alkanes
Paul M Zimmerman1, Diana C Tranca, Joseph Gomes
1Department of Chemical and Biomolecular Engineering, University of California Berkeley, California 94720-1462, USA.

Abstract:
Product selectivity of alkane cracking catalysis in the H-MFI zeolite is investigated using both static and dynamic first-principles quantum mechanics/molecular mechanics simulations. These simulations account for the electrostatic- and shape-selective interactions in the zeolite and provide enthalpic barriers that are closely comparable to experiment. Cracking transition states for n-pentane lead to a metastable intermediate (a local minimum with relatively small barriers to escape to deeper minima) where the proton is shared between two hydrocarbon fragments. The zeolite strongly stabilizes these carbocations compared to the gas phase, and the conversion of this intermediate to more stable species determines the product selectivity. Static reaction pathways on the potential energy surface starting from the metastable intermediate include a variety of possible conversions into more stable products. One-picosecond quasiclassical trajectory simulations performed at 773 K indicate that dynamic paths are substantially more diverse than the potential energy paths. Vibrational motion that is dynamically sampled after the cracking transition state causes spilling of the metastable intermediate into a variety of different products. A nearly 10-fold change in the branching ratio between C2/C3 cracking channels is found upon inclusion of post-transition-state dynamics, relative to static electronic structure calculations. Agreement with experiment is improved by the same factor. Because dynamical effects occur soon after passing through the rate-limiting transition state, it is the dynamics, and not only the potential energy barriers, that determine the catalytic selectivity. This study suggests that selectivity in zeolite catalysis is determined by high temperature pathways that differ significantly from 0 K potential surfaces.
More Related Videos
Related Concept Videos
 02:17
Regioselectivity of Electrophilic Additions to Alkenes: Markovnikov's Rule
The hydrohalogenation of an unsymmetrical alkene can yield two haloalkane products, depending on which vinylic carbon takes up the halogen. However, one product usually predominates, where hydrogen adds to the vinylic carbon bearing the...
 01:20
Radical Reactivity: Concentration Effects
 02:13
Reduction of Alkenes: Catalytic Hydrogenation
Metals like palladium, platinum, and nickel are commonly used in their solid forms — fine powder on an inert surface. As these catalysts remain insoluble in the reaction mixture, they are referred to as heterogeneous catalysts.
The hydrogenation process takes place on the surface of...
 01:23
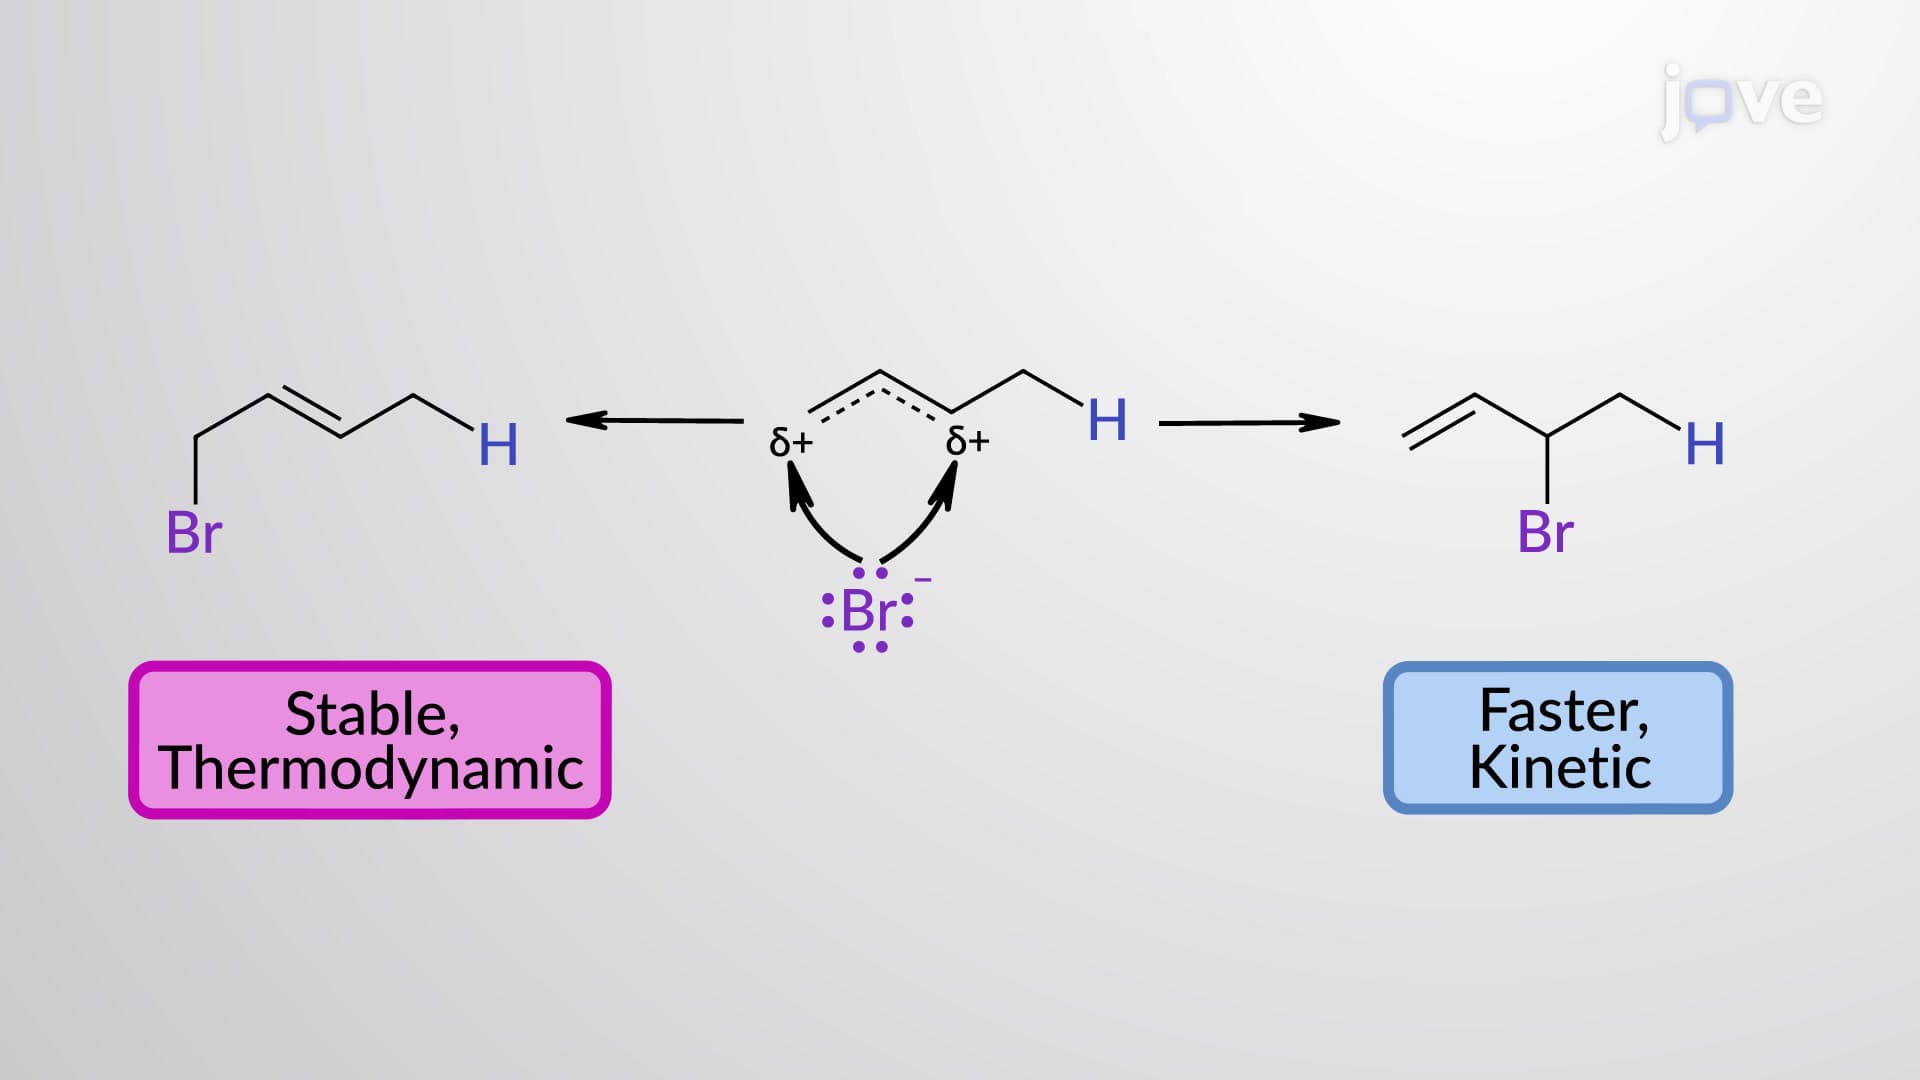
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
 02:24
Reduction of Alkynes to cis-Alkenes: Catalytic Hydrogenation
Like alkenes, alkynes can be reduced to alkanes in the presence of transition metal catalysts such as Pt, Pd, or Ni. The reaction involves two sequential syn additions of hydrogen via a cis-alkene intermediate.
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration