Related Experiment Video
Updated: May 16, 2026
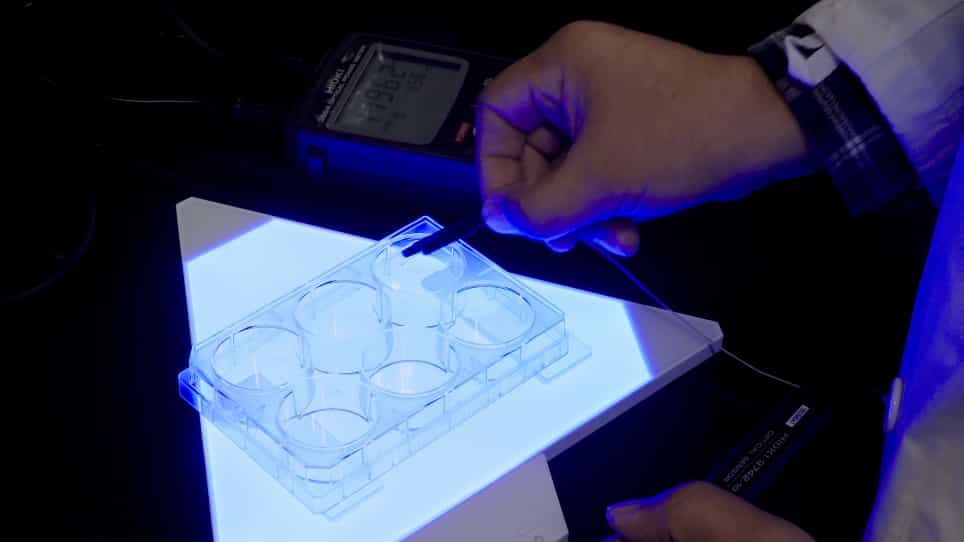

Optogenetic Phase Transition of TDP-43 in Spinal Motor Neurons of Zebrafish Larvae
Published on: February 25, 2022
Accelerated disease onset with stabilized familial amyotrophic lateral sclerosis (ALS)-linked mutant TDP-43 proteins
Shoji Watanabe1, Kumi Kaneko, Koji Yamanaka
1Laboratory for Motor Neuron Disease, RIKEN Brain Science Institute, Wako, Saitama 351-0198, Japan.

Abstract:
Abnormal protein accumulation is a pathological hallmark of neurodegenerative diseases, including accumulation of TAR DNA-binding protein 43 (TDP-43) in amyotrophic lateral sclerosis (ALS). Dominant mutations in the TDP-43 gene are causative for familial ALS; however, the relationship between mutant protein biochemical phenotypes and disease course and their significance to disease pathomechanism are not known. Here, we found that longer half-lives of mutant proteins correlated with accelerated disease onset. Based on our findings, we established a cell model in which chronic stabilization of wild-type TDP-43 protein provoked cytotoxicity and recapitulated pathogenic protein cleavage and insolubility to the detergent Sarkosyl, TDP-43 properties that have been observed in sporadic ALS lesions. Furthermore, these cells showed proteasomal impairment and dysregulation of their own mRNA levels. These results suggest that chronically increased stability of mutant or wild-type TDP-43 proteins results in a gain of toxicity through abnormal proteostasis.
Related Concept Videos
 01:29
Alzheimer Disease l: Introduction
 01:23
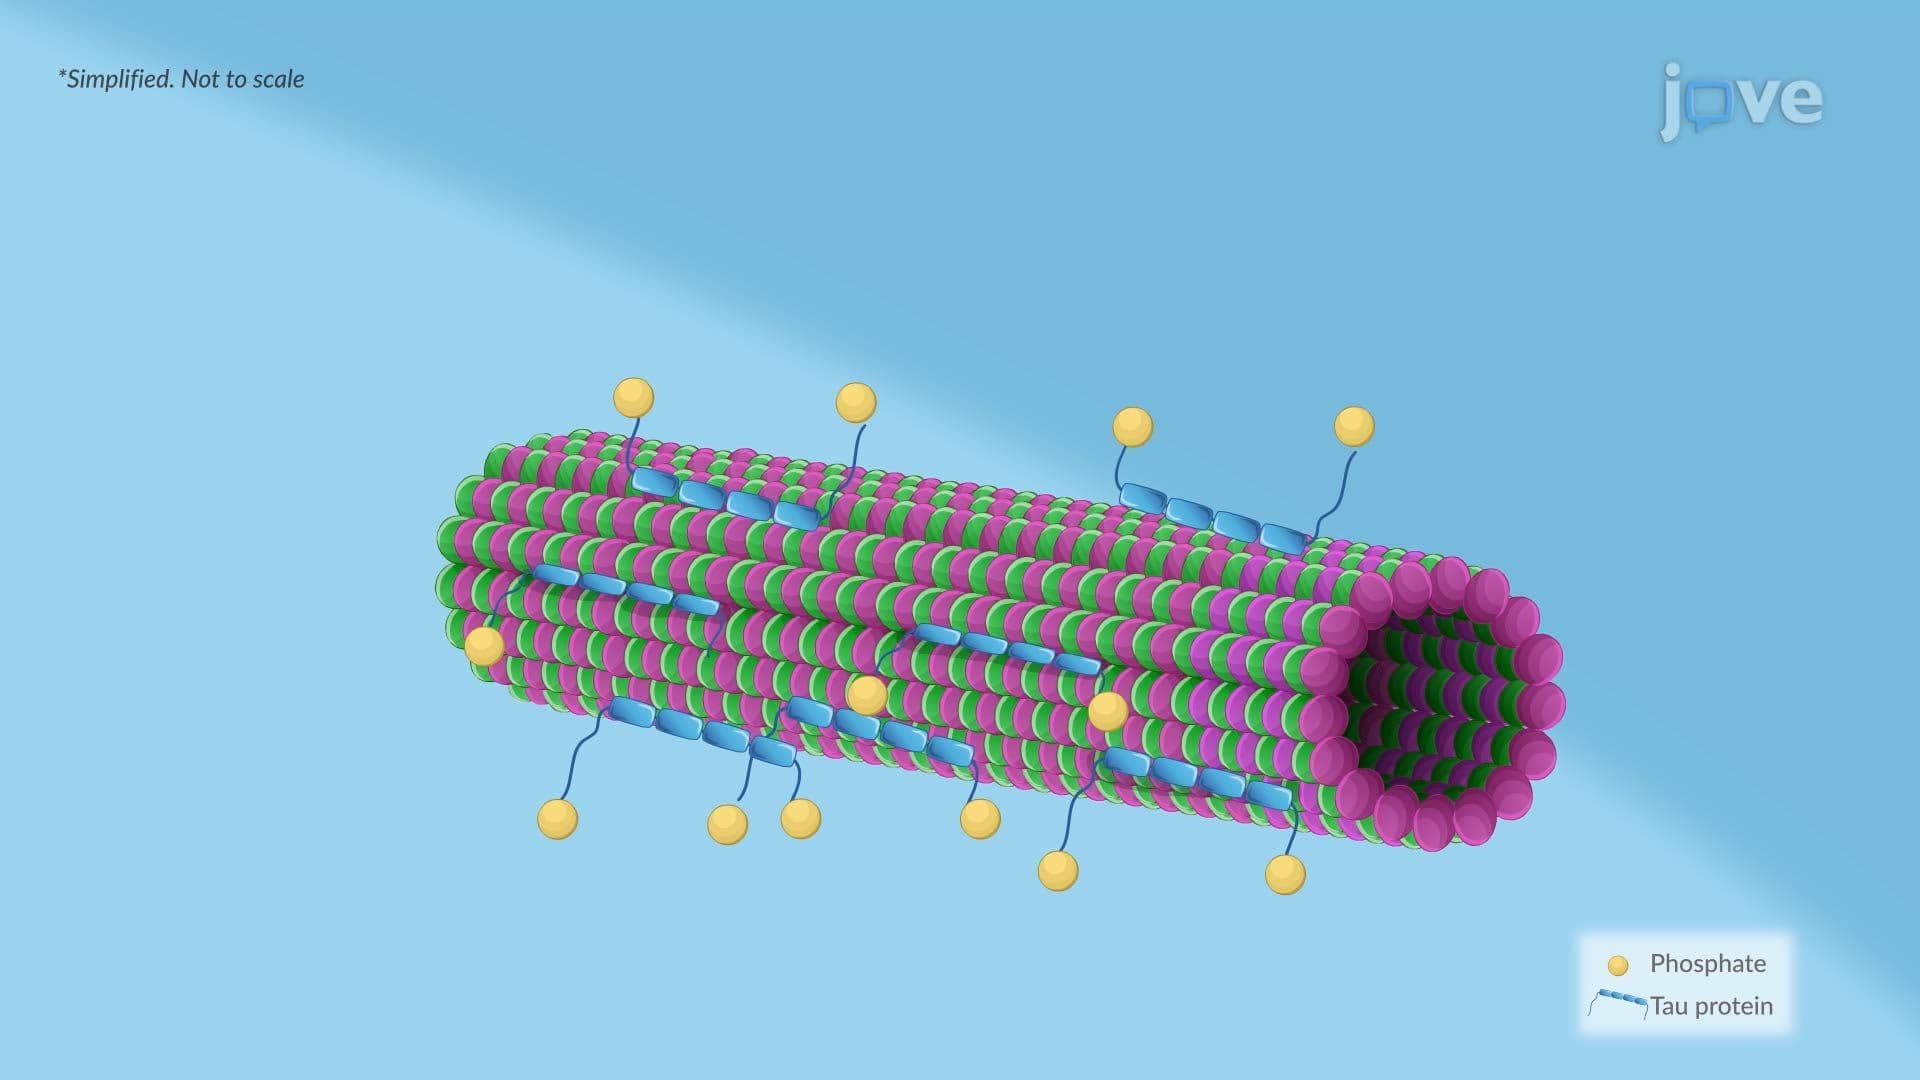
Alzheimer Disease ll: Pathophysiology
 01:21
Huntington Disease l: Introduction
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
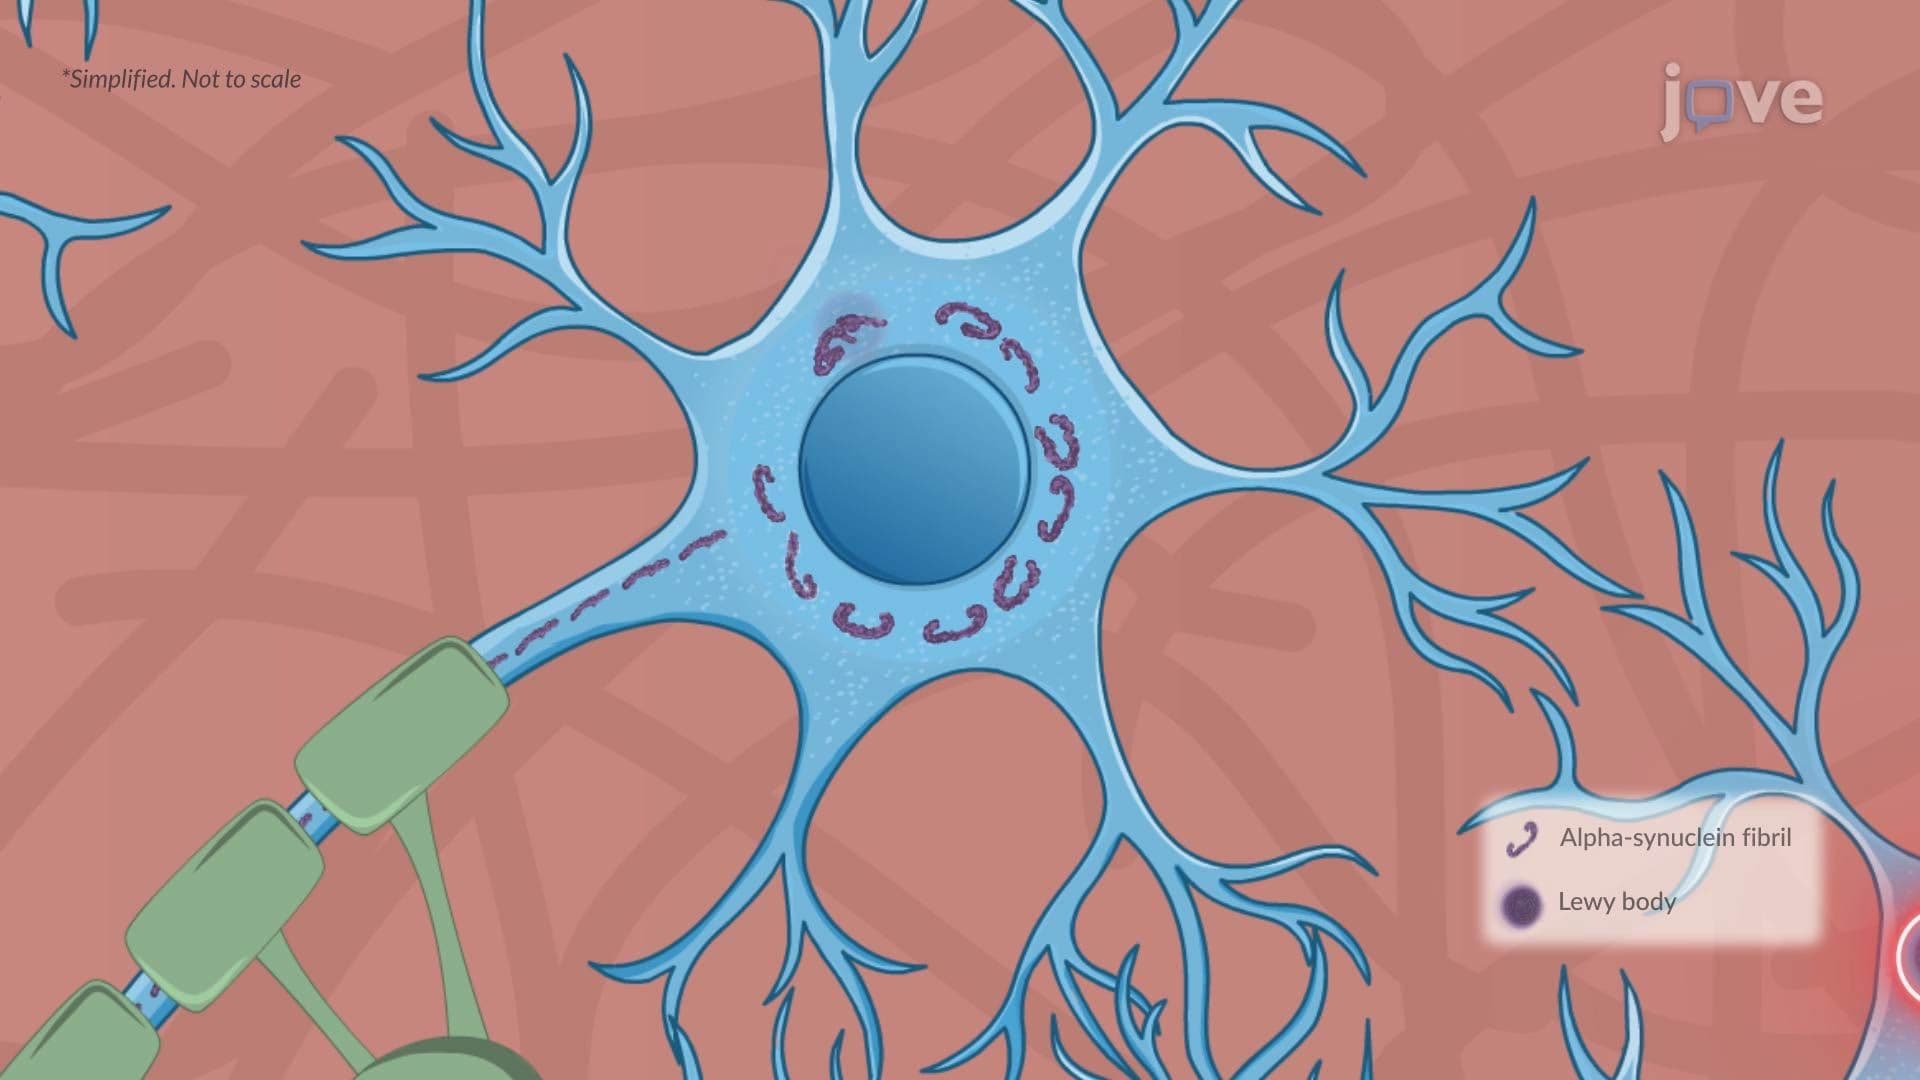
Parkinson Disease ll: Pathophysiology
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...