
[The study of infrared spectra of 2-pyridinemethanol by density functional theory]
Xiao-Ming Li1, Lai-Bin Zhang, Liu-Zhu Zhou
1College of Physics and Engineering, Qufu Normal University, Qufu 273165, China. gh1058@163.com

Abstract:
Using density functional theory (DFT), geometry optimizations and frequencies calculation of 2-pyridinemethanol were performed at B3LYP/6-311G(d, p) level, the stable structure and complete vibrational modes of 2-pyridinemethanol were attained. The calculated geometric parameters are in good agreement with the reported experimental measurement of pyridine and the reported data of pertinent literature. When comparing the calculated IR data with those reported by experiments, it was found that the calculated results are in good agreement with the experimental results. Finally, the vibrational modes of 2-pyridinemethanol molecule were assigned.
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:09
Infrared (IR) Spectroscopy: Overview
Different compounds display unique properties due to their...
 01:29
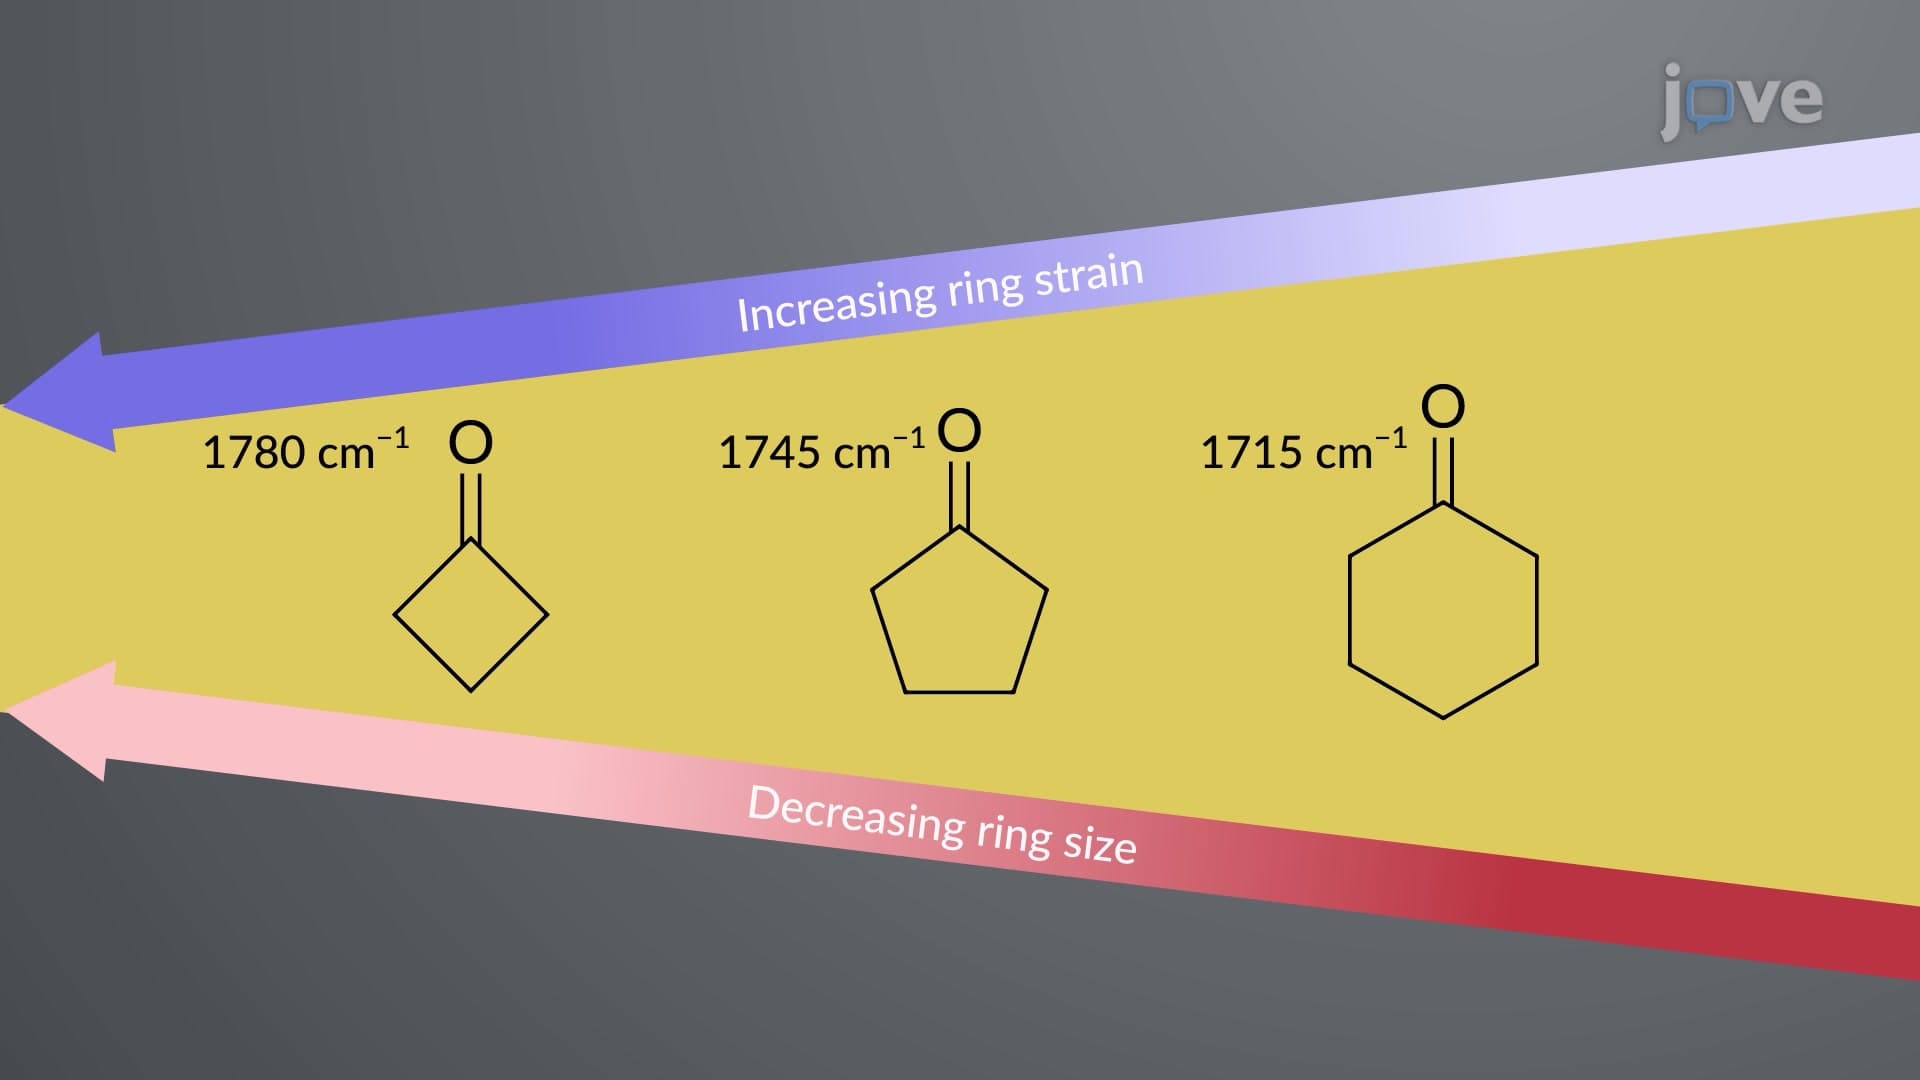
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
 01:29
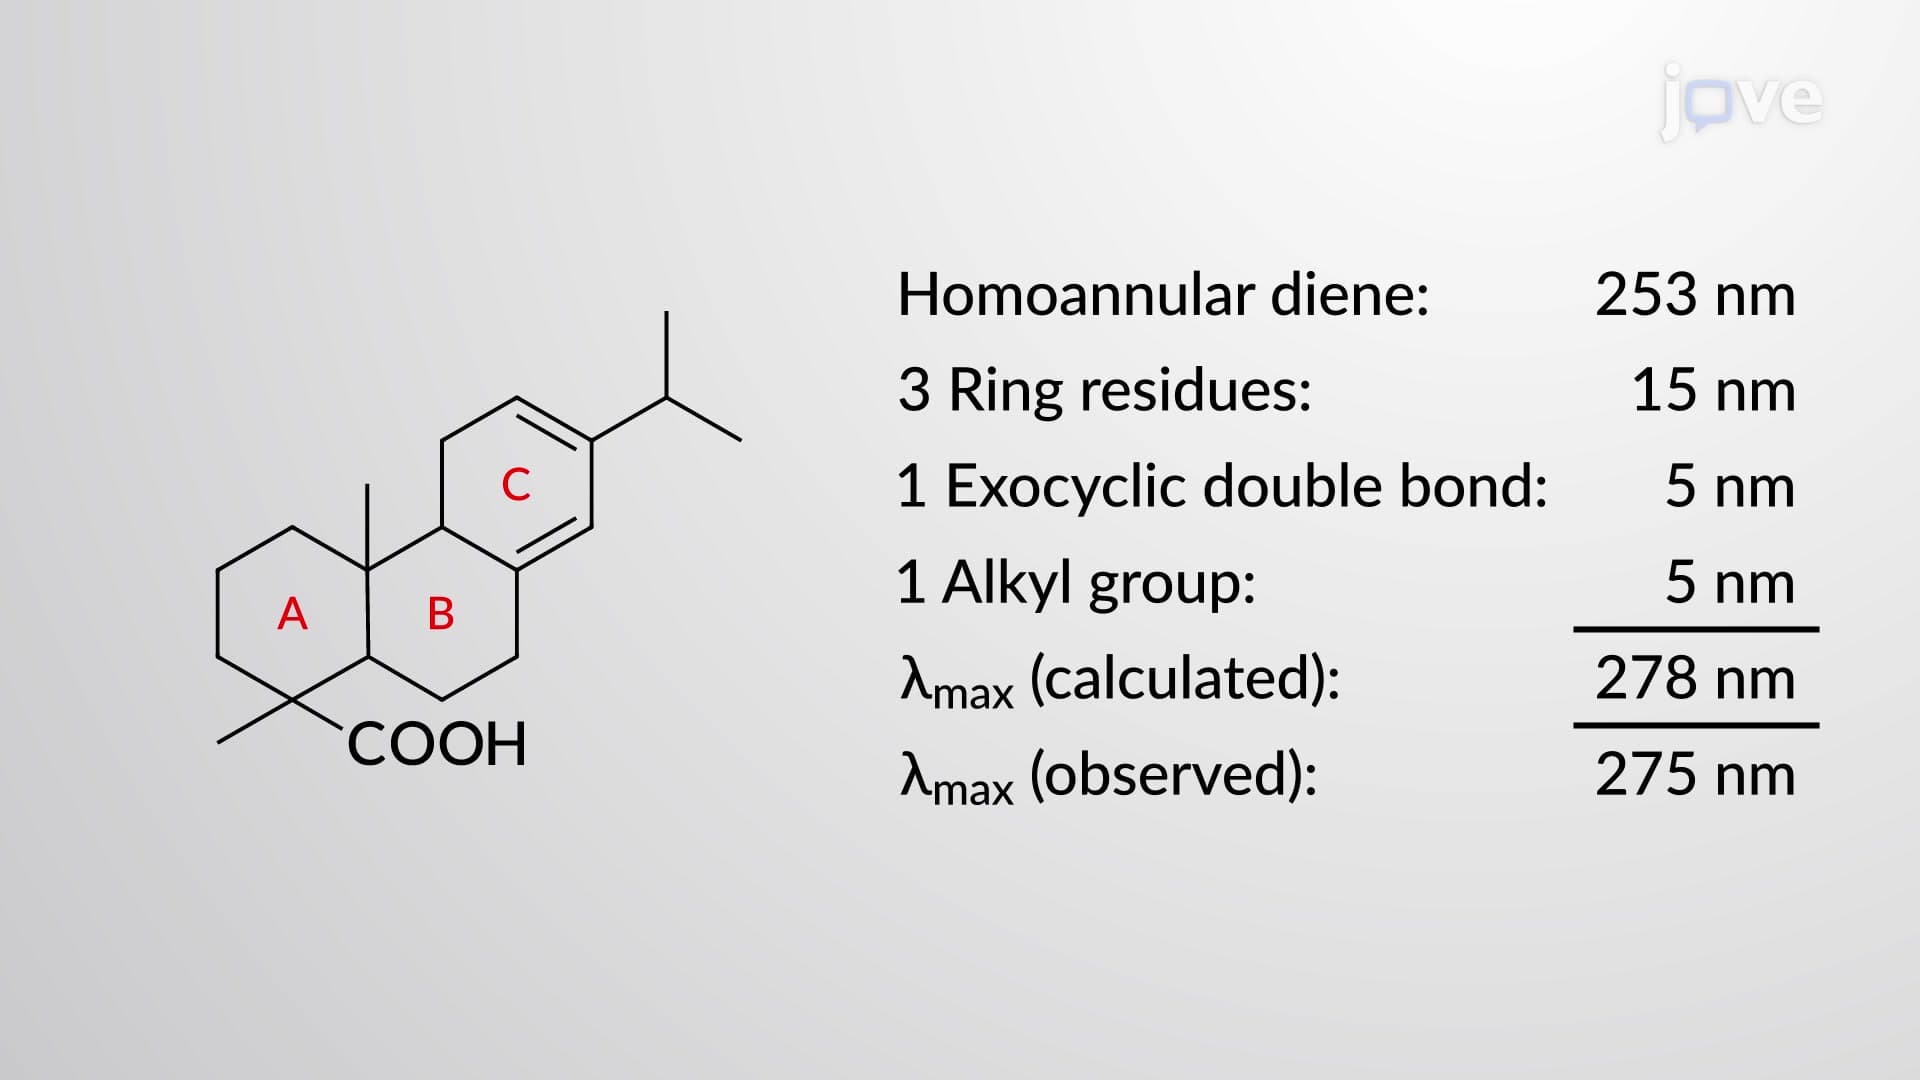
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:23
IR Spectrum Peak Broadening: Hydrogen Bonding
However, the extent of hydrogen bonding influences the observed stretching frequency and band broadening. Intermolecular or intramolecular hydrogen bonding...