

Performance of recent and high-performance approximate density functionals for time-dependent density functional
Miho Isegawa1, Roberto Peverati, Donald G Truhlar
1Department of Chemistry and Supercomputing Institute, University of Minnesota, Minneapolis, Minnesota 55455-0431, USA.

Abstract:
We report a test of 30 density functionals, including several recent ones, for their predictions of 69 singlet-to-singlet excitation energies of 11 molecules. The reference values are experimental results collected by Caricato et al. for 30 valence excitations and 39 Rydberg excitations. All calculations employ time-dependent density functional theory in the adiabatic, linear-response approximation. As far as reasonable, all of the assignments are performed by essentially the same protocol as used by Caricato et al., and this allows us to merge our mean unsigned errors (MUEs) with the ones they calculated for both density functional and wave function methods. We find 21 of the 30 density functionals calculated here have smaller MUEs for the 30 valence states than what they obtained (0.47 eV) for the state-of-the-art EOM-CCSD wave function. In contrast, for all of density functionals the MUE for 39 Rydberg states is larger than that (0.11 eV) of EOM-CCSD. Merging the 30 density functionals calculated here with the 26 calculated by Caricato et al. makes a set of 56 density functionals. Averaging the unsigned errors over both the valence excitations and the Rydberg excitations, none of the 56 density functionals shows a lower mean unsigned error than that (0.27 eV) of EOM-CCSD. Nevertheless, two functionals are successful in having an overall mean unsigned error of 0.30 eV, and another nine are moderately successful in having overall mean unsigned errors in the range 0.32-0.36 eV. Successful or moderately successful density functionals include seven hybrid density functionals with 41% to 54% Hartree-Fock exchange, and four range-separated hybrid density functionals in which the percentage of Hartree-Fock exchange increases from 0% to 19% at small interelectronic separation to 65%-100% at long range.
More Related Videos


Related Concept Videos
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:45
The Quantum-Mechanical Model of an Atom
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:12
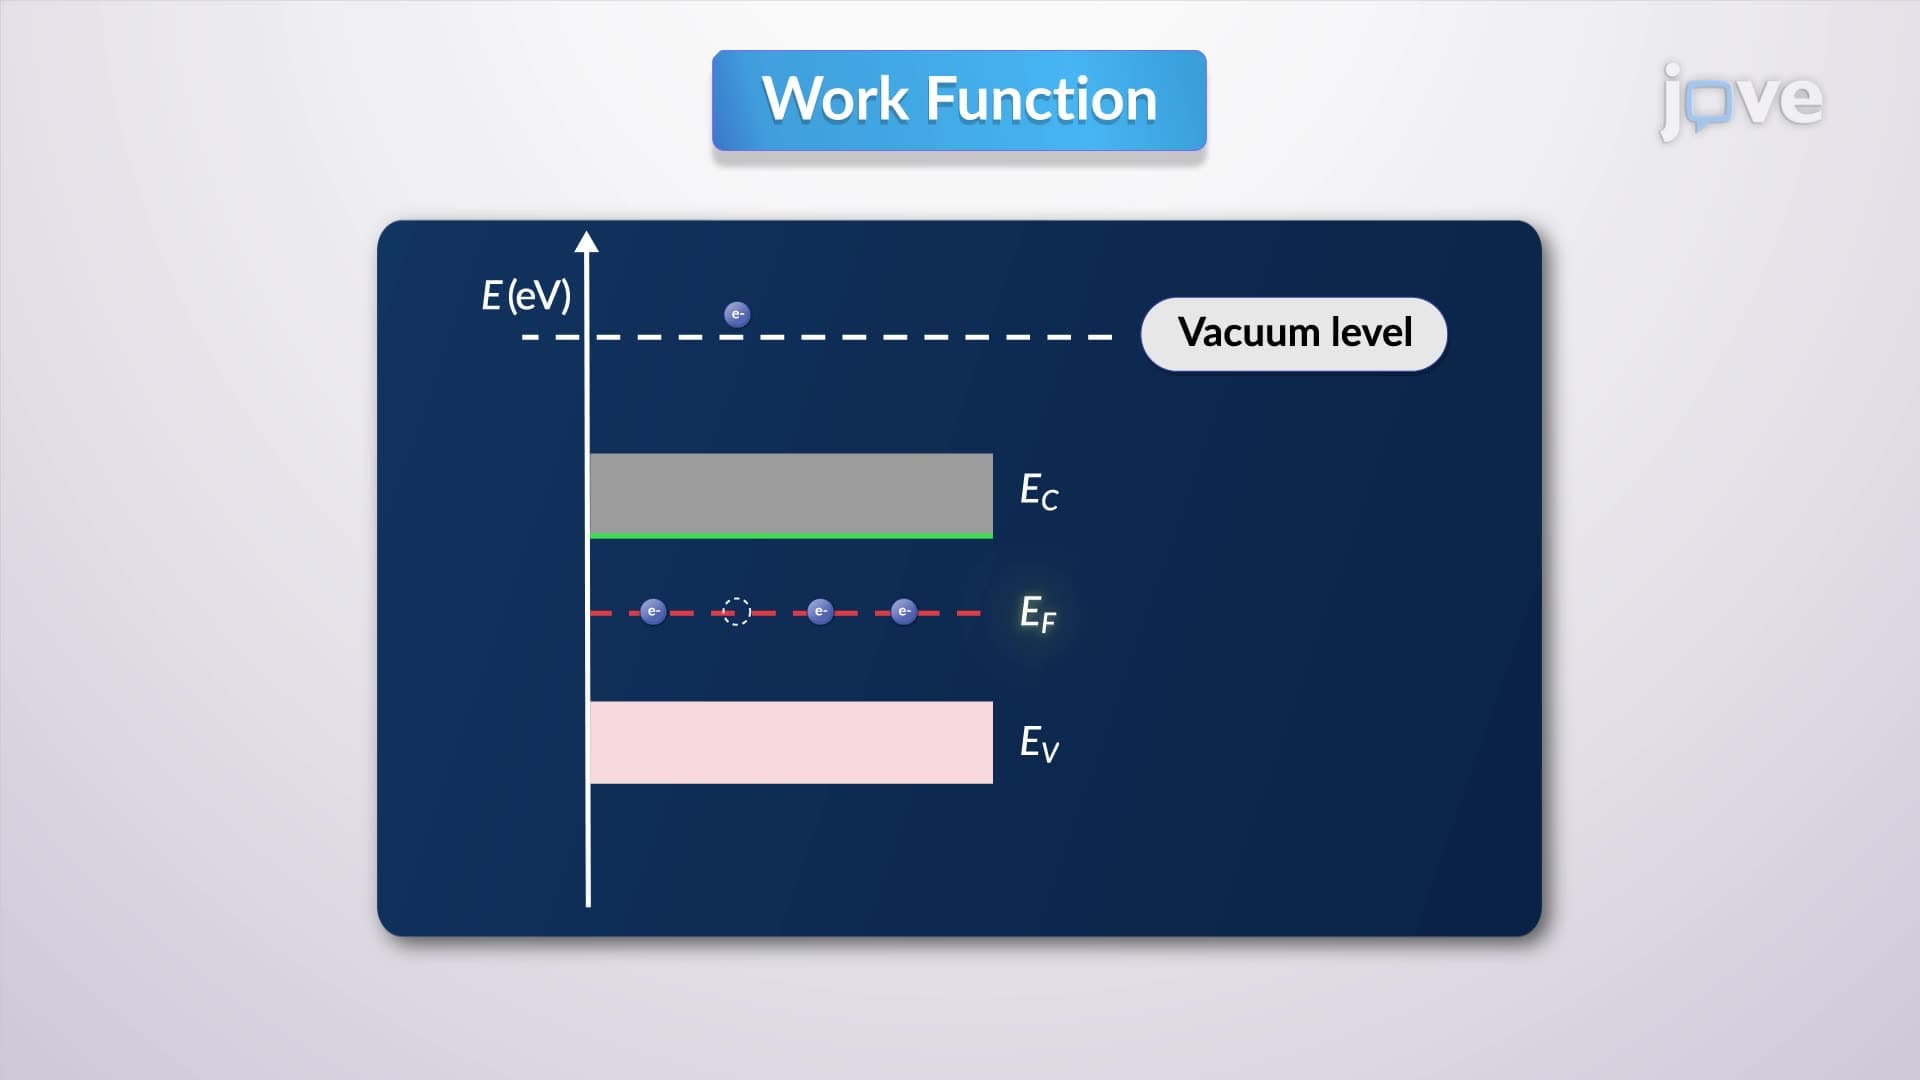
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...