

Simple pair-wise interactions for hybrid Monte Carlo-molecular dynamics simulations of titania/yttria-doped iron
Karl D Hammond1, Hyon-Jee Lee Voigt, Lauren A Marus
1Department of Nuclear Engineering, University of Tennessee, Knoxville, TN 37996, USA.

Abstract:
We present pair-wise, charge-neutral model potentials for an iron-titanium-yttrium-oxygen system. These simple models are designed to provide a tractable method of simulating nanostructured ferritic alloys (NFAs) using off-lattice Monte Carlo and molecular dynamics techniques without deviating significantly from the formalism employed in existing Monte Carlo simulations. The model is fitted to diamagnetic density functional theory (DFT) calculations of the various species over a range of densities and concentrations. The resulting model potentials provide reasonable and in some cases even excellent mechanical and thermodynamic properties for the pure metals. The model replicates the qualitative trends in formation energy predicted by DFT, though the energies of formation do not agree as well for dilute systems as they do for more concentrated systems. We find that on-lattice models will consistently favor tetrahedral oxygen interstitial sites over octahedral interstitial sites, while relaxed systems typically favor octahedral sites. This emphasizes the need for the off-lattice simulations for which this potential was designed.

Related Concept Videos
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:24
Van der Waals Interactions
 01:26
EDTA: Auxiliary Complexing Reagents
 00:43
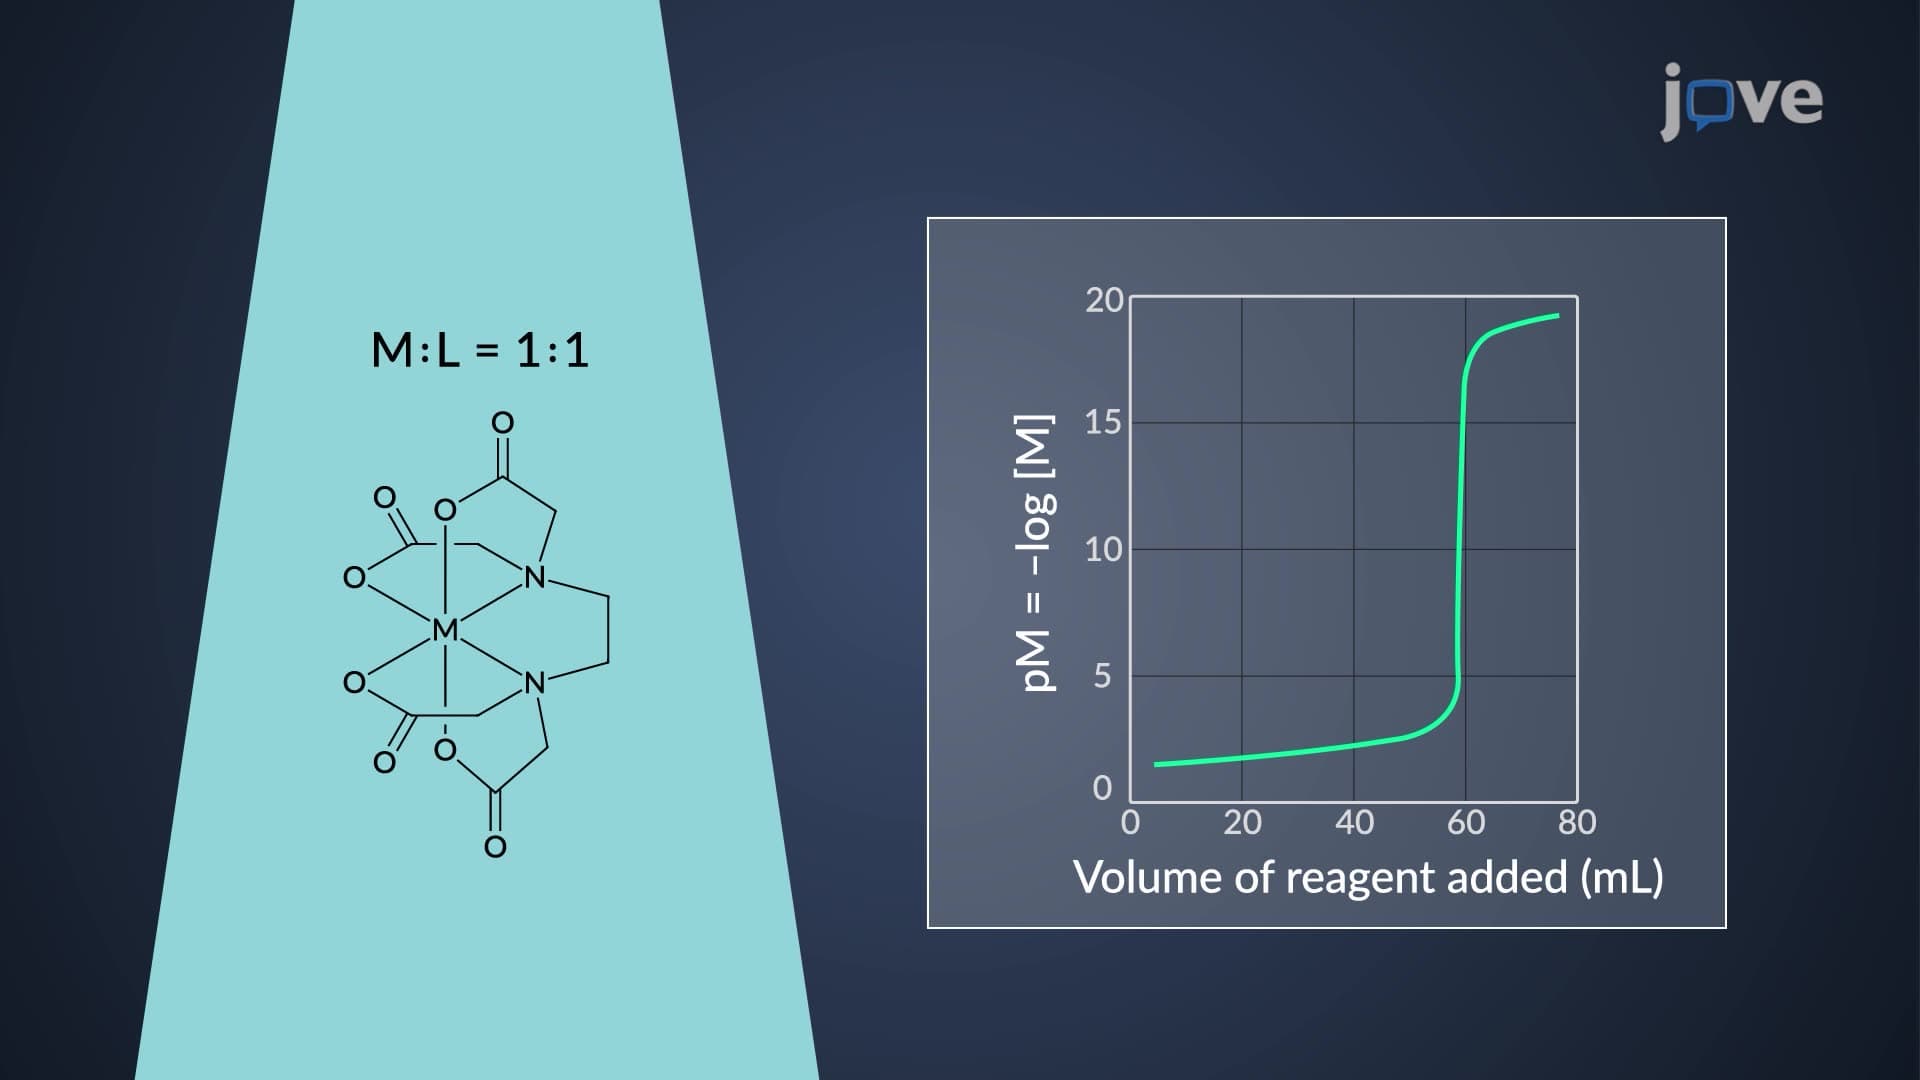
Complexometric Titration: Ligands
 01:22
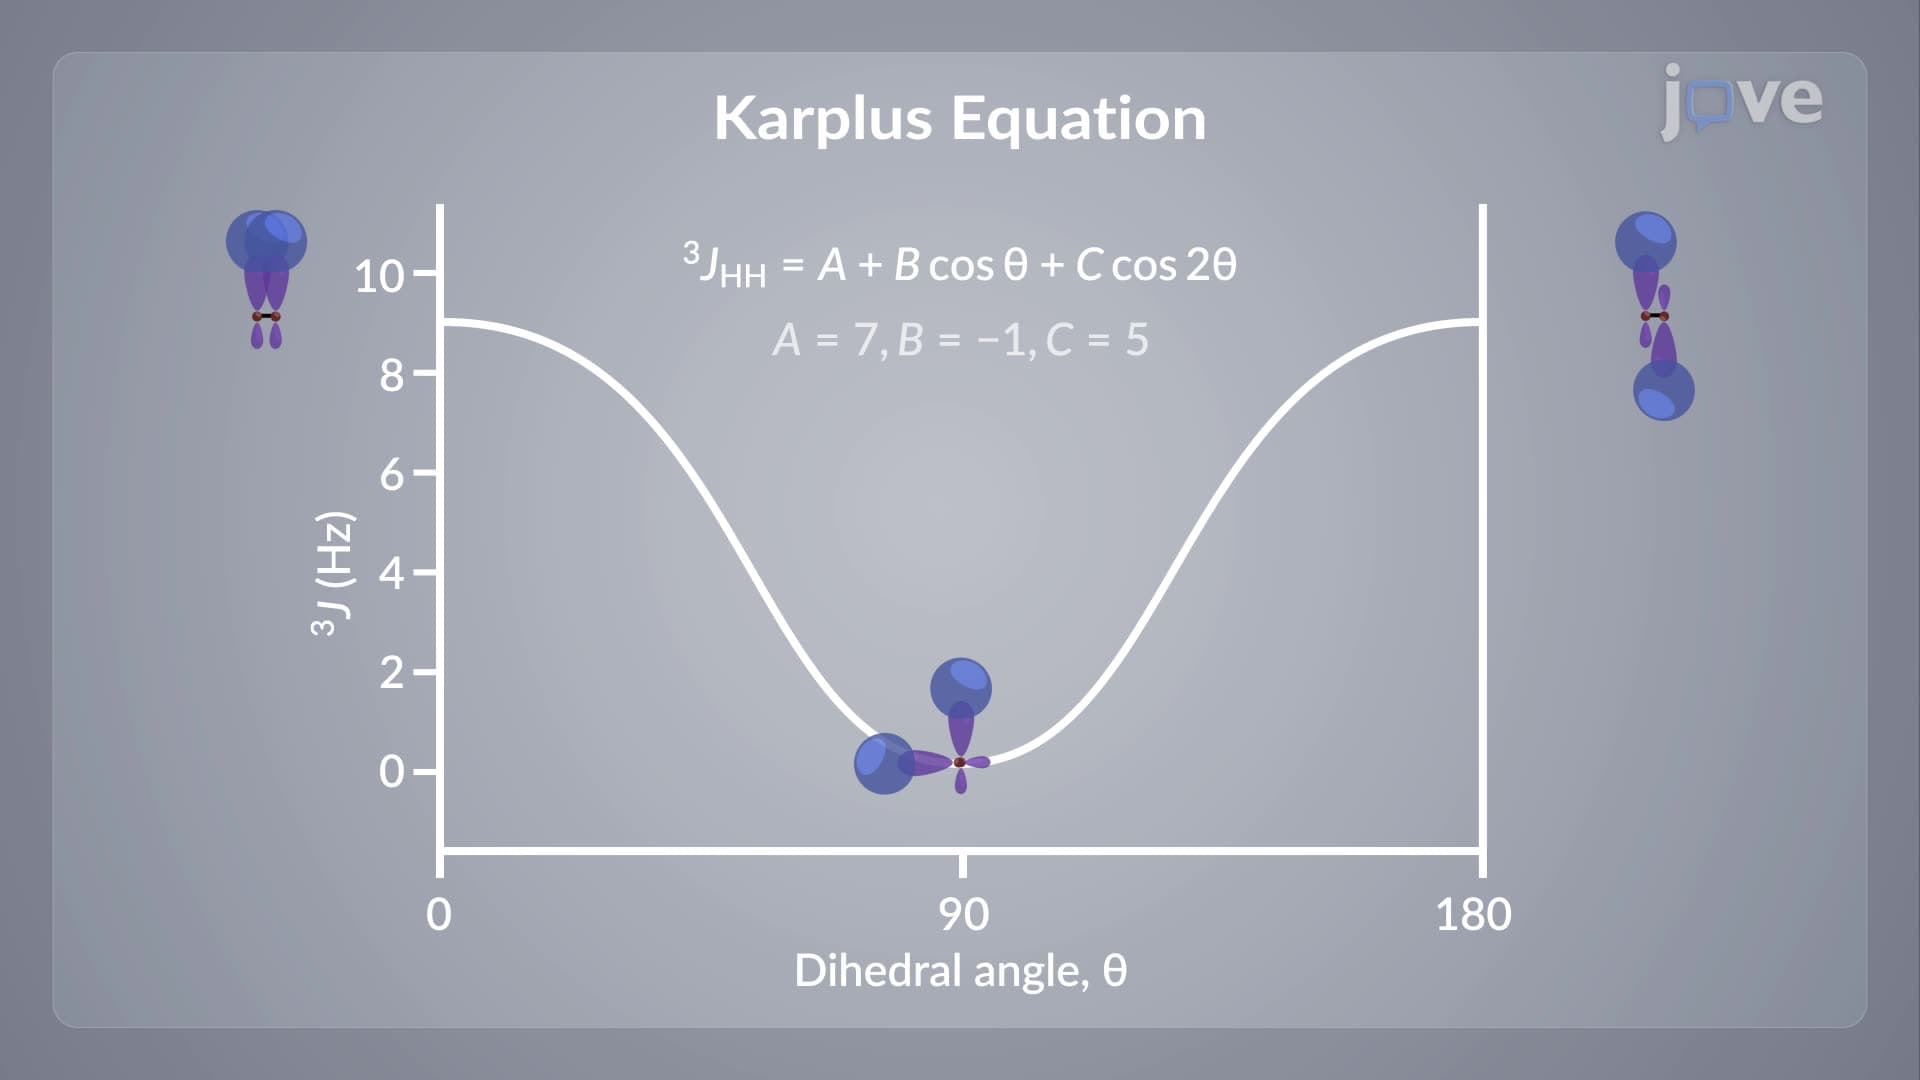
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...