Related Experiment Video
Updated: May 15, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ab initio calculations of thermochemical properties of methanol clusters
1Lehrstuhl für Technische Thermodynamik, RWTH Aachen University, Schinkelstrasse 8, 52062 Aachen, Germany.

Abstract:
Highly accurate ab initio calculations of binding enthalpies and entropies of gas phase clusters of methanol have been performed, yielding uncertainties smaller than 1 kJ/mol per hydrogen bond in the Gibbs free energy of reaction. This requires quantum chemical RIMP2 and CCSD(T) post-Hartree-Fock methods with basis sets up to aug-cc-pV5Z for energy calculations. An analysis of topological symmetry and hindered rotor effects proves necessary for reliable entropies. This approach goes beyond the rigid rotor plus harmonic oscillator method implemented in standard quantum mechanics software tools. The results demonstrate that (1) thermochemical methanol cluster properties can nowadays be obtained by ab initio methods with an accuracy comparable to or even better than that of the experimental data available, especially for larger species that cannot be studied directly by experiments and (2) cooperativity effects and state-dependent cluster distributions cause a strongly varying average enthalpy and entropy per bond as a function of temperature and density for methanol.

Related Concept Videos
 02:37
Standard Enthalpy of Formation
 01:10
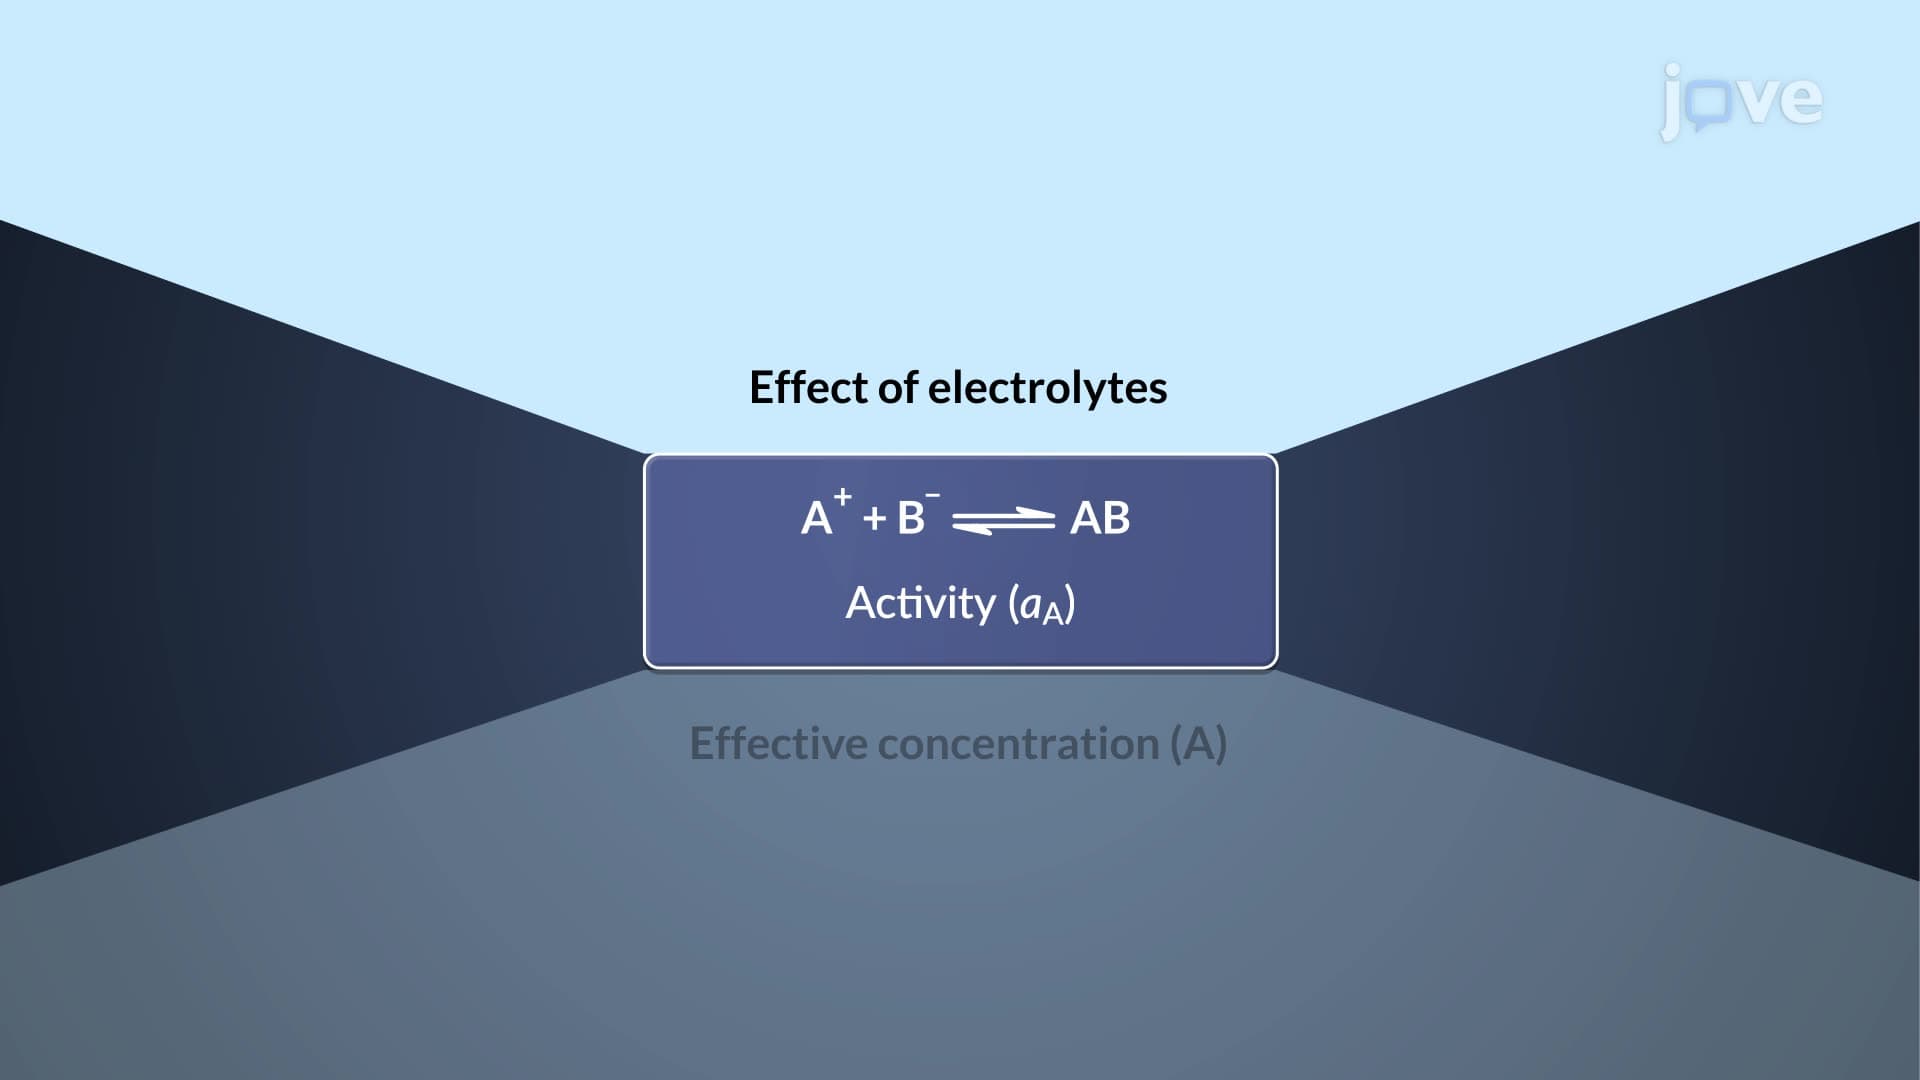
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:55
Thermochemical Equations
 01:26
Thermodynamic Potentials
 01:19
Thermodynamic Properties of Ideal Solutions
 01:32
Radical Anti-Markovnikov Addition to Alkenes: Thermodynamics