Related Experiment Video
Updated: May 15, 2026

A Rapid and Quantitative Fluorimetric Method for Protein-Targeting Small Molecule Drug Screening
Published on: October 16, 2015
Combined multispectroscopic and molecular docking investigation on the interaction between strictosamide and human
Hanlin Pu1, Hua Jiang, Rongrong Chen
1Department of Cell Biology, College of Life Science and Technology, Jinan University, Guangzhou 510632, People's Republic of China. tphl@jnu.edu.cn

Abstract:
The interaction between strictosamide (STM) and human serum albumin (HSA) was investigated by fluorescence spectroscopy, synchronous fluorescence spectroscopy, three-dimensional fluorescence spectroscopy, ultraviolet-visible absorption spectroscopy, circular dichroism spectroscopy and molecular modeling under physiological pH 7.4. STM effectively quenched the intrinsic fluorescence of HSA via static quenching. The binding site number n and apparent binding constant K(a) were determined at different temperatures by fluorescence quenching. The thermodynamic parameters, enthalpy change (ΔH) and entropy change (ΔS) for the reaction were calculated as -3.01 kJ/mol and 77.75 J/mol per K, respectively, which suggested that the hydrophobic force played major roles in stabilizing the HSA-STM complex. The distance r between donor and acceptor was obtained to be 4.10 nm according to Förster's theory. After the addition of STM, the synchronous fluorescence and three-dimensional fluorescence spectral results showed that the hydrophobicity of amino acid residues increased and the circular dichroism spectral results showed that the α-helix content of HSA decreased (from 61.48% to 57.73%). These revealed that the microenvironment and conformation of HSA were changed in the binding reaction. Furthermore, the study of molecular modeling indicated that STM could bind to site I of HSA and the hydrophobic interaction was the major acting force, which was in agreement with the binding mode study.
More Related Videos

Related Concept Videos
 01:29
Drug Distribution: Plasma Protein Binding
 01:30
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:20
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be bound by...
 01:18
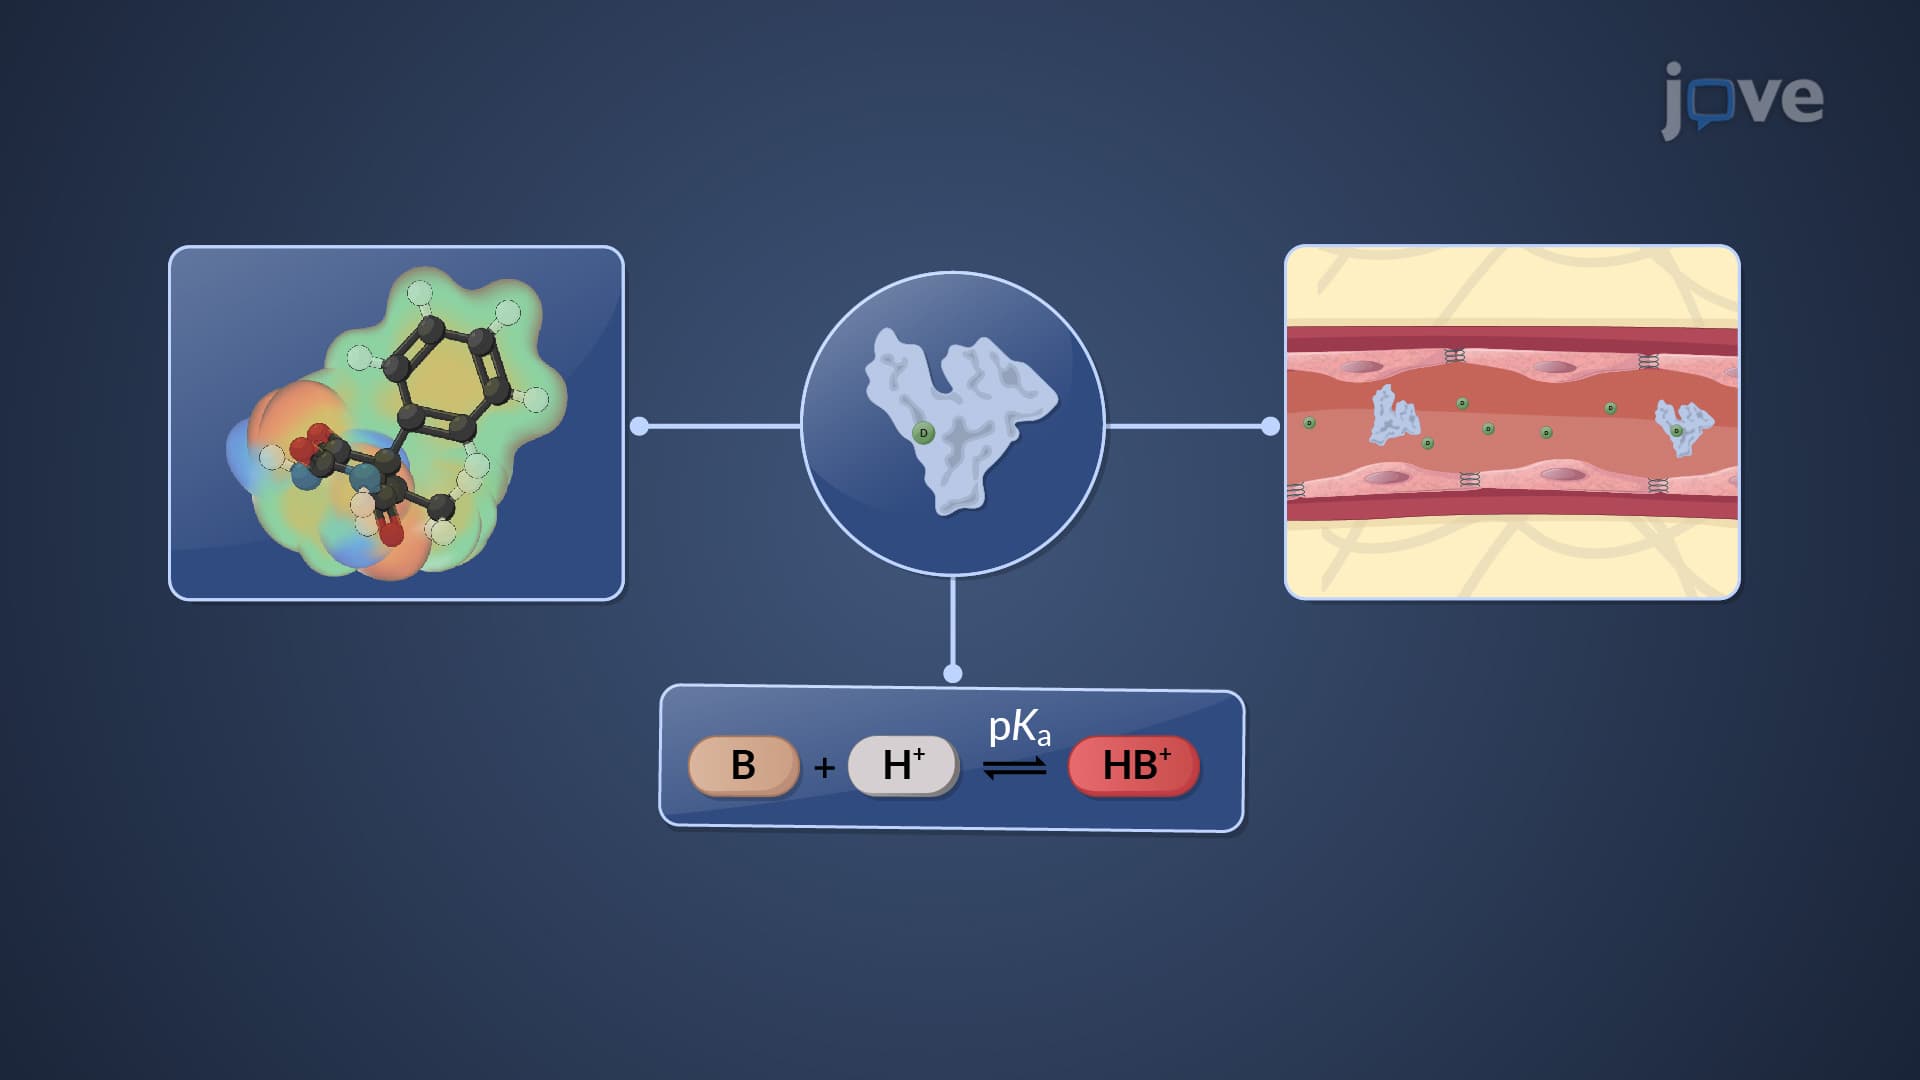
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In contrast,...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...