Advanced techniques for constrained internal coordinate molecular dynamics
Related Experiment Video
Updated: May 14, 2026
New Features in Visual Dynamics 3.0
Published on: August 9, 2024
Jeffrey R Wagner1, Gouthaman S Balaraman, Michiel J M Niesen
1Division of Immunology, Beckman Research Institute of the City of Hope, Duarte, California 91010, USA.

Abstract:
Internal coordinate molecular dynamics (ICMD) methods provide a more natural description of a protein by using bond, angle, and torsional coordinates instead of a Cartesian coordinate representation. Freezing high-frequency bonds and angles in the ICMD model gives rise to constrained ICMD (CICMD) models. There are several theoretical aspects that need to be developed to make the CICMD method robust and widely usable. In this article, we have designed a new framework for (1) initializing velocities for nonindependent CICMD coordinates, (2) efficient computation of center of mass velocity during CICMD simulations, (3) using advanced integrators such as Runge-Kutta, Lobatto, and adaptive CVODE for CICMD simulations, and (4) cancelling out the "flying ice cube effect" that sometimes arises in Nosé-Hoover dynamics. The Generalized Newton-Euler Inverse Mass Operator (GNEIMO) method is an implementation of a CICMD method that we have developed to study protein dynamics. GNEIMO allows for a hierarchy of coarse-grained simulation models based on the ability to rigidly constrain any group of atoms. In this article, we perform tests on the Lobatto and Runge-Kutta integrators to determine optimal simulation parameters. We also implement an adaptive coarse-graining tool using the GNEIMO Python interface. This tool enables the secondary structure-guided "freezing and thawing" of degrees of freedom in the molecule on the fly during molecular dynamics simulations and is shown to fold four proteins to their native topologies. With these advancements, we envision the use of the GNEIMO method in protein structure prediction, structure refinement, and in studying domain motion.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 03:35
Hybridization of Atomic Orbitals II
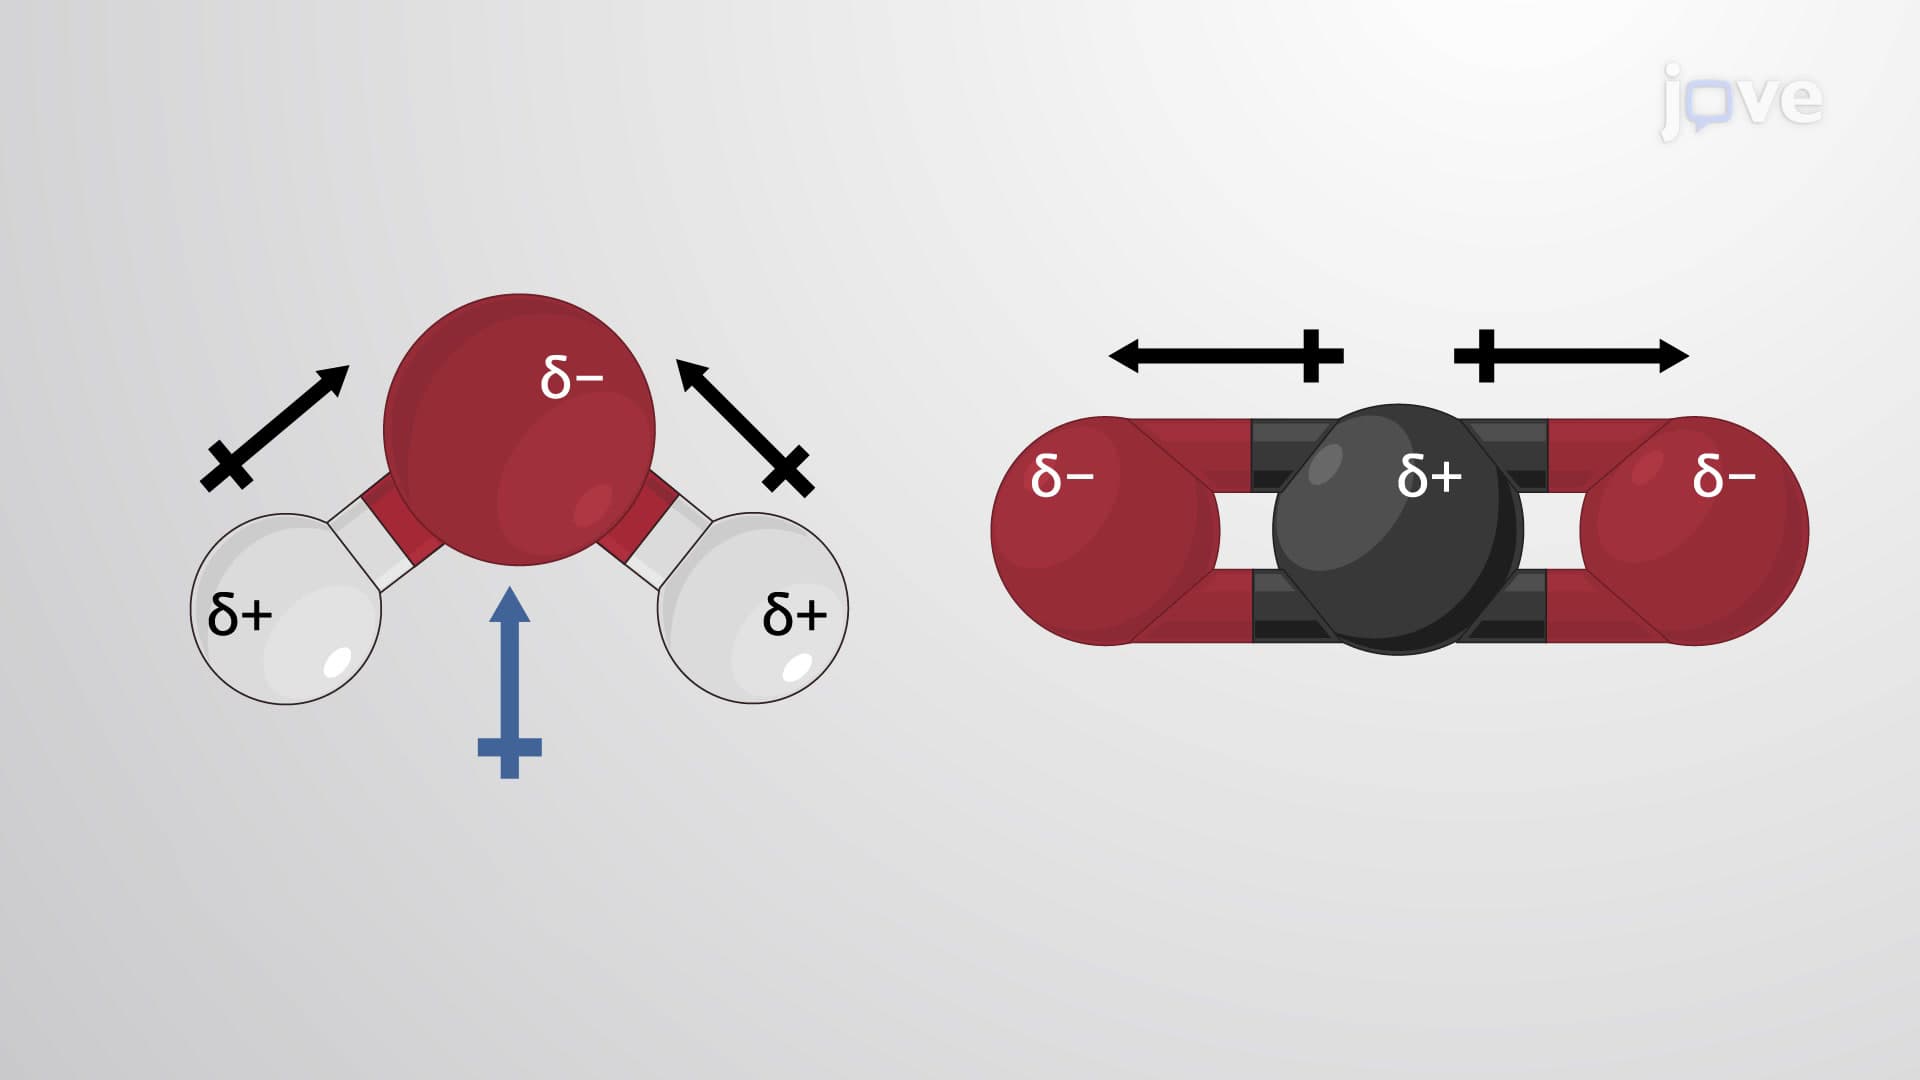 02:36
Molecular Geometry and Dipole Moments
 03:13
Intermolecular Forces