Related Experiment Video
Updated: May 14, 2026

Stepwise Dosing Protocol for Increased Throughput in Label-Free Impedance-Based GPCR Assays
Published on: February 21, 2020
Detailed analysis of biased histamine H₄ receptor signalling by JNJ 7777120 analogues
S Nijmeijer1, H F Vischer, F Sirci
1Faculty of Sciences, Amsterdam Institute for Molecules, Medicines and Systems, VU University Amsterdam, The Netherlands.

Background And Purpose:
The histamine H₄ receptor, originally thought to signal merely through Gαi proteins, has recently been shown to also recruit and signal via β-arrestin2. Following the discovery that the reference antagonist indolecarboxamide JNJ 7777120 appears to be a partial agonist in β-arrestin2 recruitment, we have identified additional biased hH₄R ligands that preferentially couple to Gαi or β-arrestin2 proteins. In this study, we explored ligand and receptor regions that are important for biased hH₄R signalling.
Experimental Approach:
We evaluated a series of 48 indolecarboxamides with subtle structural differences for their ability to induce hH₄R-mediated Gαi protein signalling or β-arrestin2 recruitment. Subsequently, a Fingerprints for Ligands and Proteins three-dimensional quantitative structure-activity relationship analysis correlated intrinsic activity values with structural ligand requirements. Moreover, a hH₄R homology model was used to identify receptor regions important for biased hH₄R signalling.
Key Results:
One indolecarboxamide (75) with a nitro substituent on position R7 of the aromatic ring displayed an equal preference for the Gαi and β-arrestin2 pathway and was classified as unbiased hH₄R ligand. The other 47 indolecarboxamides were β-arrestin2-biased agonists. Intrinsic activities of the unbiased as well as β-arrestin2-biased indolecarboxamides to induce β-arrestin2 recruitment could be correlated with different ligand features and hH₄R regions.
Conclusion And Implications:
Small structural modifications resulted in diverse intrinsic activities for unbiased (75) and β-arrestin2-biased indolecarboxamides. Analysis of ligand and receptor features revealed efficacy hotspots responsible for biased-β-arrestin2 recruitment. This knowledge is useful for the design of hH₄R ligands with biased intrinsic activities and aids our understanding of the mechanism of H₄R activation.
Related Concept Videos
 01:28
Acid Suppressive Drugs for Peptic Ulcer Disease: Histamine H2-Receptor Antagonists
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:31
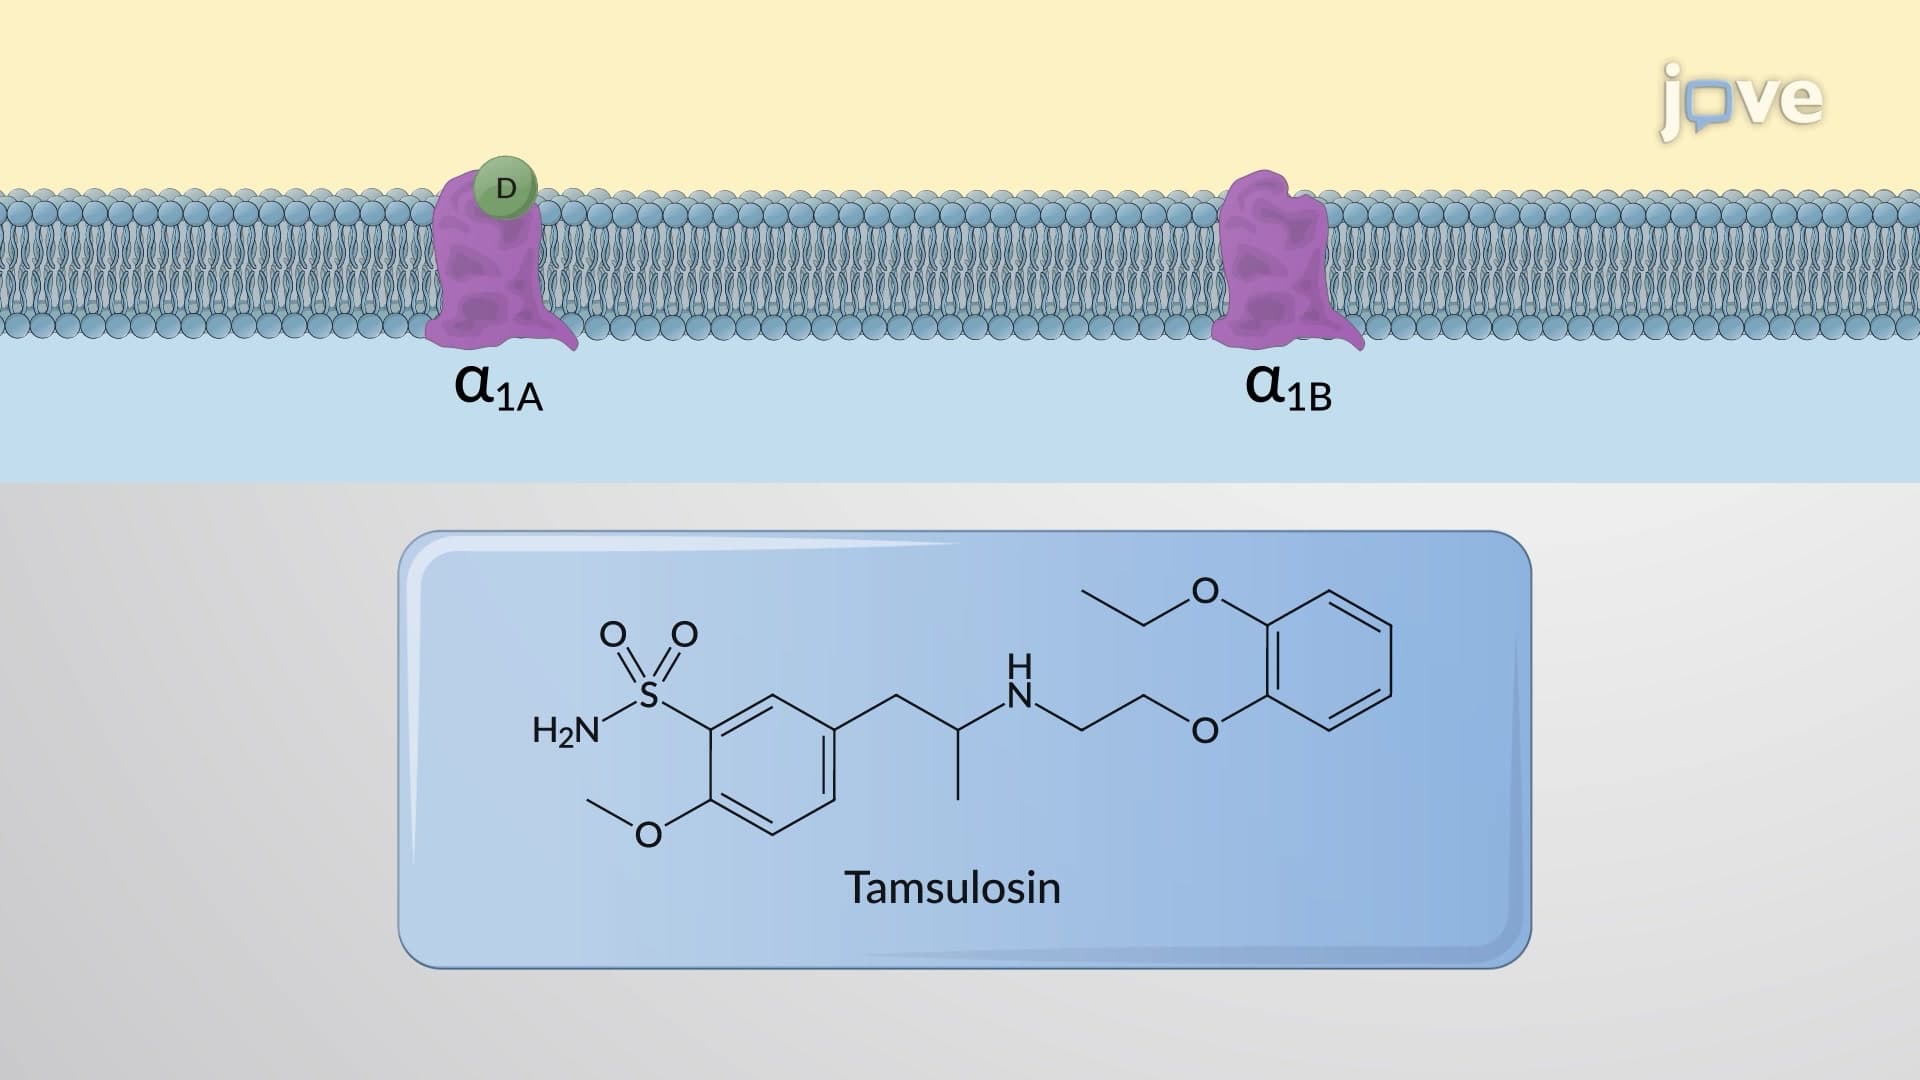
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:26
Adrenergic Receptors: β Subtype
Isoprenaline > Adrenaline > Noradrenaline
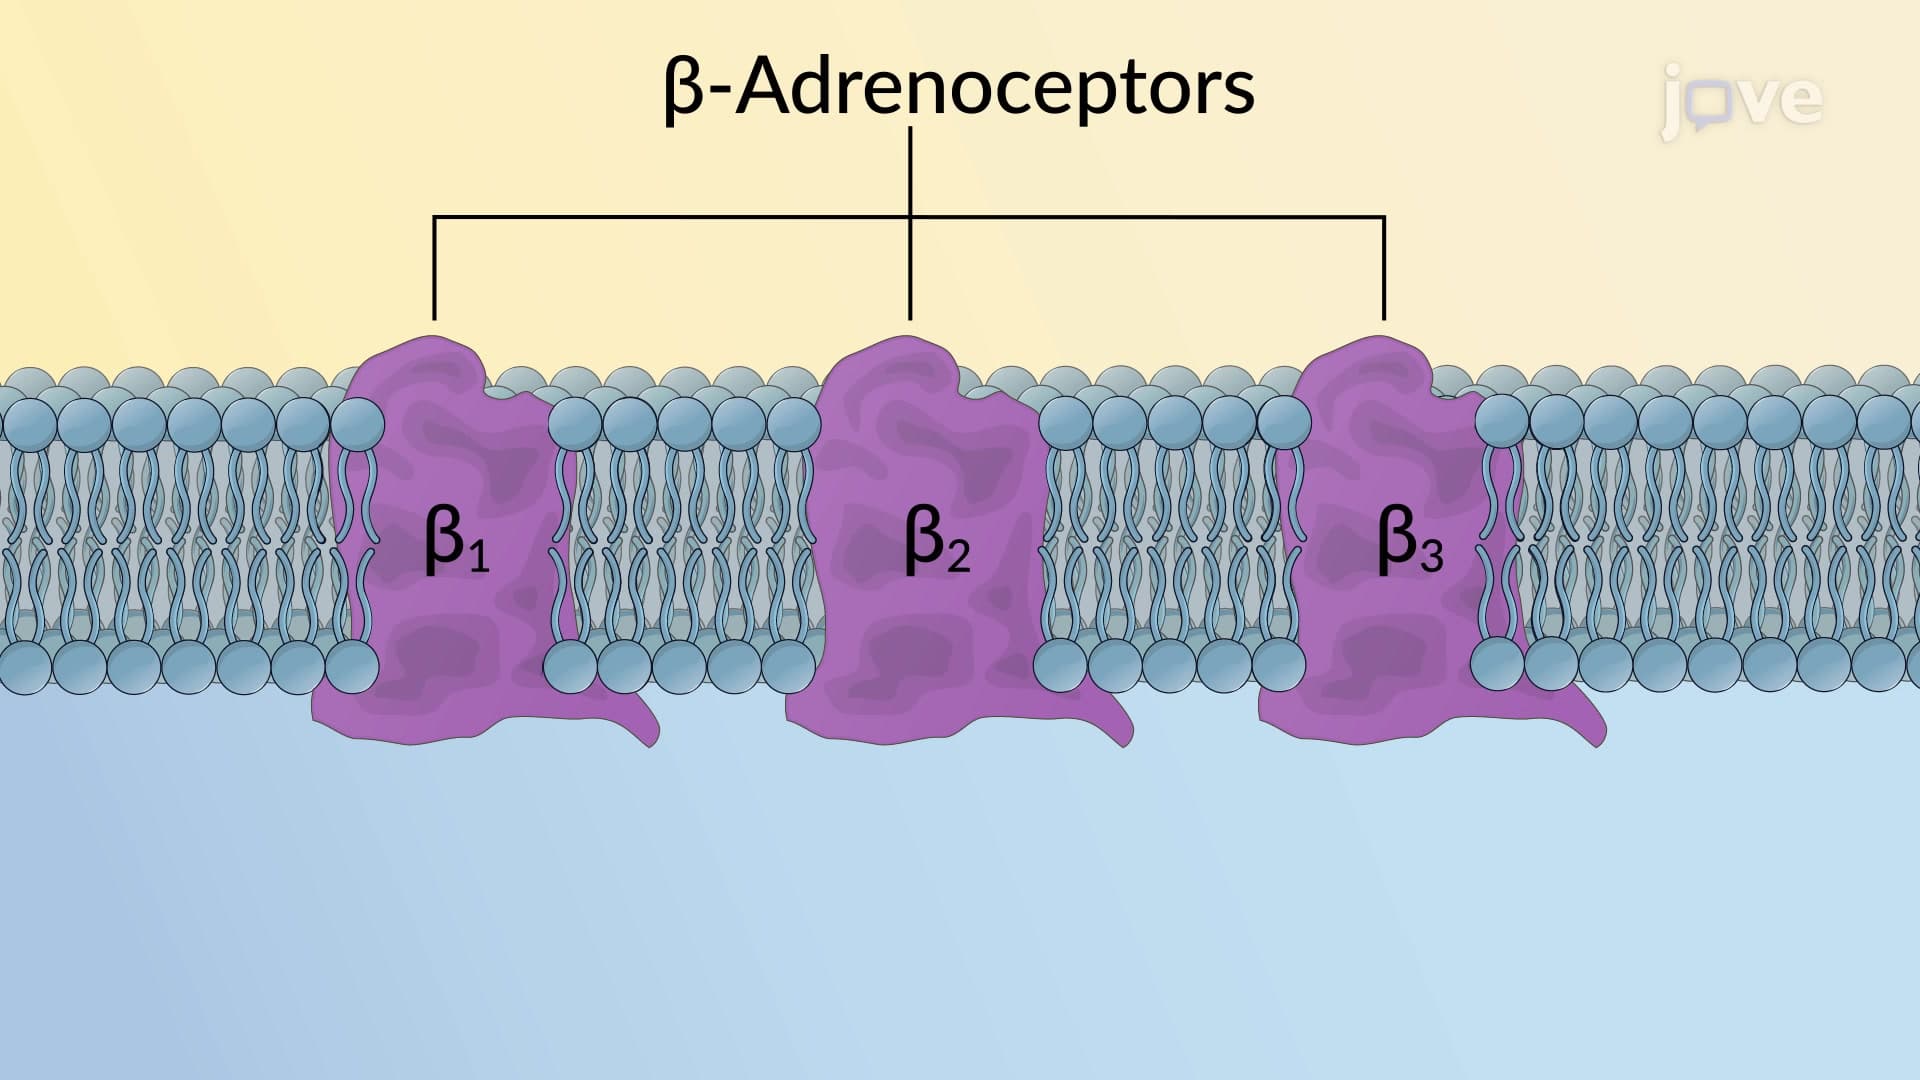
Neurotransmitter binding to these receptors causes activation of adenylyl cyclase resulting in increased concentrations of cAMP and modulation of calcium ion channels within the cell. They are further classified into β1, β2, and β3 subtypes.
β1-adrenoceptors: β1-adrenoceptors have equal affinities for...