Related Experiment Video
Updated: May 14, 2026

Functionalized Spirocyclic Heterocycle Synthesis and Cytotoxicity Assay
Published on: February 9, 2021
Experimental and quantum chemical studies of a novel synthetic prenylated chalcone
José C Espinoza-Hicks1, Alejandro A Camacho-Dávila, Norma R Flores-Holguín
1Facultad de Ciencias Químicas, Universidad Autónoma de Chihuahua, Circuito No, 1, Nuevo Campus Universitario, Chih, Mexico. lmrodrig@uach.mx.

Background:
Chalcones are ubiquitous natural compounds with a wide variety of reported biological activities, including antitumoral, antiviral and antimicrobial effects. Furthermore, chalcones are being studied for its potential use in organic electroluminescent devices; therefore the description of their spectroscopic properties is important to elucidate the structure of these molecules. One of the main techniques available for structure elucidation is the use of Nuclear Magnetic Resonance Spectroscopy (NMR). Accordingly, the prediction of the NMR spectra in this kind of molecules is necessary to gather information about the influence of substituents on their spectra.
Results:
A novel substituted chalcone has been synthetized. In order to identify the functional groups present in the new synthesized compound and confirm its chemical structure, experimental and theoretical 1H-NMR and 13C-NMR spectra were analyzed. The theoretical molecular structure and NMR spectra were calculated at both the Hartree-Fock and Density Functional (meta: TPSS; hybrid: B3LYP and PBE1PBE; hybrid meta GGA: M05-2X and M06-2X) levels of theory in combination with a 6-311++G(d,p) basis set. The structural parameters showed that the best method for geometry optimization was DFT:M06-2X/6-311++G(d,p), whereas the calculated bond angles and bond distances match experimental values of similar chalcone derivatives. The NMR calculations were carried out using the Gauge-Independent Atomic Orbital (GIAO) formalism in a DFT:M06-2X/6-311++G(d,p) optimized geometry.
Conclusion:
Considering all HF and DFT methods with GIAO calculations, TPSS and PBE1PBE were the most accurate methods used for calculation of 1H-NMR and 13C-NMR chemical shifts, which was almost similar to the B3LYP functional, followed in order by HF, M05-2X and M06-2X methods. All calculations were done using the Gaussian 09 software package. Theoretical calculations can be used to predict and confirm the structure of substituted chalcones with good correlation with the experimental data.
More Related Videos
 06:31
Highly Stereoselective Synthesis of 1,6-Ketoesters Mediated by Ionic Liquids: A Three-component Reaction Enabling Rapid Access to a New Class of Low Molecular Weight Gelators
Published on: November 27, 2015
 14:11
Synthesis of pH Dependent Pyrazole, Imidazole, and Isoindolone Dipyrrinone Fluorophores using a Claisen-Schmidt Condensation Approach
Published on: June 10, 2021
Related Concept Videos
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 02:05
Prochirality
 02:34
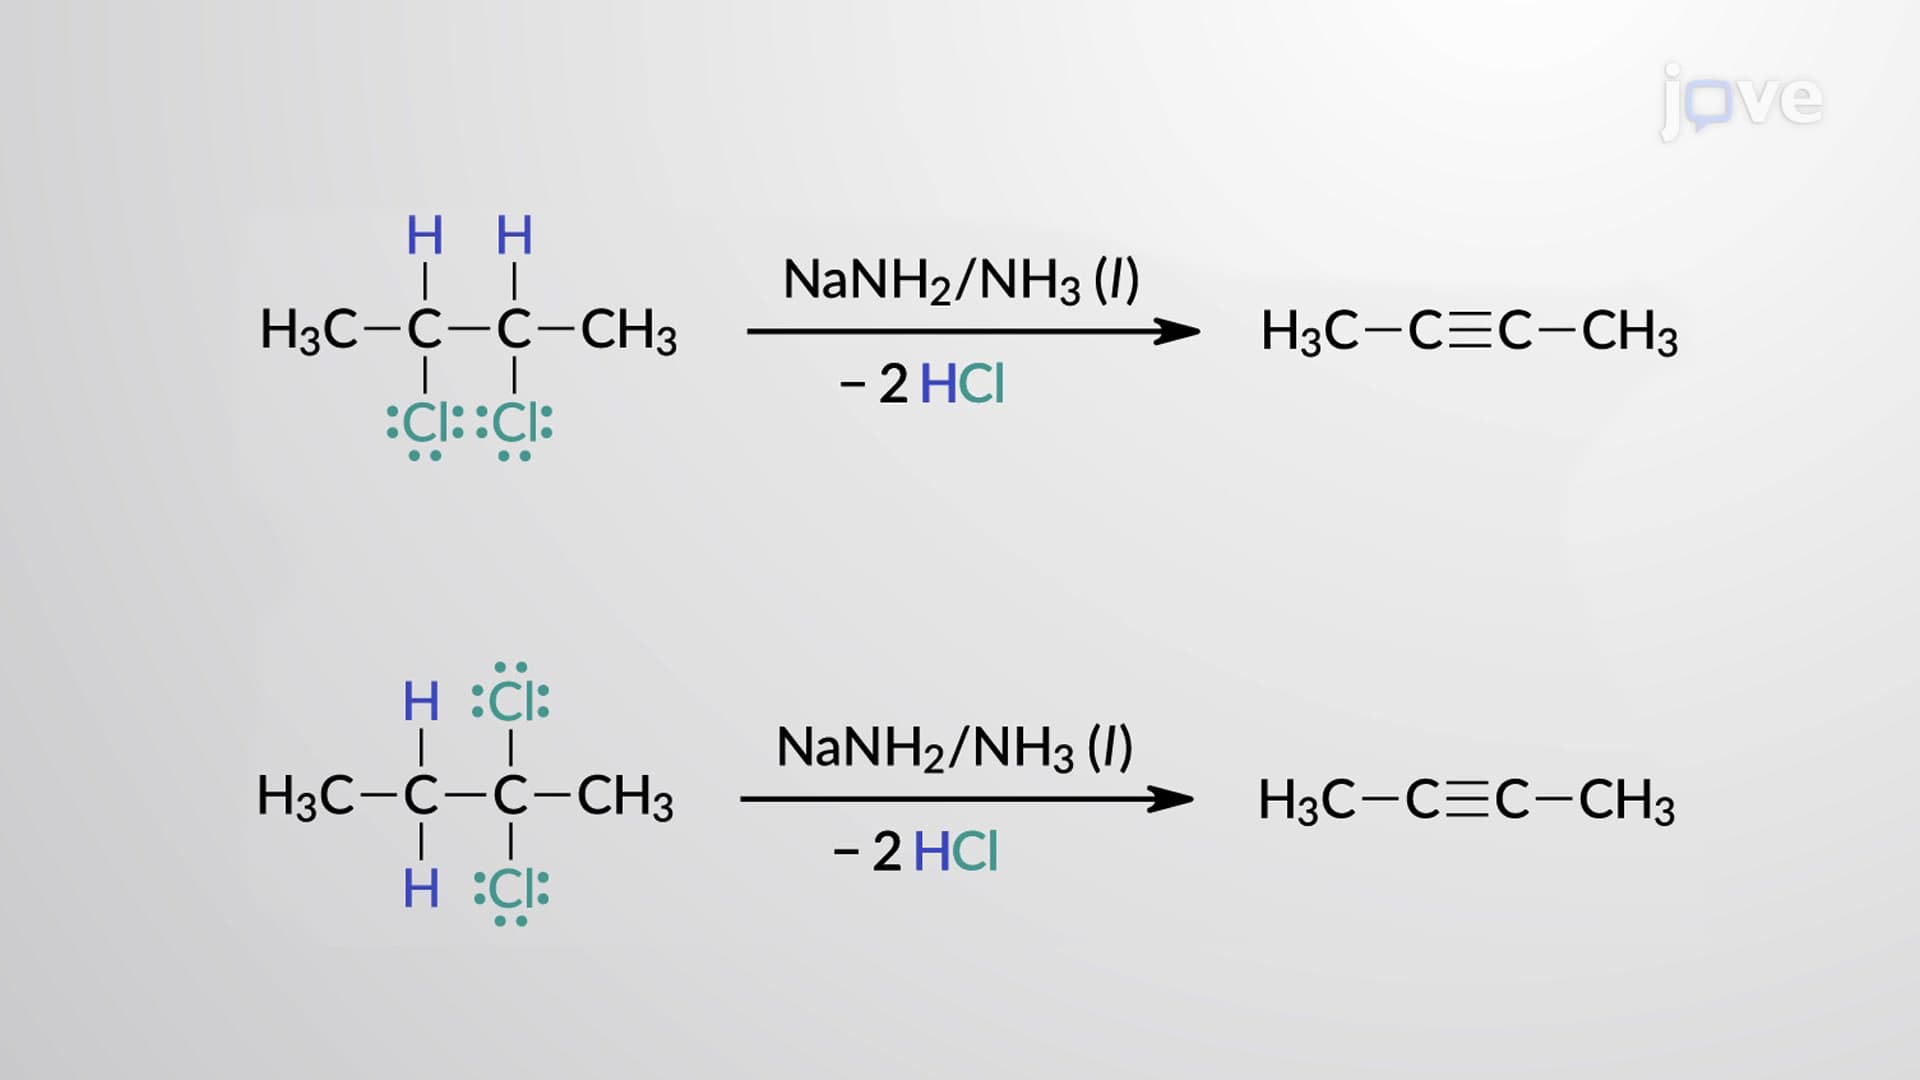
Preparation of Alkynes: Dehydrohalogenation
Alkynes can be prepared by dehydrohalogenation of vicinal or geminal dihalides in the presence of a strong base like sodium amide in liquid ammonia. The reaction proceeds with the loss of two equivalents of hydrogen halide (HX) via two successive E2 elimination reactions.
 01:26
Synthesis of α-Substituted Carbonyl Compounds: The Stork Enamine Reaction
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:29
Diels–Alder Reaction: Characteristics of Dienes
Characteristics of the diene
Conformation
The simplest example of a diene is 1,3-butadiene, an acyclic conjugated π system. At room temperature, the molecule exists as a mixture of s-cis and s-trans conformers by virtue of rotation around the carbon–carbon single bond. Although the s-trans isomer is more stable, the...