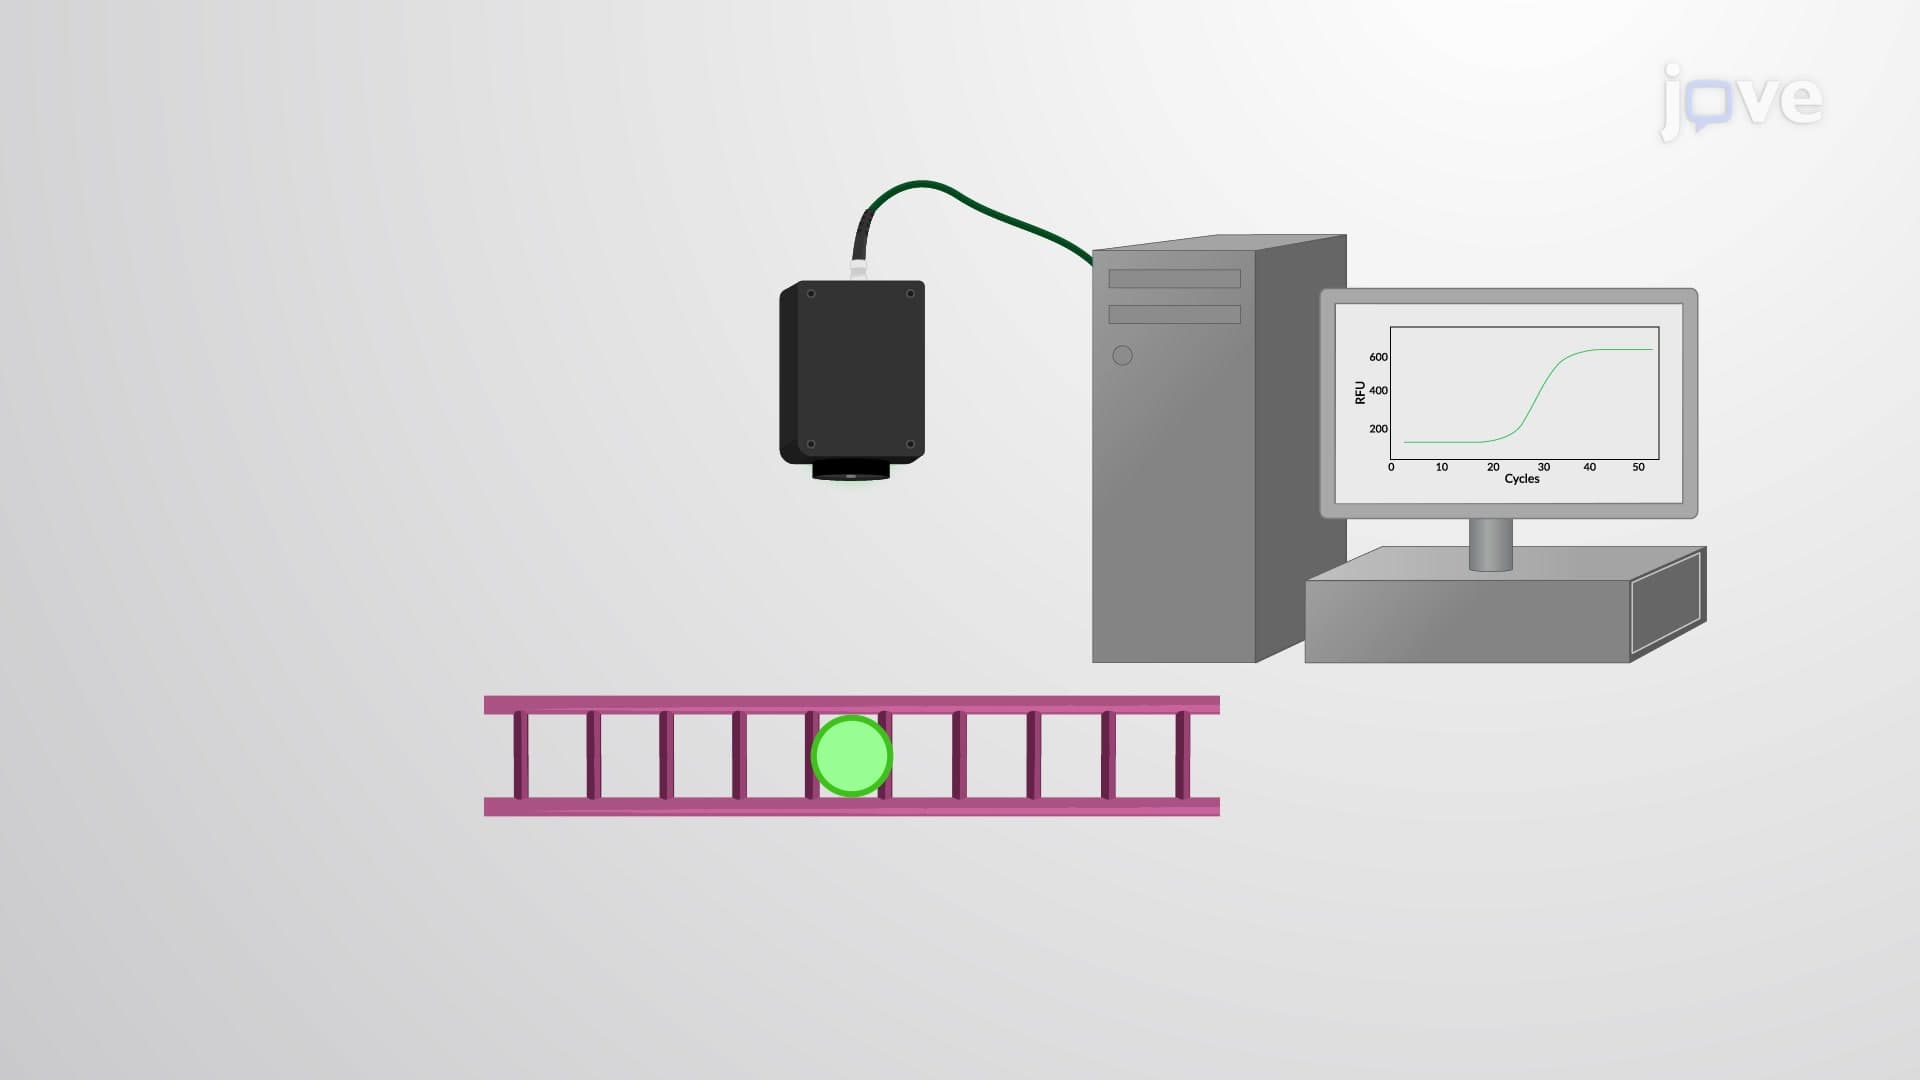
A novel method for the normalization of microRNA RT-PCR data
Related Experiment Video
Updated: May 14, 2026
A Complete Pipeline for Isolating and Sequencing MicroRNAs, and Analyzing Them Using Open Source Tools
Published on: August 21, 2019
1Center for integrated Bioinformatics, School of Biomedical Engineering, Science and Health System, Drexel University, 3120 Market Street, Philadelphia, PA 19104, USA.

Background:
MicroRNAs (miRNAs) are short non-coding RNA molecules that regulate mRNA transcript levels and translation. Deregulation of microRNAs is indicated in a number of diseases and microRNAs are seen as a promising target for biomarker identification and drug development. miRNA expression is commonly measured by microarray or real-time polymerase chain reaction (RT-PCR). The findings of RT-PCR data are highly dependent on the normalization techniques used during preprocessing of the Cycle Threshold readings from RT-PCR. Some of the commonly used endogenous controls themselves have been discovered to be differentially expressed in various conditions such as cancer, making them inappropriate internal controls.
Methods:
We demonstrate that RT-PCR data contains a systematic bias resulting in large variations in the Cycle Threshold (CT) values of the low-abundant miRNA samples. We propose a new data normalization method that considers all available microRNAs as endogenous controls. A weighted normalization approach is utilized to allow contribution from all microRNAs, weighted by their empirical stability.
Results:
The systematic bias in RT-PCR data is illustrated on a microRNA dataset obtained from primary cutaneous melanocytic neoplasms. We show that through a single control parameter, this method is able to emulate other commonly used normalization methods and thus provides a more general approach. We explore the consistency of RT-PCR expression data with microarray expression by utilizing a dataset where both RT-PCR and microarray profiling data is available for the same miRNA samples.
Conclusions:
A weighted normalization method allows the contribution of all of the miRNAs, whether they are highly abundant or have low expression levels. Our findings further suggest that the normalization of a particular miRNA should rely on only miRNAs that have comparable expression levels.
More Related Videos
 10:28
Probe-based Real-time PCR Approaches for Quantitative Measurement of microRNAs
Published on: April 14, 2015
 11:13
Enrichment of Native Lipoprotein Particles with microRNA and Subsequent Determination of Their Absolute/Relative microRNA Content and Their Cellular Transfer Rate
Published on: May 9, 2019