Related Experiment Video
Updated: May 14, 2026

Development of Organoids from Mouse Pituitary as In Vitro Model to Explore Pituitary Stem Cell Biology
Published on: February 25, 2022
Variations in PROKR2, but not PROK2, are associated with hypopituitarism and septo-optic dysplasia
Mark J McCabe1, Carles Gaston-Massuet, Louise C Gregory
1Developmental Endocrinology Research Group, Clinical and Molecular Genetics Unit, University College London (UCL)-Institute of Child Health, London WC1N 1EH, United Kingdom.

Context:
Loss-of-function mutations in PROK2 and PROKR2 have been implicated in Kallmann syndrome (KS), characterized by hypogonadotropic hypogonadism and anosmia. Recent data suggest overlapping phenotypes/genotypes between KS and congenital hypopituitarism (CH), including septo-optic dysplasia (SOD).
Objective:
We screened a cohort of patients with complex forms of CH (n = 422) for mutations in PROK2 and PROKR2.
Results:
We detected 5 PROKR2 variants in 11 patients with SOD/CH: novel p.G371R and previously reported p.A51T, p.R85L, p.L173R, and p.R268C-the latter 3 being known functionally deleterious variants. Surprisingly, 1 patient with SOD was heterozygous for the p.L173R variant, whereas his phenotypically unaffected mother was homozygous for the variant. We sought to clarify the role of PROKR2 in hypothalamopituitary development through analysis of Prokr2(-/-) mice. Interestingly, these revealed predominantly normal hypothalamopituitary development and terminal cell differentiation, with the exception of reduced LH; this was inconsistent with patient phenotypes and more analogous to the healthy mother, although she did not have KS, unlike the Prokr2(-/-) mice.
Conclusions:
The role of PROKR2 in the etiology of CH, SOD, and KS is uncertain, as demonstrated by no clear phenotype-genotype correlation; loss-of-function variants in heterozygosity or homozygosity can be associated with these disorders. However, we report a phenotypically normal parent, homozygous for p.L173R. Our data suggest that the variants identified herein are unlikely to be implicated in isolation in these disorders; other genetic or environmental modifiers may also impact on the etiology. Given the phenotypic variability, genetic counseling may presently be inappropriate.
More Related Videos
 08:03
Midface Hypoplasia and Cranial Base Morphology in Syndromic Craniosynostosis: A Comparative Analysis Study Using a Predictive Regression Model
Published on: November 4, 2025
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
Related Concept Videos
 01:17
The Pituitary Gland
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...
 01:23
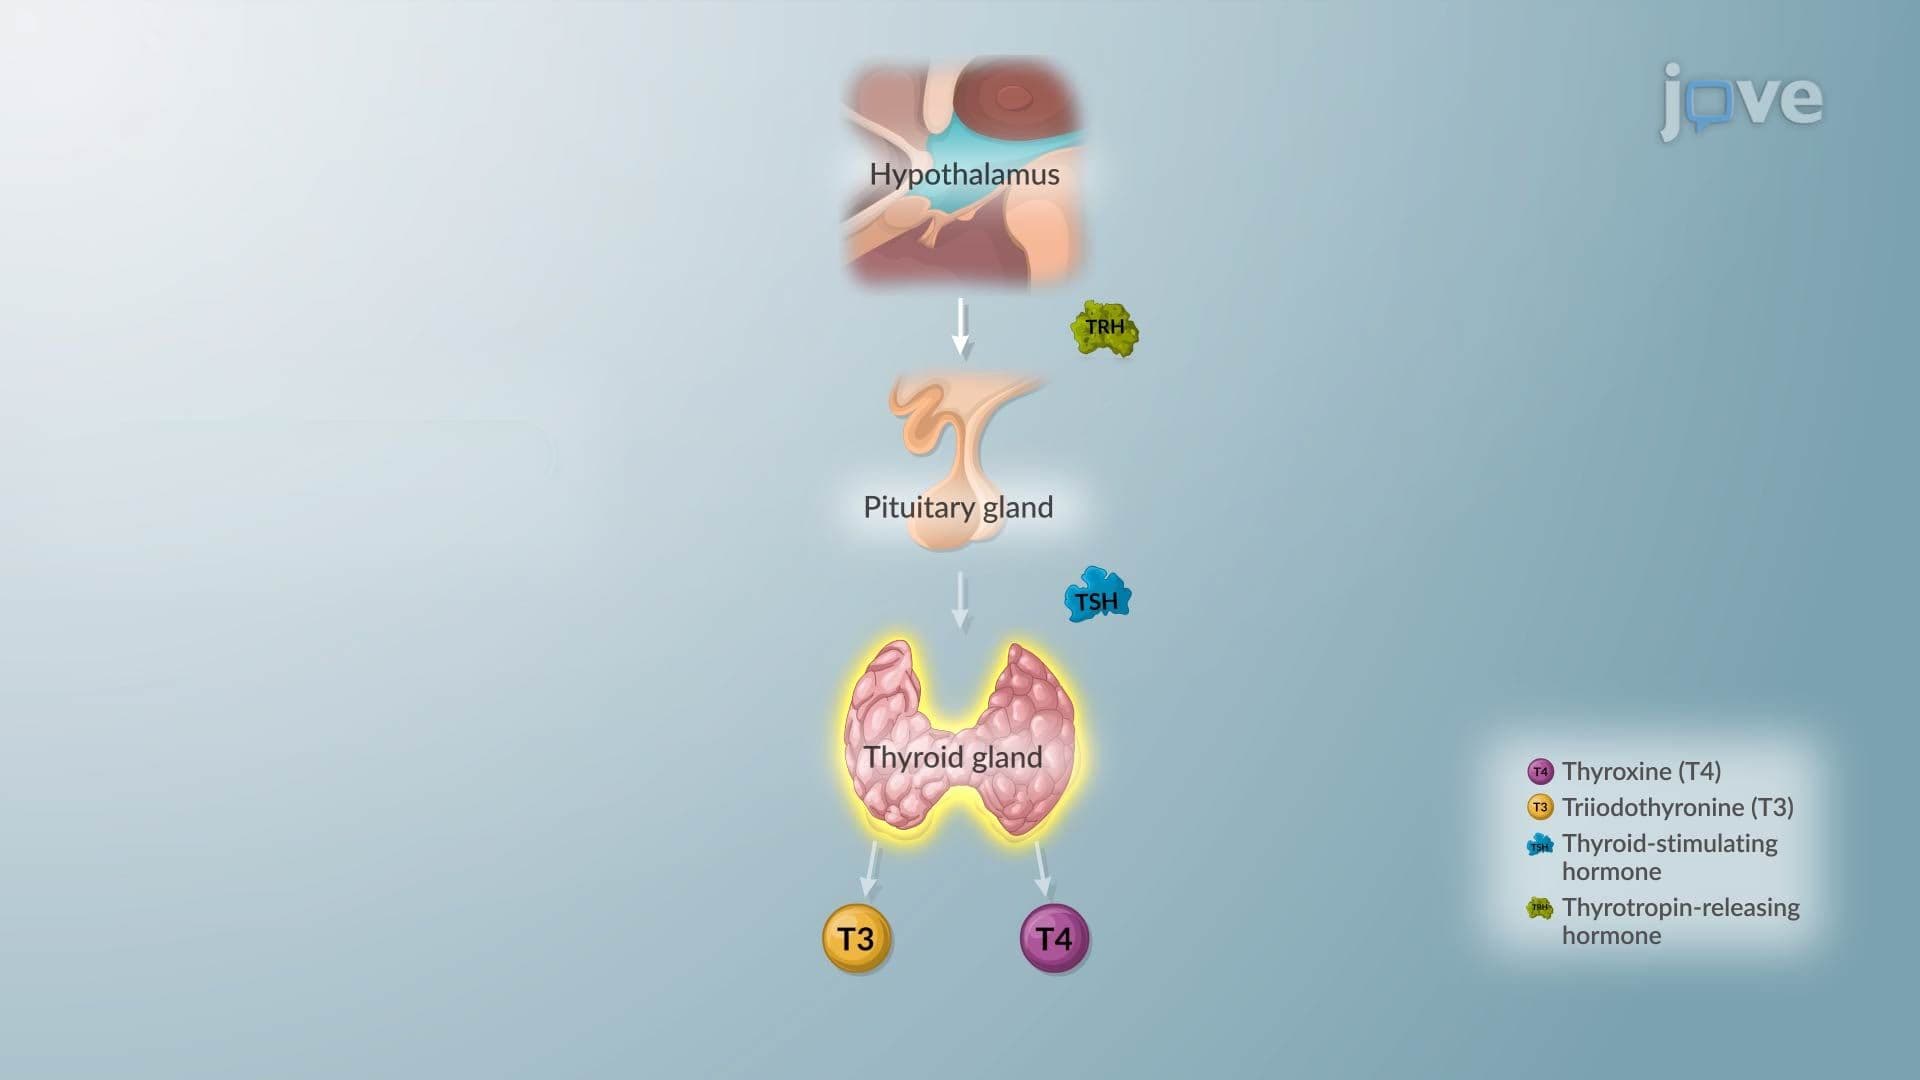
Hypothyroidism II: Pathophysiology
 01:33
Pleiotropy
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...