Related Experiment Video
Updated: May 14, 2026


Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Anharmonic zero point vibrational energies: tipping the scales in accurate thermochemistry calculations?
Florian Pfeiffer1, Guntram Rauhut, David Feller
1Institut für Theoretische Chemie, Universität Stuttgart, D-70550 Stuttgart, Germany.

Abstract:
Anharmonic zero point vibrational energies (ZPVEs) calculated using both conventional CCSD(T) and MP2 in combination with vibrational second-order perturbation theory (VPT2) are compared to explicitly correlated CCSD(T)-F12 and MP2-F12 results that utilize vibrational configuration interaction (VCI) theory for 26 molecules of varying size. Sequences of correlation consistent basis sets are used throughout. It is found that the explicitly correlated methods yield results close to the basis set limit even with double-zeta quality basis sets. In particular, the anharmonic contributions to the ZPVE are accurately recovered at just the MP2 (or MP2-F12) level of theory. Somewhat surprisingly, the best vibrational CI results agreed with the VPT2 values with a mean unsigned deviation of just 0.09 kJ/mol and a standard deviation of just 0.11 kJ/mol. The largest difference was observed for C(4)H(4)O (0.34 kJ/mol). A simplified version of the vibrational CI procedure that limited the modal expansion to at most 2-mode coupling yielded anharmonic corrections generally within about 0.1 kJ/mol of the full 3- or 4-mode results, except in the cases of C(3)H(8) and C(4)H(4)O where the contributions were underestimated by 1.3 and 0.8 kJ/mol, respectively (34% and 40%, respectively). For the molecules considered in this work, accurate anharmonic ZPVEs are most economically obtained by combining CCSD(T)-F12a/cc-pVDZ-F12 harmonic frequencies with either MP2/aug-cc-pVTZ/VPT2 or MP2-F12/cc-pVDZ-F12/VCI anharmonic corrections.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:27
The Molecular Nature of Internal Energy
 01:14
Zeroth Law of Thermodynamics
 02:38
Third Law of Thermodynamics
 03:40
Hess's Law
 01:14
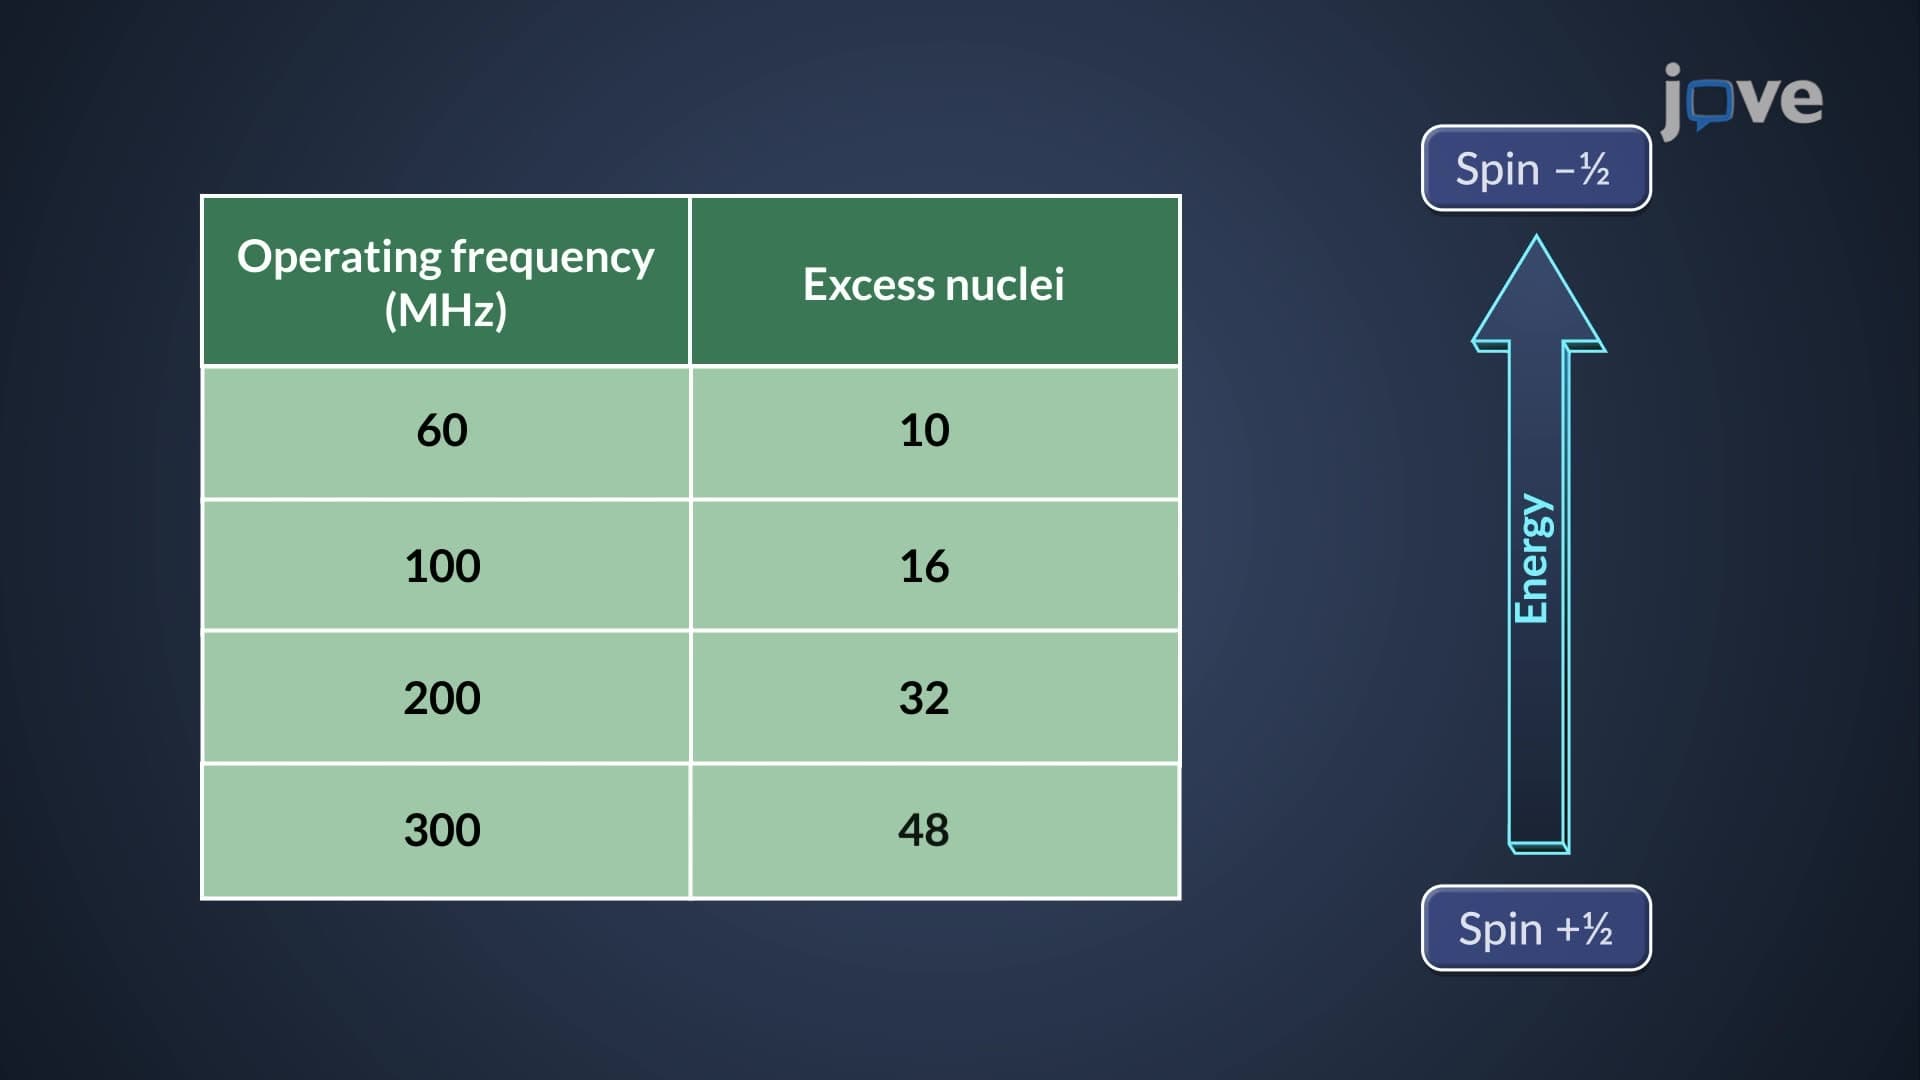
Atomic Nuclei: Nuclear Spin State Population Distribution